Related Experiment Video
Updated: Jun 20, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Large-scale parallel uncontracted multireference-averaged quadratic coupled cluster: the ground state of the chromium
1Institute of Advanced Simulation, Jülich Supercomputing Center, Research Centre Jülich, Germany. th.mueller@fz-juelich.de

Abstract:
The accurate prediction of the potential energy function of the X1Sigmag+ state of Cr2 is a remarkable challenge; large differential electron correlation effects, significant scalar relativistic contributions, the need for large flexible basis sets containing g functions, the importance of semicore valence electron correlation, and its multireference nature pose considerable obstacles. So far, the only reasonable successful approaches were based on multireference perturbation theory (MRPT). Recently, there was some controversy in the literature about the role of error compensation and systematic defects of various MRPT implementations that cannot be easily overcome. A detailed basis set study of the potential energy function is presented, adopting a variational method. The method of choice for this electron-rich target with up to 28 correlated electrons is fully uncontracted multireference-averaged quadratic coupled cluster (MR-AQCC), which shares the flexibility of the multireference configuration interaction (MRCI) approach and is, in addition, approximately size-extensive (0.02 eV in error as compared to the MRCI value of 1.37 eV for two noninteracting chromium atoms). The best estimate for De arrives at 1.48 eV and agrees well with the experimental data of 1.47 +/- 0.056 eV. At the estimated CBS limit, the equilibrium bond distance (1.685 A) and vibrational frequency (459 cm-1) are in agreement with experiment (1.679 A, 481 cm-1). Large basis sets and reference configuration spaces invariably result in huge wave function expansions (here, up to 2.8 billion configuration state functions), and efficient parallel implementations of the method are crucial. Hence, relevant details on implementation and general performance of the parallel program code are discussed as well.
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:22
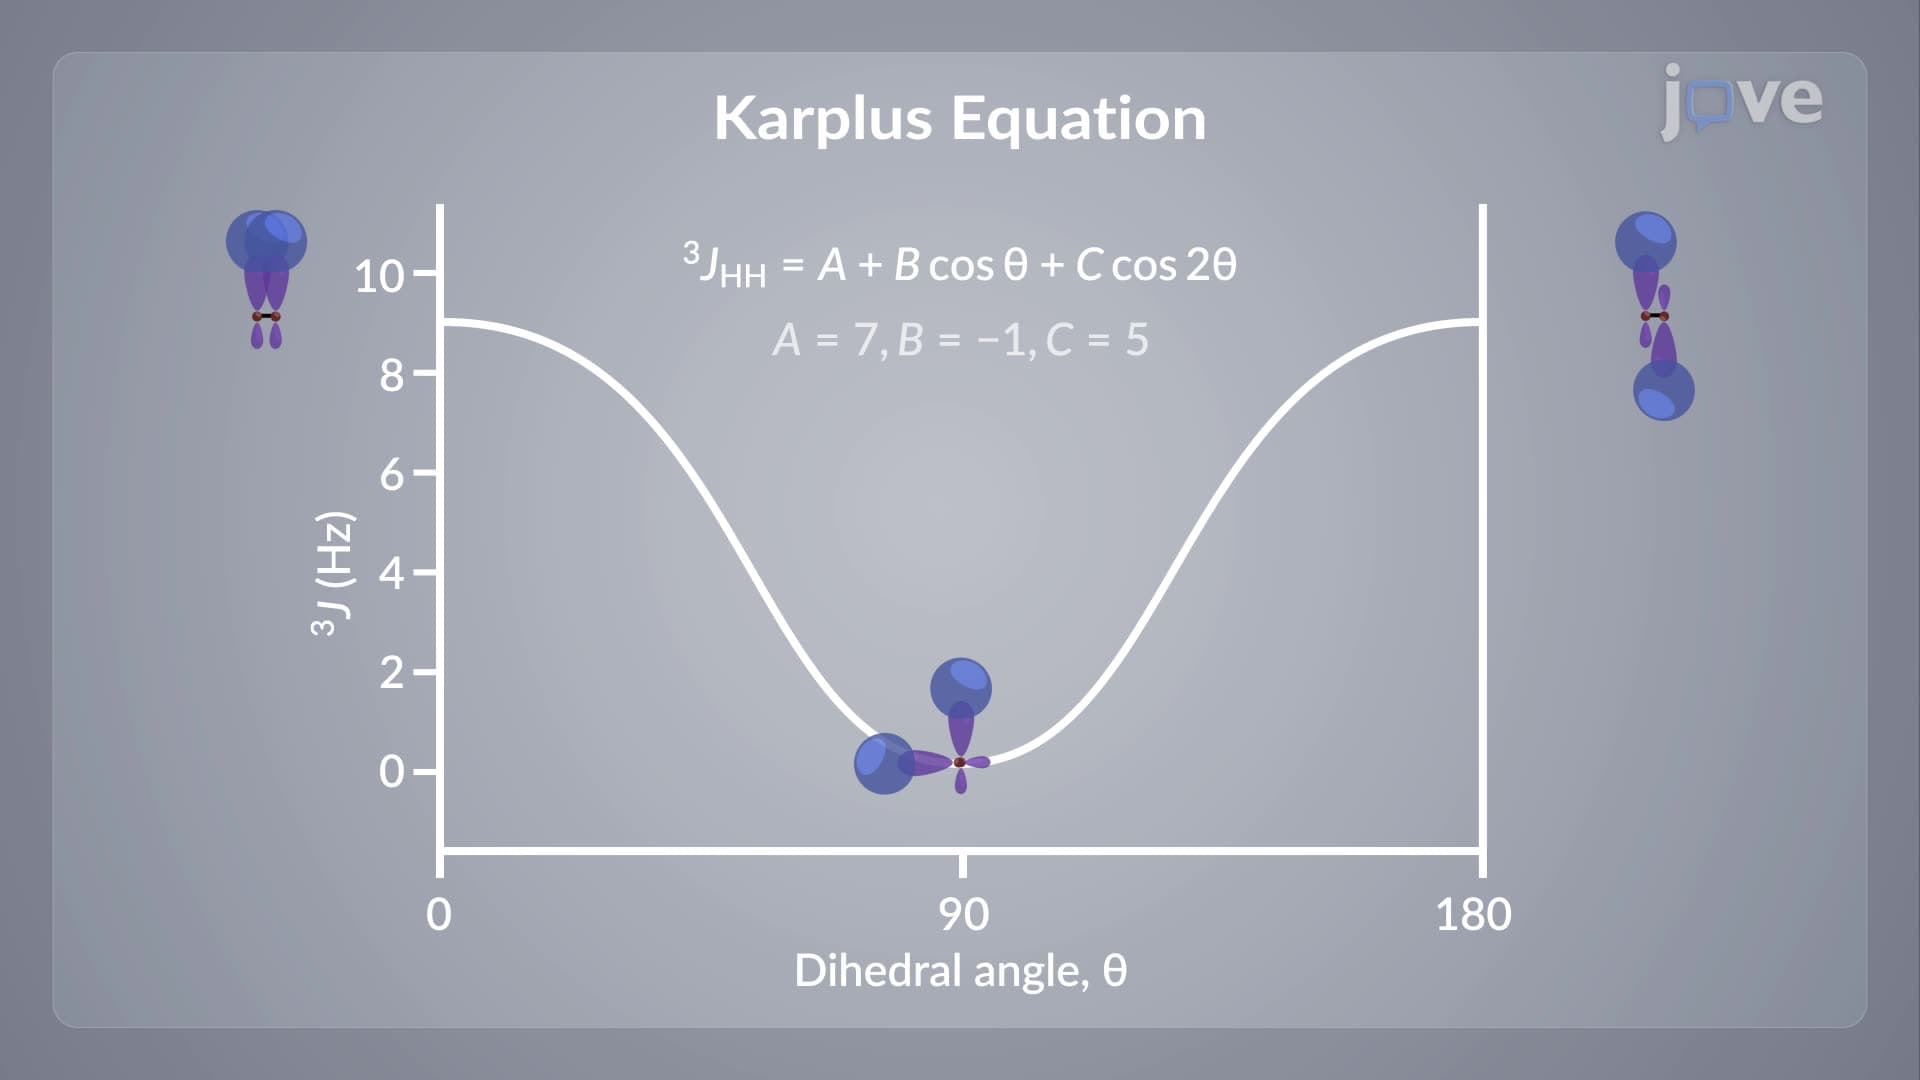
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...
 01:13
¹H NMR: Complex Splitting
Splitting diagrams or splitting tree diagrams are routinely used to depict such complex couplings. While drawing splitting diagrams, the splitting with the larger coupling constant is usually applied first.
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene π orbitals.