Related Experiment Video
Updated: Jun 20, 2026
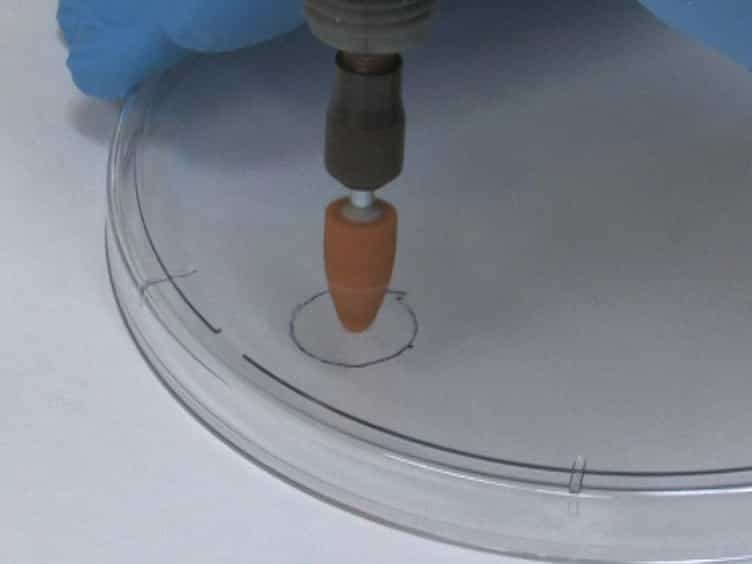

The Assembly and Application of 'Shear Rings': A Novel Endothelial Model for Orbital, Unidirectional and Periodic Fluid Flow and Shear Stress
Published on: October 31, 2016
Shear stress inhibits homocysteine-induced stromal cell-derived factor-1 expression in endothelial cells
Mao-Lin Sung1, Chia-Ching Wu, Hsin-I Chang
1Department of Cardiology, St Martin De Porres Hospital, Chiayi, Taiwan.

Insights
High homocysteine levels increase stromal cell-derived factor-1 (SDF-1) in endothelial cells via the JNK pathway. Shear stress protects against this by activating nitric oxide (NO) signaling.
Area of Science:
- Vascular Biology
- Molecular Cardiology
- Biochemistry
Background:
- Hyperhomocysteinemia is linked to vascular dysfunction and cardiovascular disease risk.
- Stromal cell-derived factor-1 (SDF-1), a chemokine in endothelial cells (ECs), is prevalent in advanced atherosclerotic lesions.
- The interplay between homocysteine, chemokines, and shear stress in regulating endothelial function remains unclear.
Purpose of the Study:
- To investigate the mechanisms by which homocysteine and shear stress modulate EC SDF-1 expression.
- To elucidate the role of specific signaling pathways, including mitogen-activated protein kinases (MAPKs) and nitric oxide (NO), in this process.
Main Methods:
- Endothelial cells were stimulated with homocysteine and subjected to varying shear stress conditions.
- Investigated SDF-1 expression, MAPK phosphorylation (ERK, JNK, p38), and transcription factor activity (Sp1, AP-1) using inhibitors, siRNA, and dominant-negative mutants.
- Assessed the role of the nitric oxide (NO) pathway via NO donors and endothelial NO synthase (eNOS) inhibition.
Main Results:
- Homocysteine dose- and time-dependently increased SDF-1 expression and JNK phosphorylation.
- JNK pathway activation was critical for homocysteine-induced SDF-1 expression.
- Homocysteine enhanced Sp1 and AP-1 DNA binding, which were blocked by siRNA inhibition.
- Preshearing inhibited homocysteine-induced JNK phosphorylation, Sp1/AP-1 activation, and SDF-1 expression.
- NO donors suppressed homocysteine-induced SDF-1, and eNOS inhibition abolished shear stress protection.
Conclusions:
- Elucidated the molecular mechanisms of homocysteine-induced SDF-1 expression in ECs.
- Demonstrated that shear stress protects against homocysteine effects via NO signaling.
- Findings highlight the complex regulation of SDF-1 in vascular disease pathogenesis.
Rationale:
Hyperhomocysteinemia contributes to vascular dysfunction and risks of cardiovascular diseases. Stromal cell-derived factor (SDF)-1, a chemokine expressed by endothelial cells (ECs), is highly expressed in advanced atherosclerotic lesions. The interplays among homocysteine, chemokines, and shear stress in regulating vascular endothelial function are not clearly understood.
Objective:
To investigate the mechanisms for modulations of EC SDF-1 expression by homocysteine and shear stress.
Methods And Results:
Homocysteine stimulation induced dose- and time-dependent SDF-1 expression and phosphorylation of mitogen-activated protein kinases extracellular signal-regulated kinase (ERK), c-Jun N-terminal kinase (JNK), and p38. By using specific inhibitors, small interfering (si)RNA, and dominant negative mutants, we demonstrated that activation of JNK pathway is critical for the homocysteine-induced SDF-1 expression. Transcription factor ELISA and chromatin immunoprecipitation assays showed that homocysteine increased Sp1- and AP-1-DNA binding activities in ECs. Inhibition of Sp1 and AP-1 activations by specific siRNA blocked the homocysteine-induced SDF-1 promoter activity and expression. Preshearing of ECs for 1 to 4 hours at 20 dyn/cm2 inhibited the homocysteine-induced JNK phosphorylation, Sp1 and AP-1 activation, and SDF-1 expression. The homocysteine-induced SDF-1 expression was suppressed by NO donor. Inhibitor or siRNA for endothelial NO synthase abolished the shear inhibition of SDF-1 expression.
Conclusions:
Our findings serve to elucidate the molecular mechanisms underlying the homocysteine induction of SDF-1 expression in ECs and the shear stress protection against this induction.
More Related Videos
Related Concept Videos
 01:01
Regulation of Hematopoietic Stem Cells
 01:24
Regulation of Angiogenesis and Blood Supply