Related Experiment Video
Updated: Jun 20, 2026

Synthesis of Non-uniformly Pr-doped SrTiO3 Ceramics and Their Thermoelectric Properties
Published on: August 15, 2015
Energetic and electronic properties of X- (Si, Ge, Sn, Pb) doped TiO2 from first-principles
1School of Physics, State Key Laboratory of Crystal Materials, Shandong University, Jinan 250100, P. R. China.

Abstract:
First-principles density functional theory calculations were performed on the electronic properties of substitutional Si, Ge, Sn, and Pb-doped rutile and anatase TiO(2), where Ti was replaced by dopants, to explore the effect of the dopants on the band edges and their possible efficacy for the visible light photocatalysis and solar energy conversion. Based on the optimized structures of X-doped TiO(2), the defect formation energies and the density of states were calculated to analyze the energetic properties, band edges, and the band gap states. Our calculated results show that dopants substituting for Ti is energetically unfavorable in the order Si < Sn < Ge < Pb under both O-rich and Ti-rich growth conditions. The electronic structures show that there is a reduction of band gap about 0.1 approximately 0.55 eV in X-doped rutile TiO(2). Si- and Ge-doped anatase TiO(2) also decrease by 0.20 and 0.15 eV, but broadening by 0.06 and 0.02 eV occurs in Sn- and Pb-doped systems. The calculated results explain the experimental observations. Furthermore, it can be predicted that Ge-doped TiO(2) can be active for overall water splitting under visible light photons whereas Si-, Sn-, and Pb-doped TiO(2) are only suitable for photo-oxidation.
More Related Videos
 07:24
Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
 09:23
Solution-Processed, Surface-Engineered, Polycrystalline CdSe-SnSe Exhibiting Low Thermal Conductivity
Published on: May 17, 2024
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:54
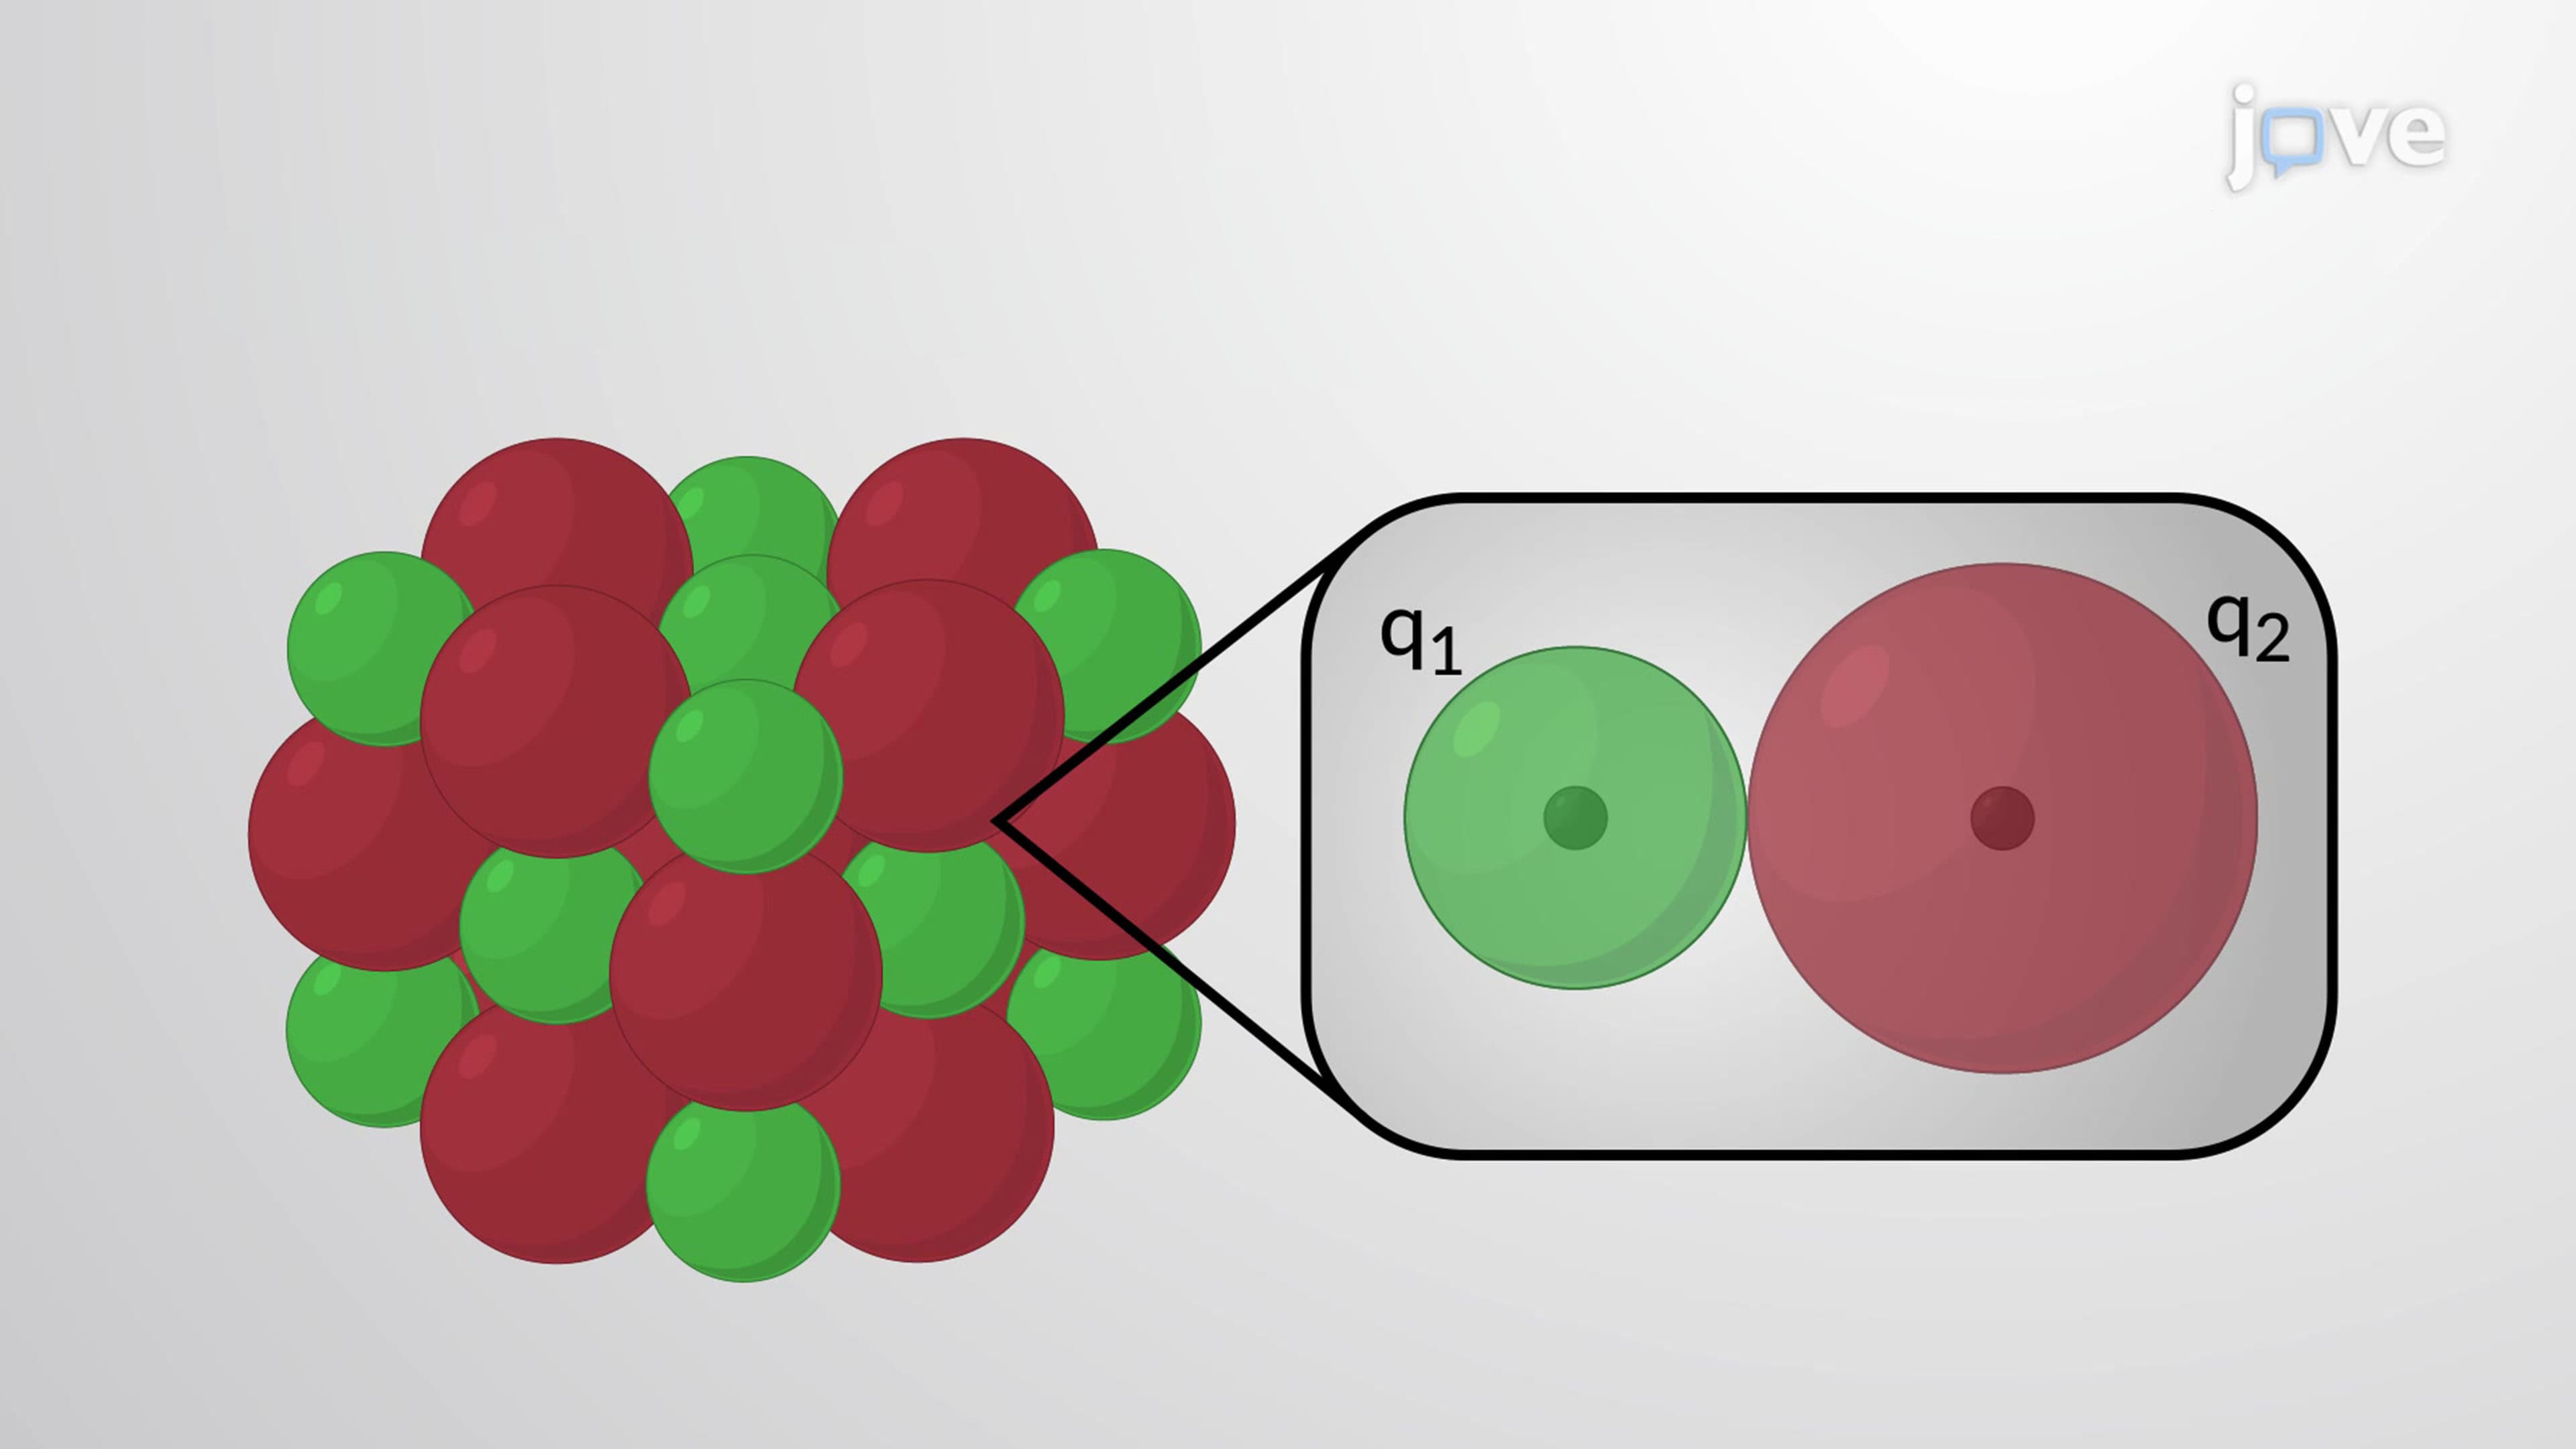
Trends in Lattice Energy: Ion Size and Charge
 01:22
Semiconductors
Metals such as copper (Cu), zinc (Zn), or lead (Pb) have low resistivity and feature conduction bands that are either not fully occupied or overlap with the valence band, making a bandgap non-existent. This allows electrons in the highest energy levels of the valence band to easily transition to the conduction band upon gaining...
 01:01
Energy Bands in Solids
Band Formation:
When atoms are brought close together, as in a solid, these discrete energy levels begin to split due to the overlap of electron orbitals from adjacent atoms. This split occurs because of the Pauli exclusion principle, which states that no two...
 02:27
Predicting Molecular Geometry