Evaluation of exchange-correlation functionals for time-dependent density functional theory calculations on metal complexes
Related Experiment Video
Updated: Jun 20, 2026
Thermochemical Studies of Ni(II) and Zn(II) Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Jason P Holland1, Jennifer C Green
1Department of Chemistry, Chemistry Research Laboratory, University of Oxford, 12 Mansfield Road, Oxford OX1 3TA, United Kingdom. hollanj3@mskcc.org

Abstract:
The electronic absorption spectra of a range of copper and zinc complexes have been simulated by using time-dependent density functional theory (TD-DFT) calculations implemented in Gaussian03. In total, 41 exchange-correlation (XC) functionals including first-, second-, and third-generation (meta-generalized gradient approximation) DFT methods were compared in their ability to predict the experimental electronic absorption spectra. Both pure and hybrid DFT methods were tested and differences between restricted and unrestricted calculations were also investigated by comparison of analogous neutral zinc(II) and copper(II) complexes. TD-DFT calculated spectra were optimized with respect to the experimental electronic absorption spectra by use of a Matlab script. Direct comparison of the performance of each XC functional was achieved both qualitatively and quantitatively by comparison of optimized half-band widths, root-mean-squared errors (RMSE), energy scaling factors (epsilon(SF)), and overall quality-of-fit (Q(F)) parameters. Hybrid DFT methods were found to outperform all pure DFT functionals with B1LYP, B97-2, B97-1, X3LYP, and B98 functionals providing the highest quantitative and qualitative accuracy in both restricted and unrestricted systems. Of the functionals tested, B1LYP gave the most accurate results with both average RMSE and overall Q(F) < 3.5% and epsilon(SF) values close to unity (>0.990) for the copper complexes. The XC functional performance in spin-restricted TD-DFT calculations on the zinc complexes was found to be slightly worse. PBE1PBE, mPW1PW91 and B1LYP gave the most accurate results with typical RMSE and Q(F) values between 5.3 and 7.3%, and epsilon(SF) around 0.930. These studies illustrate the power of modern TD-DFT calculations for exploring excited state transitions of metal complexes.
More Related Videos


Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Valence Bond Theory
 01:23
Complexation Equilibria: Overview
The equilibrium constant of the complexation reaction is represented as the formation constant...
 01:09
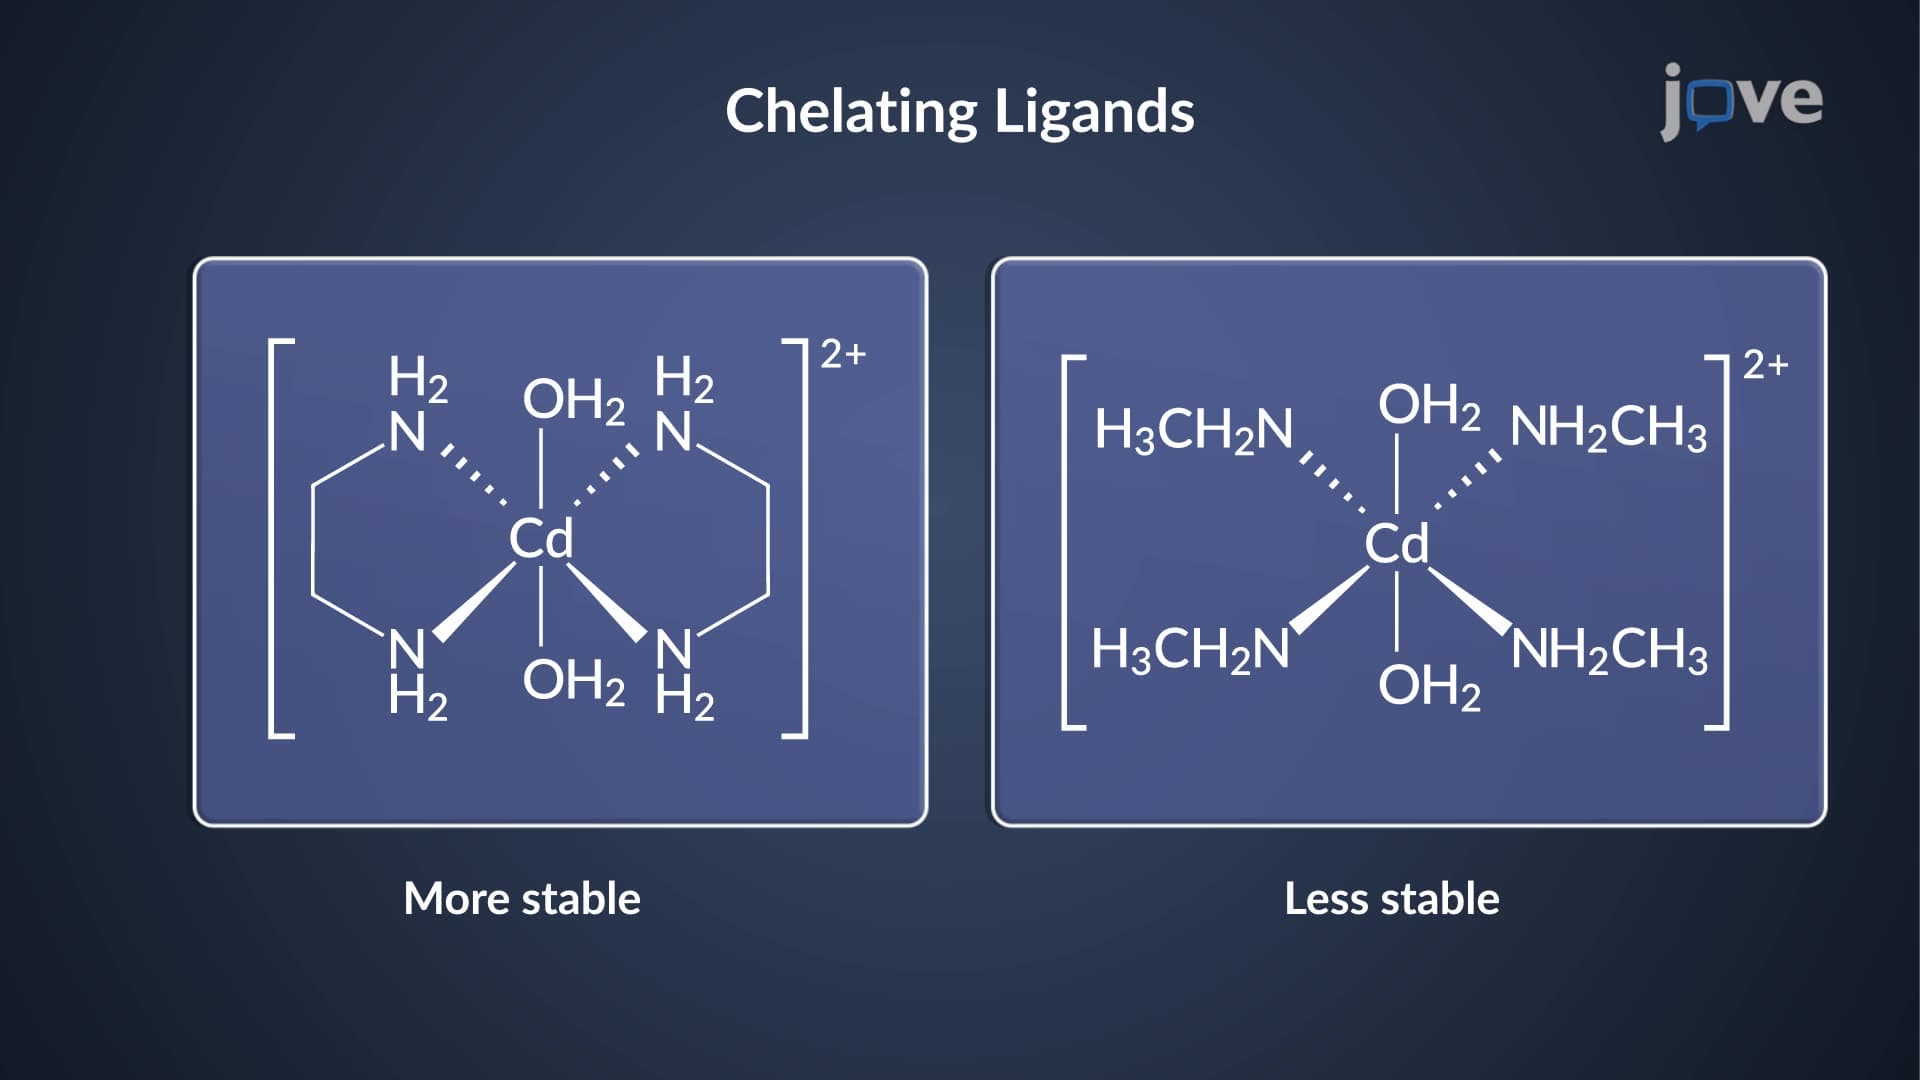
Complexation Equilibria: Factors Influencing Stability of Complexes
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...