Related Experiment Video
Updated: Jun 19, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Performance of ab initio and density functional methods for conformational equilibria of C(n)H(2n+2) alkane isomers
David Gruzman1, Amir Karton, Jan M L Martin
1Department of Organic Chemistry, Weizmann Institute of Science, IL-76100 Rechovot, Israel.

Abstract:
Conformational energies of n-butane, n-pentane, and n-hexane have been calculated at the CCSD(T) level and at or near the basis set limit. Post-CCSD(T) contributions were considered and found to be unimportant. The data thus obtained were used to assess the performance of a variety of density functional methods. Double-hybrid functionals like B2GP-PLYP and B2K-PLYP, especially with a small Grimme-type empirical dispersion correction, are capable of rendering conformational energies of CCSD(T) quality. These were used as a "secondary standard" for a larger sample of alkanes, including isopentane and the branched hexanes as well as key isomers of heptane and octane. Popular DFT functionals like B3LYP, B3PW91, BLYP, PBE, and PBE0 tend to overestimate conformer energies without dispersion correction, while the M06 family severely underestimates GG interaction energies. Grimme-type dispersion corrections for these overcorrect and lead to qualitatively wrong conformer orderings. All of these functionals also exhibit deficiencies in the conformer geometries, particularly the backbone torsion angles. The PW6B95 and, to a lesser extent, BMK functionals are relatively free of these deficiencies. Performance of these methods is further investigated to derive conformer ensemble corrections to the enthalpy function, H(298) - H(0), and the Gibbs energy function, gef(T) identical with -[G(T) - H(0)]/T, of these alkanes. These are essential for accurate computed heats of formation of especially the larger species as the corrections for these are several times the expected uncertainty in modern computational thermochemistry methods such as W4 theory. While H(298) - H(0) is only moderately sensitive to the level of theory, gef(T) exhibits more pronounced sensitivity. Once again, double hybrids acquit themselves very well. The effects of zero-point energy and nonfactorizable rovibrational partition functions have been considered.
Related Concept Videos
 02:18
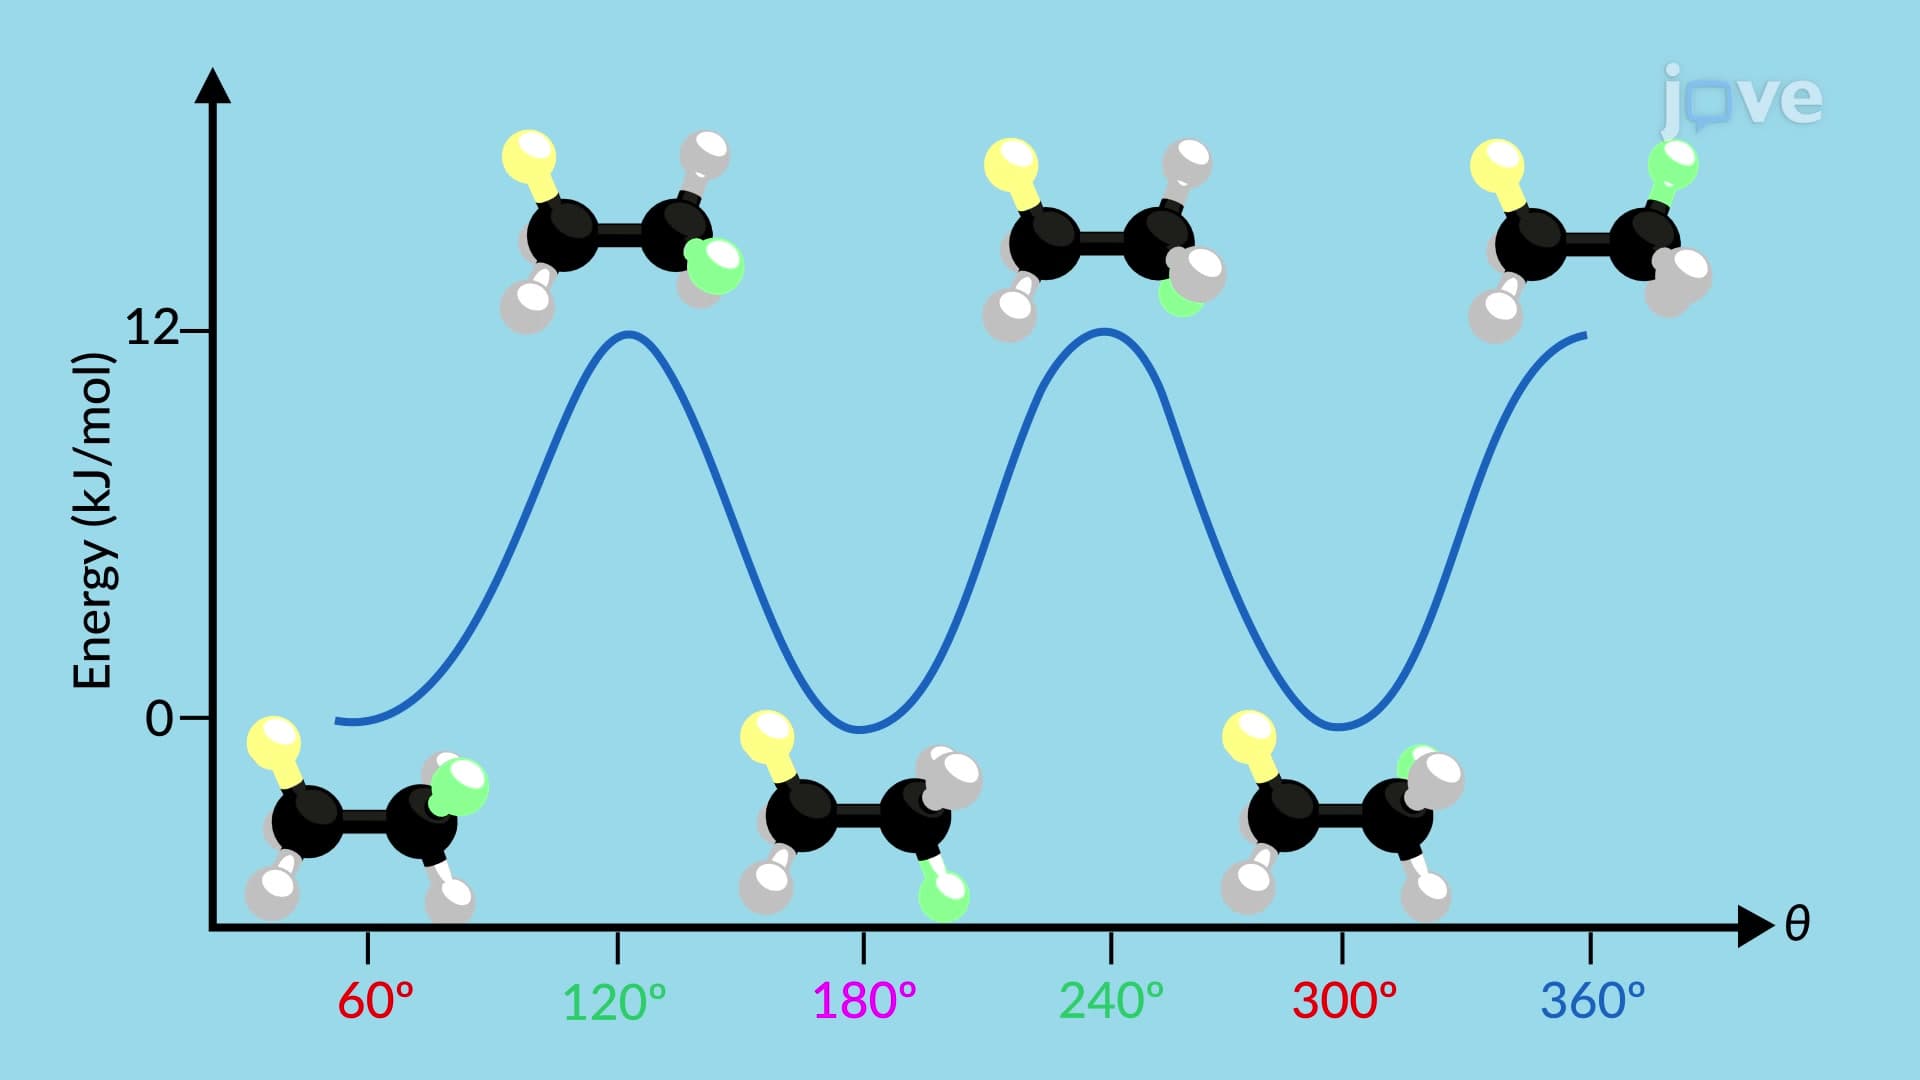
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered ethane, the...
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 02:18
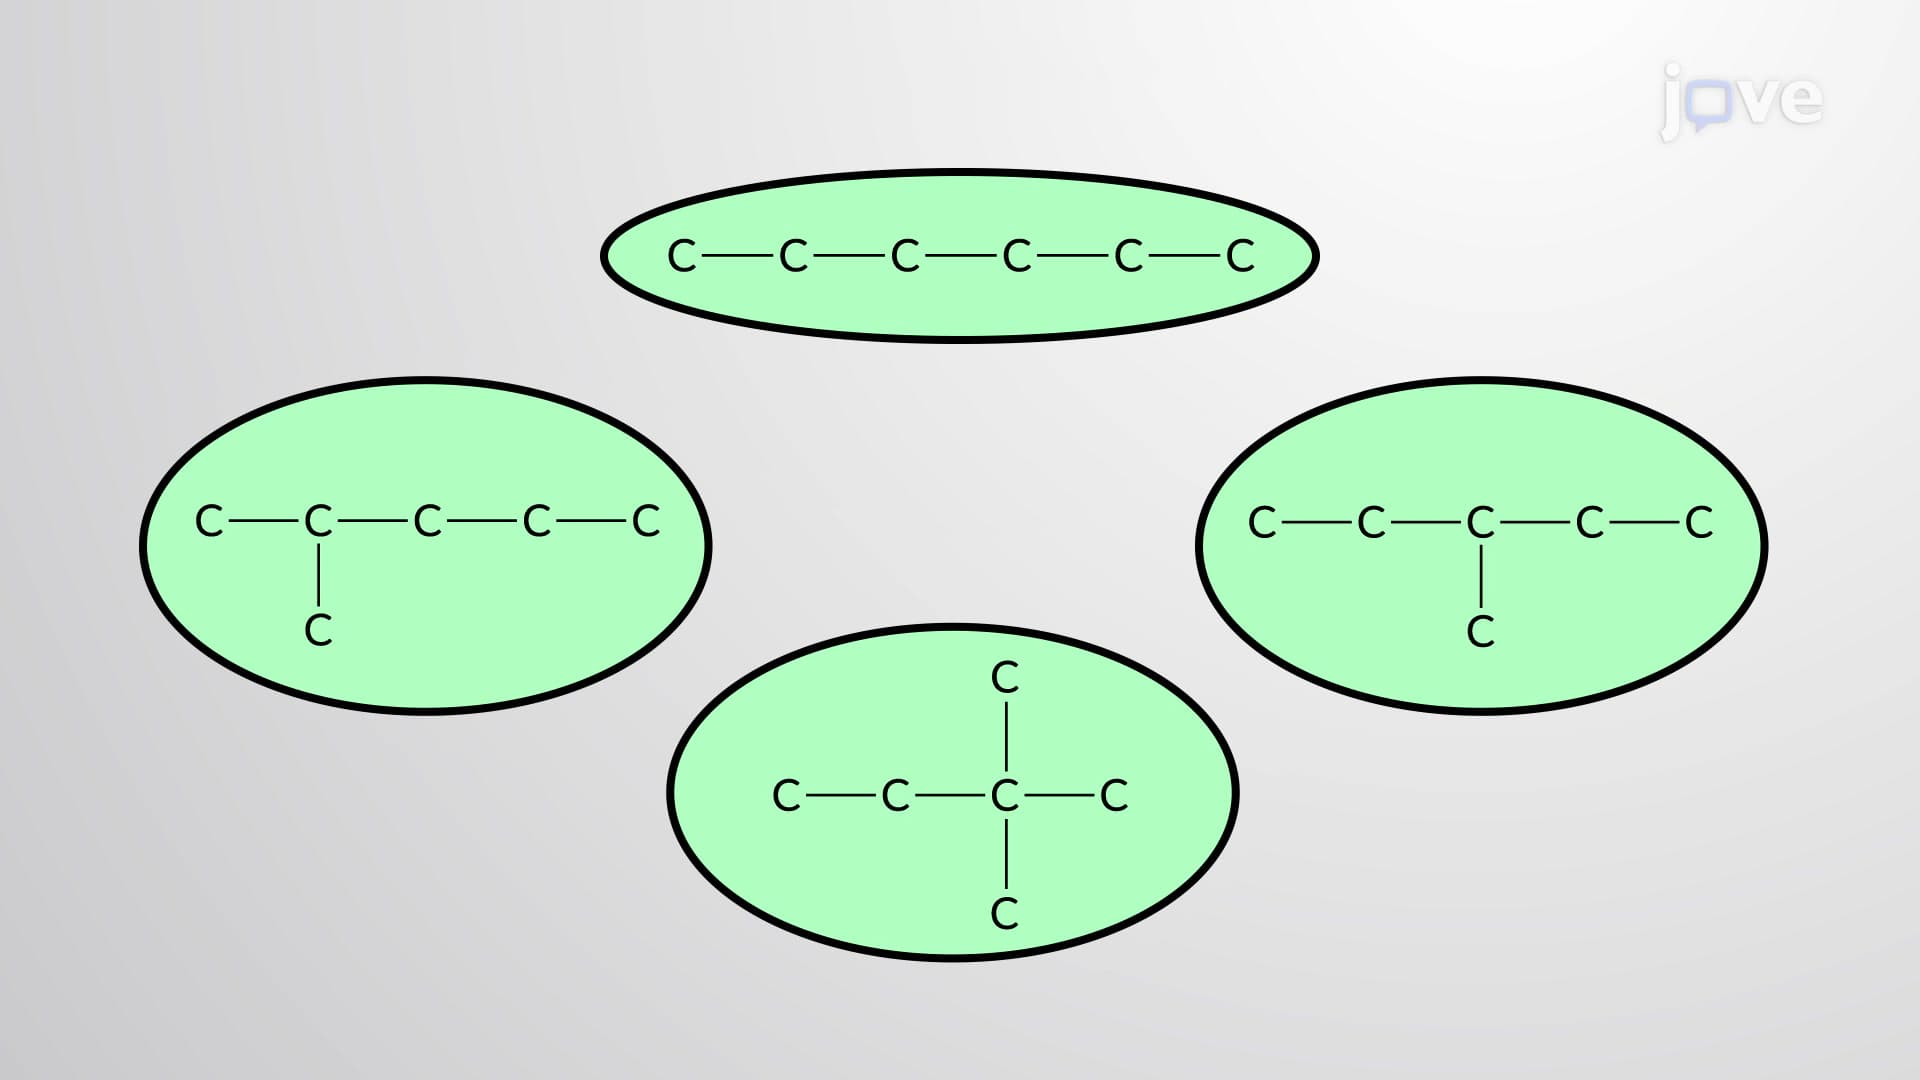
Constitutional Isomers of Alkanes
The linear isomer of an alkane is prefixed by the term “n”; hence a linear isomer of pentane is known as n-pentane. Based on the type of branching, some of the branched isomers are given...
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
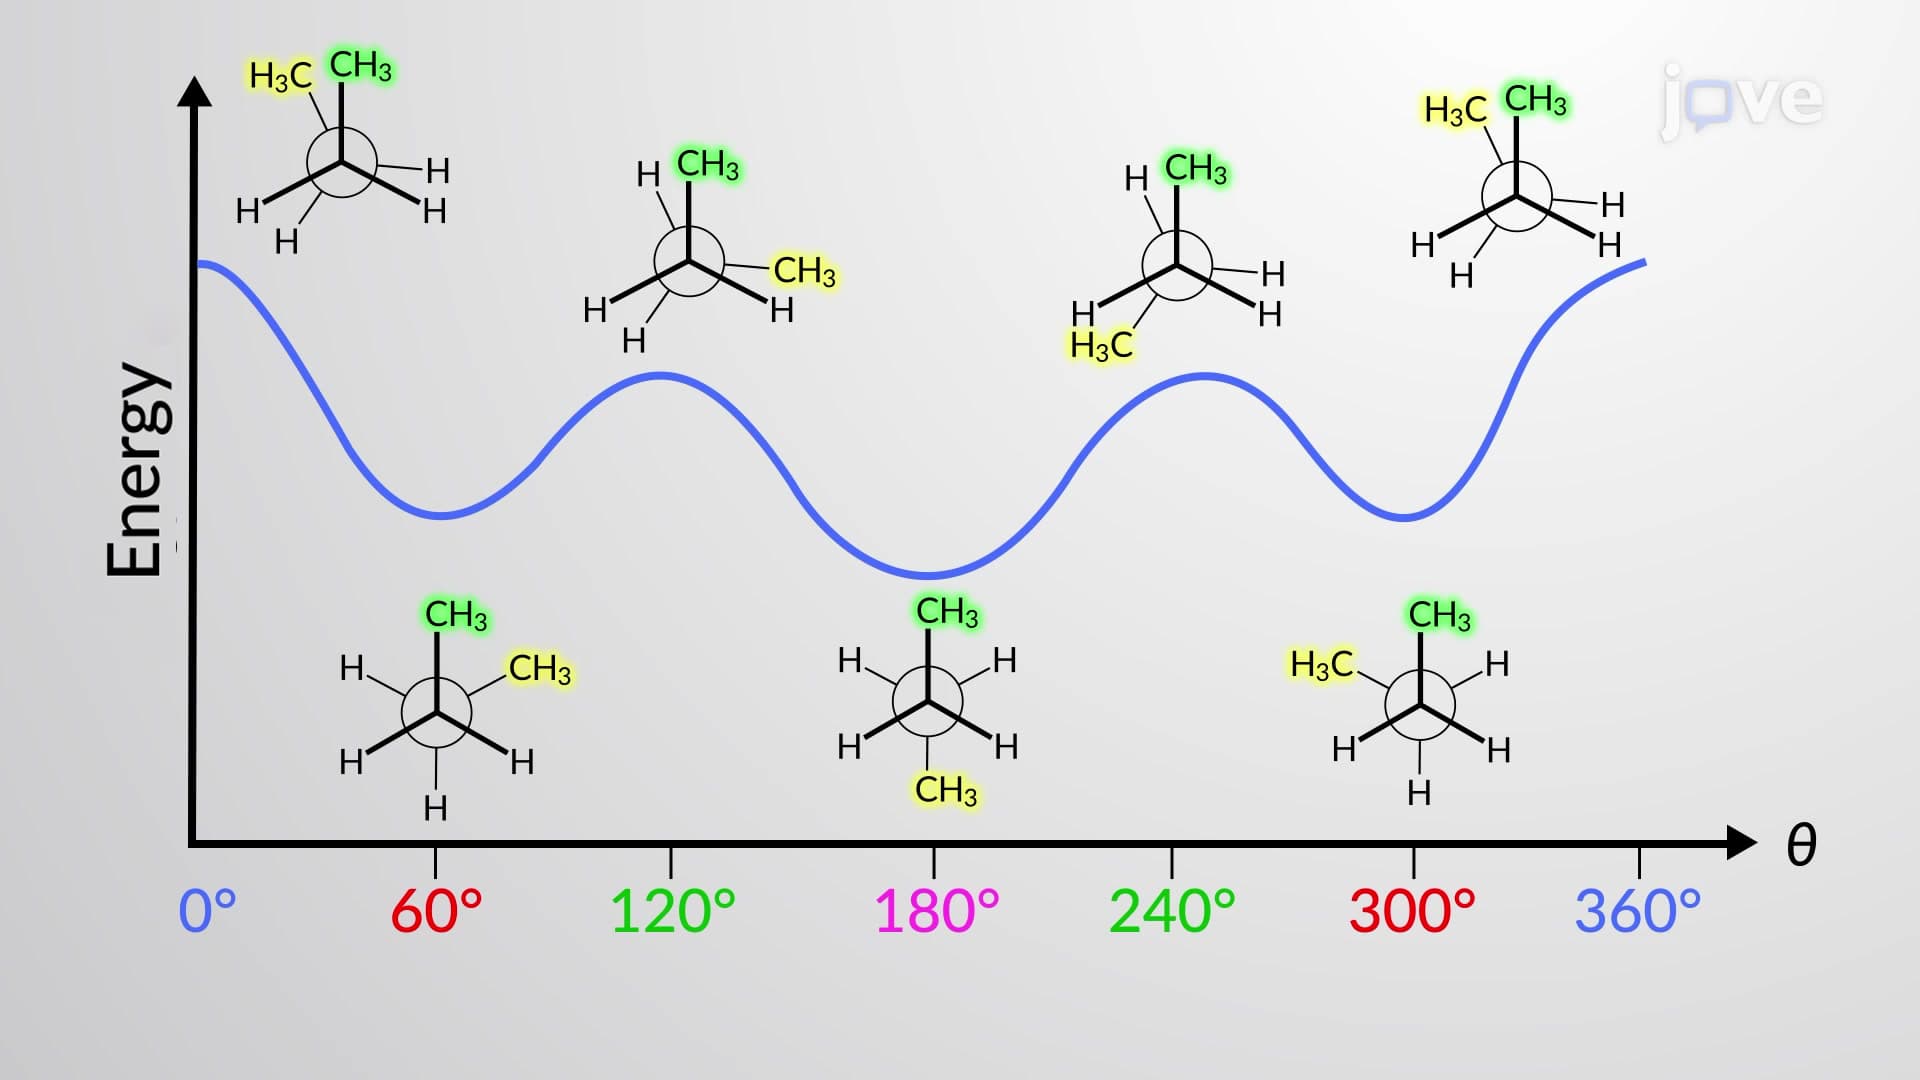 02:20
Conformations of Butane
 02:02
Chair Conformation of Cyclohexane
The hydrogen atoms linked to carbons are arranged in two different axial and equatorial orientations to achieve this staggered...