Related Experiment Video
Updated: Jun 19, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Benchmarking density-functional-theory calculations of rotational g tensors and magnetizabilities using accurate
Ola B Lutnaes1, Andrew M Teale, Trygve Helgaker
1Department of Chemistry, Centre for Theoretical and Computational Chemistry, University of Oslo, P.O.B. 1033 Blindern, Oslo N-0315, Norway. o.b.lutnas@kjemi.uio.no

Abstract:
An accurate set of benchmark rotational g tensors and magnetizabilities are calculated using coupled-cluster singles-doubles (CCSD) theory and coupled-cluster single-doubles-perturbative-triples [CCSD(T)] theory, in a variety of basis sets consisting of (rotational) London atomic orbitals. The accuracy of the results obtained is established for the rotational g tensors by careful comparison with experimental data, taking into account zero-point vibrational corrections. After an analysis of the basis sets employed, extrapolation techniques are used to provide estimates of the basis-set-limit quantities, thereby establishing an accurate benchmark data set. The utility of the data set is demonstrated by examining a wide variety of density functionals for the calculation of these properties. None of the density-functional methods are competitive with the CCSD or CCSD(T) methods. The need for a careful consideration of vibrational effects is clearly illustrated. Finally, the pure coupled-cluster results are compared with the results of density-functional calculations constrained to give the same electronic density. The importance of current dependence in exchange-correlation functionals is discussed in light of this comparison.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:36
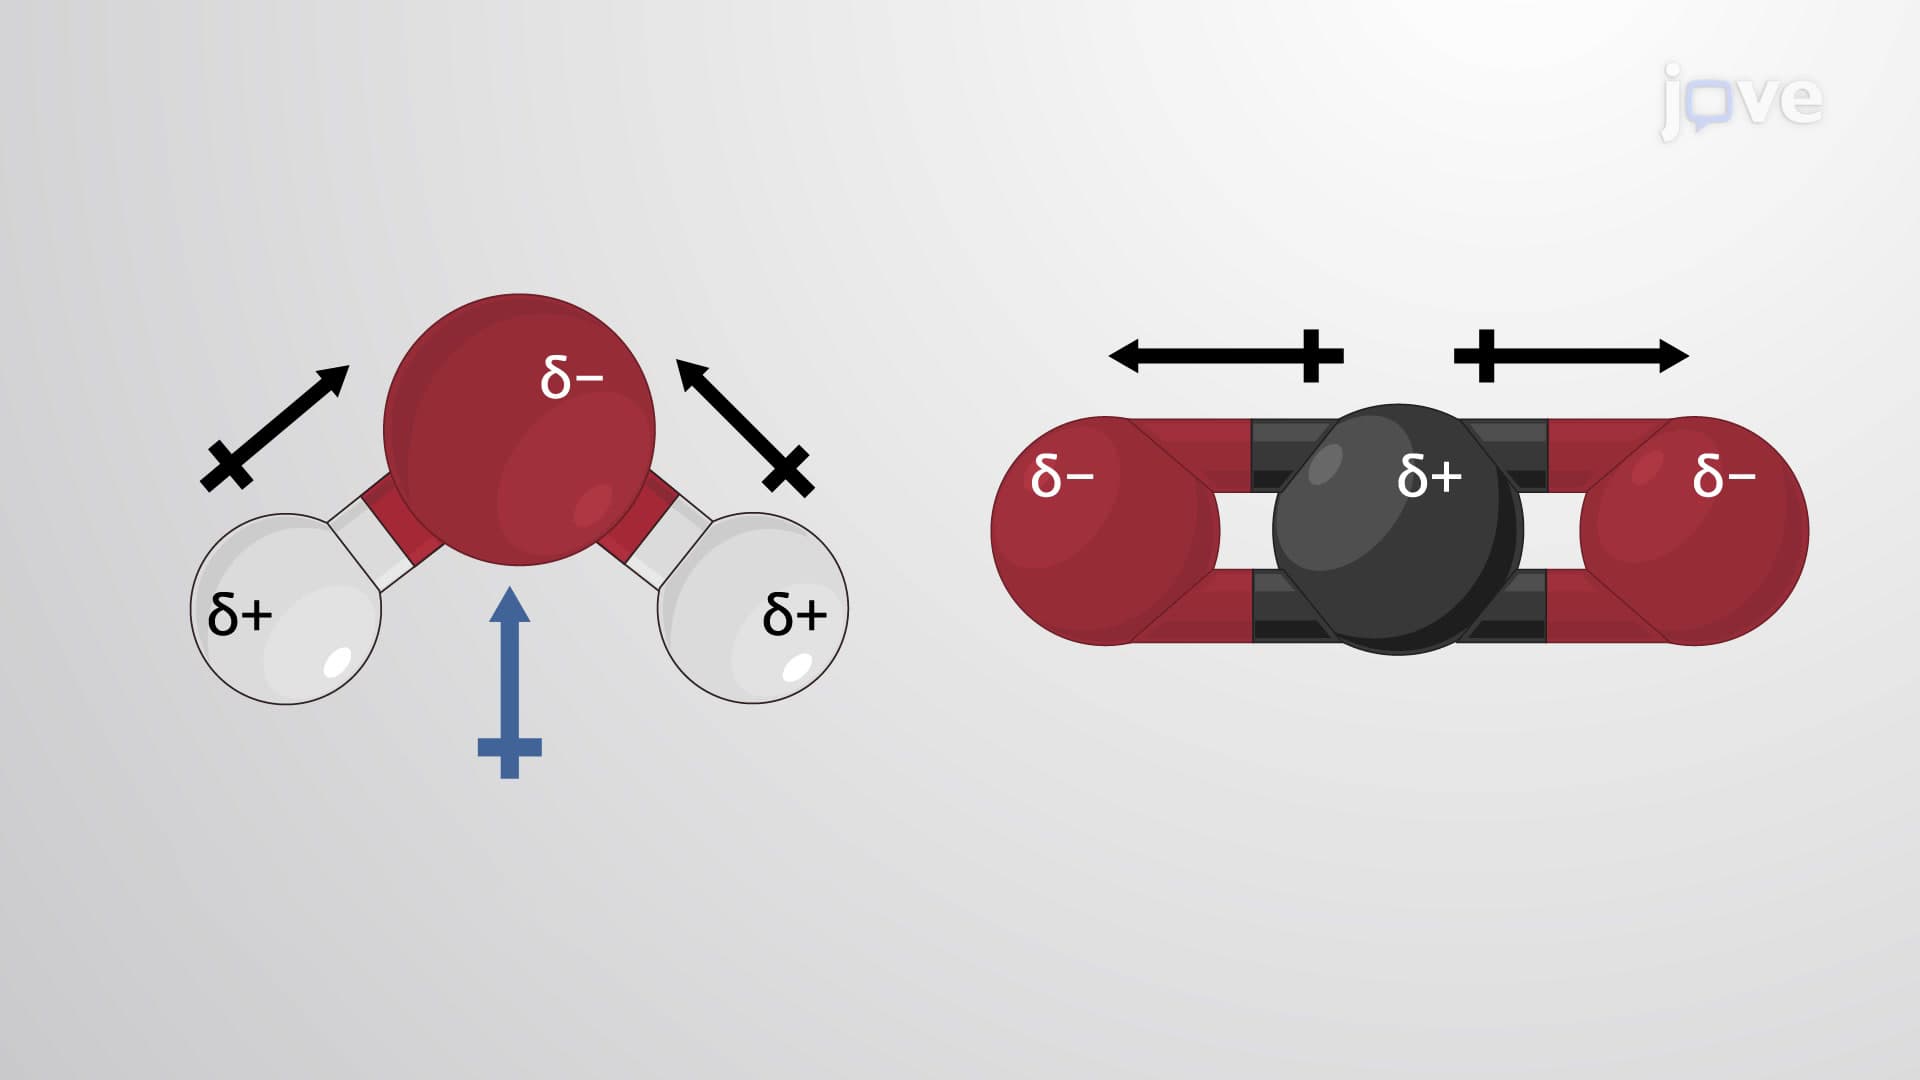
Molecular Geometry and Dipole Moments
 01:08
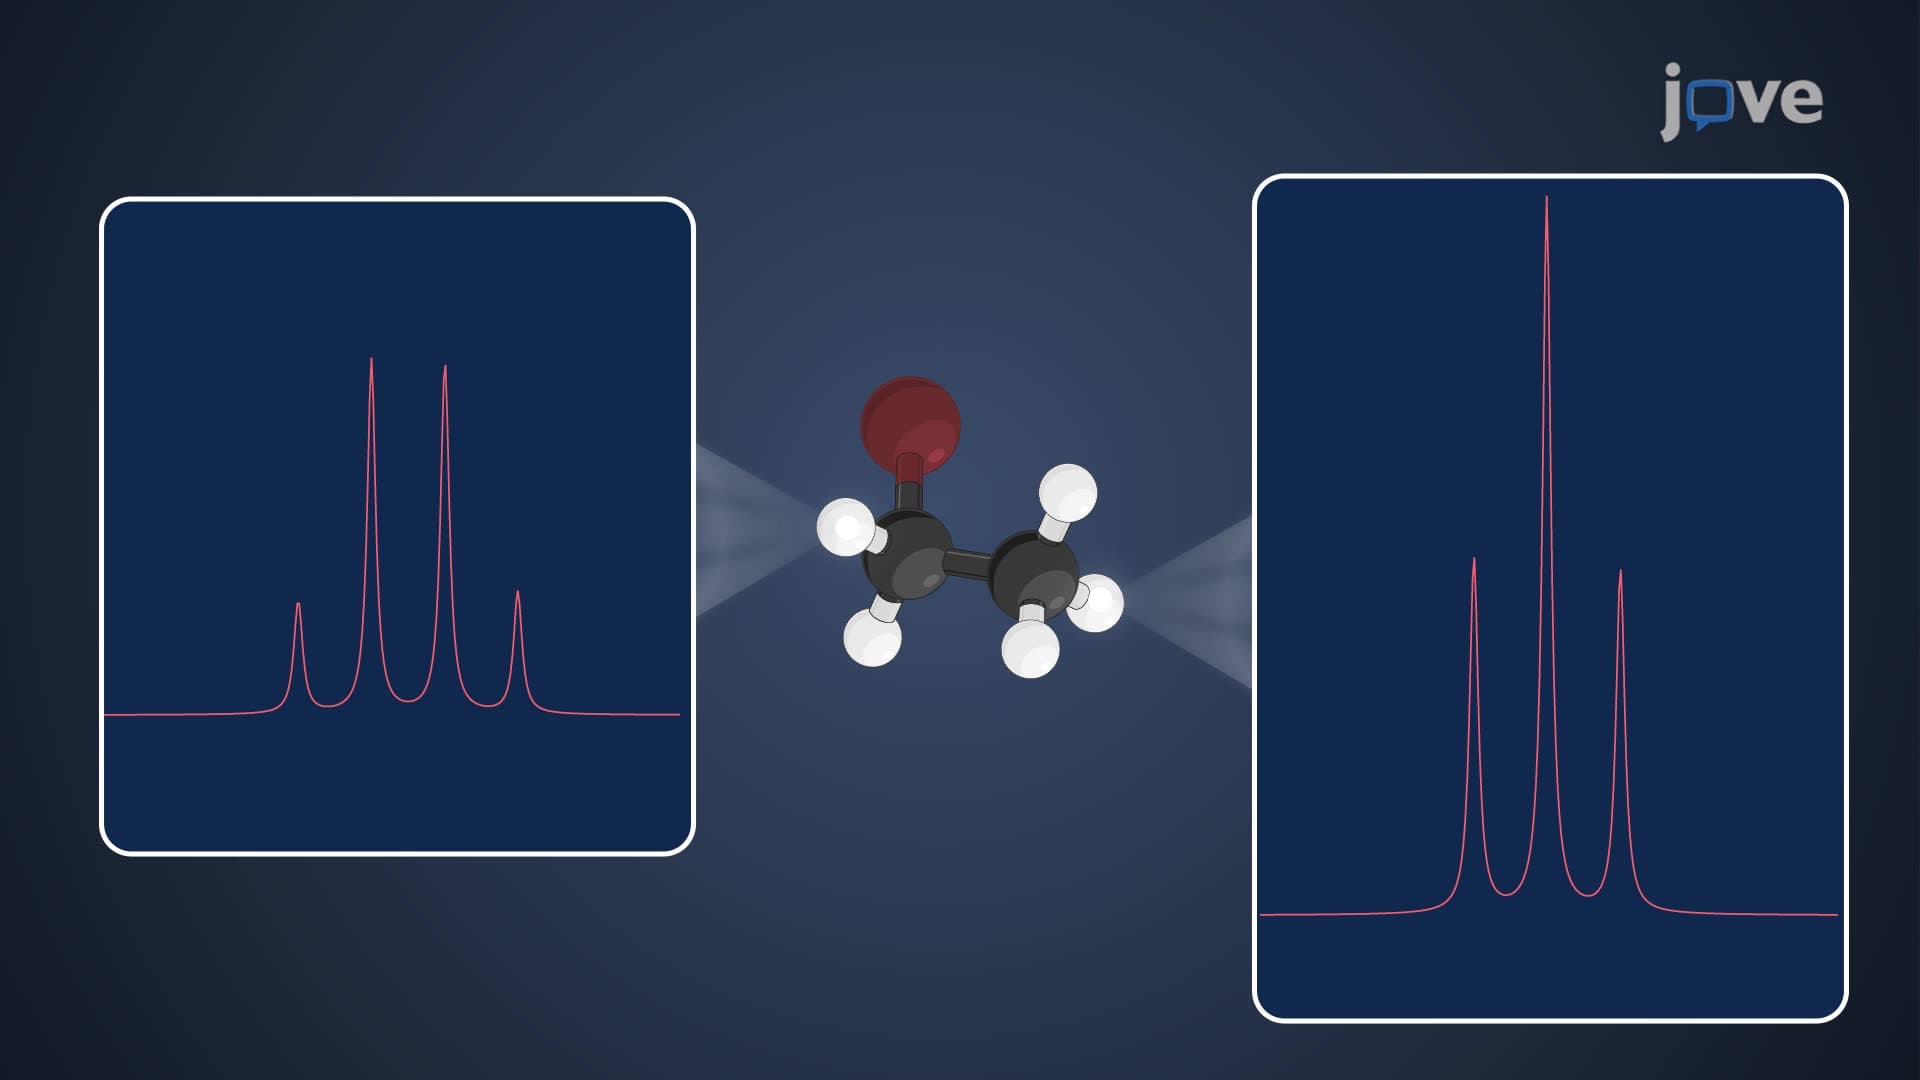
Spin–Spin Coupling Constant: Overview
Qualitatively, any spin plus-half nucleus polarizes the spins of its electrons to the minus-half state. Consequently, the paired electron in the hydrogen–carbon bond must have a...
 01:24
Potential Due to a Magnetized Object
The vector...
 02:42
Valence Bond Theory