Related Experiment Video
Updated: Jun 19, 2026

Modeling Mitochondrial Disease Using Brain Organoids: A Focus on Mitochondrial Encephalomyopathy, Lactic Acidosis, and Stroke-like Episodes
Published on: October 10, 2025
Menkes disease
Zeynep Tümer1, Lisbeth B Møller
1Kennedy Centre, Glostrup, Denmark. zet@kennedy.dk

Abstract:
Menkes disease (MD) is a lethal multisystemic disorder of copper metabolism. Progressive neurodegeneration and connective tissue disturbances, together with the peculiar 'kinky' hair are the main manifestations. MD is inherited as an X-linked recessive trait, and as expected the vast majority of patients are males. MD occurs due to mutations in the ATP7A gene and the vast majority of ATP7A mutations are intragenic mutations or partial gene deletions. ATP7A is an energy dependent transmembrane protein, which is involved in the delivery of copper to the secreted copper enzymes and in the export of surplus copper from cells. Severely affected MD patients die usually before the third year of life. A cure for the disease does not exist, but very early copper-histidine treatment may correct some of the neurological symptoms.
Insights
Menkes disease (MD) is a lethal genetic disorder affecting copper metabolism, causing neurodegeneration and connective tissue issues. Early copper-histidine treatment shows potential for mitigating neurological symptoms in affected males.
Area of Science:
- Genetics and Molecular Biology
- Biochemistry
- Pediatric Medicine
Background:
- Menkes disease (MD) is a fatal multisystem disorder impacting copper metabolism.
- Key manifestations include progressive neurodegeneration, connective tissue abnormalities, and distinctive 'kinky' hair.
- MD is an X-linked recessive condition primarily affecting males, stemming from mutations in the ATP7A gene.
Purpose of the Study:
- To summarize the key features and genetic basis of Menkes disease.
- To discuss the role of the ATP7A gene in copper transport and its implications in MD.
- To review current therapeutic approaches, including the potential of early copper-histidine treatment.
Main Methods:
- Literature review of Menkes disease.
- Analysis of ATP7A gene mutations and their impact on copper metabolism.
- Evaluation of treatment strategies for Menkes disease.
Main Results:
- Menkes disease is characterized by severe neurodegeneration and connective tissue problems.
- Mutations in the ATP7A gene disrupt cellular copper transport, leading to enzyme deficiencies.
- Most severely affected patients succumb before age three; however, early copper-histidine intervention may ameliorate neurological deficits.
Conclusions:
- Menkes disease is a severe genetic disorder with no definitive cure.
- Understanding ATP7A's function is crucial for comprehending MD pathogenesis.
- Prompt therapeutic intervention, such as copper-histidine treatment, offers a potential avenue for managing symptoms.
Related Concept Videos
 01:20
Inborn Errors of Metabolism
 01:49
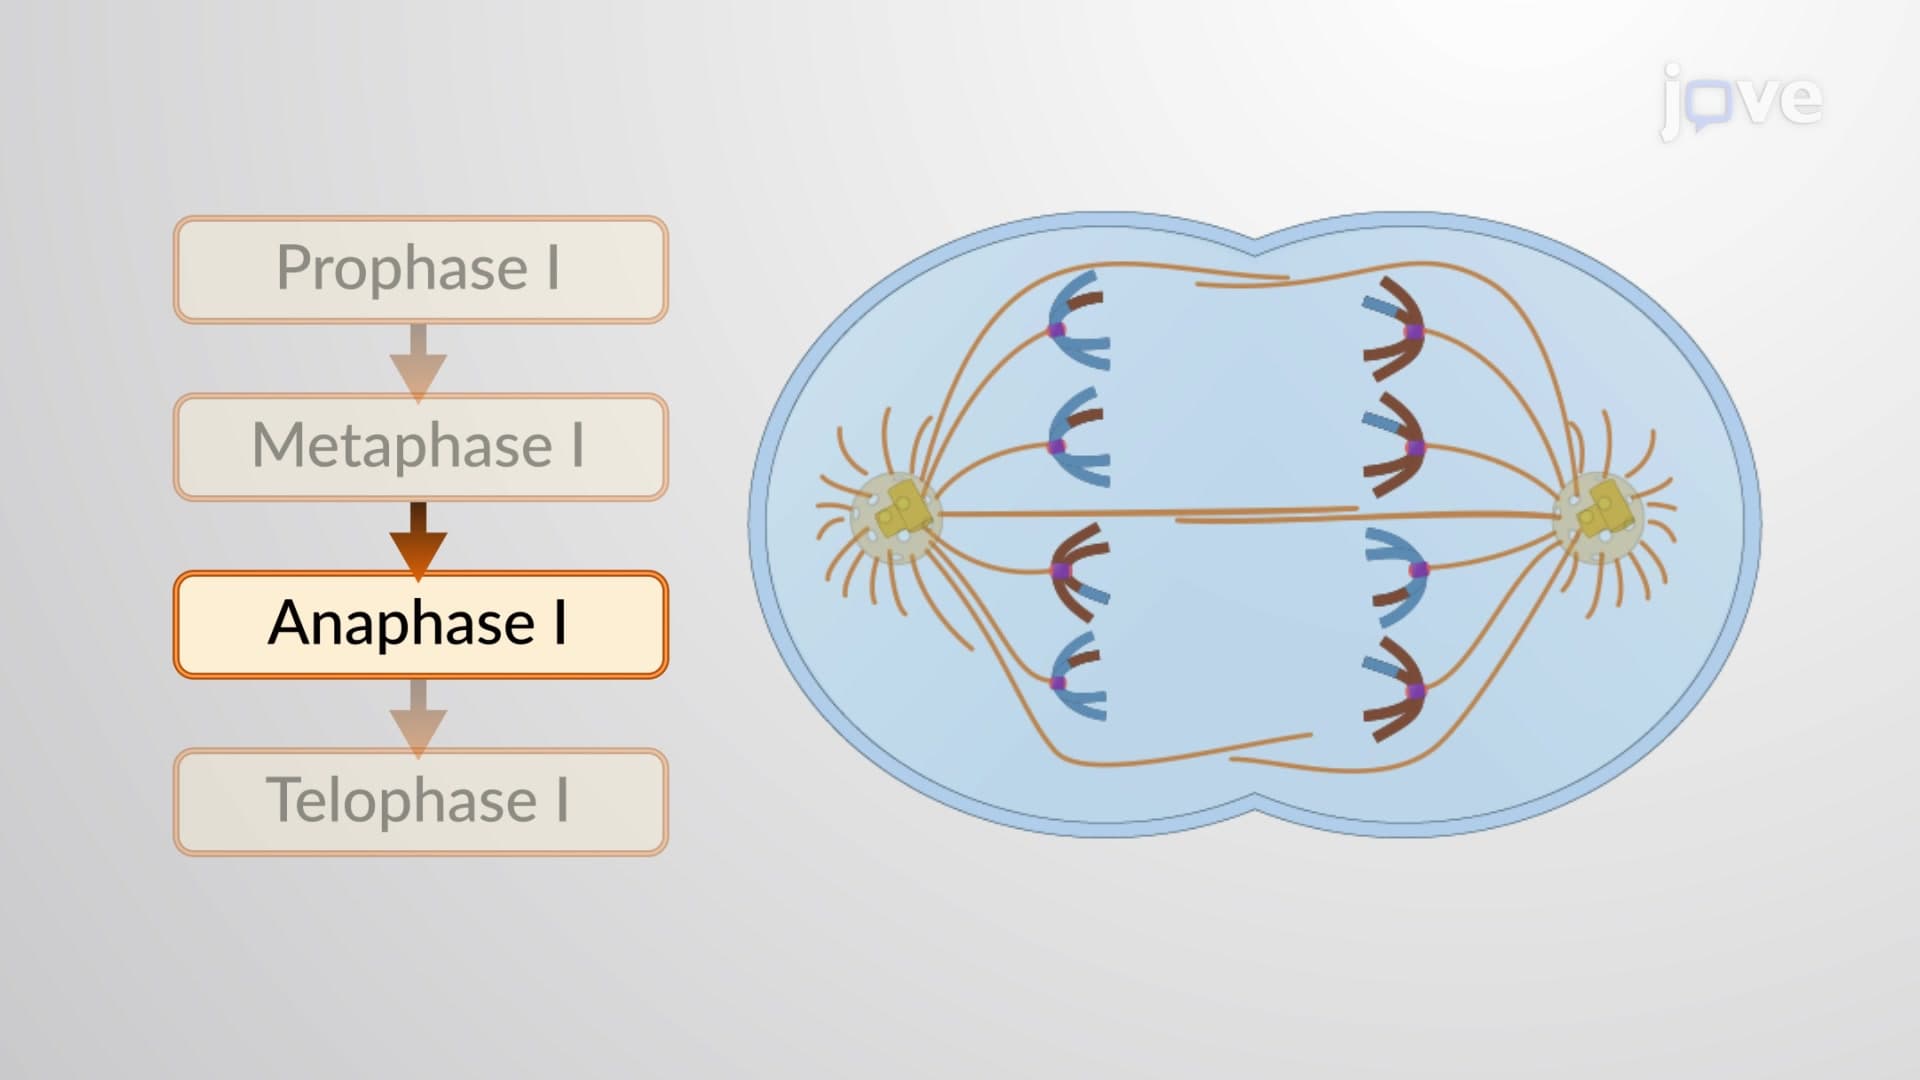
Meiosis I
 03:09
Meiosis I
Prophase I is the most extended and complex step of meiosis I characterized by synapsis, chromosome pairing, and recombination of the homologous chromosomes. This process is facilitated by a proteinaceous structure called the...
 01:22
Lysosomal Hydrolases
 01:21
Huntington Disease l: Introduction
 01:21
Overview of Protein Metabolism
Amino acids play various roles in the body once they are absorbed into cells. They are restructured...