Related Experiment Video
Updated: Jun 17, 2026

Experimental Model to Evaluate Resolution of Pneumonia
Published on: February 17, 2023
Respiratory disease in Niemann-Pick type C2 is caused by pulmonary alveolar proteinosis
M Griese1, F Brasch, V R Aldana
1Dr von Haunersches Kinderspital, University of Munich, Munich, Germany. Matthias.griese@med.uni-muenchen.de

Abstract:
Niemann-Pick diseases are hereditary neurovisceral lysosomal lipid storage disorders, of which the rare type C2 almost uniformly presents with respiratory distress in early infancy. In the patient presented here, the NPC2 exon 4 frameshift mutation c.408_409delAA caused reduced NPC2 protein levels in serum and lung lavage fluid and the synthesis of an aberrant, larger sized protein of around 28 kDa. Protein expression was strongly reduced also in alveolar macrophages. The infant developed failure to thrive and tachypnea. Lung lavage, computer tomography, and histology showed typical signs of pulmonary alveolar proteinosis with an abnormal intraalveolar accumulation of surfactant as well as macrophages. An NPC2-hypomorph animal model also showed pulmonary alveolar proteinosis and accumulation of macrophages in the lung, liver, and spleen long before the mice died. Due to the elevation of cholesterol, the surfactant had an abnormal composition and function. Despite the removal of large amounts of surfactant from the lungs by therapeutic lung lavages, this treatment was only temporarily successful and the infant died of respiratory failure. Our data indicate that respiratory distress in NPC2 disease is associated with a loss of normal NPC2 protein expression in alveolar macrophages and the accumulation of functionally inactive surfactant rich in cholesterol.
More Related Videos
Related Concept Videos
 01:20
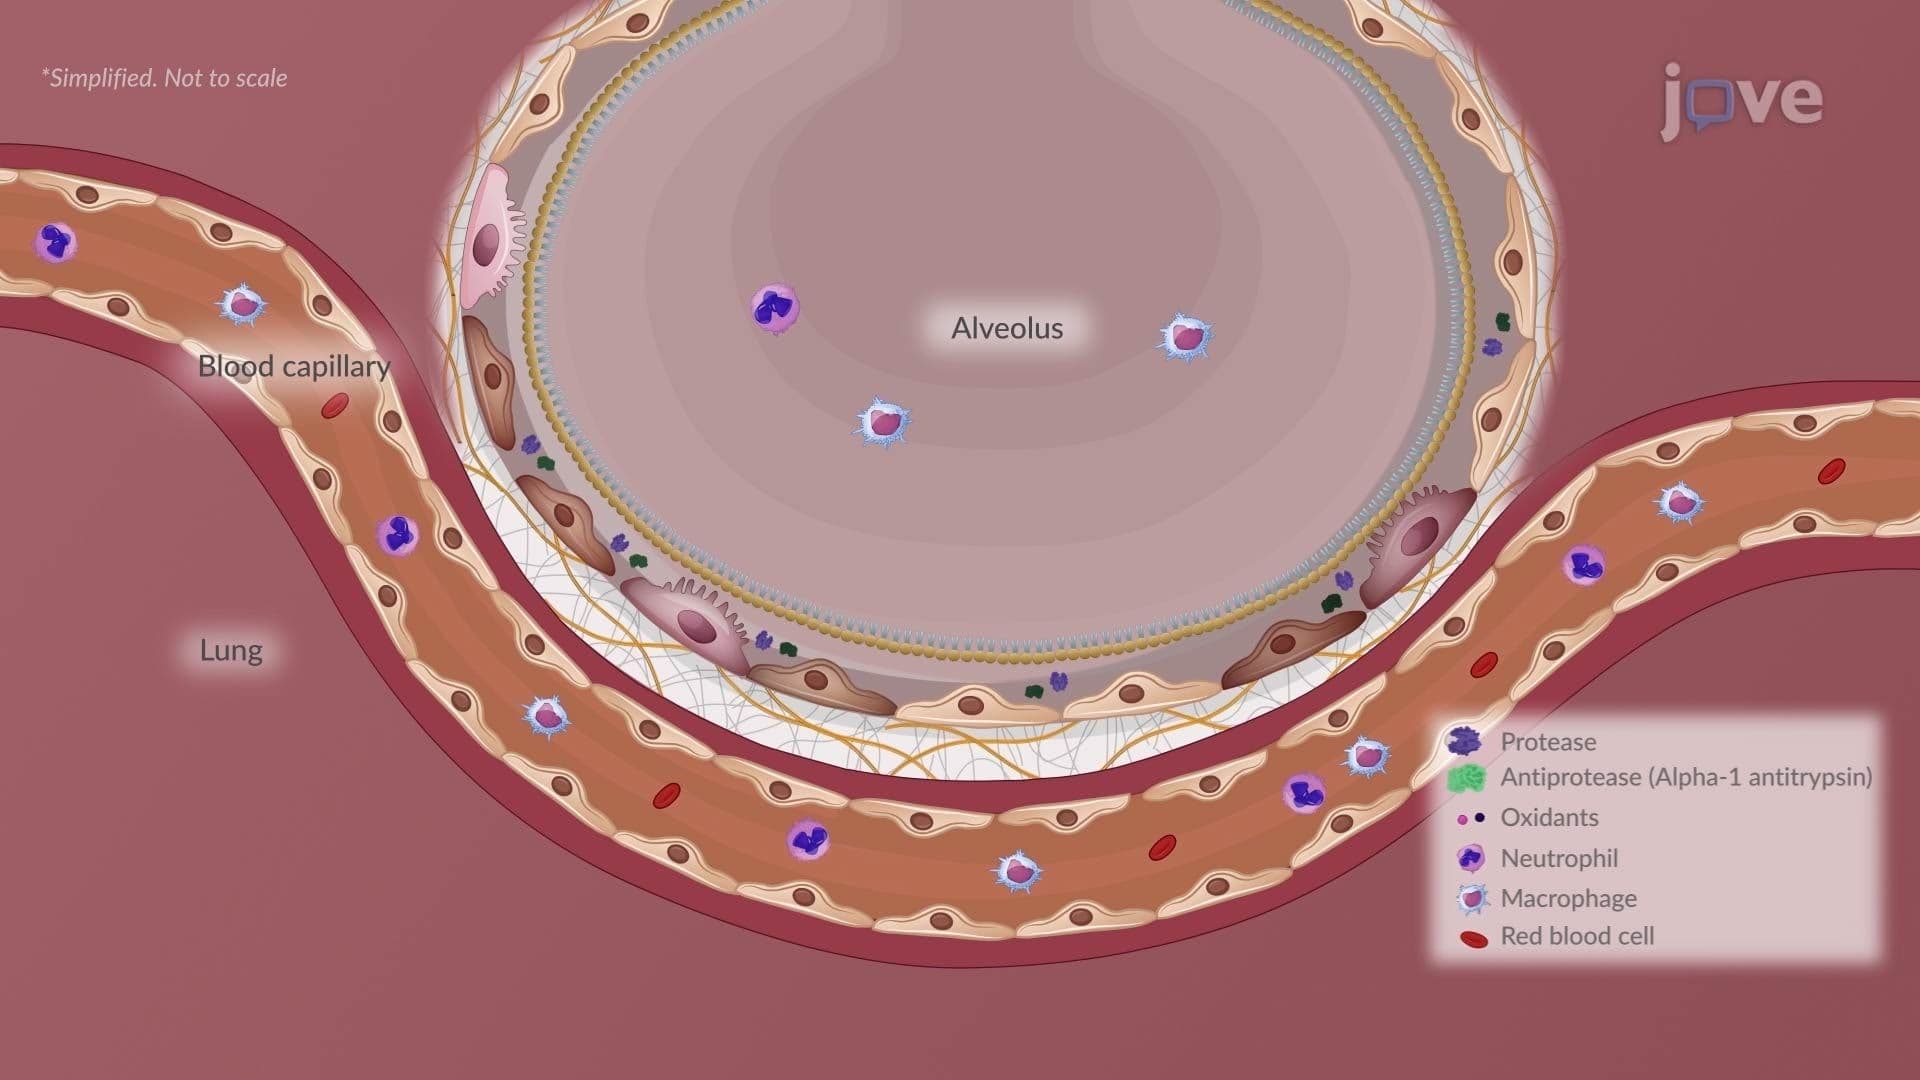
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
Chronic Inflammation
 01:23
Chronic Obstructive Pulmonary Disease II: Emphysema
 01:30
Acute Respiratory Failure-III
 01:29
Pneumonia II: Pathophysiology
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:29
Respiratory Syncytial Virus Disease