Related Experiment Video
Updated: Jun 17, 2026
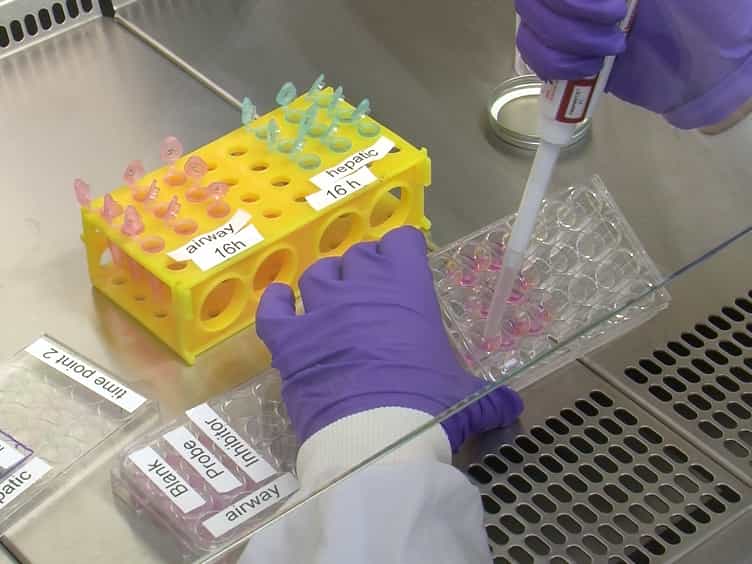

Mass Spectrometry and Luminogenic-based Approaches to Characterize Phase I Metabolic Competency of In Vitro Cell Cultures
Published on: March 28, 2017
Current cytochrome P450 phenotyping methods applied to metabolic drug-drug interaction prediction in dogs
Beth Miskimins Mills1, Matthew J Zaya, Rodney R Walters
1University of Iowa, Division of Medicinal and Natural Products Chemistry, Iowa City, Iowa, USA.

Abstract:
Recombinant cytochrome P450 (P450) phenotyping, different approaches for estimating fraction metabolized (f(m)), and multiple measures of in vivo inhibitor exposure were tested for their ability to predict drug interaction magnitude in dogs. In previous reports, midazolam-ketoconazole interaction studies in dogs have been attributed to inhibition of CYP3A pathways. However, in vitro phenotyping studies demonstrated higher apparent intrinsic clearances (CL(int,app)) of midazolam with canine CYP2B11 and CYP2C21. Application of activity correction factors and isoform hepatic abundance to liver microsome CL(int,app) values further implicated CYP2B11 (f(m) >or= 0.89) as the dog enzyme responsible for midazolam- and temazepam-ketoconazole interactions in vivo. Mean area under the curve (AUC) in the presence of the inhibitor/AUC ratios from intravenous and oral midazolam interaction studies were predicted well with unbound K(i) and estimates of unbound hepatic inlet inhibitor concentrations and intestinal metabolism using the AUC-competitive inhibitor relationship. No interactions were observed in vivo with bufuralol, although significant interactions with bufuralol were predicted with fluoxetine via CYP2D and CYP2C pathways (>2.45-fold) but not with clomipramine (<2-fold). The minor caffeine-fluvoxamine interaction (1.78-fold) was slightly higher than predicted values based on determination of a moderate f(m) value for CYP1A1, although CYP1A2 may also be involved in caffeine metabolism. The findings suggest promise for in vitro approaches to drug interaction assessment in dogs, but they also highlight the need to identify improved substrate and inhibitor probes for canine P450s.
Related Concept Videos
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:27
Pharmacogenetics of Drug Metabolism: Overview
 01:17
Drug Metabolism: Phase I Reactions
 01:30
Drug toxicity: Drug–Drug Interaction
 01:16
Drug toxicity: Idiosyncratic Reactions
 01:29
Pharmacogenetic Phenotypes: Alterations in Pharmacokinetics, Drug Targets and Biologic Milieu