Related Experiment Video
Updated: Jun 17, 2026
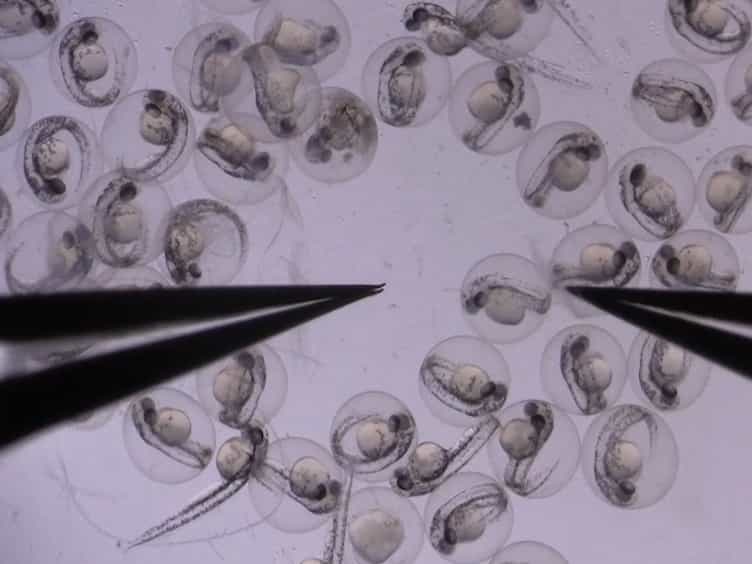
Using Zebrafish Models of Human Influenza A Virus Infections to Screen Antiviral Drugs and Characterize Host Immune Cell Responses
Published on: January 20, 2017
Reconstructing the initial global spread of a human influenza pandemic: A Bayesian spatial-temporal model for the
Philippe Lemey1, Marc Suchard, Andrew Rambaut
1UCLA and Institute of Evolutionary Biology, University of Edinburgh, Edinburgh & The Fogarty International Center, NIH.

Abstract:
Here, we present an analysis of the H1N1pdm genetic data sampled over the initial stages in the epidemic. To infer phylodynamic spread in time and space we employ a recently developed Bayesian statistical inference framework (Lemey et al., in press). We model spatial diffusion as a continuous-time Markov chain process along time-measured genealogies. In this analysis, we consider 40 locations for which sequence data were available on 06-Aug-2009. The sampling time interval of the 242 sequences spans from 30-Mar-2009 to 12-Jul-2009. The Bayesian inference typically results in a posterior distribution of phylogenetic trees, each having an estimate of the epidemic locations at the ancestral nodes in the tree. We summarize these trees using the most representative clustering pattern and annotate these clusters with the most probable location states. We can visualize this information as tree that grows over time, seeding locations each time an ancestral node is inferred to exist at a different location. A Bayes factor test provides statistical support for epidemiological linkage throughout the evolutionary history. We demonstrate how our full probabilistic approach efficiently tracks an epidemic based on viral genetic data as it unfolds across the globe.
More Related Videos
Related Concept Videos
 01:18
Steps in Outbreak Investigation
 01:28
Infectious Diseases and Their Occurrence
 00:57
Viral Recombination
 00:36
Viral Mutations
 01:21
Causality in Epidemiology
 01:27
Influenza