
Structural parameters and vibrational spectra of a series of zinc meso-phenylporphyrins: a DFT and experimental study
Ying-Hui Zhang1, Wei Zhao, Ping Jiang
1Department of Chemistry, Nankai University, Weijin Road 94, Tianjin 300071, People's Republic of China. zhangyhi@nankai.edu.cn

Abstract:
The influences of meso-phenyl substitution on the geometric structure and vibrational spectra have been studied by DFT calculation (B3LYP/6-31G(d)) and experiment on a series of zinc porphyrins (ZnTPP: zinc 5,10,15,20-tetraphenylporphyrin; ZnTrPP: zinc 5,10,15-triphenylporphryin; ZnDPP: zinc 5,15-dipenylporphyirn; ZnMPP: zinc 5-monophenylporphyrin; ZnP: zinc porphine). Calculation indicates that meso-phenyl substitution gives rise to slight out-of-plane distortion but significant in-plane distortion, especially for the configuration around C(m) atom, to zinc porphyrin. The assignment of experimental vibrational spectra was proposed mainly on the basis of calculation. Different shifting tendency upon meso-phenyl substitution is observed for different structure-sensitive bands, such as the shifting of nu(2), nu(3), nu(6), and nu(8) modes toward higher frequencies and nu(4) and nu(28) modes toward lower frequencies, upon meso-phenyl substitution. This is attributed primarily to in-plane nuclear reorganization effect (IPNR), though the contribution from out-of-plane distortion cannot be excluded completely. Analysis on vibrational structure reveals that asymmetric meso-phenyl substitution, especially the 5,15-diphenyl substitution of ZnDPP, brings about asymmetric vibrational displacement, or even splitting of vibrational structure to normal modes involving mainly the motion of meso-C(m). This is ascribed to the reduction of symmetry of porphyrin skeleton caused by asymmetric meso-phenyl substitution.
Related Concept Videos
 01:29
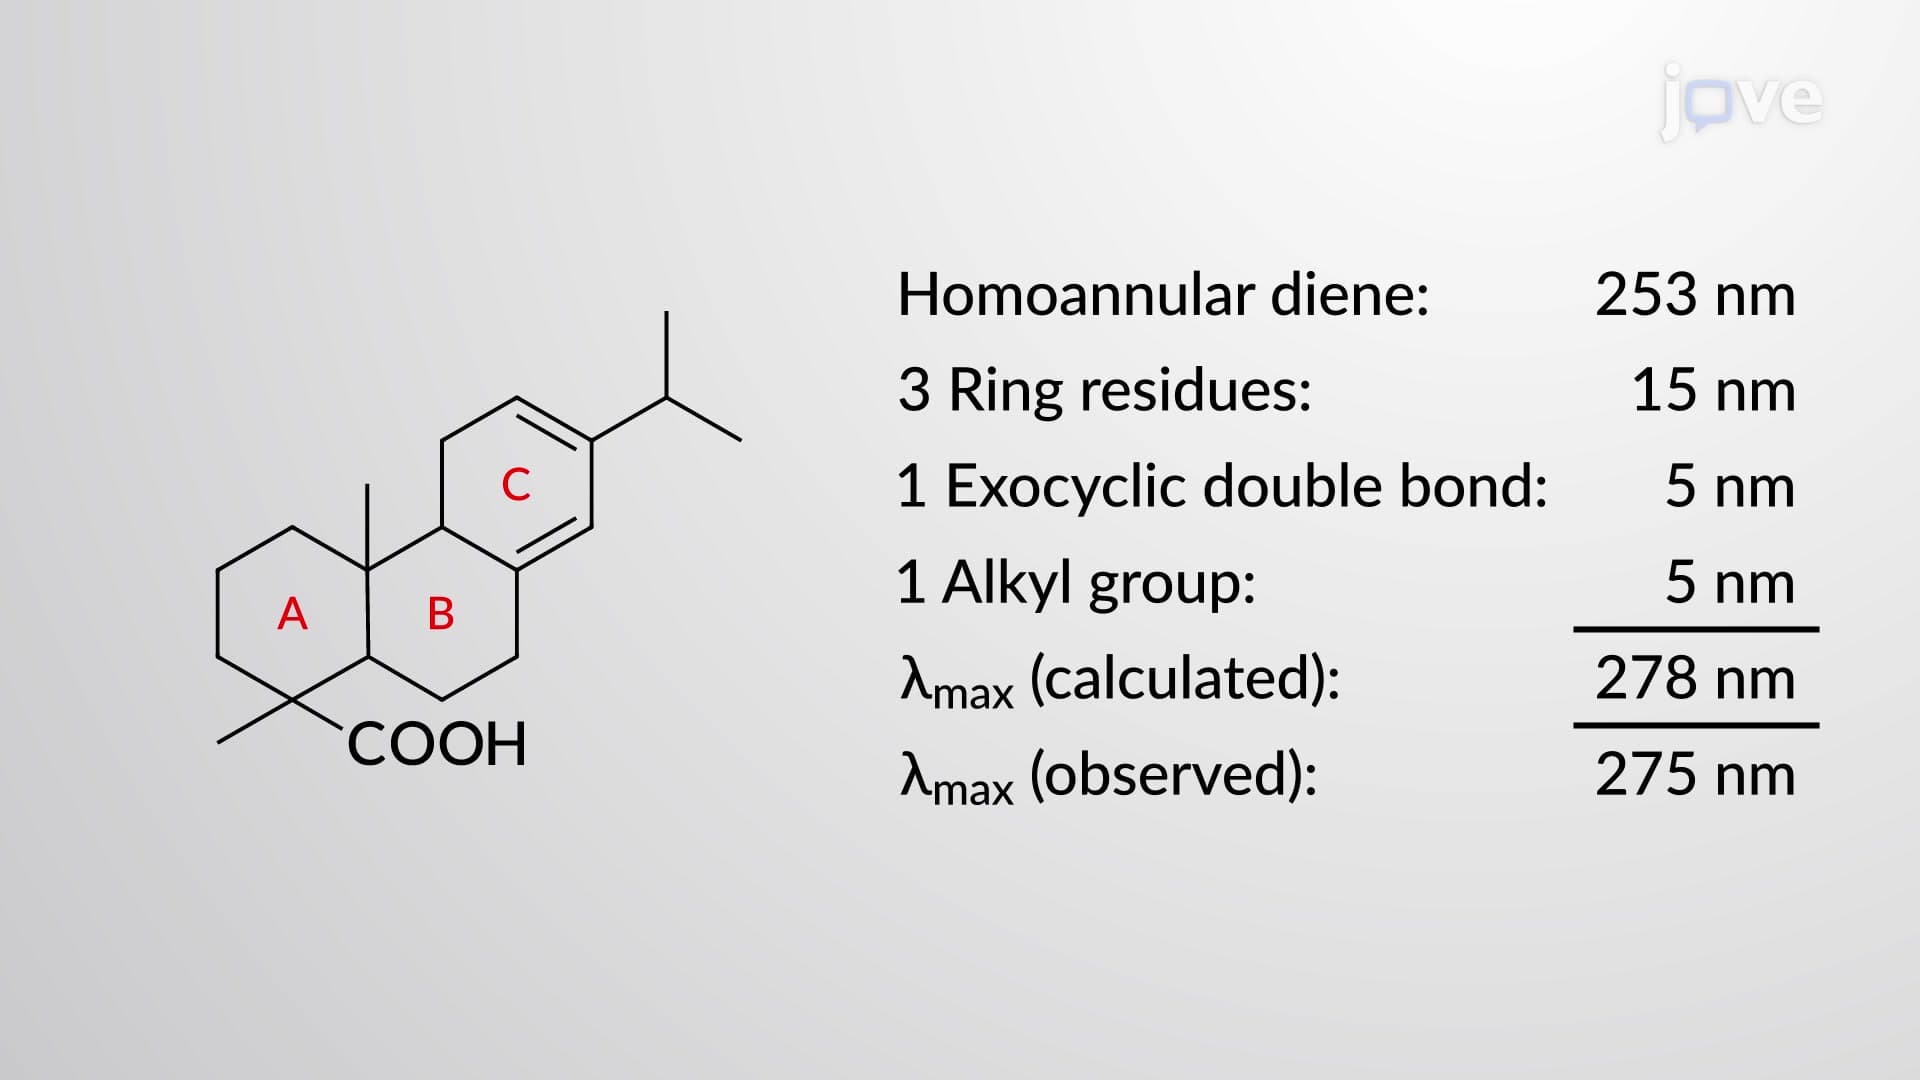
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:18
Structure of Benzene: Molecular Orbital Model
 01:37
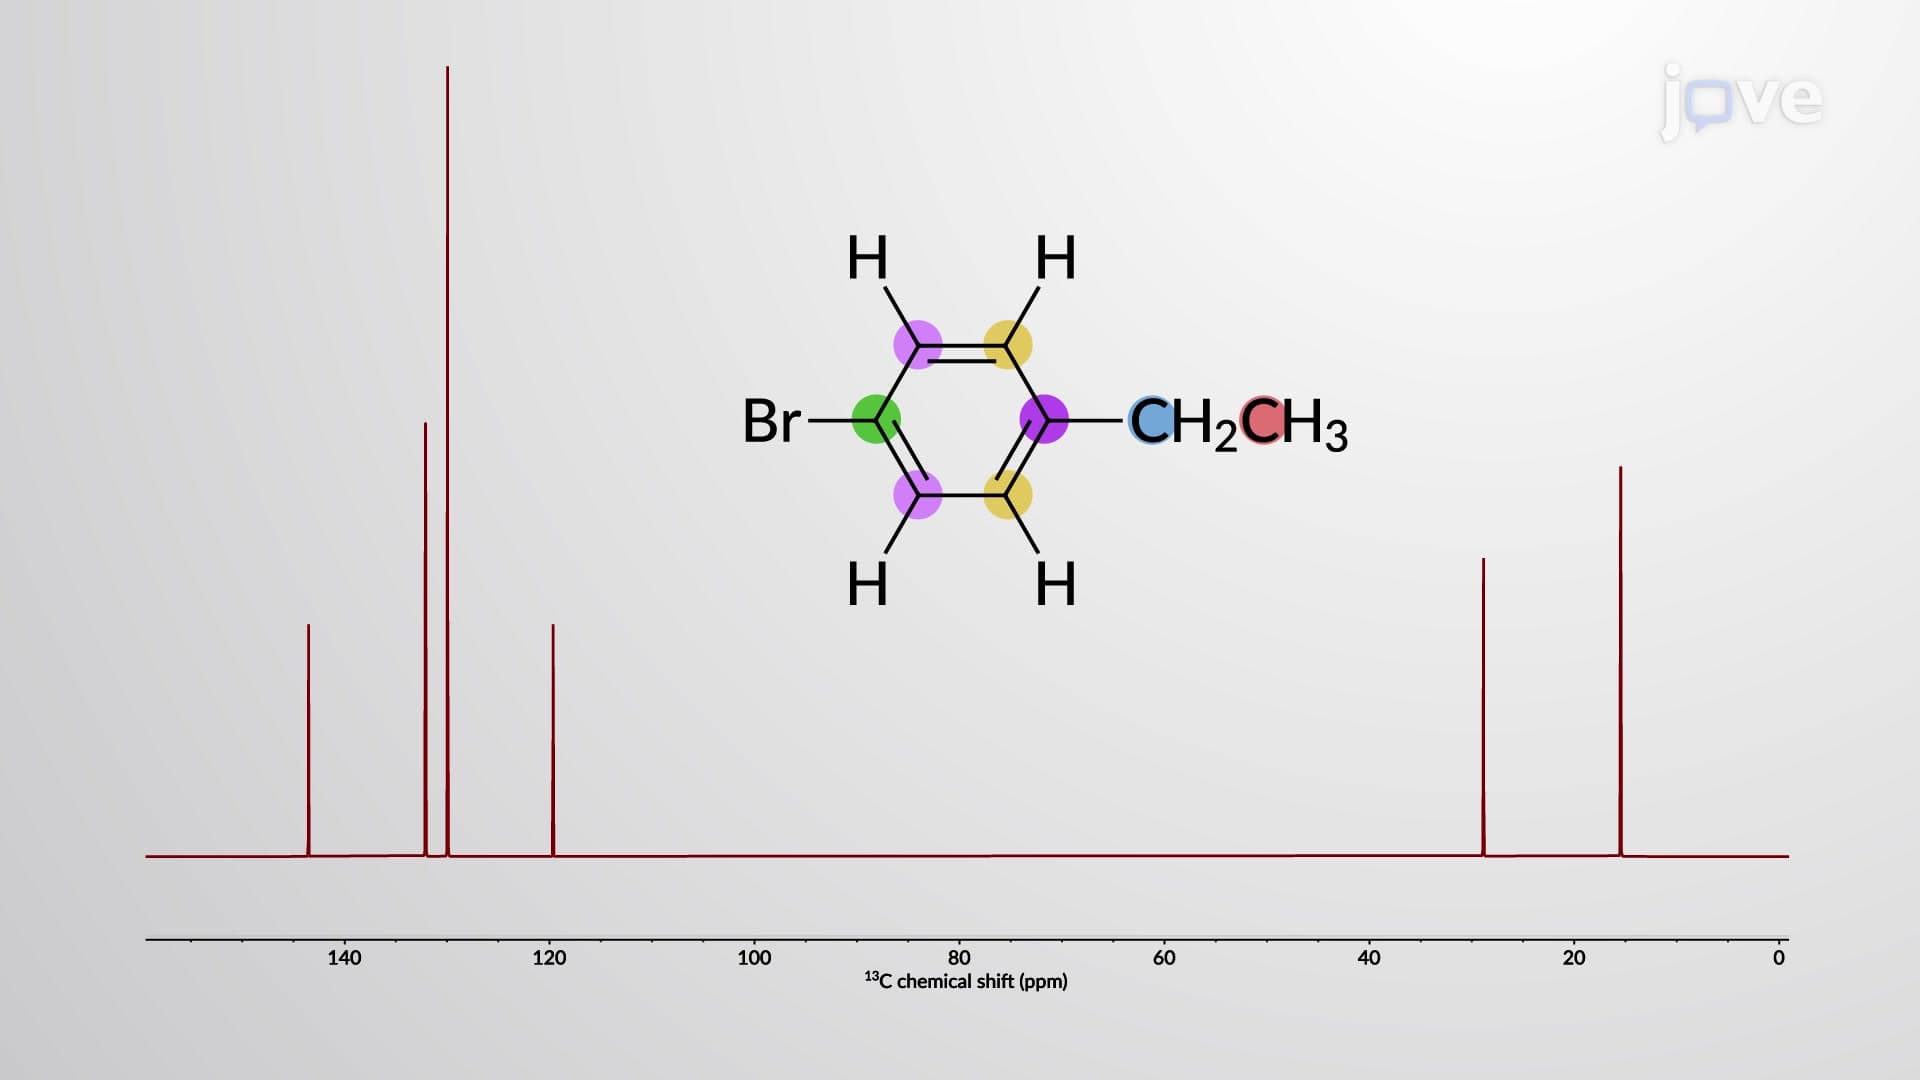
NMR Spectroscopy of Benzene Derivatives
 01:26
Variables Affecting Phosphorescence and Fluorescence
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:32
UV–Vis Spectroscopy of Conjugated Systems
One of the factors influencing λmax is the extent of conjugation in the...