Related Experiment Video
Updated: Jun 17, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Theoretical study of one- and two-photon absorption spectra of azoaromatic compounds
D L Silva1, P Krawczyk, W Bartkowiak
1Instituto de Física de São Carlos, Universidade de São Paulo, CP 369, 13560-970 São Carlos, São Paulo, Brazil. dlsilva@ifsc.usp.br

Abstract:
In this study, the one- and two-photon absorption spectra of seven azoaromatic compounds (five pseudostilbenes-type and two aminoazobenzenes) were theoretically investigated using the density functional theory combined with the response functions formalism. The equilibrium molecular structure of each compound was obtained at three different levels of theory: Hartree-Fock, density functional theory (DFT), and Møller-Plesset 2. The effect of solvent on the equilibrium structure and the electronic transitions of the compounds were investigated using the polarizable continuum model. For the one-photon absorption, the allowed pi-->pi(*) transition energy showed to be dependent on the molecular structures and the effect of solvent, while the n-->pi(*) and pi-->pi(*)(n) transition energies exhibited only a slight dependence. An inversion between the bands corresponding to the pi-->pi(*) and n-->pi(*) states due to the effect of solvent was observed for the pseudostilbene-type compounds. To characterize the allowed two-photon absorption transitions for azoaromatic compounds, the response functions formalism combined with DFT using the hybrid B3LYP and PBE0 functionals and the long-range corrected CAM-B3LYP functional was employed. The theoretical results support the previous findings based on the three-state model. The model takes into account the ground and two electronic excited states and has already been used to describe and interpret the two-photon absorption spectrum of azoaromatic compounds. The highest energy two-photon allowed transition for the pseudostilbene-type compounds shows to be more effectively affected (approximately 20%) by the torsion of the molecular structure than the lowest allowed transition (approximately 10%). In order to elucidate the effect of the solvent on the two-photon absorption spectra, the lowest allowed two-photon transition (dipolar transition) for each compound was analyzed using a two-state approximation and the polarizable continuum model. The results obtained reveal that the effect of solvent increases drastically the two-photon cross-section of the dipolar transition of the pseudostilbene-type compounds. In general, the features of both one- and two-photon absorption spectra of the azoaromatic compounds are well reproduced by the theoretical calculations.
Related Concept Videos
 01:23
Atomic Spectroscopy: Absorption, Emission, and Fluorescence
 01:14
Molecular Spectroscopy: Absorption and Emission
 01:11
Aryldiazonium Salts to Azo Dyes: Diazo Coupling
 01:14
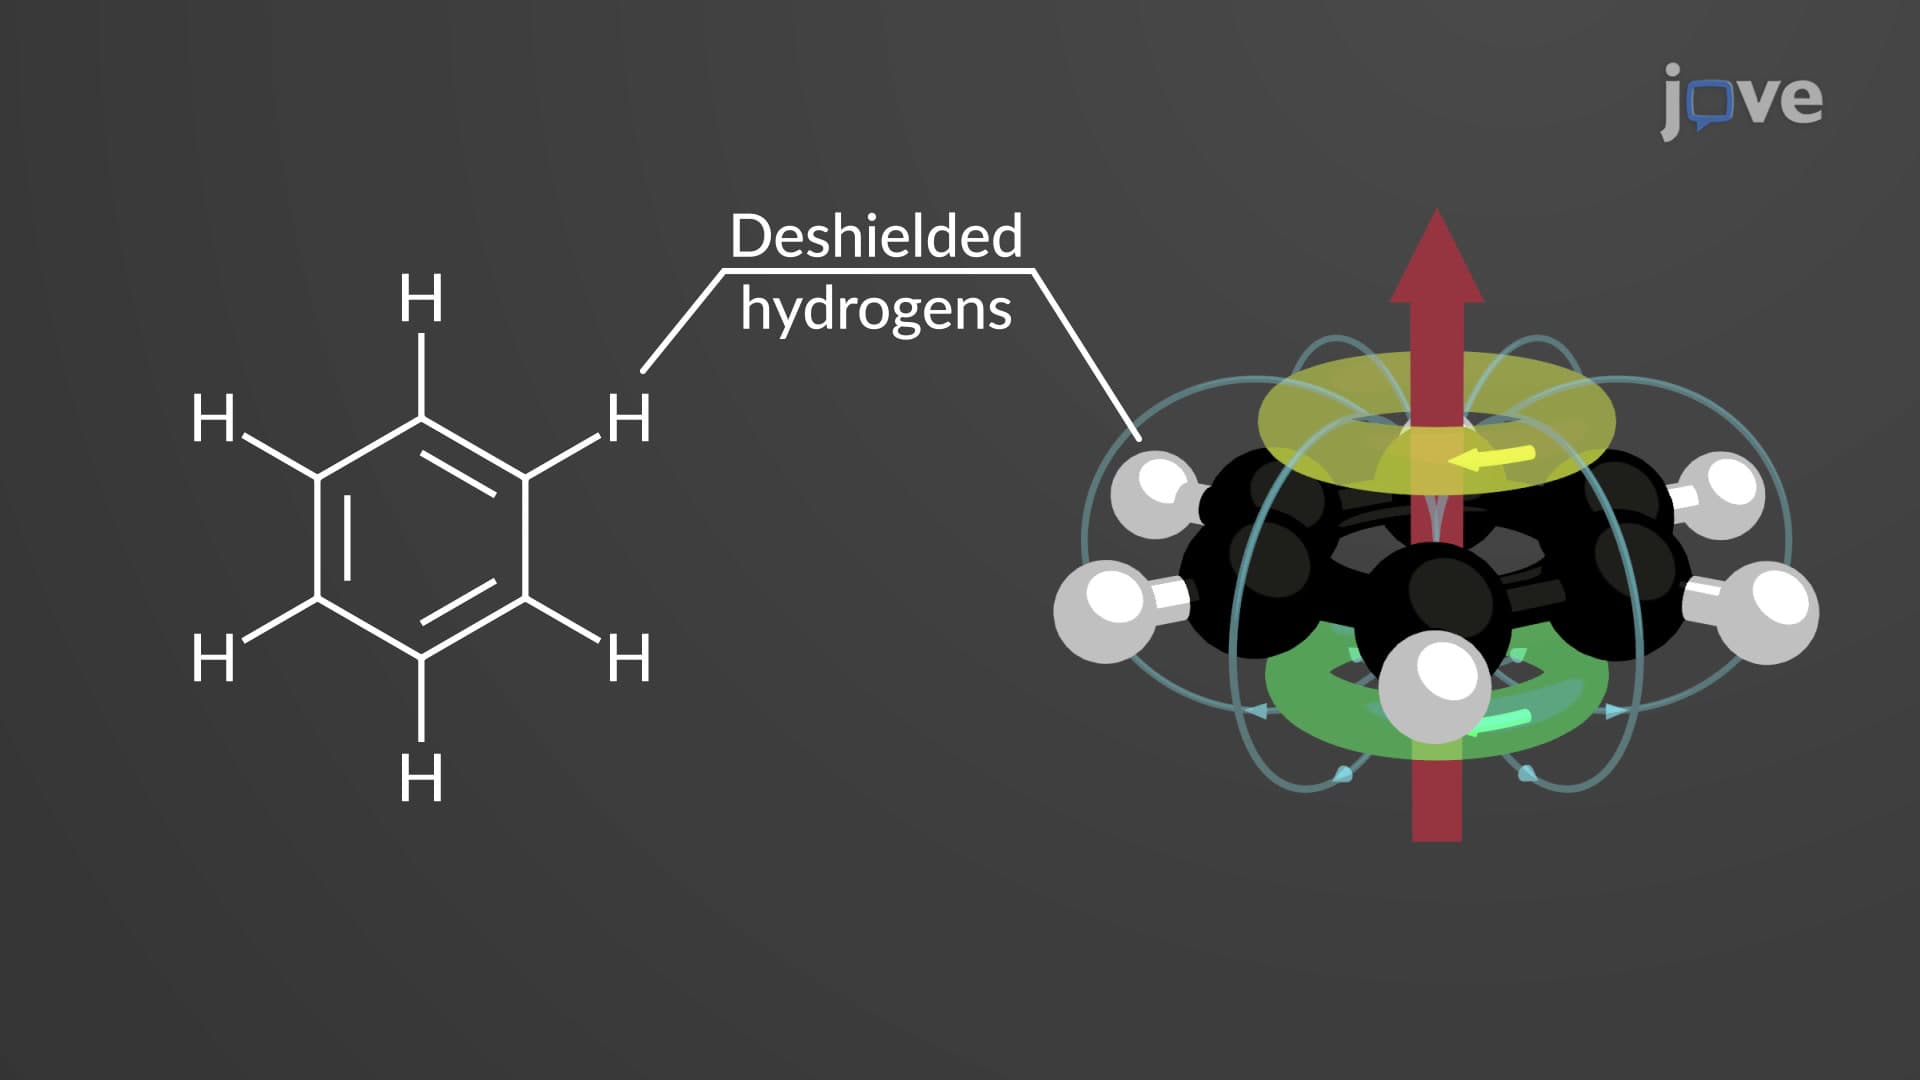
NMR Spectroscopy of Aromatic Compounds
 01:32
UV–Vis Spectroscopy of Conjugated Systems
One of the factors influencing λmax is the extent of conjugation in the...
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds