Related Experiment Video
Updated: Jun 17, 2026

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
On the SmCo dimer: a detailed density functional theory analysis
1BK21 Physics Program and Department of Physics, Chungbuk National University, Cheongju 361-763, Korea. hoymak@atilim.edu.tr

Abstract:
Making use of 21 different exchange-correlation functionals, we performed density functional theory calculations, within the effective core potential level, to investigate some spectroscopic and electronic features of the SmCo dimer in its ground state. A particular emphasis was placed on the (spin) multiplicity of SmCo. Most of the functionals under discussion unanimously agreed that the multiplicity of SmCo should be 10. It was observed that the nature of interaction between Sm and Co atoms to form the SmCo dimer can be described, to a good approximation, by a Lennard-Jones curve. For the multiplicity value 10, the binding energy D(e) was seen to be in the range 1.08-1.77 eV, while the equilibrium separation distance and the fundamental frequency were found to be r(e) = 2.975 +/- 0.035 A and omega(e) = 120 +/- 10 cm(-1), respectively.
Related Concept Videos
 02:40
MO Theory and Covalent Bonding
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:35
Molecular Orbital Theory I
 02:36
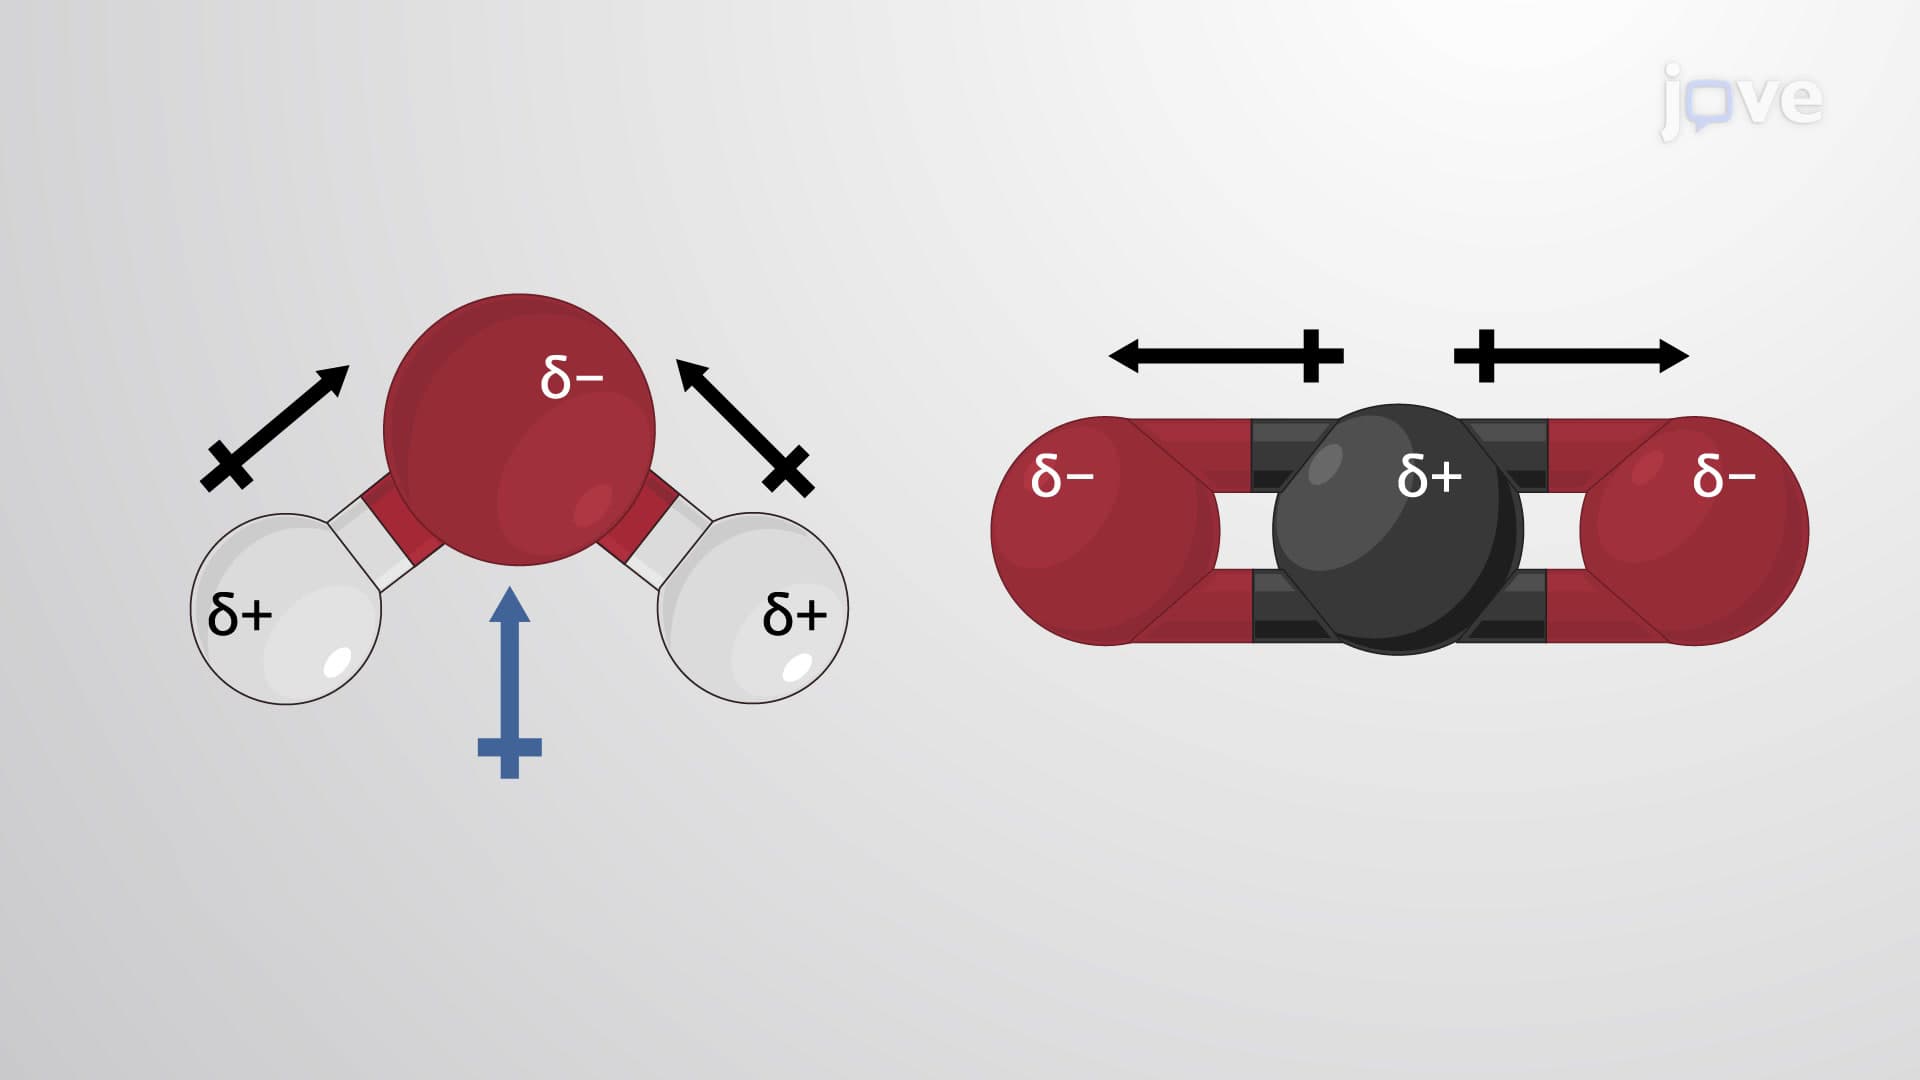
Molecular Geometry and Dipole Moments
 02:42
Valence Bond Theory
 03:51
Molecular Orbital Theory II