Reducing phylogenetic bias in correlated mutation analysis
Related Experiment Video
Updated: Jun 17, 2026
Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
Haim Ashkenazy1, Yossef Kliger
1Compugen Ltd, 72 Pinchas Rosen, Tel Aviv 69512, Israel.

Abstract:
Correlated mutation analysis (CMA) is a sequence-based approach for ab initio protein contact map prediction. The basis of this approach is the observed correlation between mutations in interacting amino acid residues. These correlations are often estimated by either calculating the Pearson's correlation coefficient (PCC) or the mutual information (MI) between columns in a multiple sequence alignment (MSA) of the protein of interest and its homologs. A major challenge of CMA is to filter out the background noise originating from phylogenetic relatedness between sequences included in the MSA. Recently, a procedure to reduce this background noise was demonstrated to improve an MI-based predictor. Herein, we tested whether a similar approach can also improve the performance of the classical PCC-based method. Indeed, performance improvements were achieved for all four major SCOP classes. Furthermore, the results reveal that the improved PCC-based method is superior to MI-based methods for proteins having MSAs of up to 100 sequences.
Related Concept Videos
 02:54
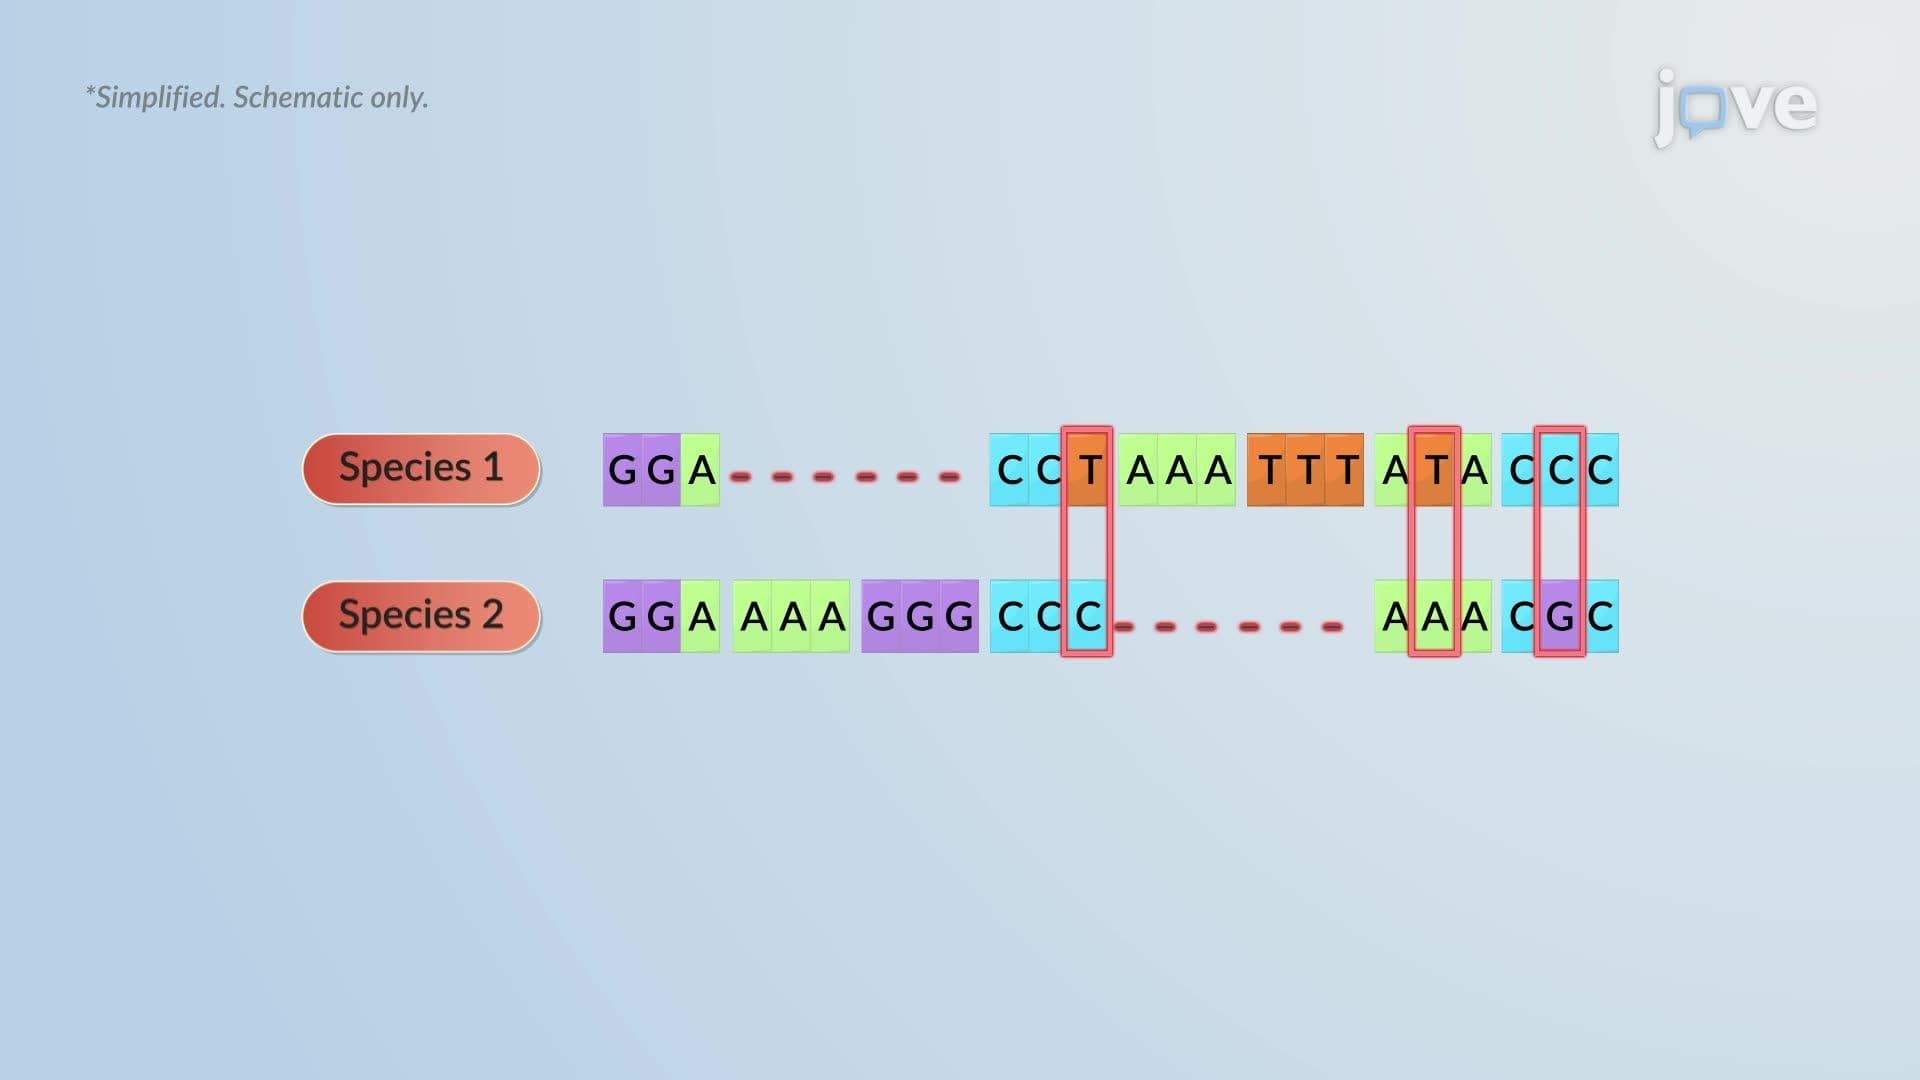
Evolutionary Relationships through Genome Comparisons
 01:28
Microbial Phylogeny
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...
02:05Gene Evolution - Fast or Slow?
In contrast, regions which code...
 01:09
Mutation, Gene Flow, and Genetic Drift
 01:20
Mismatch Repair
The Mutator Protein Family Plays a Key Role in DNA Mismatch Repair
The human genome has more than 3 billion base pairs of DNA per cell. Prior to cell division, that vast amount of genetic...