Related Experiment Video
Updated: Jun 17, 2026
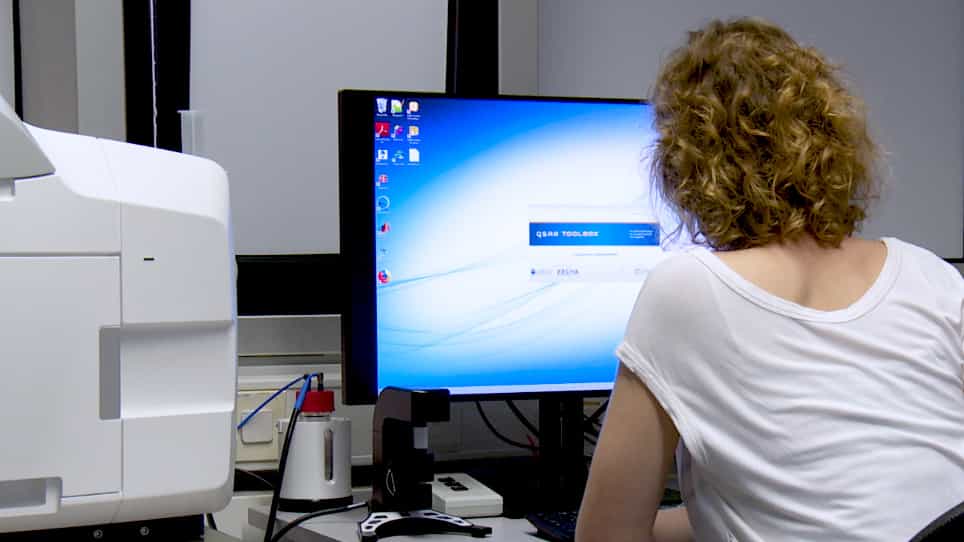

In Silico Modeling Method for Computational Aquatic Toxicology of Endocrine Disruptors: A Software-Based Approach Using QSAR Toolbox
Published on: August 28, 2019
Quantitative structure-activity relationship (QSAR) modeling of EC50 of aquatic toxicities for Daphnia magna
Alan R Katritzky1, Svetoslav H Slavov, Iva S Stoyanova-Slavova
1Center for Heterocyclic Compounds, Department of Chemistry, University of Florida, Gainesville, Florida, USA. katritzky@chem.ufl.edu

Abstract:
The experimental EC(50) toxicities toward Daphnia magna for a series of 130 benzoic acids, benzaldehydes, phenylsulfonyl acetates, cycloalkane-carboxylates, benzanilides, and other esters were studied using the Best multilinear regression algorithm (BMLR) implemented in CODESSA. A modified quantitative structure-activity relationships (QSAR) procedure was applied guaranteeing the stability and reproducibility of the results. Separating the initial data set into training and test subsets generated three independent models with an average R(2) of .735. A five-descriptor general model including all 130 compounds, constructed using the descriptors found effective for the independent subsets, was characterized by the following statistical parameters: R(2) = .712; R(2)(cv) = .676; F = 61.331; s(2) = 0.6. The removal of two extreme outliers improved significantly the statistical parameters: R(2) = .759; R(2)(cv) = .728; F = 77.032; s(2) = 0.499. The sensitivity of the general model to chance correlations was estimated by applying a scrambling procedure involving 20 randomizations of the original property values. The resulting R(2) = .192 demonstrated the high robustness of the model proposed. The descriptors appearing in the obtained models are related to the biochemical nature of the adverse effects. An additional study of the EC(50)/LC(50) relationship for a series of 28 compounds (part of our general data set) revealed that these endpoints correlated with R(2) = .98.
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:15
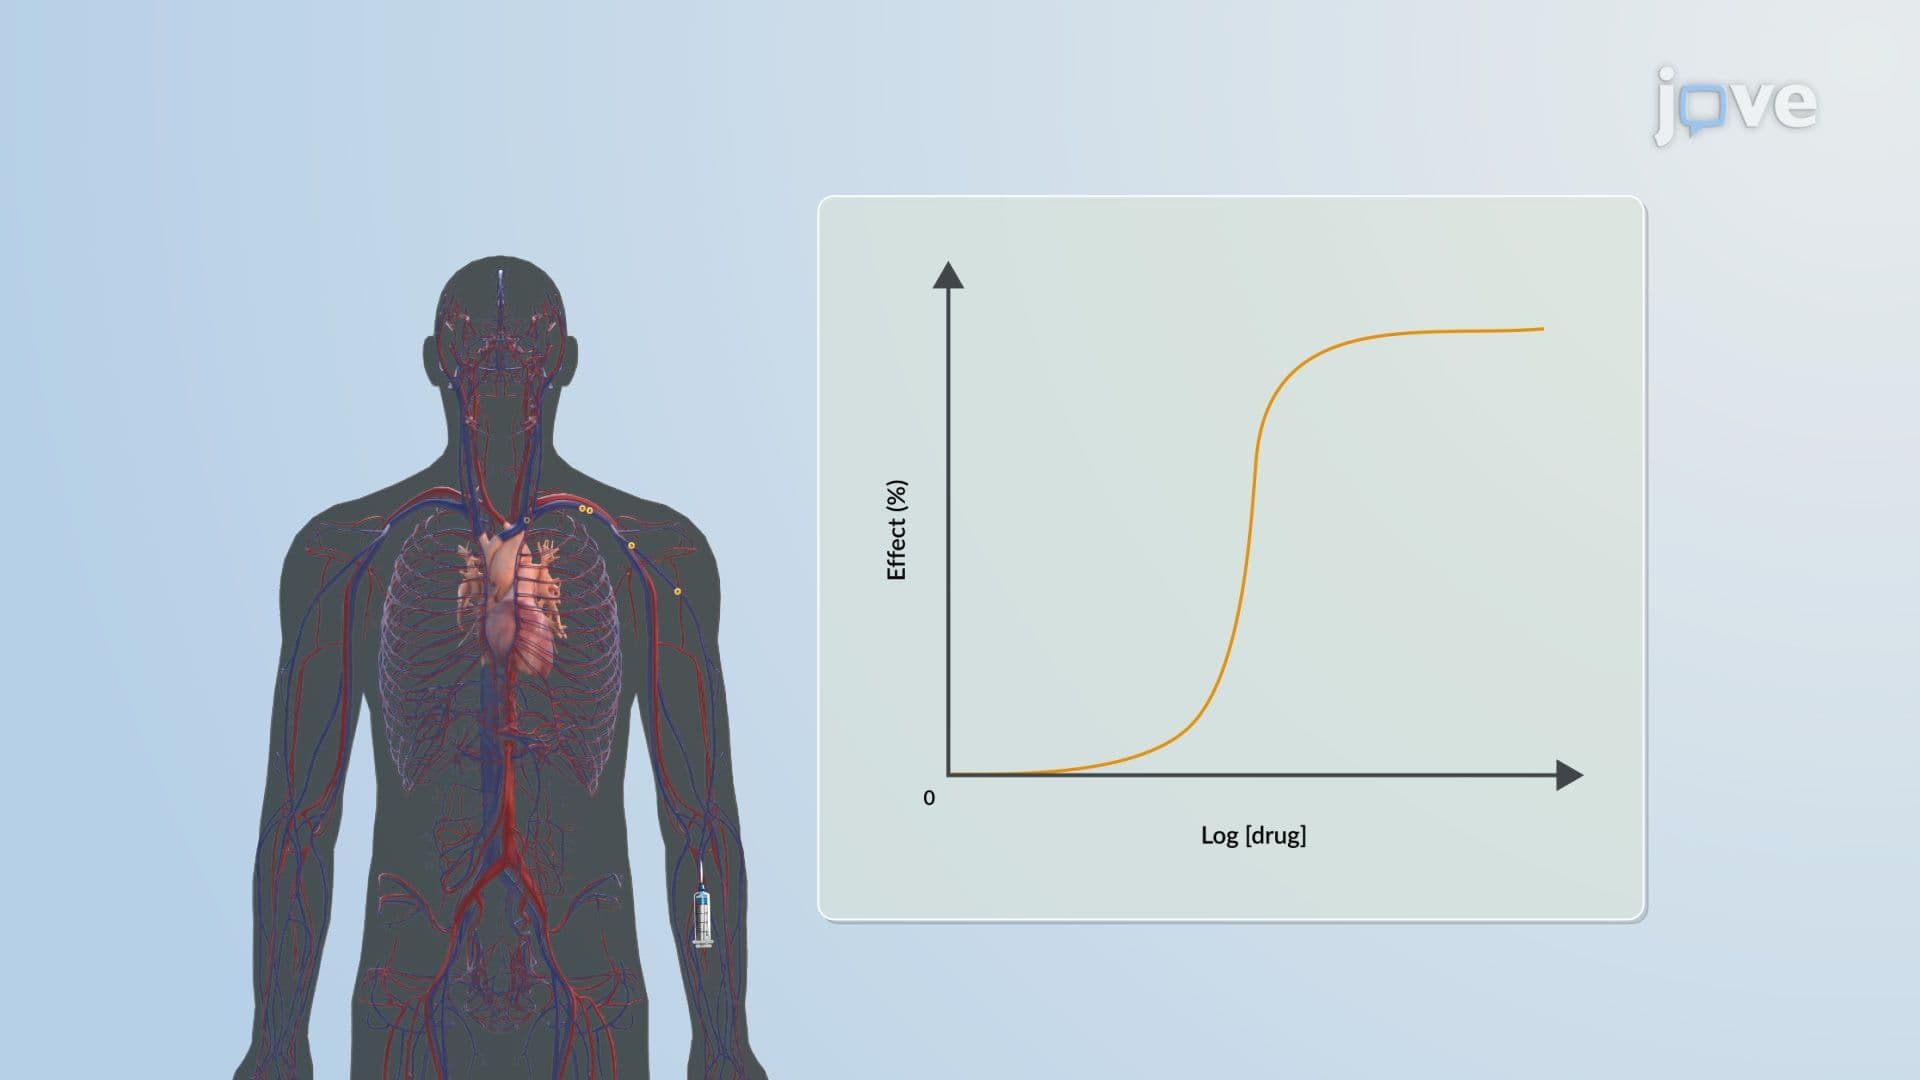
Pharmacodynamic Models: Linear Concentration–Effect Model
 01:23
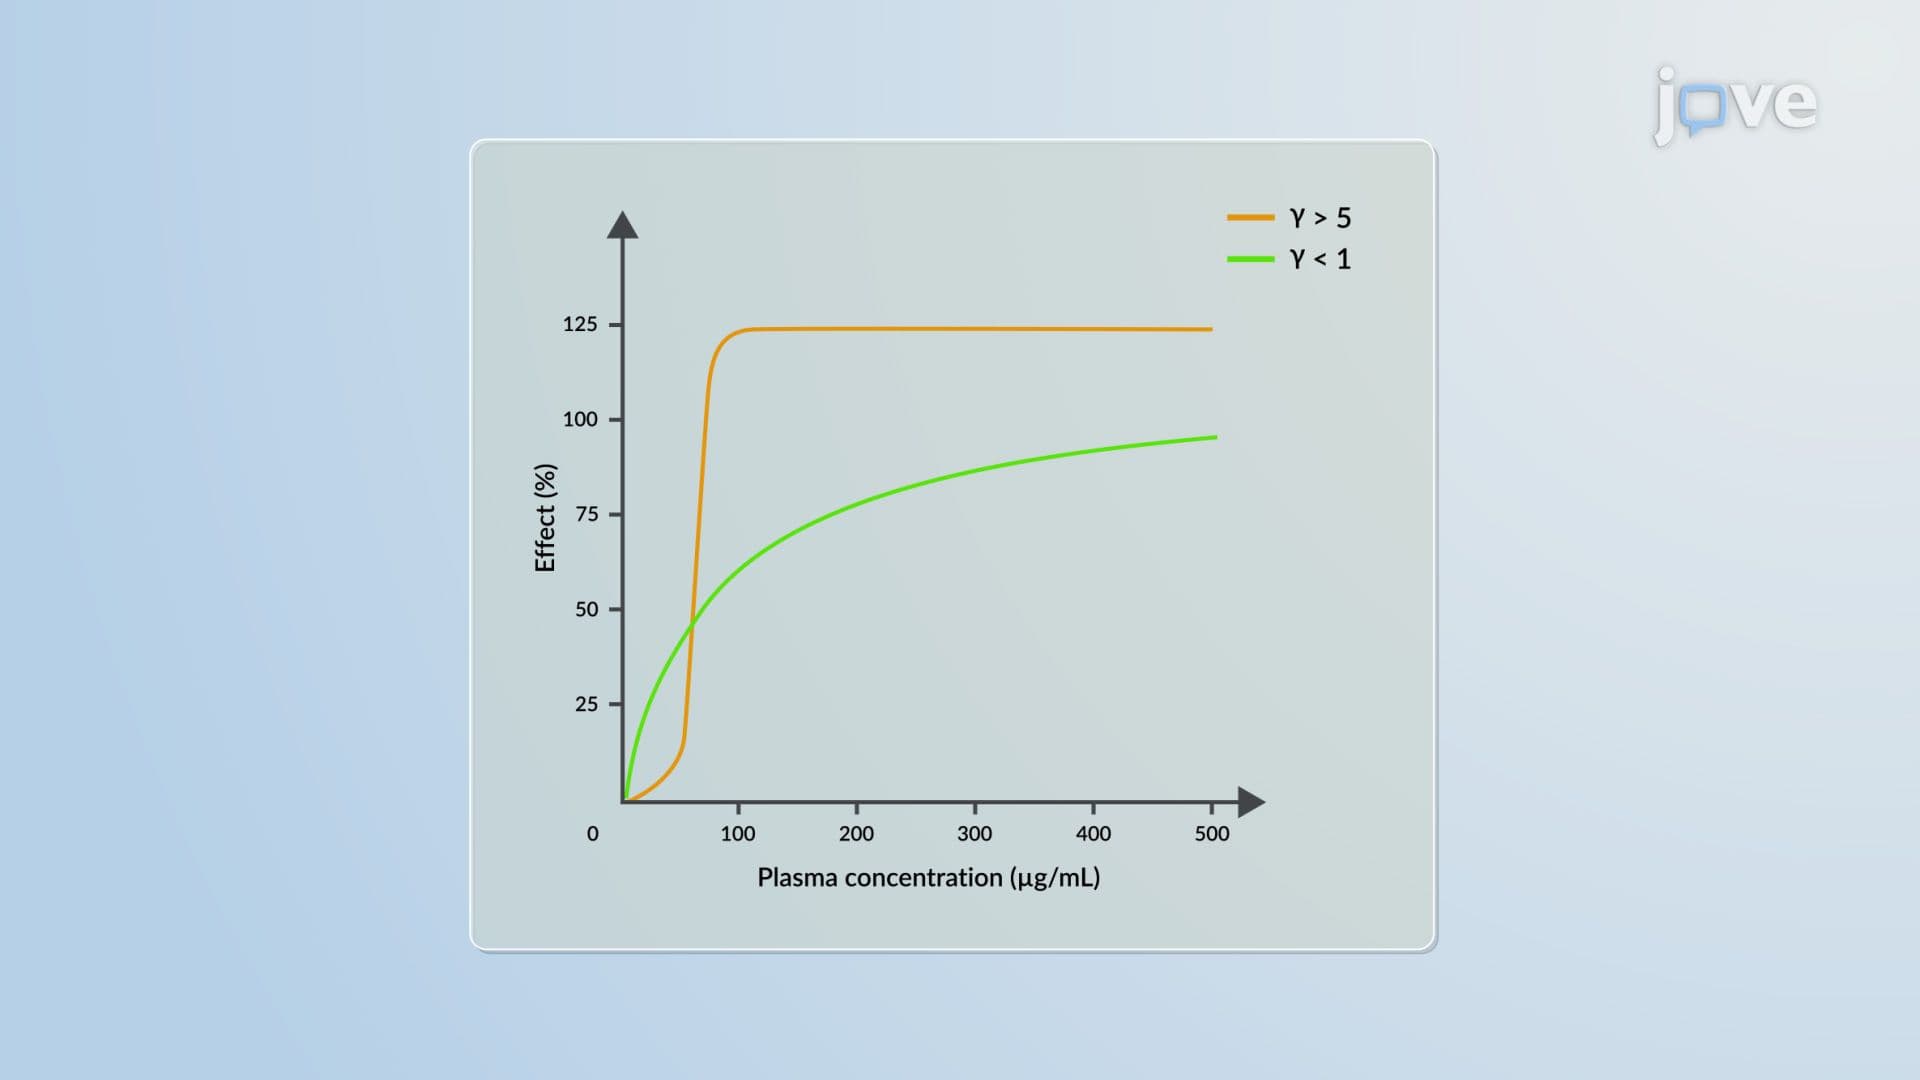
Pharmacokinetic–Pharmacodynamic Relationship: Intensity of Dose-Effect Relationship
 01:18
Pharmacodynamic Models: Emax Drug–Concentration Effect Model
 01:23
Toxicity Testing in Animals
 01:30
Quantitative Aspects of Drug-Receptor Interaction