Related Experiment Video
Updated: Jun 17, 2026
Anaerobic Protein Purification and Kinetic Analysis via Oxygen Electrode for Studying DesB Dioxygenase Activity and Inhibition
Published on: October 3, 2018
Structural basis for cofactor-independent dioxygenation of N-heteroaromatic compounds at the alpha/beta-hydrolase
Roberto A Steiner1, Helge J Janssen, Pietro Roversi
1Randall Division of Cell and Molecular Biophysics, King's College London, New Hunt's House, Guy's Campus, London SE1 1UL, England. roberto.steiner@kcl.ac.uk

Abstract:
Enzymatic catalysis of oxygenation reactions in the absence of metal or organic cofactors is a considerable biochemical challenge. The CO-forming 1-H-3-hydroxy-4-oxoquinaldine 2,4-dioxygenase (HOD) from Arthrobacter nitroguajacolicus Rü61a and 1-H-3-hydroxy-4-oxoquinoline 2,4-dioxygenase (QDO) from Pseudomonas putida 33/1 are homologous cofactor-independent dioxygenases involved in the breakdown of N-heteroaromatic compounds. To date, they are the only dioxygenases suggested to belong to the alpha/beta-hydrolase fold superfamily. Members of this family typically catalyze hydrolytic processes rather than oxygenation reactions. We present here the crystal structures of both HOD and QDO in their native state as well as the structure of HOD in complex with its natural 1-H-3-hydroxy-4-oxoquinaldine substrate, its N-acetylanthranilate reaction product, and chloride as dioxygen mimic. HOD and QDO are structurally very similar. They possess a classical alpha/beta-hydrolase fold core domain additionally equipped with a cap domain. Organic substrates bind in a preorganized active site with an orientation ideally suited for selective deprotonation of their hydroxyl group by a His/Asp charge-relay system affording the generation of electron-donating species. The "oxyanion hole" of the alpha/beta-hydrolase fold, typically employed to stabilize the tetrahedral intermediate in ester hydrolysis reactions, is utilized here to host and control oxygen chemistry, which is proposed to involve a peroxide anion intermediate. Product release by proton back transfer from the catalytic histidine is driven by minimization of intramolecular charge repulsion. Structural and kinetic data suggest a nonnucleophilic general-base mechanism. Our analysis provides a framework to explain cofactor-independent dioxygenation within a protein architecture generally employed to catalyze hydrolytic reactions.
More Related Videos
 08:02
Benchtop Immobilized Metal Affinity Chromatography, Reconstitution and Assay of a Polyhistidine Tagged Metalloenzyme for the Undergraduate Laboratory
Published on: August 23, 2018
 12:08
Monitoring the Reductive and Oxidative Half-Reactions of a Flavin-Dependent Monooxygenase using Stopped-Flow Spectrophotometry
Published on: March 18, 2012
Related Concept Videos
 01:11
ortho–para-Directing Activators: –CH3, –OH, –NH2, –OCH3
 01:24
Cofactors and Coenzymes
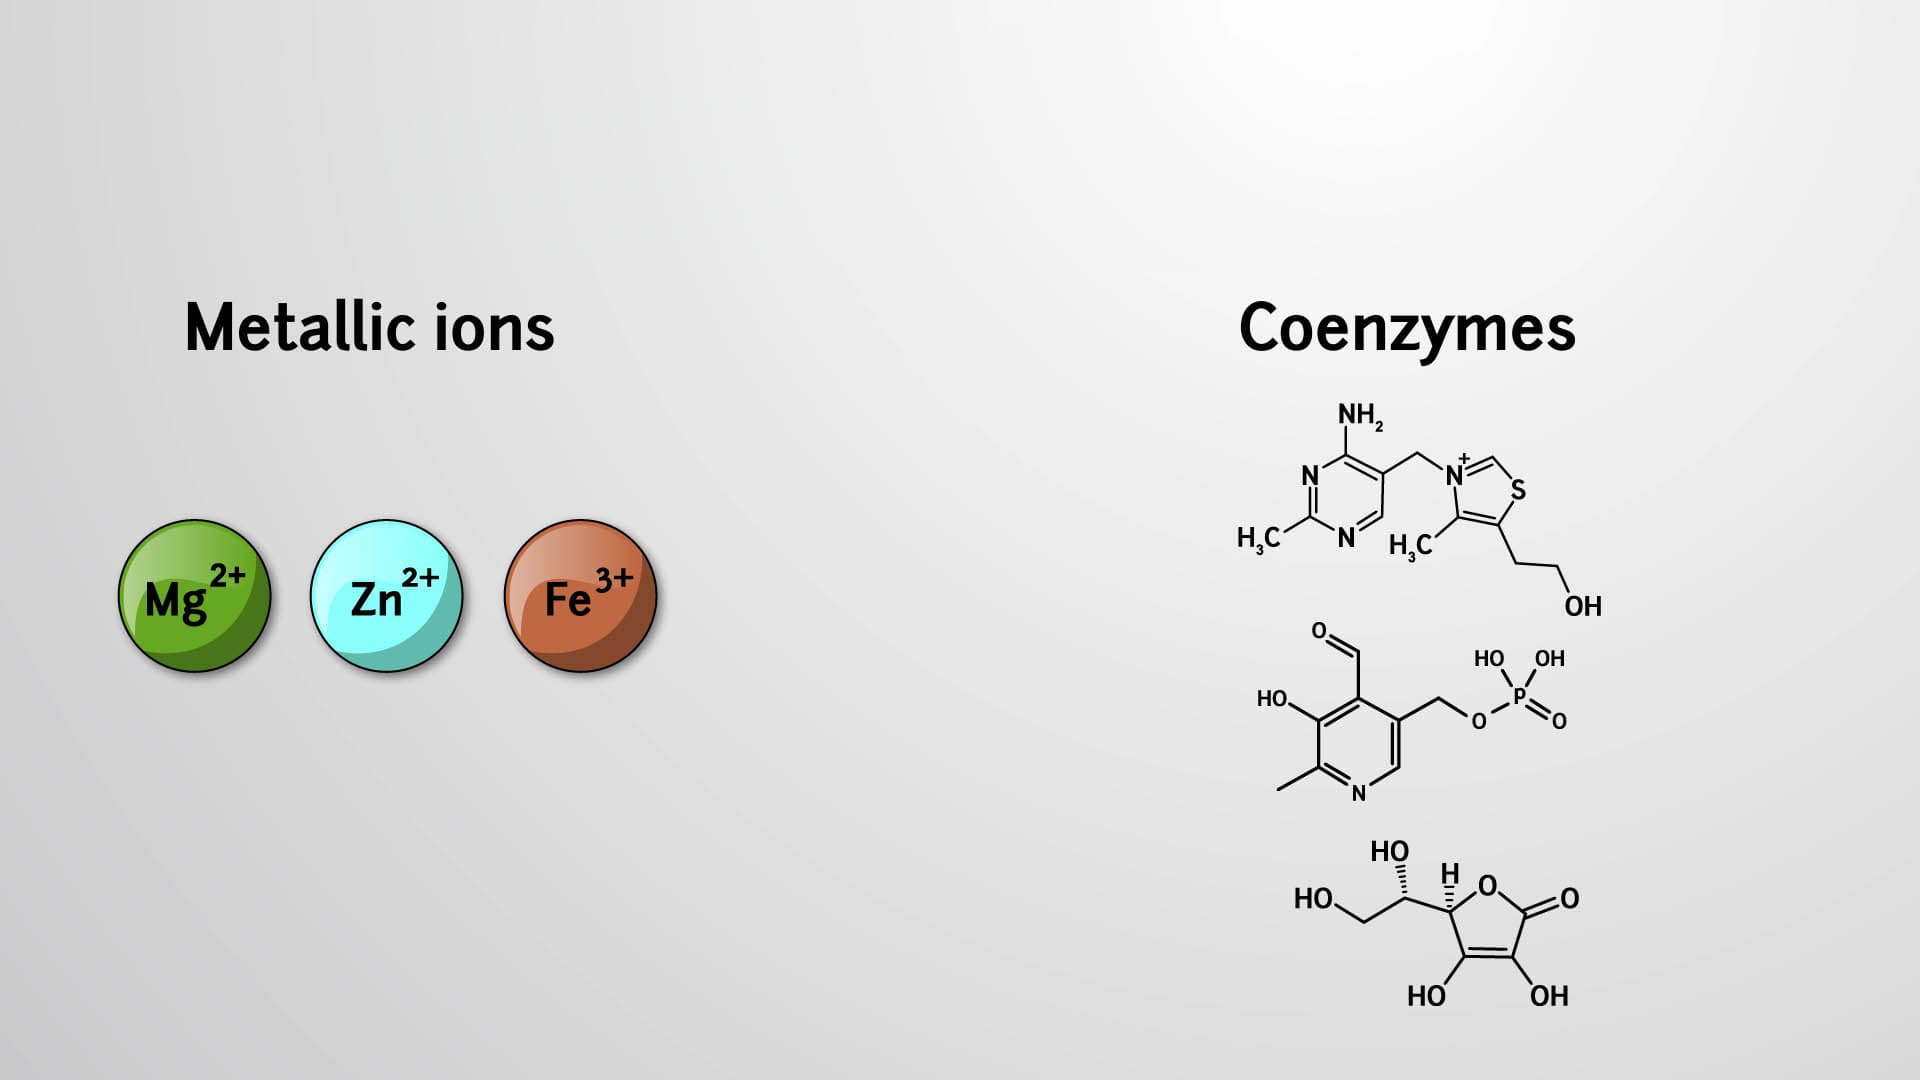
Cofactors can be metallic ions or organic molecules called coenzymes. These types of helper...
 01:27
Cofactors and Coenzymes
01:27Cofactors and Coenzymes
 01:18
Aromatic Hydrocarbon Anions: Structural Overview
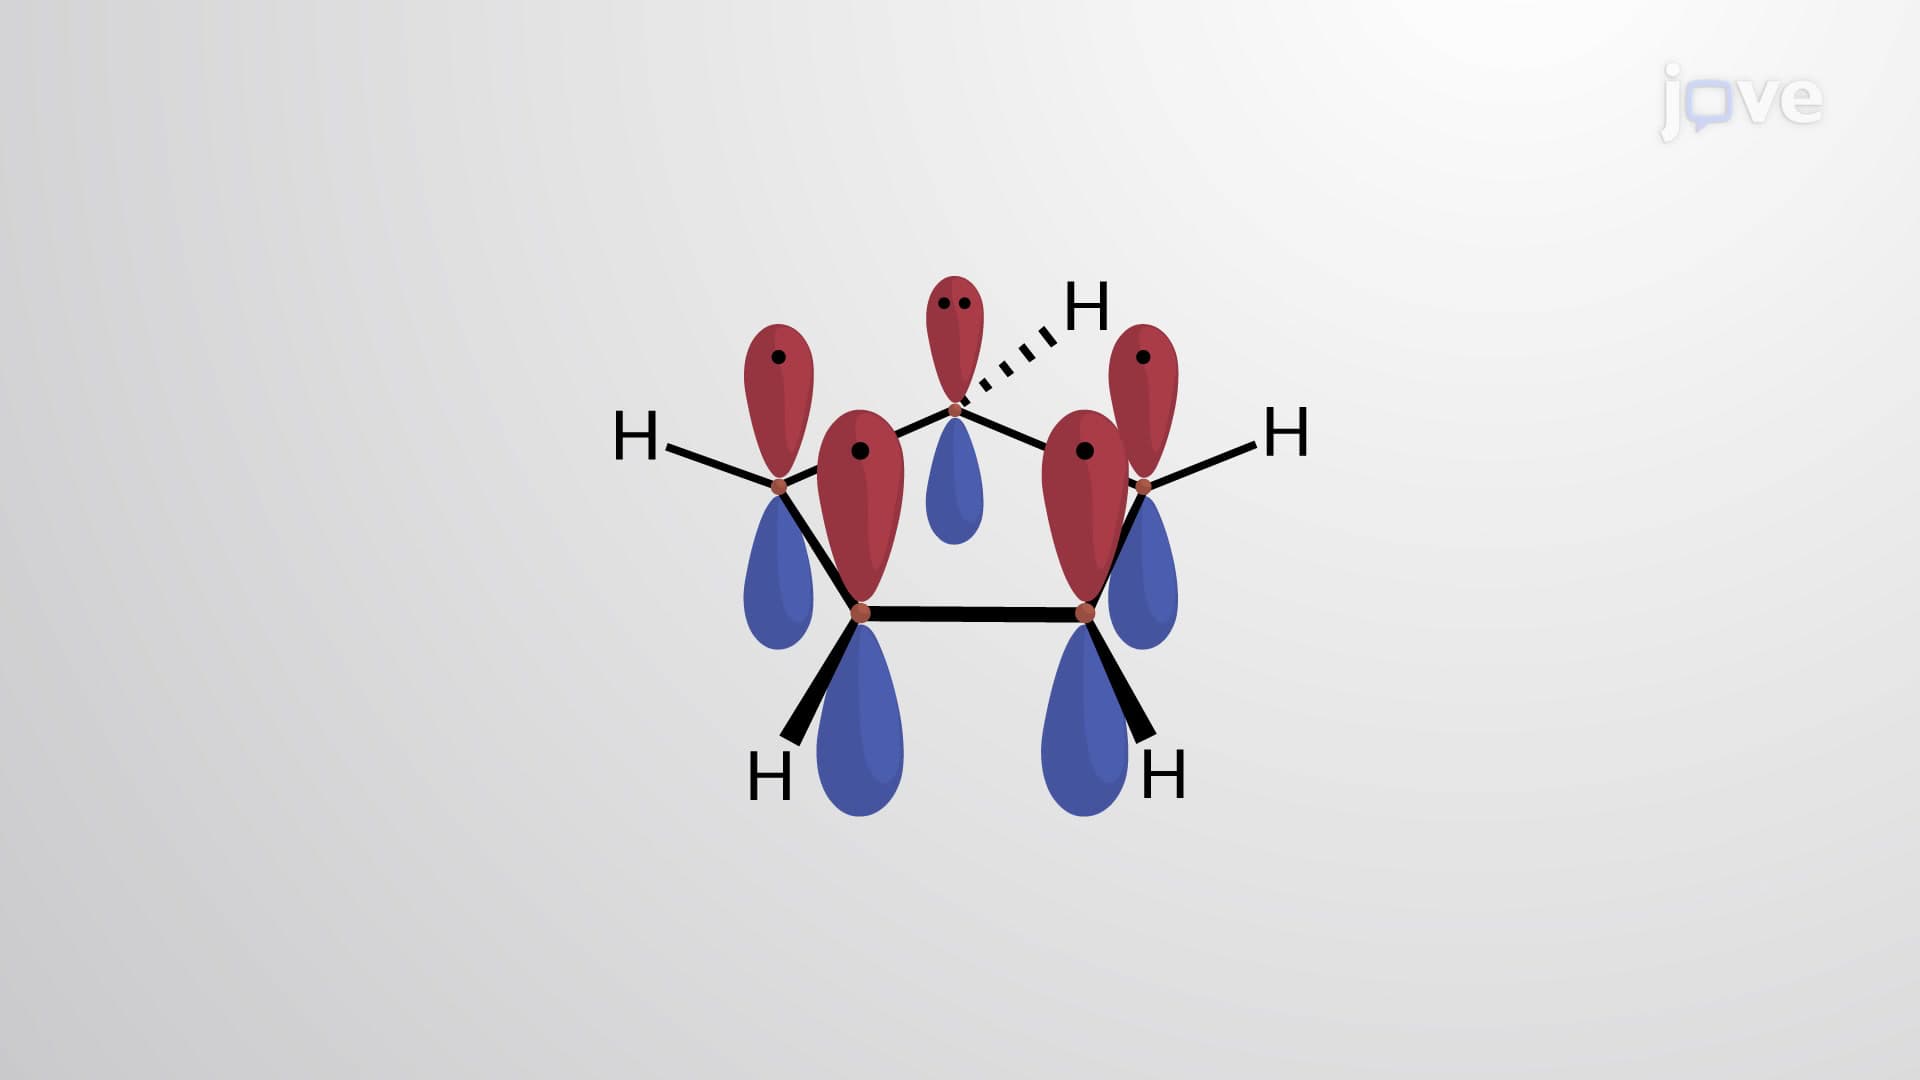
Due to the absence of continuous overlap of p...
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group with both...