Related Experiment Video
Updated: Jun 16, 2026

A High Throughput MHC II Binding Assay for Quantitative Analysis of Peptide Epitopes
Published on: March 25, 2014
Epitope discovery with phylogenetic hidden Markov models
Miguel Lacerda1, Konrad Scheffler, Cathal Seoighe
1School of Mathematics, Statistics and Applied Mathematics, National University of Ireland, Galway, Ireland.

Existing methods for the prediction of immunologically active T-cell epitopes are based on the amino acid sequence or structure of pathogen proteins. Additional information regarding the locations of epitopes may be acquired by considering the evolution of viruses in hosts with different immune backgrounds. In particular, immune-dependent evolutionary patterns at sites within or near T-cell epitopes can be used to enhance epitope identification. We have developed a mutation-selection model of T-cell epitope evolution that allows the human leukocyte antigen (HLA) genotype of the host to influence the evolutionary process. This is one of the first examples of the incorporation of environmental parameters into a phylogenetic model and has many other potential applications where the selection pressures exerted on an organism can be related directly to environmental factors. We combine this novel evolutionary model with a hidden Markov model to identify contiguous amino acid positions that appear to evolve under immune pressure in the presence of specific host immune alleles and that therefore represent potential epitopes. This phylogenetic hidden Markov model provides a rigorous probabilistic framework that can be combined with sequence or structural information to improve epitope prediction. As a demonstration, we apply the model to a data set of HIV-1 protein-coding sequences and host HLA genotypes.
Existing methods for the prediction of immunologically active T-cell epitopes are based on the amino acid sequence or structure of pathogen proteins. Additional information regarding the locations of epitopes may be acquired by considering the evolution of viruses in hosts with different immune backgrounds. In particular, immune-dependent evolutionary patterns at sites within or near T-cell epitopes can be used to enhance epitope identification. We have developed a mutation-selection model of T-cell epitope evolution that allows the human leukocyte antigen (HLA) genotype of the host to influence the evolutionary process. This is one of the first examples of the incorporation of environmental parameters into a phylogenetic model and has many other potential applications where the selection pressures exerted on an organism can be related directly to environmental factors. We combine this novel evolutionary model with a hidden Markov model to identify contiguous amino acid positions that appear to evolve under immune pressure in the presence of specific host immune alleles and that therefore represent potential epitopes. This phylogenetic hidden Markov model provides a rigorous probabilistic framework that can be combined with sequence or structural information to improve epitope prediction. As a demonstration, we apply the model to a data set of HIV-1 protein-coding sequences and host HLA genotypes.
Related Concept Videos
 01:24
Tagging and Fusion Proteins
 01:28
Microbial Phylogeny
 02:54
Evolutionary Relationships through Genome Comparisons
 01:33
Peptide Identification Using Tandem Mass Spectrometry
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
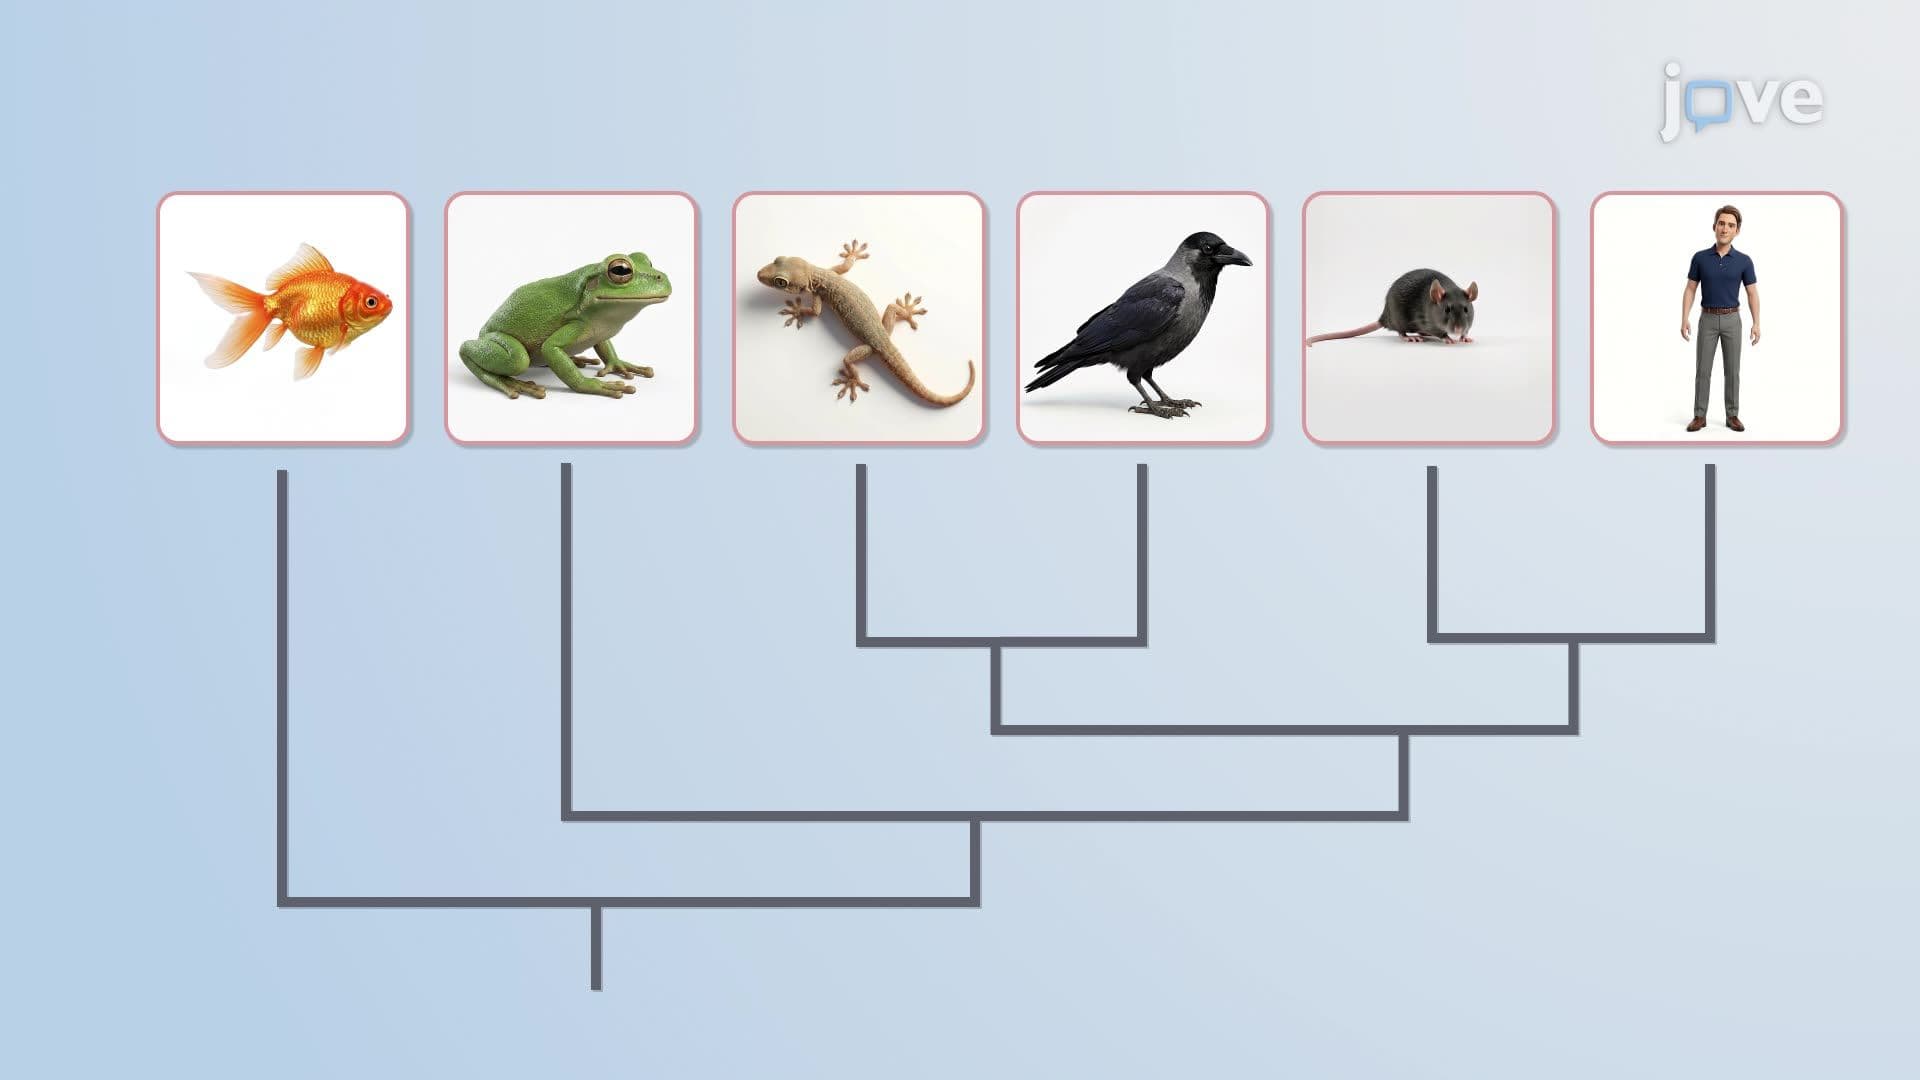 01:23
Phylogeny