

Structural and dynamical aspects of the unsymmetric hydration of Sb(III): an ab initio quantum mechanical charge
Len Herald V Lim1, Anirban Bhattacharjee, S Sikander Asam
1Theoretical Chemistry Division, Institute of General, Inorganic, and Theoretical Chemistry, University of Innsbruck, Innrain 52a, A-6020 Innsbruck, Austria.

Abstract:
An ab initio quantum mechanical charge field molecular dynamics (QMCF MD) simulation was performed to investigate the behavior of the Sb(3+) ion in aqueous solution. The simulation reveals a significant influence of the residual valence shell electron density on the solvation structure and dynamics of Sb(3+). A strong hemidirectional behavior of the ligand binding pattern is observed for the first hydration shell extending up to the second hydration layer. The apparent domain partitioned structural behavior was probed by solvent reorientational kinetics and three-body distribution functions. The three-dimensional hydration space was conveniently segmented such that domains having different properties were properly resolved. The approach afforded a fair isolation of localized solvent structural and dynamical motifs that Sb(3+) seems to induce to a remarkable degree. Most intriguing is the apparent impact of the lone pair electrons on the second hydration shell, which offers insight into the mechanistic aspects of hydrogen bonding networks in water. Such electronic effects observed in the hydration of Sb(3+) can only be studied by applying a suitable quantum mechanical treatment including first and second hydration shell as provided by the QMCF ansatz.


Related Concept Videos
 03:24
Hybridization of Atomic Orbitals I
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:18
Structure of Benzene: Molecular Orbital Model
 03:51
Molecular Orbital Theory II
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration
 01:18
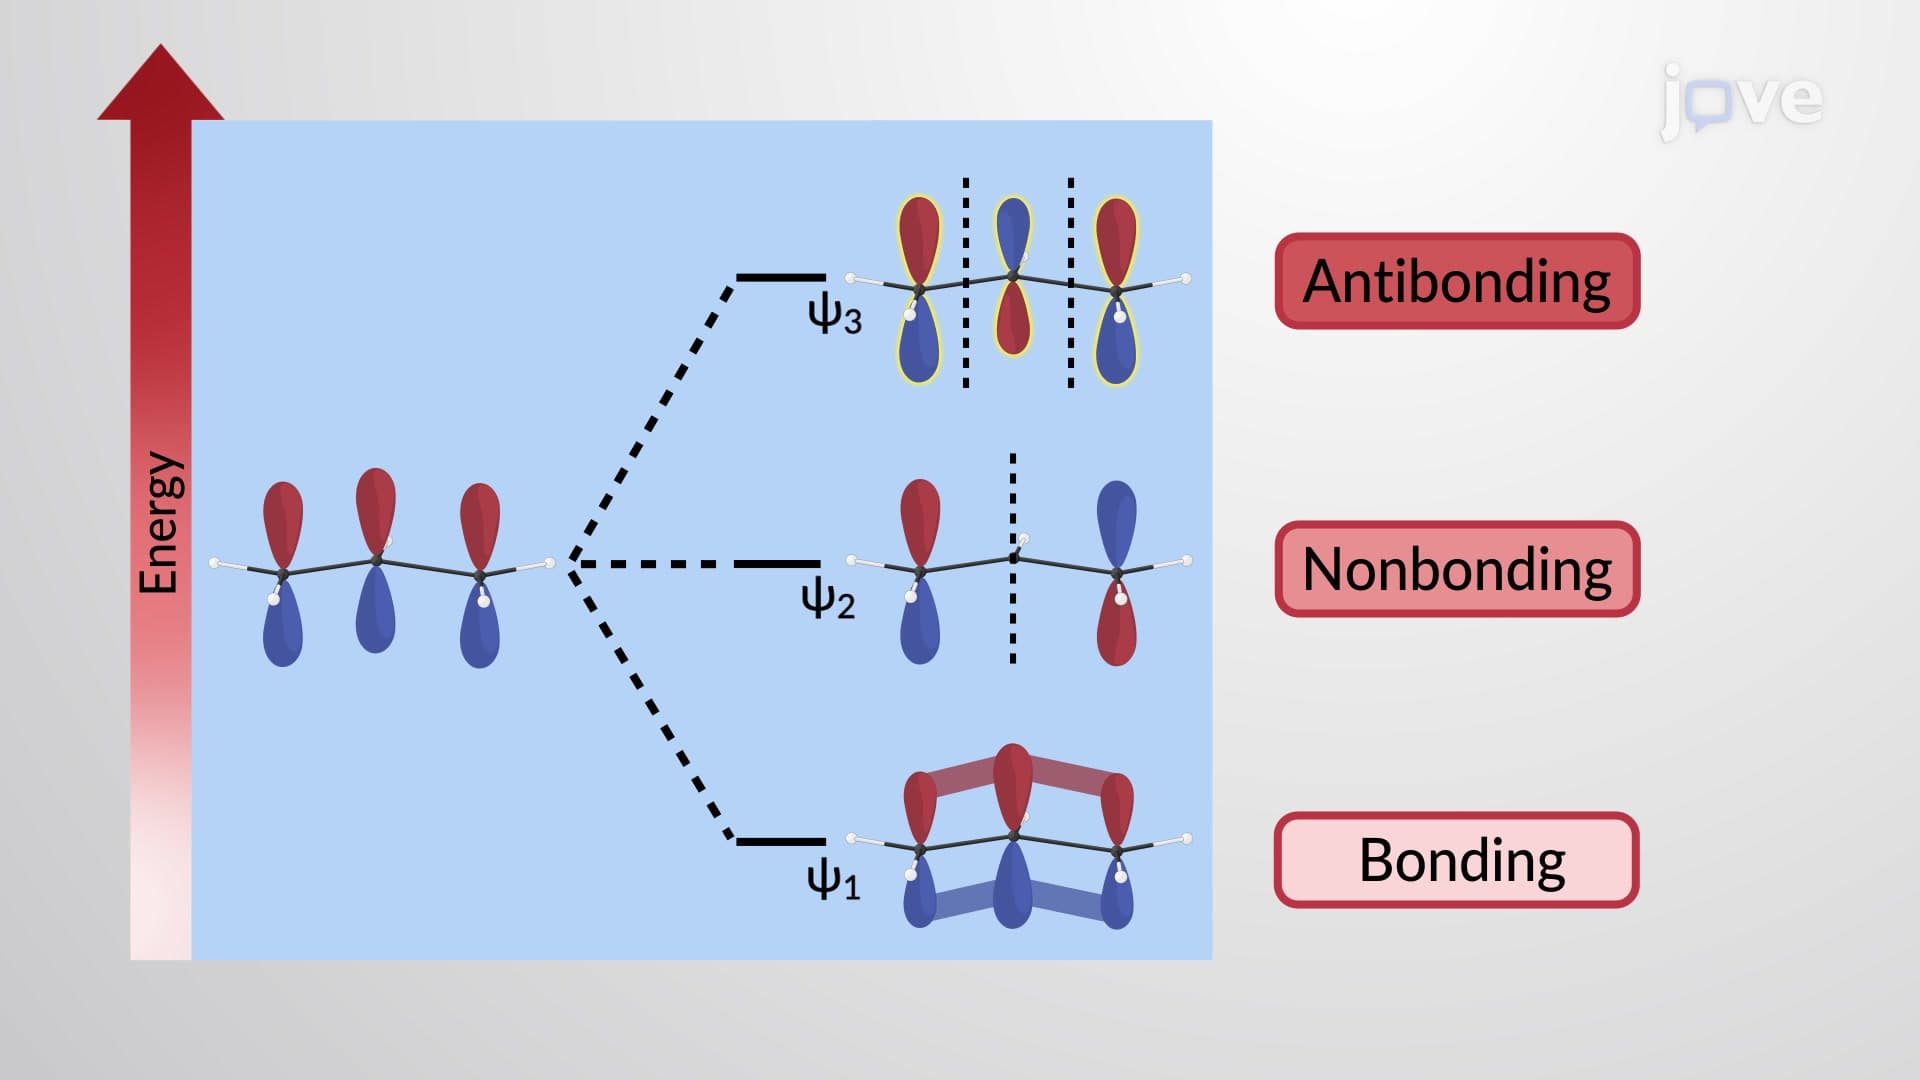
π Molecular Orbitals of the Allyl Cation and Anion