Related Experiment Video
Updated: Jun 15, 2026

A Kinetic Fluorescence-based Ca2+ Mobilization Assay to Identify G Protein-coupled Receptor Agonists, Antagonists, and Allosteric Modulators
Published on: February 20, 2018
A common intracellular allosteric binding site for antagonists of the CXCR2 receptor
K Salchow1, M E Bond, S C Evans
1Novartis Institutes for Biomedical Research, Horsham, West Sussex, UK.

Background And Purpose:
We have previously shown that SB265610 (1-(2-bromo-phenyl)-3-(7-cyano-3H-benzotriazol-4-yl)-urea) behaves as an allosteric, inverse agonist at the C-X-C chemokine (CXCR)2 receptor. The aim of this study was to determine whether SB265610, in addition to two other known antagonists, bind to either of the two putative, topographically distinct, allosteric binding sites previously reported in the Literature.
Experimental Approach:
Ten single point mutations were introduced into the CXCR2 receptor using site-directed mutagenesis. Three CXCR2 antagonists were investigated, SB265610, Pteridone-1 (2-(2,3 difluoro-benzylsulphanyl)-4-((R)-2-hydroxy-1-methyl-ethylamino)-8H-pteridin-7-one) and Sch527123 (2-hydroxy-N,N-dimethyl-3-{2-[[(R)-1-(5-methyl-furan-2-yl)-propyl]amino]-3,4-dioxo-cyclobut-1enylamino}-benzamide), and the effect of these mutations on their binding affinity and ability to inhibit interleukin-8-stimulated binding of [(35)S]GTPgammaS was examined.
Key Results:
Seven of the nine mutations introduced into the C-terminal domain and intracellular loops of the receptor produced a significant reduction in affinity at least one of the antagonists tested. Of those seven mutations, three produced a significant reduction in the affinity of all three antagonists, namely K320A, Y314A and D84N. In all but one mutation, the changes observed on antagonist affinity were matched with effects on inhibition of interleukin-8-stimulated [(35)S]GTPgammaS binding.
Conclusions And Implications:
These antagonists bind to a common intracellular, allosteric, binding site of the CXCR2 receptor, which has been further delineated. As many of these mutations are close to the site of G protein coupling or to a region of the receptor that is responsible for the transduction of the activation signal, our results suggest a molecular mechanism for the inhibition of receptor activation.
More Related Videos
 06:56
A Flow Cytometry-based Assay to Identify Compounds That Disrupt Binding of Fluorescently-labeled CXC Chemokine Ligand 12 to CXC Chemokine Receptor 4
Published on: March 10, 2018
 14:02
Optimizing the Genetic Incorporation of Chemical Probes into GPCRs for Photo-crosslinking Mapping and Bioorthogonal Chemistry in Live Mammalian Cells
Published on: April 9, 2018
Related Concept Videos
 01:09
GPCRs Regulate Adenylyl Cylase Activity
Two...
 01:28
Drug-Receptor Interaction: Antagonist
Antagonists can be classified as competitive or noncompetitive based on their...
 01:19
Allosteric Proteins-ATCase
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis pathway,...
 01:29
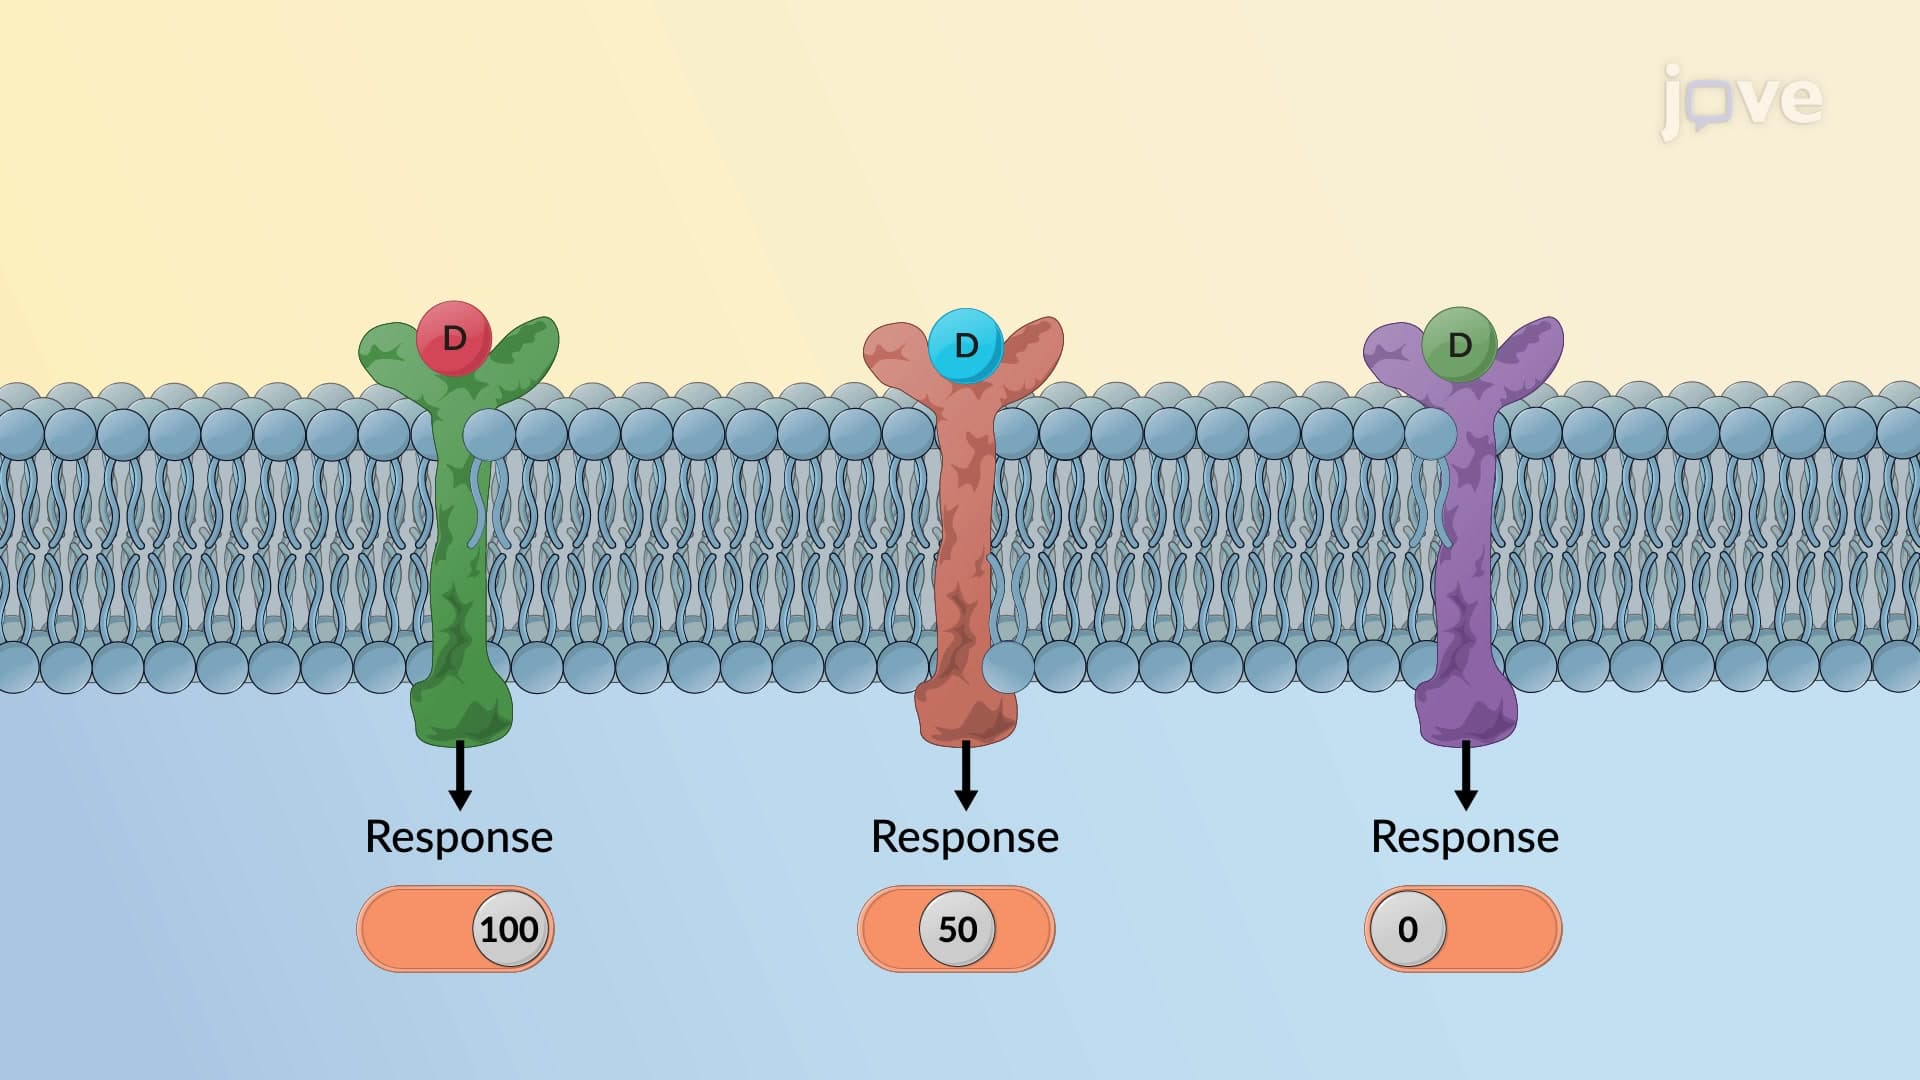
Drug-Receptor Interactions
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue.
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with one...
 00:49
Ligand Binding and Linkage