Related Experiment Video
Updated: Jun 14, 2026


Realistic Membrane Modeling Using Complex Lipid Mixtures in Simulation Studies
Published on: September 1, 2023
Density functional theory calculations of the molecular force field of L-ascorbic acid, vitamin C
Laura C Bichara1, Hernán E Lanús, Carlos G Nieto
1Cátedra de Fisicoquímica I, Instituto de Química Física, Facultad de Bioquímica, Química y Farmacia, Universidad Nacional de Tucumán, San Lorenzo 456, T4000CAN, S. M. de Tucumán, R, Argentina.

Abstract:
We have studied L-ascorbic acid and characterized it by infrared spectroscopy in solid and aqueous solution phases. The density functional theory (DFT) method together with Pople's basis set show that three stable molecules for the compound have been theoretically determined in the gas phase, and that an average of only two more stable conformations are present in the solid phase, as it was experimentally observed. The harmonic vibrational wavenumbers for the optimized geometries of both structures were calculated at B3LYP/6-31G*and B3LYP/6-311++G** levels at the proximity of the isolated molecule. For a complete assignment of the vibrational spectra in the compound solid and aqueous solution phases, DFT calculations were combined with Pulay's scaled quantum mechanics force field methodology in order to fit the theoretical wavenumber values to the experimental ones. In this way, a complete assignment of all the observed bands in the infrared spectrum for l-ascorbic acid was performed. The natural bond orbital study reveals the characteristics of the electronic delocalization of the three structures while the corresponding topological properties of electronic charge density are analyzed by employing Bader's atoms-in-molecules theory.
More Related Videos
 05:37
Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:27
The Debye–Hückel Theory of Electrolyte Solutions
 01:10
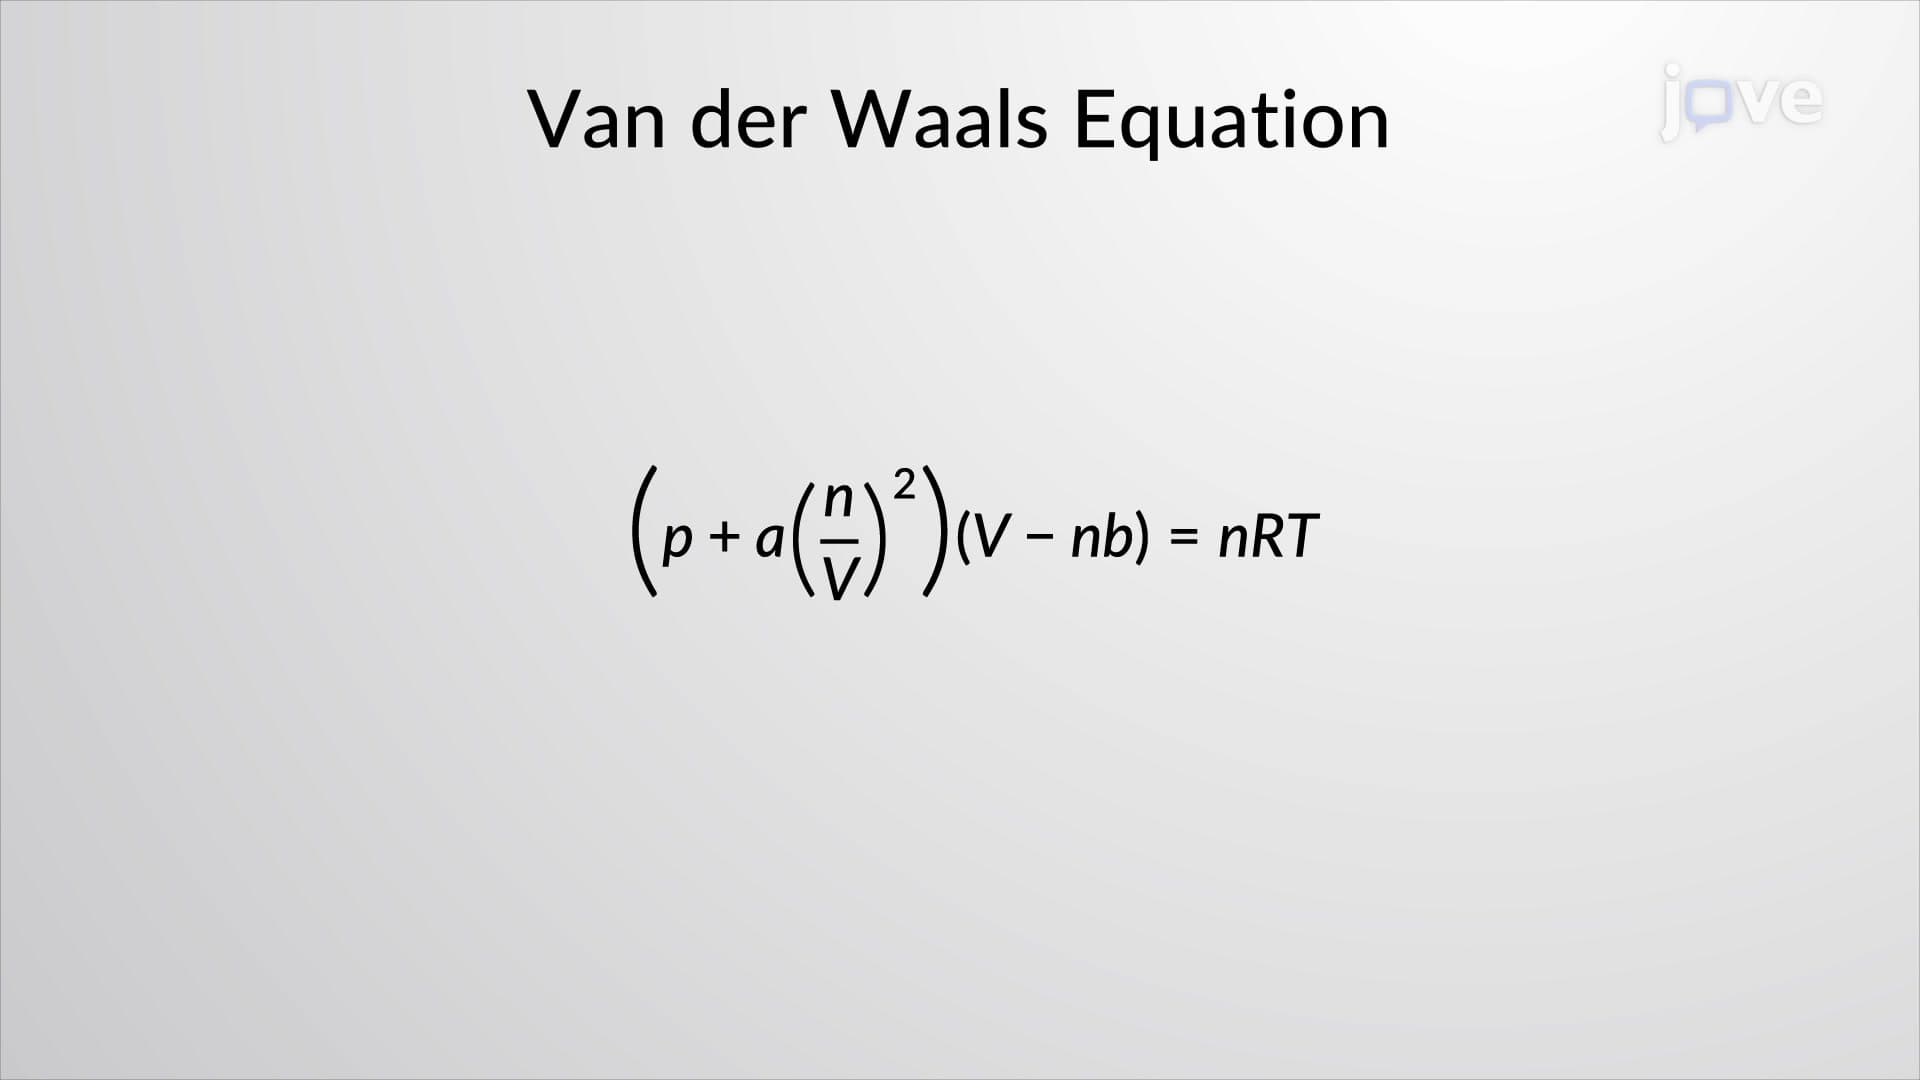
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:26
The Van der Waals Equation
 01:24
Thermodynamics: Activity Coefficient
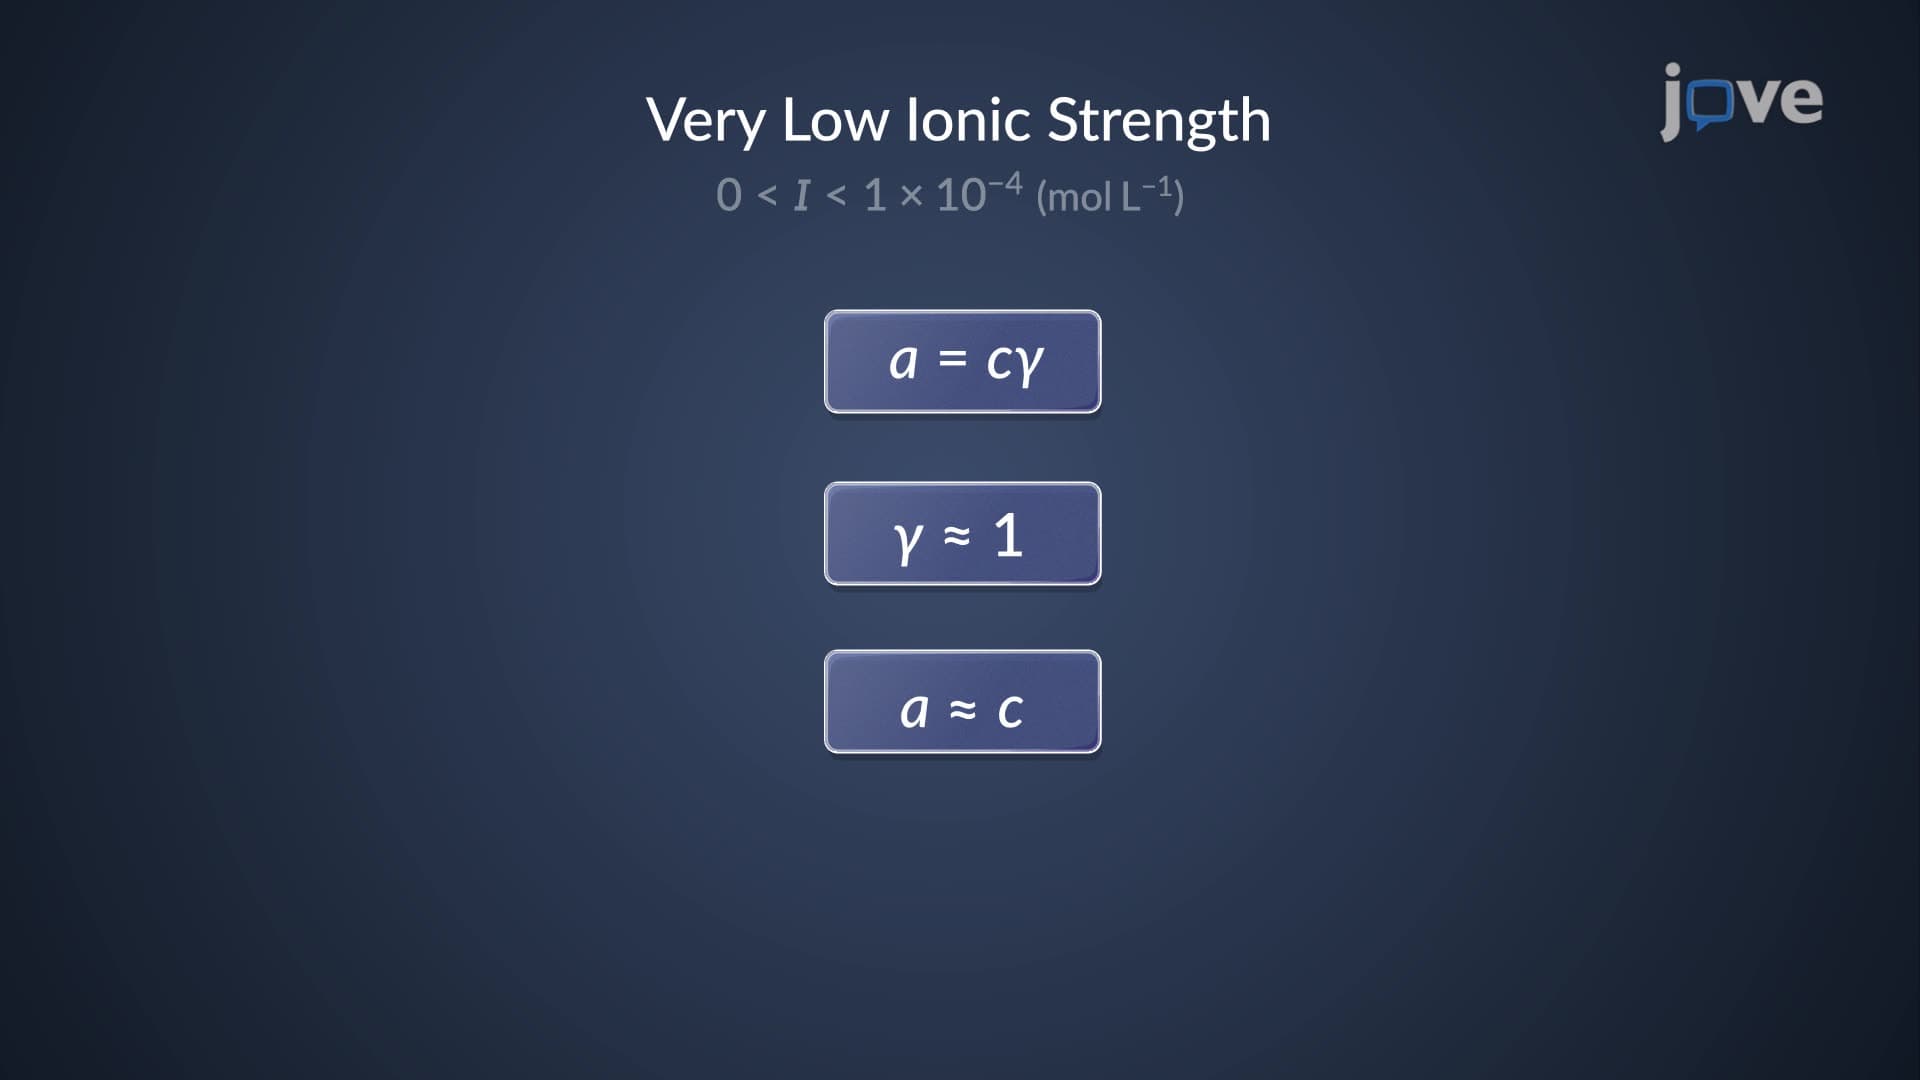
The activity coefficient is a measure of the deviation from ideal behavior. When the ionic strength of the solution is minimal, the activity coefficient of an ionic species is close to unity, making...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...