Related Experiment Video
Updated: Jun 14, 2026
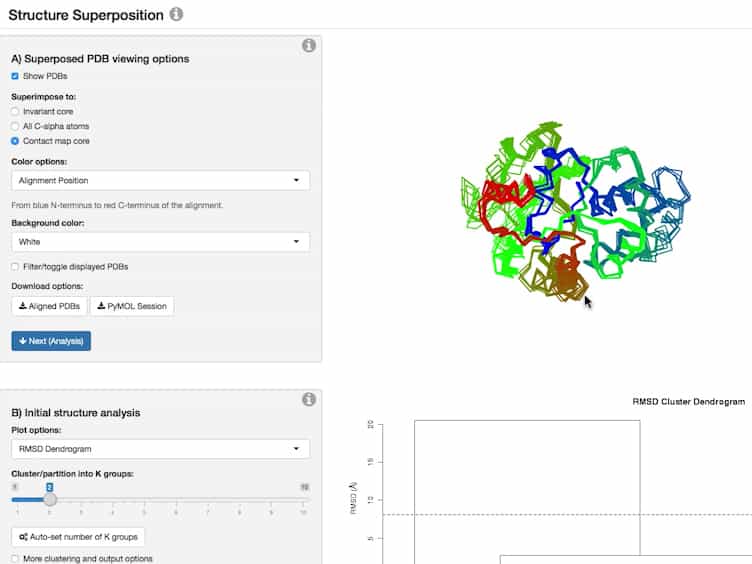

Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Metadynamics as a tool for mapping the conformational and free-energy space of peptides--the alanine dipeptide case
1Faculty of Science, Charles University in Prague, Albertov 2038/6, 128 00 Prague 2, Czech Republic.

Abstract:
There is a need for a fast, accurate, and reliable method of sampling the conformational space of peptides and proteins in order to obtain a balanced free-energy profile which can lead to our understanding of protein structure. We have utilized metadynamics for the conformational study of the solvated alanine dipeptide molecule, and our results show that the method has proven to be competent as a fast, robust, and reliable method for the conformation free-energy calculations of peptides in an explicit solvent, surpassing traditional methods such as umbrella sampling. We have also addressed the issue of the influence of different water models on the resulting free-energy profile in order to consistently decompose the setting of our simulation. All of the explicit water models for the simulation of biomolecules TIP3P, TIP4P, TIP4P/Ew, TIP5P, and SPCE have exhibited similar effects on the conformational preferences of alanine dipeptide with no significant differences. On the other hand, by comparing the potential energy surface in the gas phase and the free-energy surface in a water environment, we have shown that the interaction with water molecules is one of the most important structure-driving elements, with a great influence on the free-energy surface (FES) of the solvated peptide and the conformational preferences of the peptide backbone. All of the tested force fields (ff03, ff99SB, opls-aa, and charmm27) appreciably differ in the population of the individual conformers and the barriers between them. Significant divergence was found on both the potential energy surface (PES) as well as free-energy surface (FES) calculated by charmm27. We have therefore concluded that the differences originate dominantly from the parametrization of the peptide backbone in the given force field rather than from a noncovalent interaction with water molecules.
