Related Experiment Video
Updated: Jun 14, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Structure and energetics of azobenzene on Ag(111): benchmarking semiempirical dispersion correction approaches
G Mercurio1, E R McNellis, I Martin
1Institut für Bio- und Nanosysteme 3, Forschungszentrum Jülich, 52425 Jülich, Germany.

Abstract:
We employ normal-incidence x-ray standing wave and temperature programed desorption spectroscopy to derive the adsorption geometry and energetics of the prototypical molecular switch azobenzene at Ag(111). This allows us to assess the accuracy of semiempirical correction schemes as a computationally efficient means to overcome the deficiency of semilocal density-functional theory with respect to long-range van der Waals (vdW) interactions. The obtained agreement underscores the significant improvement provided by the account of vdW interactions, with remaining differences mainly attributed to the neglect of electronic screening at the metallic surface.


Related Concept Videos
 01:18
Structure of Benzene: Molecular Orbital Model
 03:51
Molecular Orbital Theory II
 01:07
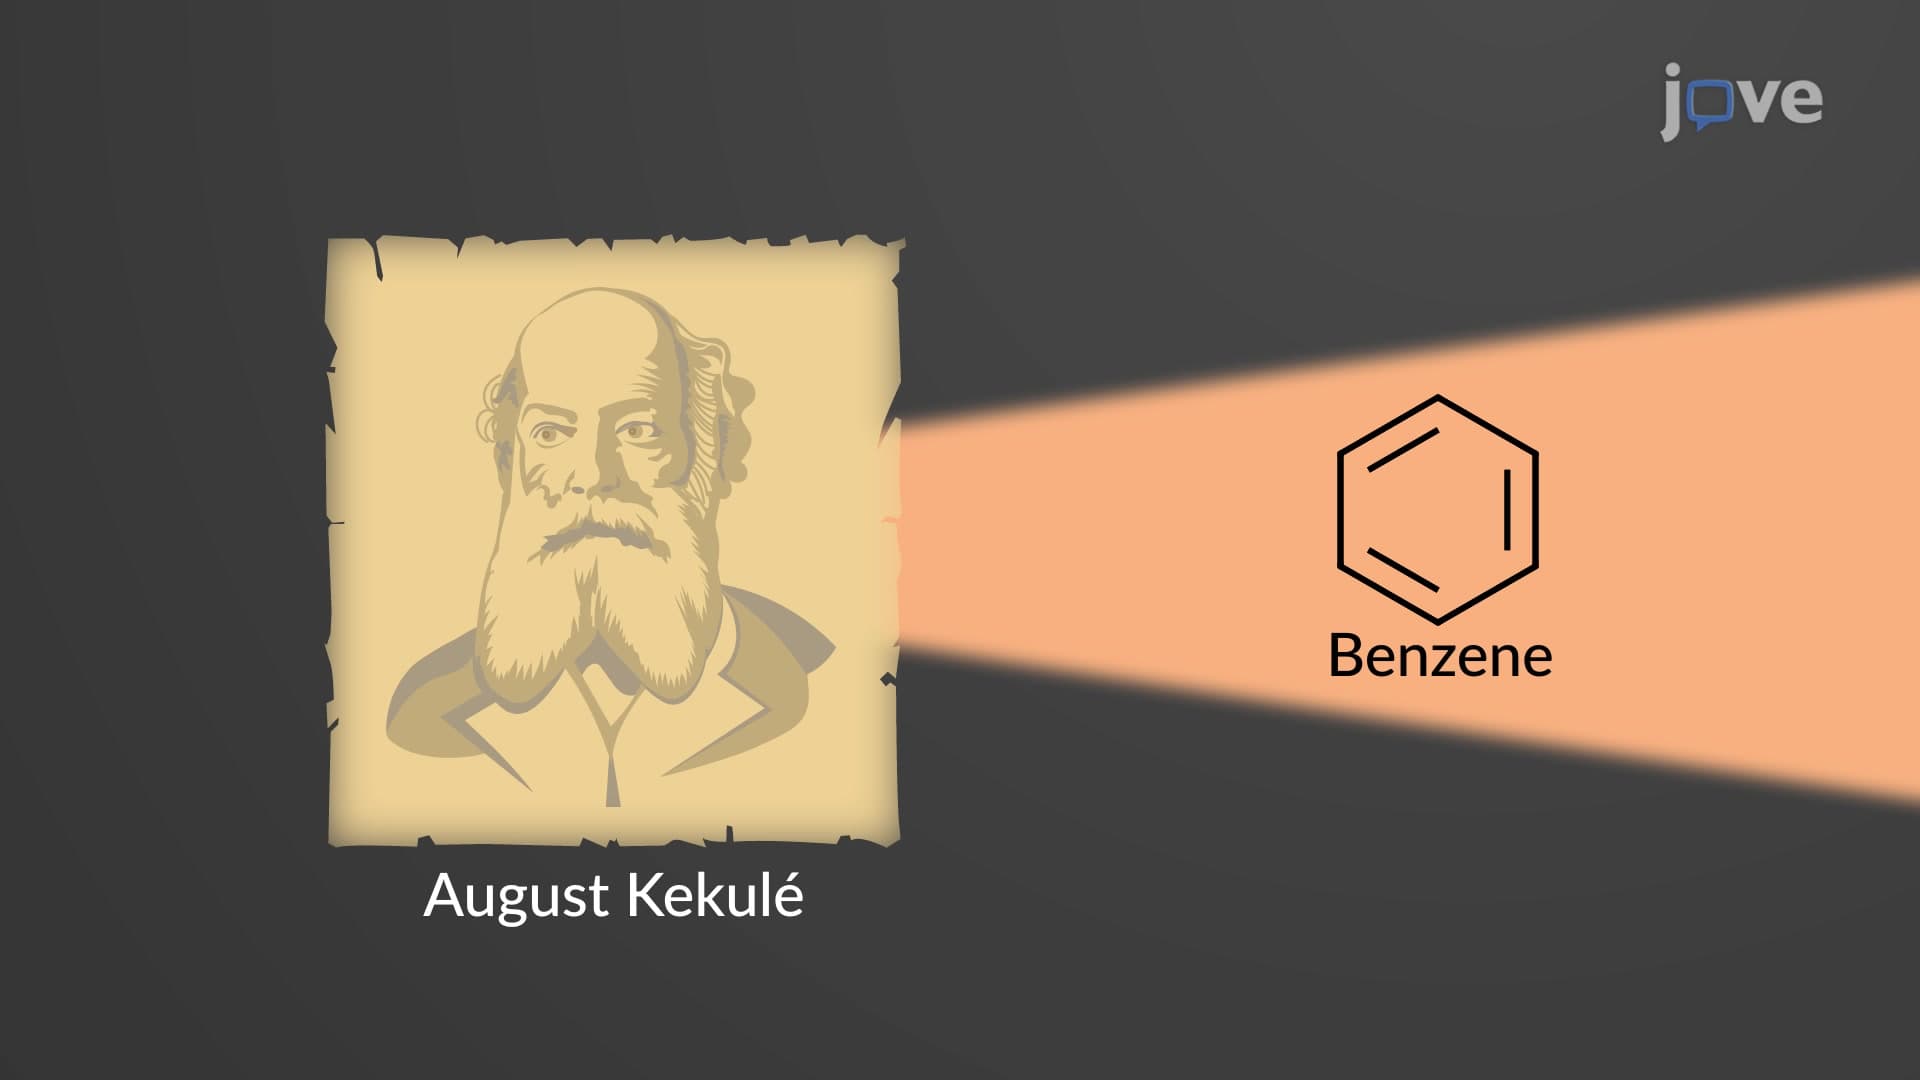
Structure of Benzene: Kekulé Model
He proposed that benzene has a cyclic structure of six carbon atoms attached to one hydrogen atom each, with three alternating pi bonds.
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 01:18
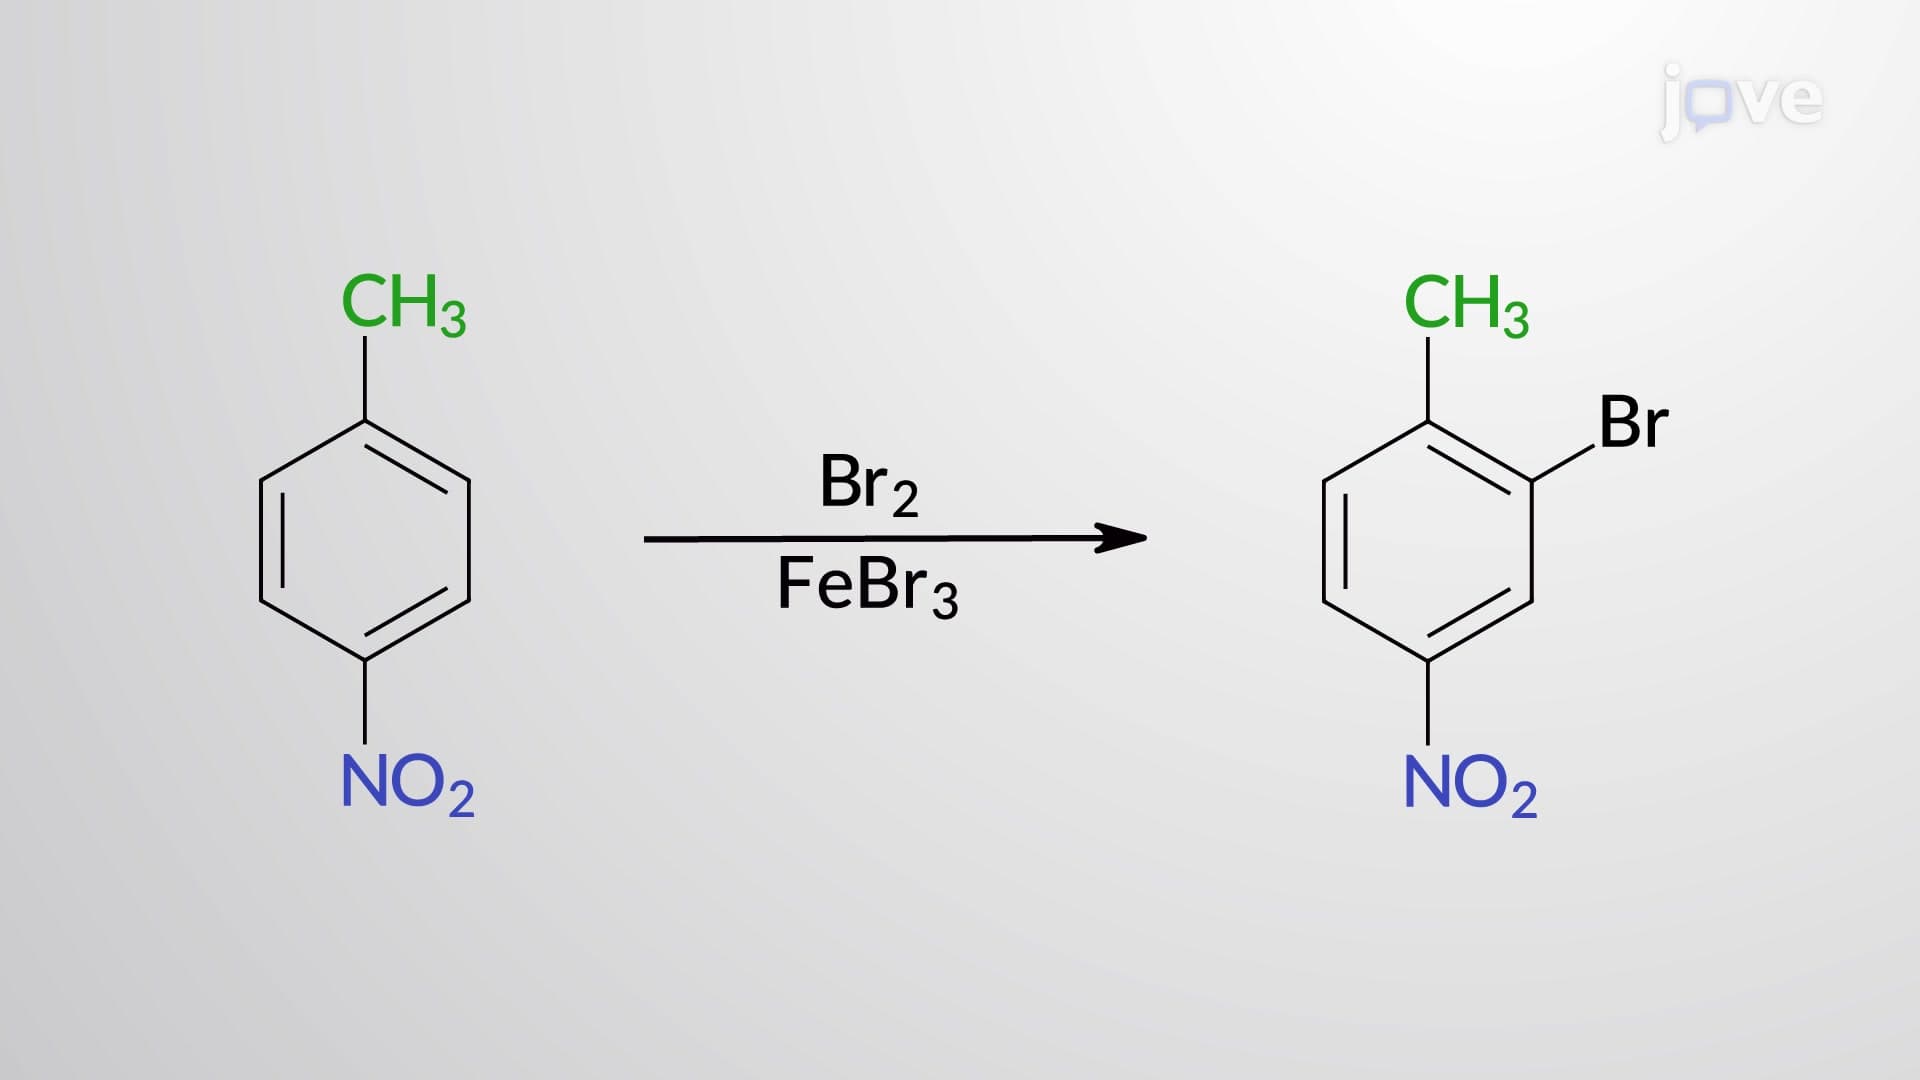
Directing and Steric Effects in Disubstituted Benzene Derivatives
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...