Related Experiment Video
Updated: Jun 14, 2026

Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
Phylogenetic assessment of alignments reveals neglected tree signal in gaps
Christophe Dessimoz1, Manuel Gil
1Department of Computer Science, ETH Zurich, Universitaetstr, 6, 8092 Zürich, Switzerland. cdessimoz@inf.ethz.ch

Background:
The alignment of biological sequences is of chief importance to most evolutionary and comparative genomics studies, yet the two main approaches used to assess alignment accuracy have flaws: reference alignments are derived from the biased sample of proteins with known structure, and simulated data lack realism.
Results:
Here, we introduce tree-based tests of alignment accuracy, which not only use large and representative samples of real biological data, but also enable the evaluation of the effect of gap placement on phylogenetic inference. We show that (i) the current belief that consistency-based alignments outperform scoring matrix-based alignments is misguided; (ii) gaps carry substantial phylogenetic signal, but are poorly exploited by most alignment and tree building programs; (iii) even so, excluding gaps and variable regions is detrimental; (iv) disagreement among alignment programs says little about the accuracy of resulting trees.
Conclusions:
This study provides the broad community relying on sequence alignment with important practical recommendations, sets superior standards for assessing alignment accuracy, and paves the way for the development of phylogenetic inference methods of significantly higher resolution.
Related Concept Videos
 03:21
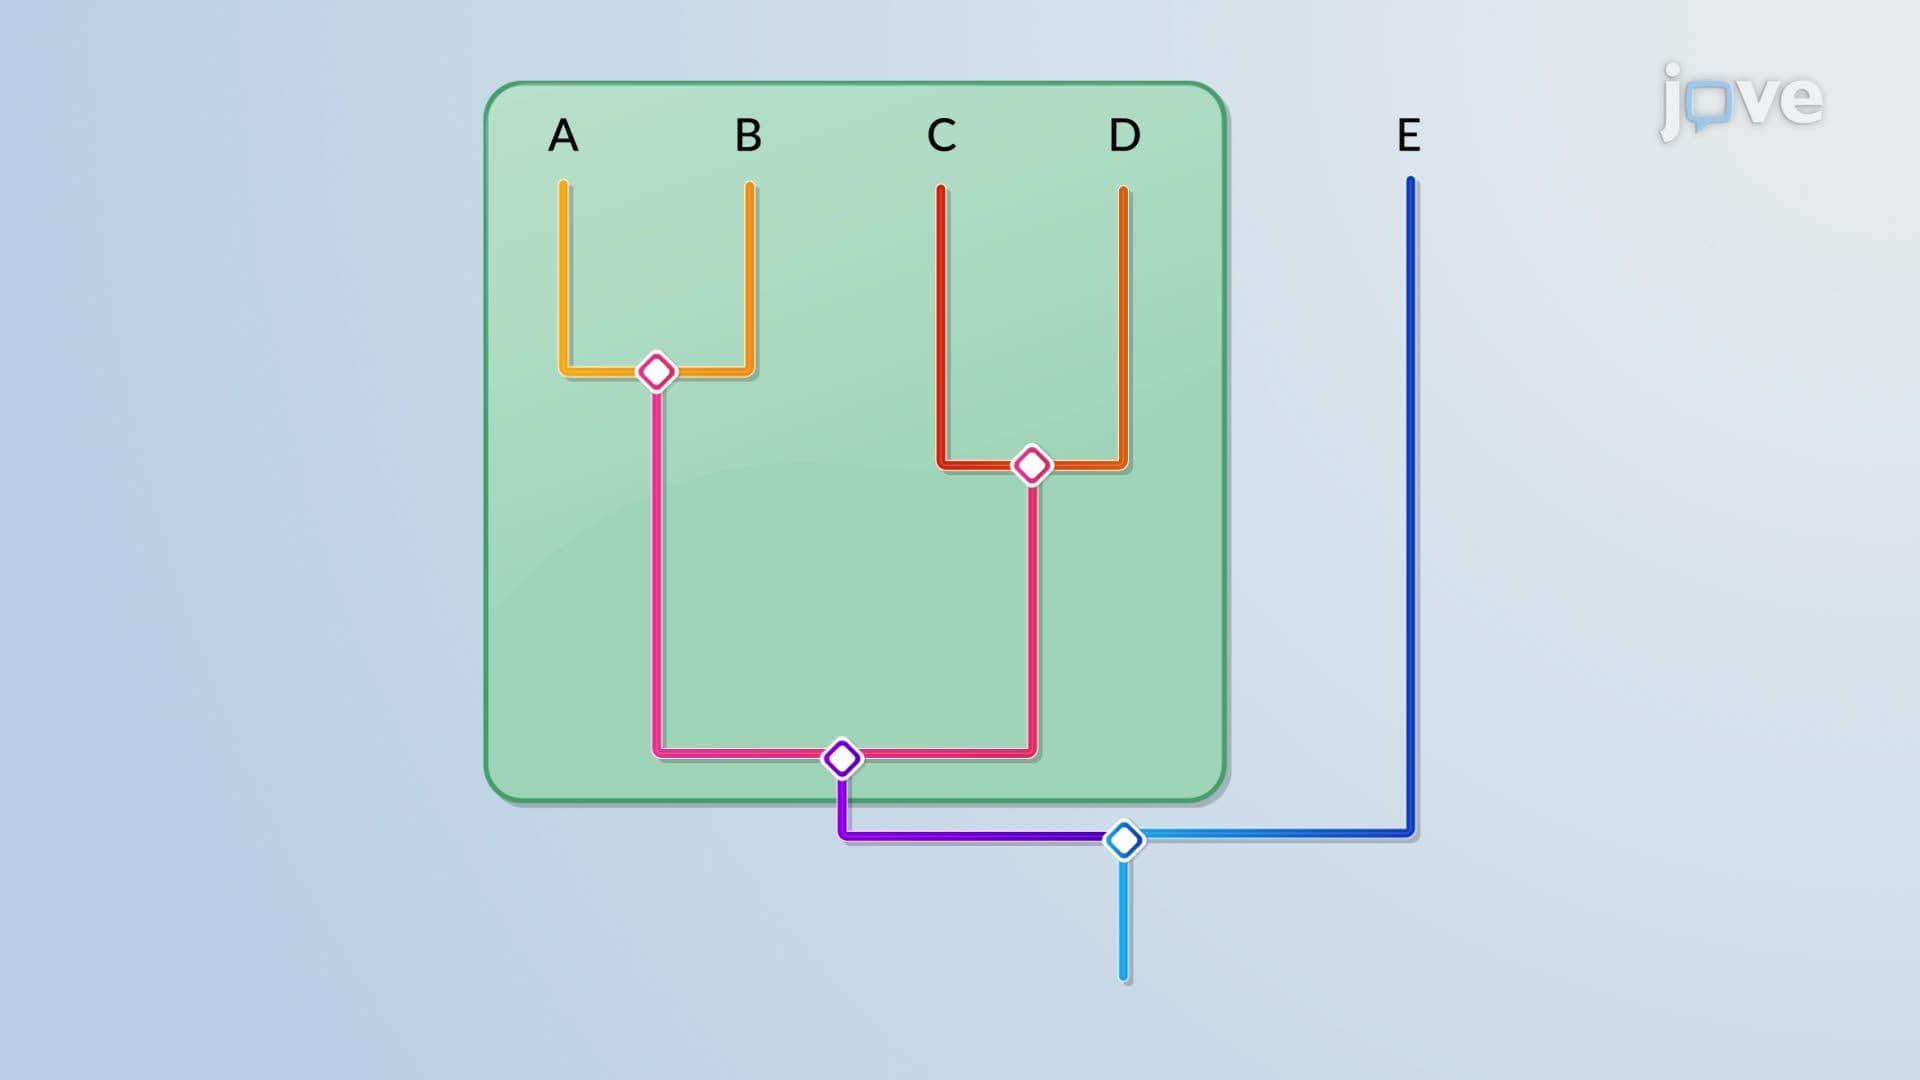
Phylogenetic Trees
03:21Phylogenetic Trees
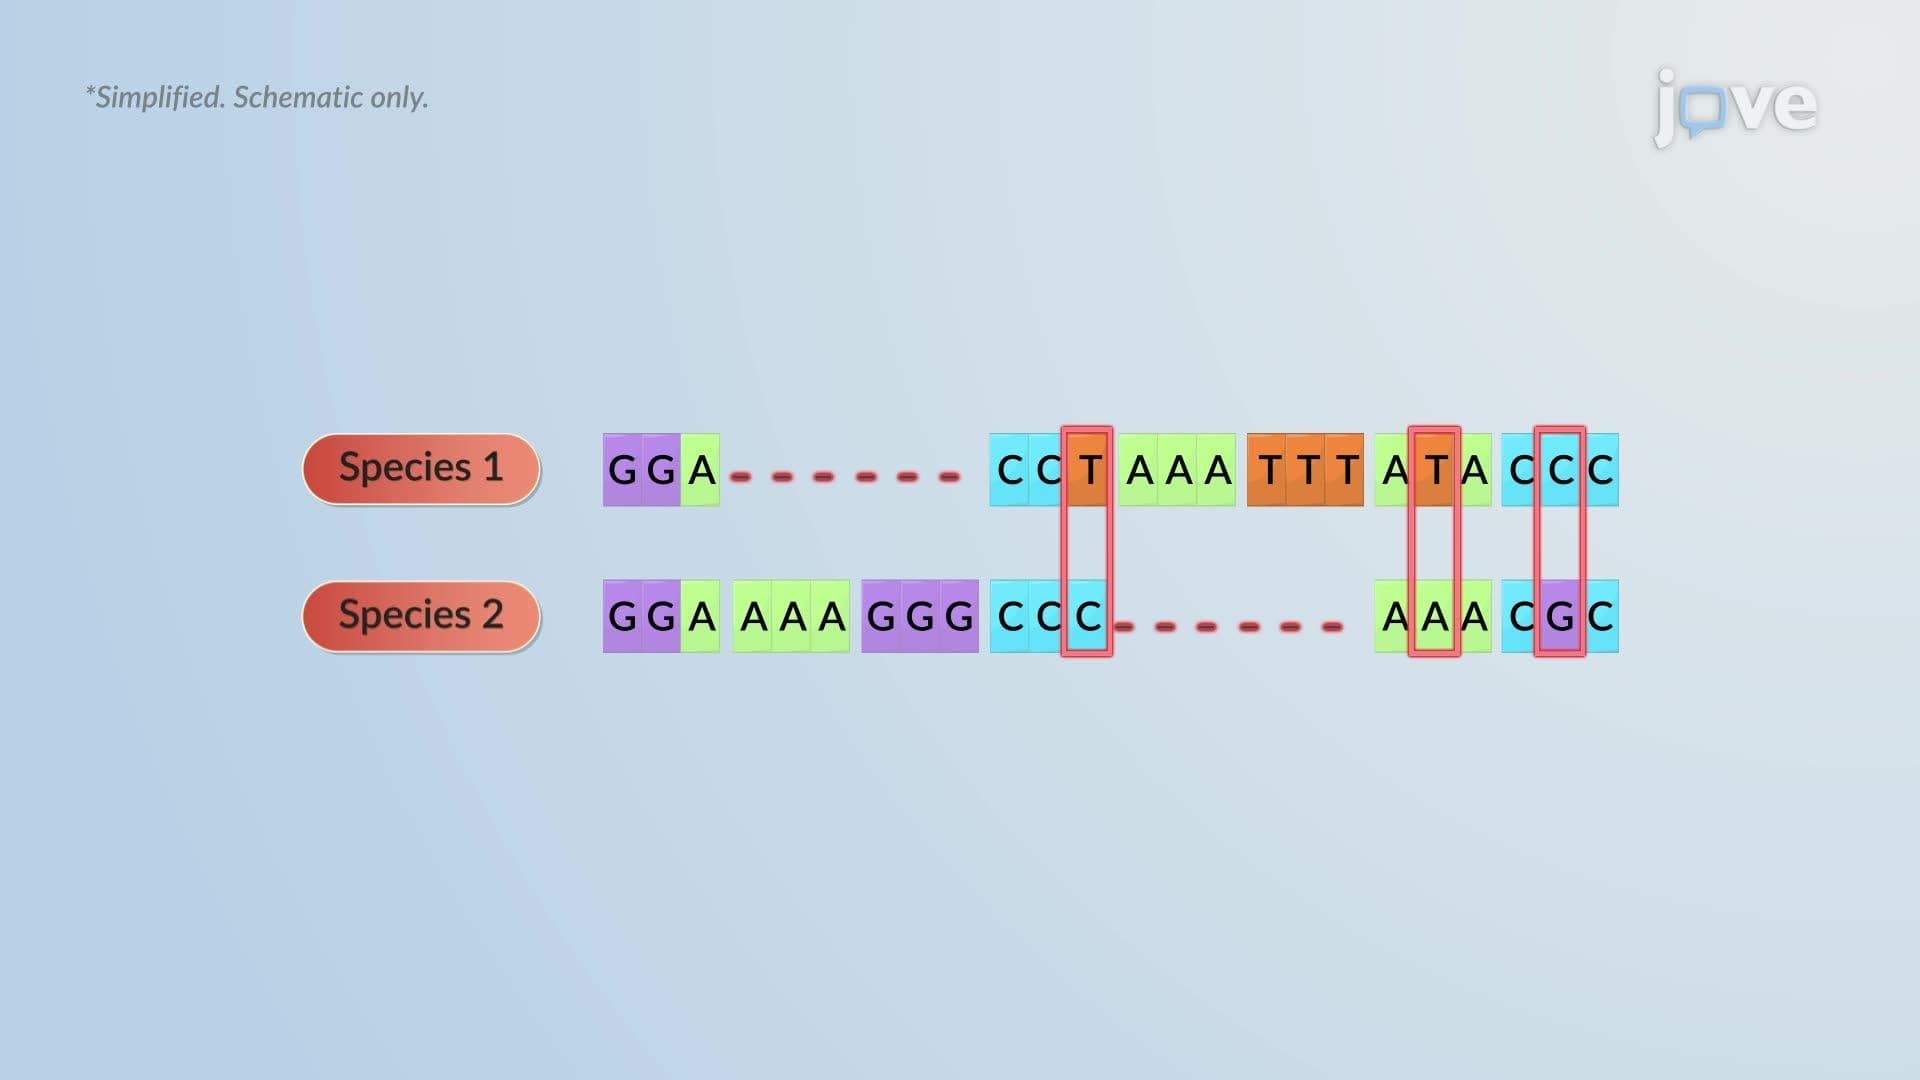 01:28
Microbial Phylogeny
 02:54
Evolutionary Relationships through Genome Comparisons
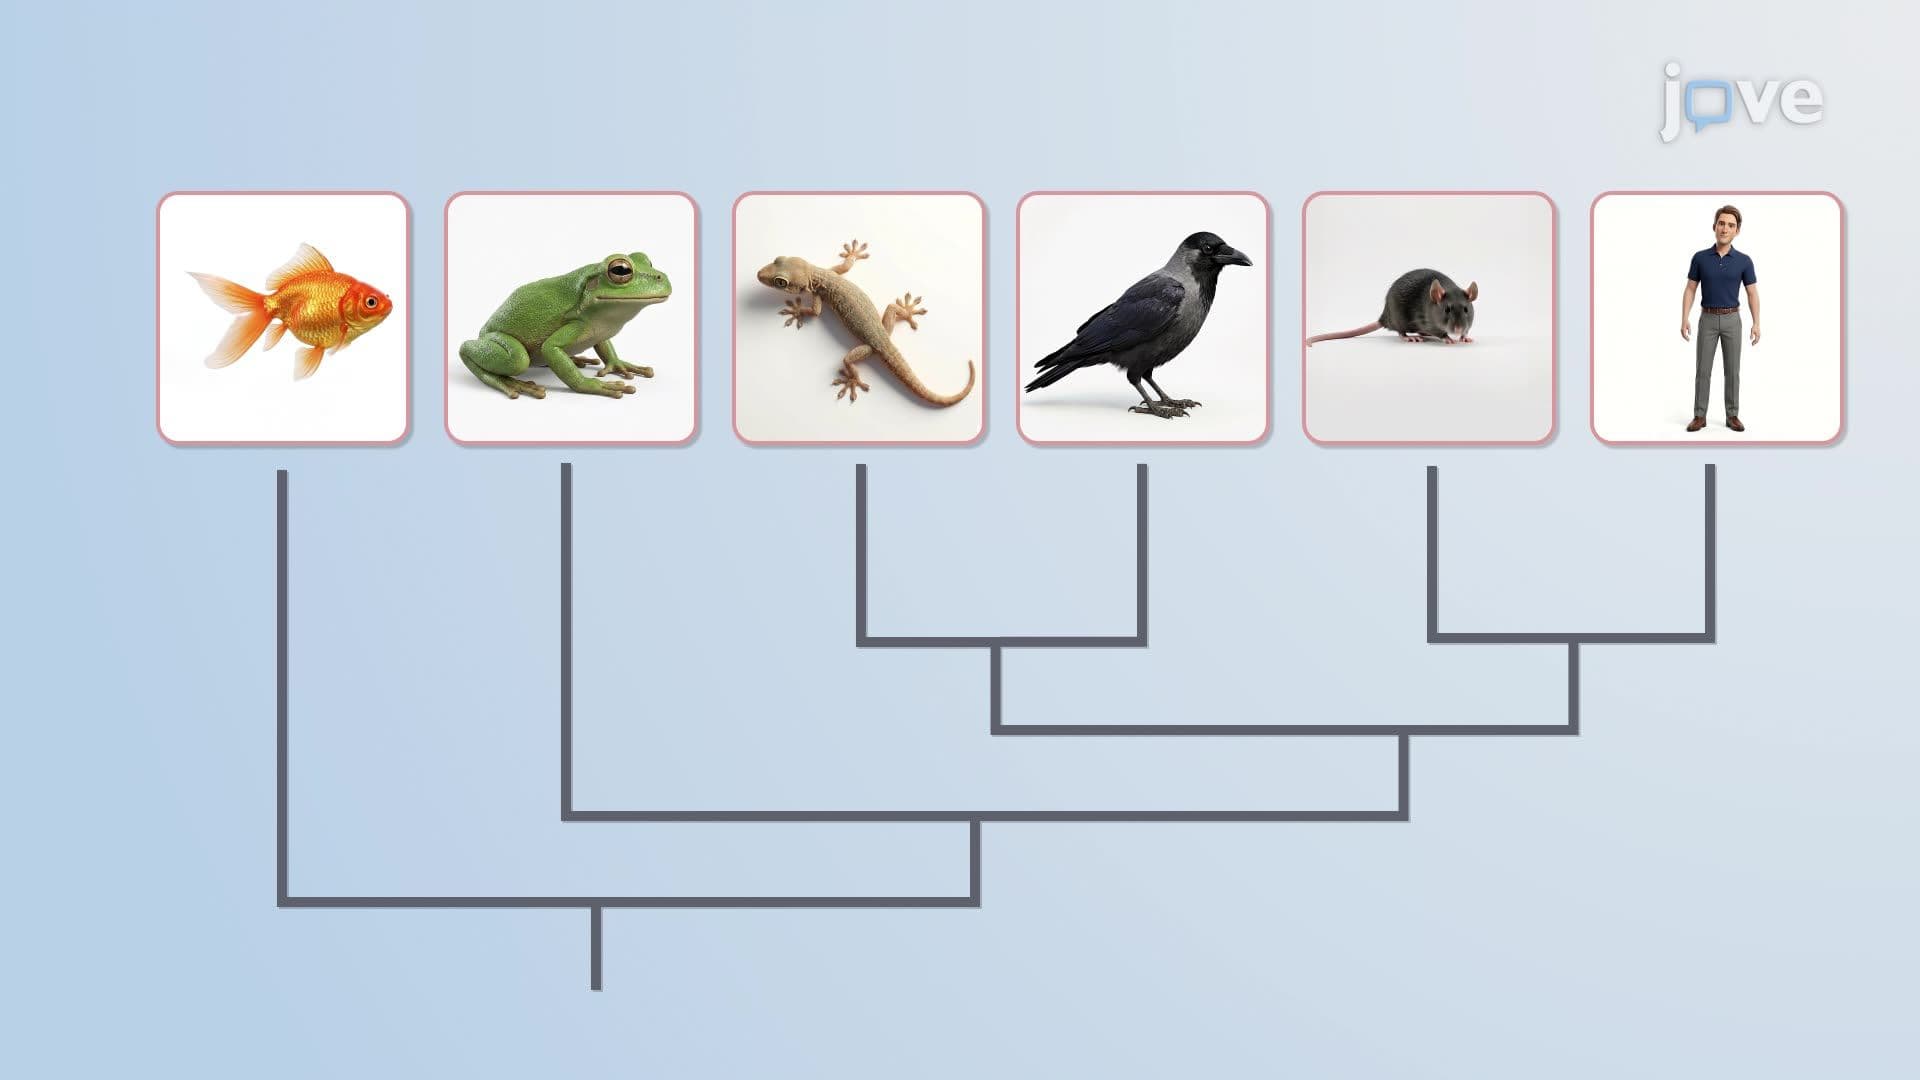 01:23
Phylogeny
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...