Related Experiment Video
Updated: Jun 13, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Accurate ensemble molecular dynamics binding free energy ranking of multidrug-resistant HIV-1 proteases
S Kashif Sadiq1, David W Wright, Owain A Kenway
1Centre for Computational Science, Department of Chemistry, University College London, London, WC1H 0AJ, United Kingdom.

Abstract:
Accurate calculation of important thermodynamic properties, such as macromolecular binding free energies, is one of the principal goals of molecular dynamics simulations. However, single long simulation frequently produces incorrectly converged quantitative results due to inadequate sampling of conformational space in a feasible wall-clock time. Multiple short (ensemble) simulations have been shown to explore conformational space more effectively than single long simulations, but the two methods have not yet been thermodynamically compared. Here we show that, for end-state binding free energy determination methods, ensemble simulations exhibit significantly enhanced thermodynamic sampling over single long simulations and result in accurate and converged relative binding free energies that are reproducible to within 0.5 kcal/mol. Completely correct ranking is obtained for six HIV-1 protease variants bound to lopinavir with a correlation coefficient of 0.89 and a mean relative deviation from experiment of 0.9 kcal/mol. Multidrug resistance to lopinavir is enthalpically driven and increases through a decrease in the protein-ligand van der Waals interaction, principally due to the V82A/I84V mutation, and an increase in net electrostatic repulsion due to water-mediated disruption of protein-ligand interactions in the catalytic region. Furthermore, we correctly rank, to within 1 kcal/mol of experiment, the substantially increased chemical potency of lopinavir binding to the wild-type protease compared to saquinavir and show that lopinavir takes advantage of a decreased net electrostatic repulsion to confer enhanced binding. Our approach is dependent on the combined use of petascale computing resources and on an automated simulation workflow to attain the required level of sampling and turn around time to obtain the results, which can be as little as three days. This level of performance promotes integration of such methodology with clinical decision support systems for the optimization of patient-specific therapy.

Related Concept Videos
 01:16
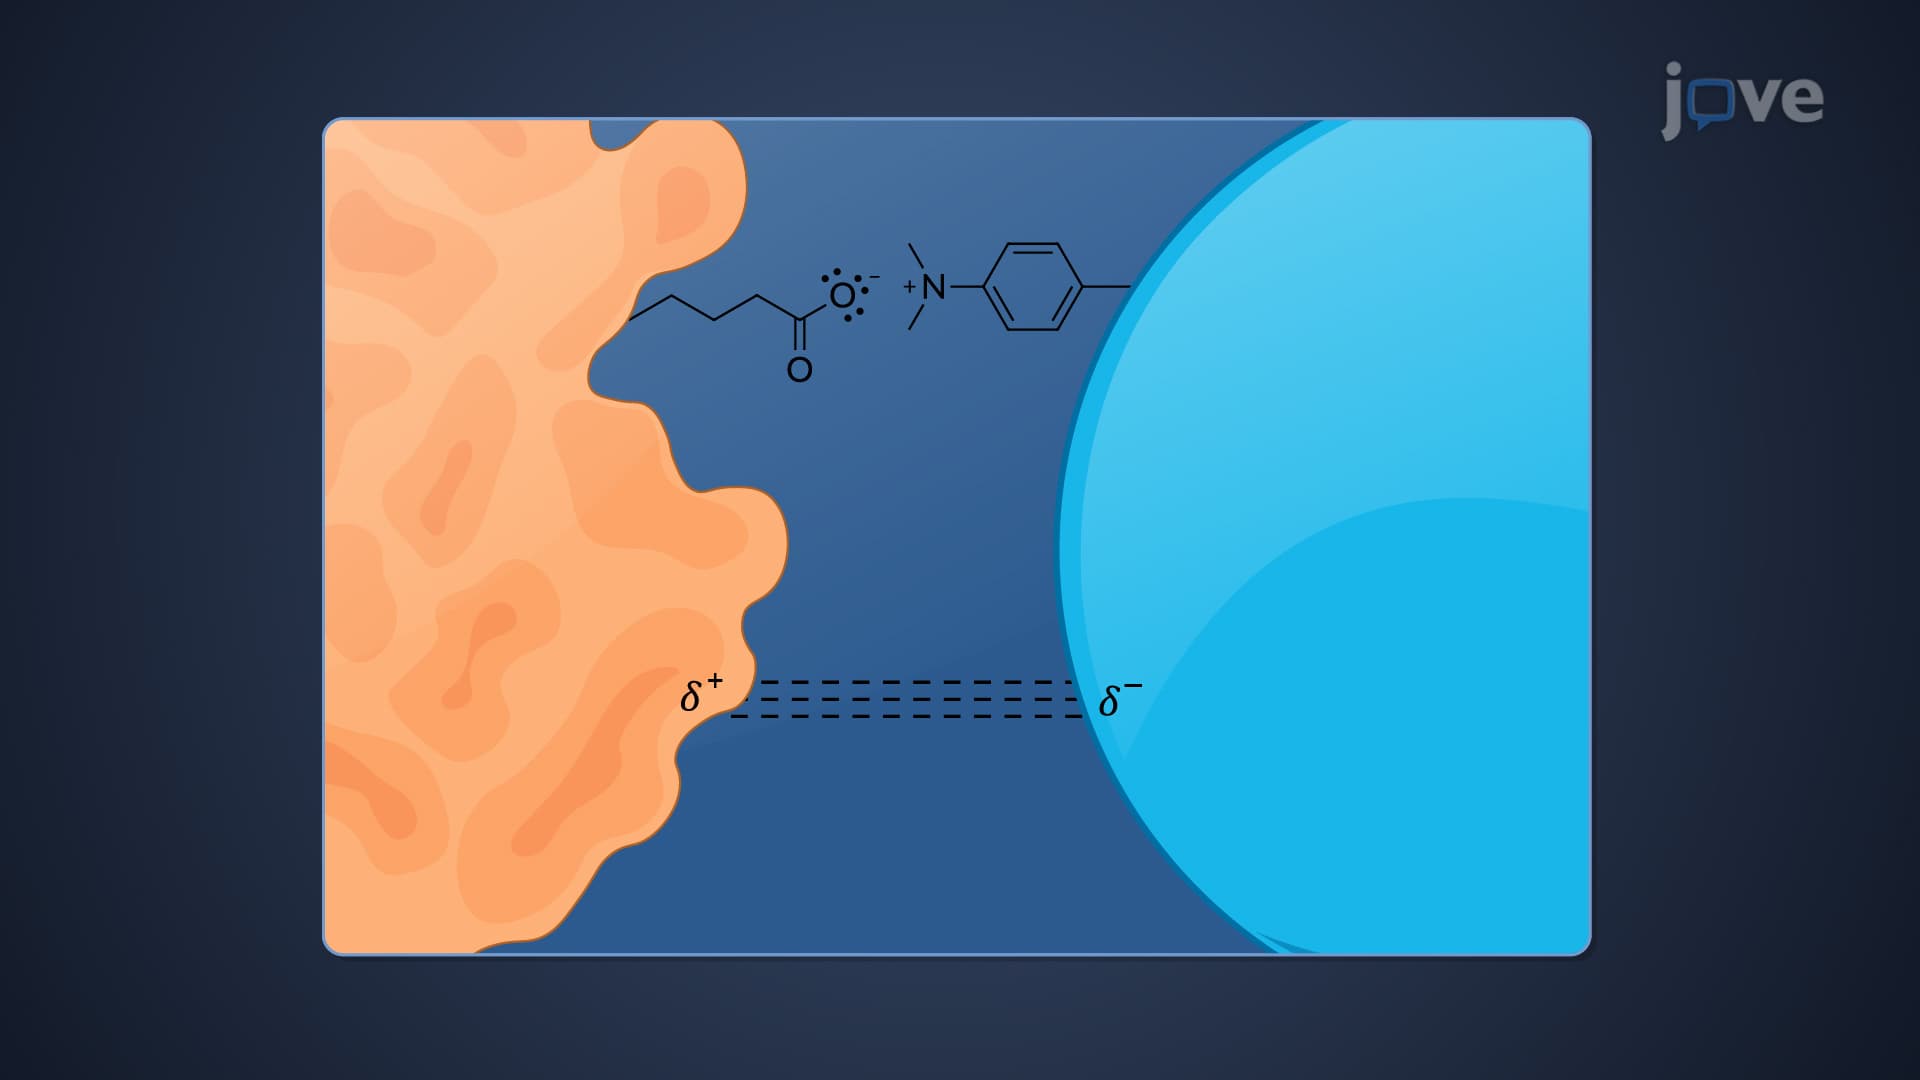
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...