Related Experiment Video
Updated: Jun 13, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Assessment of the performance of common density functional methods for describing the interaction energies of (H2O)6
F-F Wang1, G Jenness, W A Al-Saidi
1Department of Chemistry and Center for Molecular and Materials Simulations, University of Pittsburgh, Pittsburgh, Pennsylvania 15260, USA.

Abstract:
Localized molecular orbital energy decomposition analysis and symmetry-adapted perturbation theory (SAPT) calculations are used to analyze the two- and three-body interaction energies of four low-energy isomers of (H(2)O)(6) in order to gain insight into the performance of several popular density functionals for describing the electrostatic, exchange-repulsion, induction, and short-range dispersion interactions between water molecules. The energy decomposition analyses indicate that all density functionals considered significantly overestimate the contributions of charge transfer to the interaction energies. Moreover, in contrast to some studies that state that density functional theory (DFT) does not include dispersion interactions, we adopt a broader definition and conclude that for (H(2)O)(6) the short-range dispersion interactions recovered in the DFT calculations account about 75% or more of the net (short-range plus long-range) dispersion energies obtained from the SAPT calculations.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 10:28
Probing the Structure and Dynamics of Interfacial Water with Scanning Tunneling Microscopy and Spectroscopy
Published on: May 27, 2018
Related Concept Videos
 03:51
Molecular Orbital Theory II
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:24
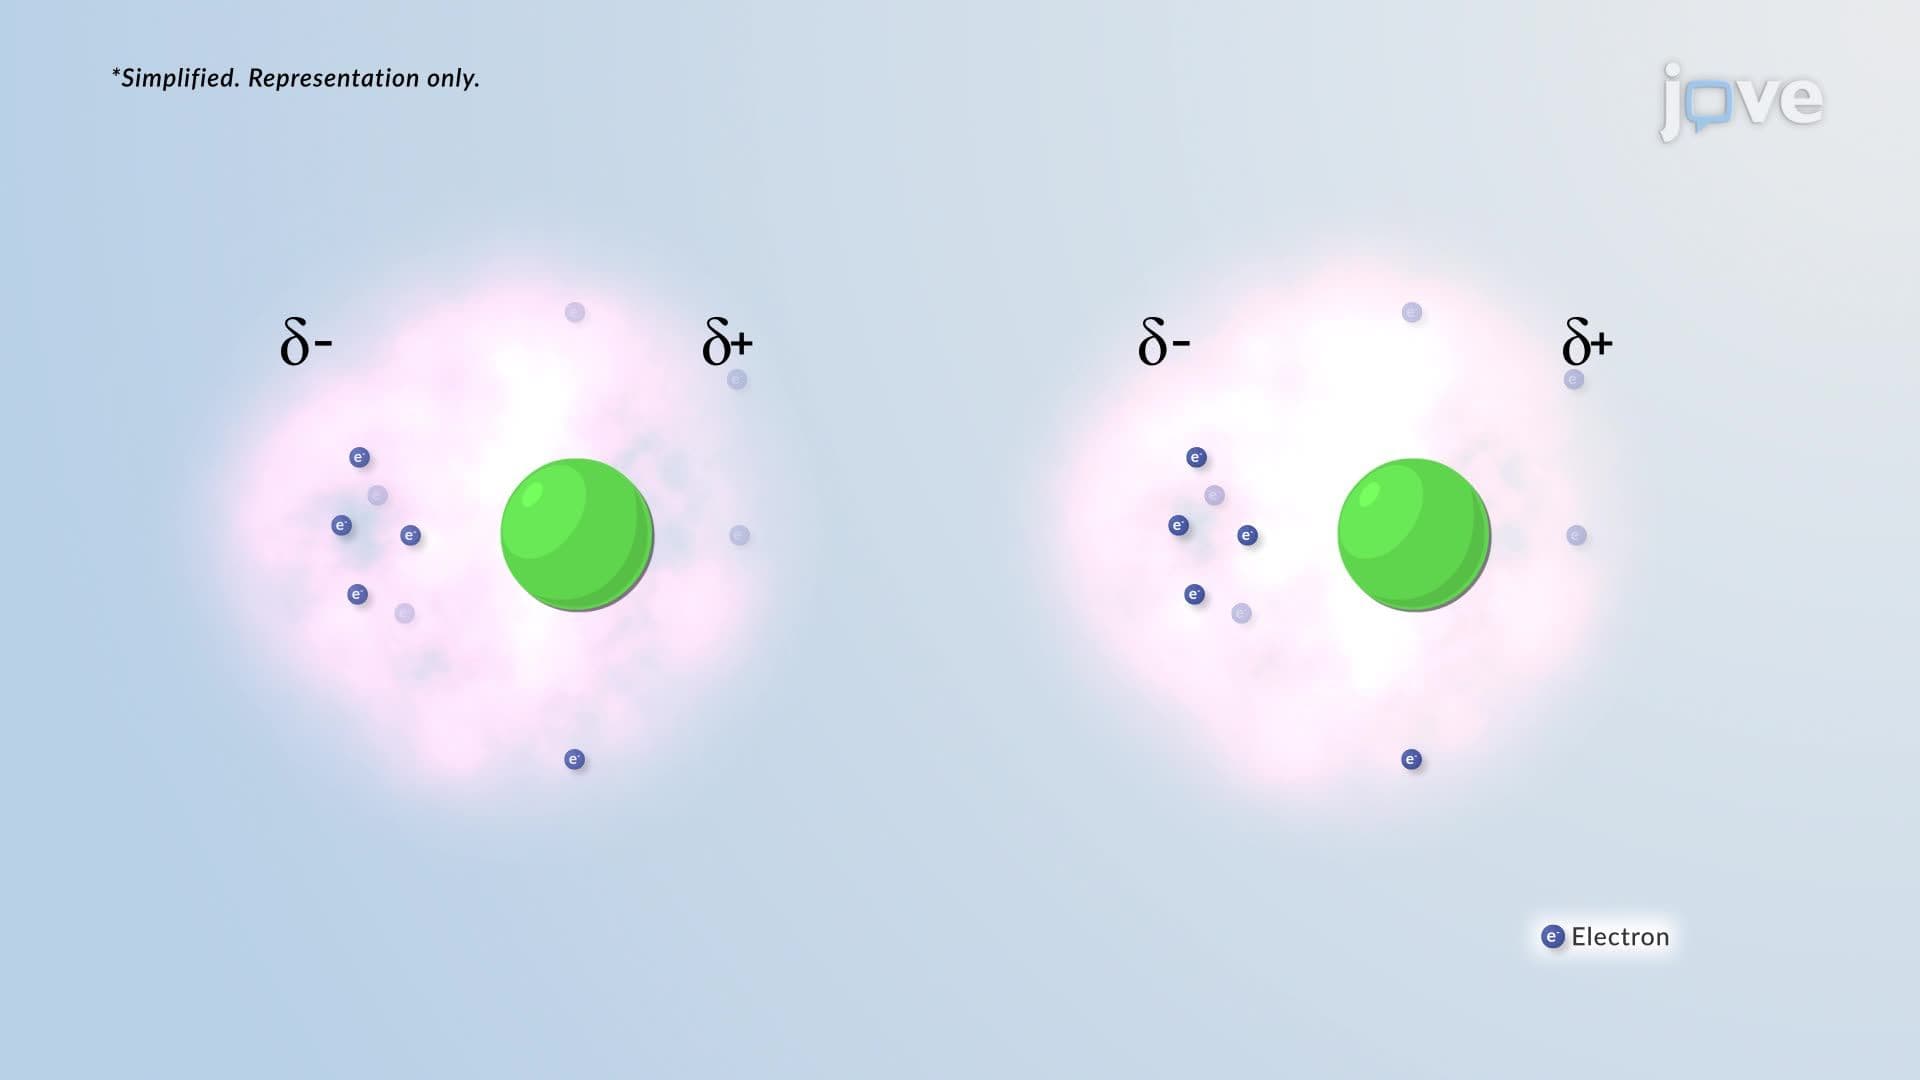
Van der Waals Interactions
 03:13
Intermolecular Forces
03:13Intermolecular Forces
 02:36
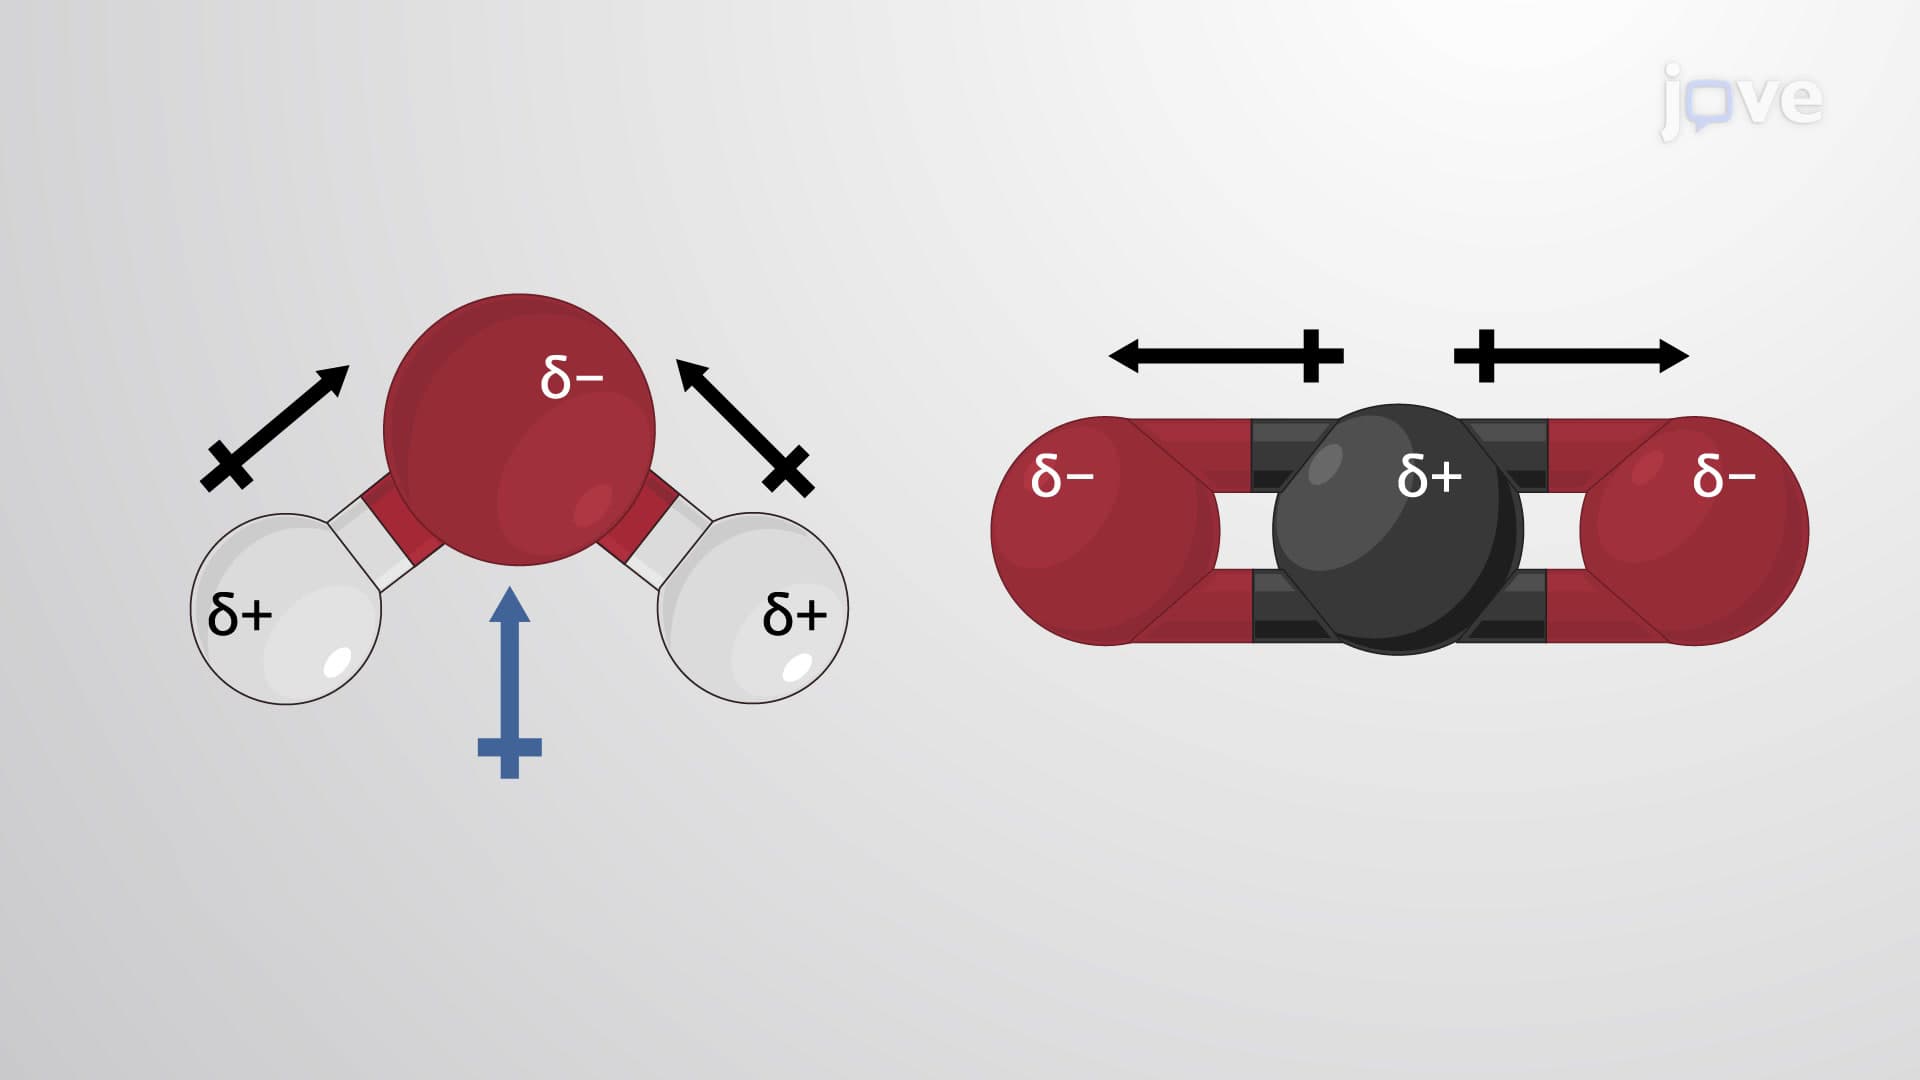
Molecular Geometry and Dipole Moments