Related Experiment Video
Updated: Jun 13, 2026


Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Density functional theoretical study of pentacene/noble metal interfaces with van der Waals corrections: vacuum level
Kenji Toyoda1, Ikutaro Hamada, Kyuho Lee
1The Institute of Scientific and Industrial Research, Osaka University, 8-1 Mihogaoka, Ibaraki, Osaka 567-0047, Japan.

Abstract:
In order to clarify factors determining the interface dipole, we have studied the electronic structures of pentacene adsorbed on Cu(111), Ag(111), and Au(111) by using first-principles density functional theoretical calculations. In the structural optimization, a semiempirical van der Waals (vdW) approach [S. Grimme, J. Comput. Chem. 27, 1787 (2006)] is employed to include long-range vdW interactions and is shown to reproduce pentacene-metal distances quite accurately. The pentacene-metal distances for Cu, Ag, and Au are evaluated to be 0.24, 0.29, and 0.32 nm, respectively, and work function changes calculated by using the theoretically optimized adsorption geometries are in good agreement with the experimental values, indicating the validity of the present approach in the prediction of the interface dipole at metal/organic interfaces. We examined systematically how the geometric factors, especially the pentacene-substrate distance (Z(C)), and the electronic properties of the metal substrates contribute to the interface dipole. We found that at Z(C) > or = 0.35 nm, the work function changes (Delta phi's) do not depend on the substrate work function (phi(m)), indicating that the interface level alignment is nearly in the Schottky limit, whereas at Z(C) < or = 0.25 nm, Delta phi's vary nearly linearly with phi(m), and the interface level alignment is in the Bardeen limit. Our results indicate the importance of both the geometric and the electronic factors in predicting the interface dipoles. The calculated electronic structure shows that on Au, the long-range vdW interaction dominates the pentacene-substrate interaction, whereas on Cu and Ag, the chemical hybridization contributes to the interaction.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 09:45
Accessing Valuable Ligand Supports for Transition Metals: A Modified, Intermediate Scale Preparation of 1,2,3,4,5-Pentamethylcyclopentadiene
Published on: March 20, 2017
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:12
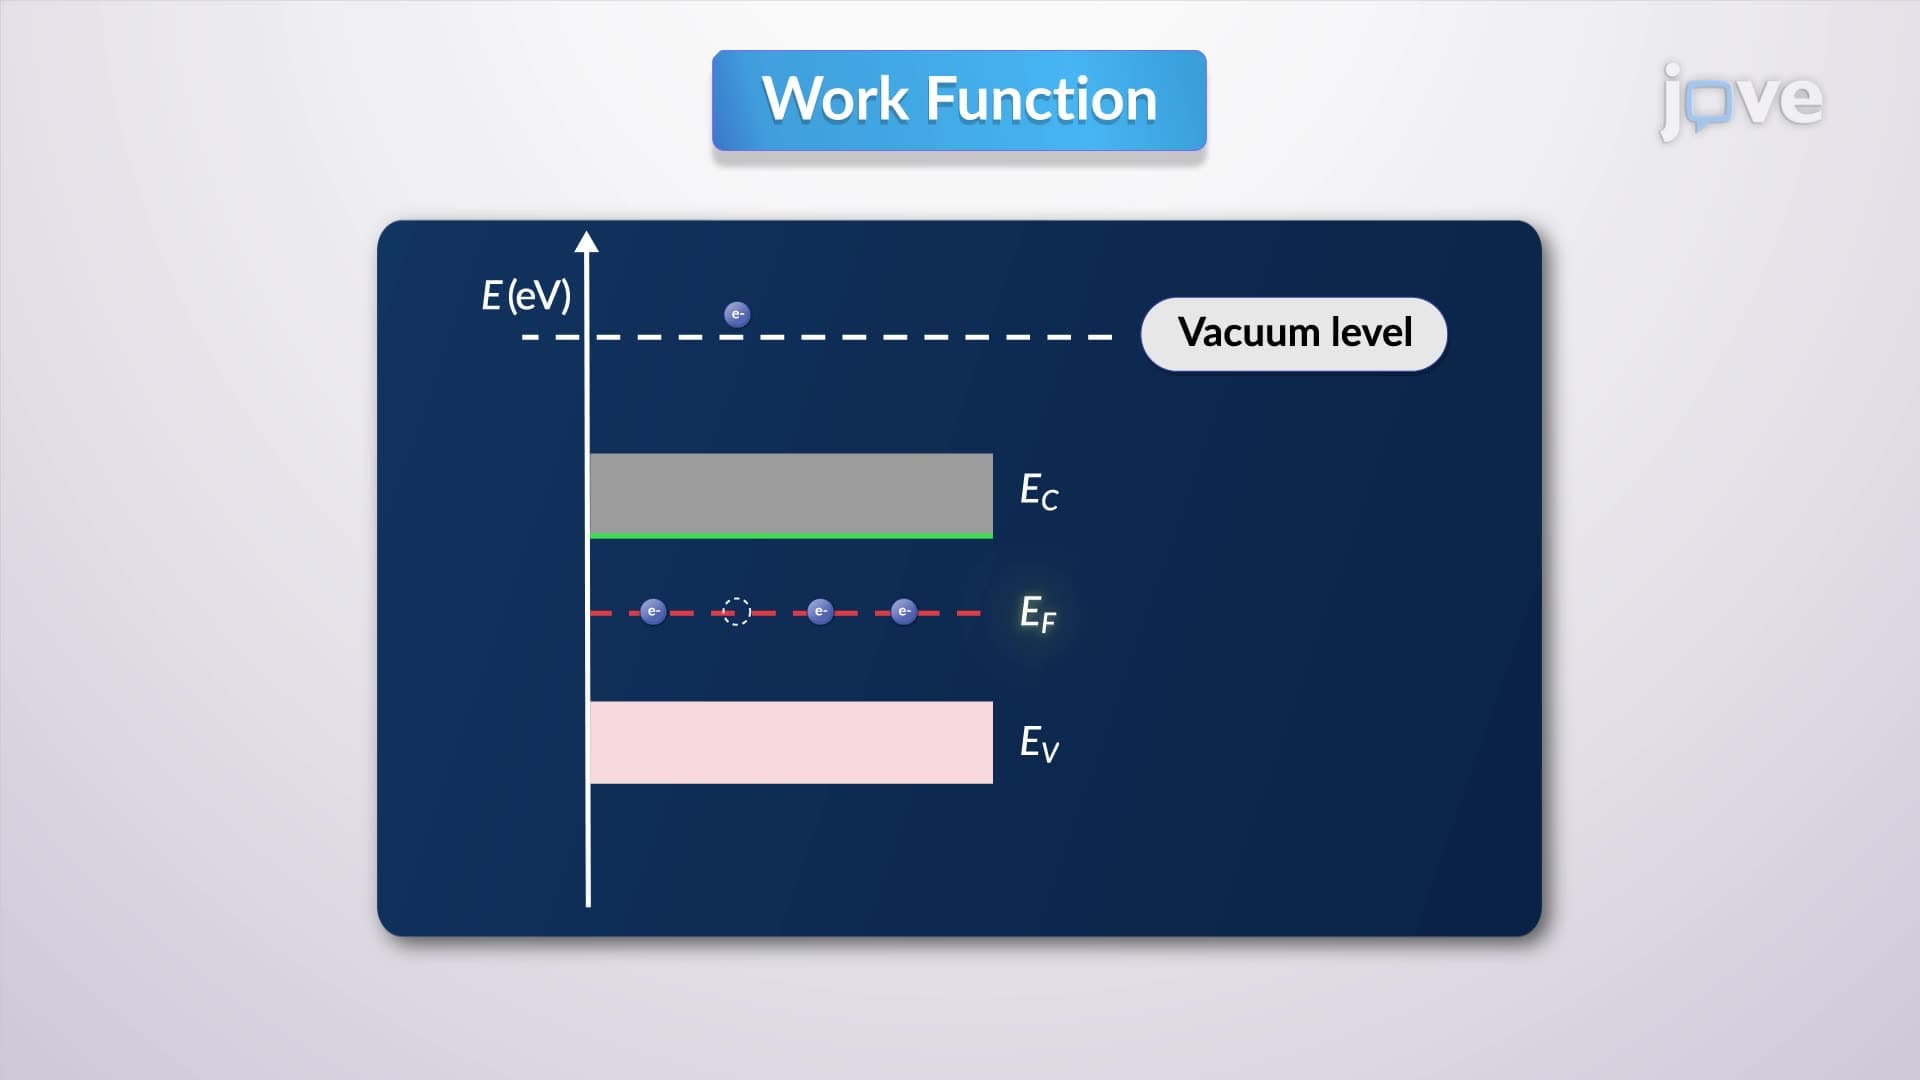
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 02:42
Valence Bond Theory
 01:18
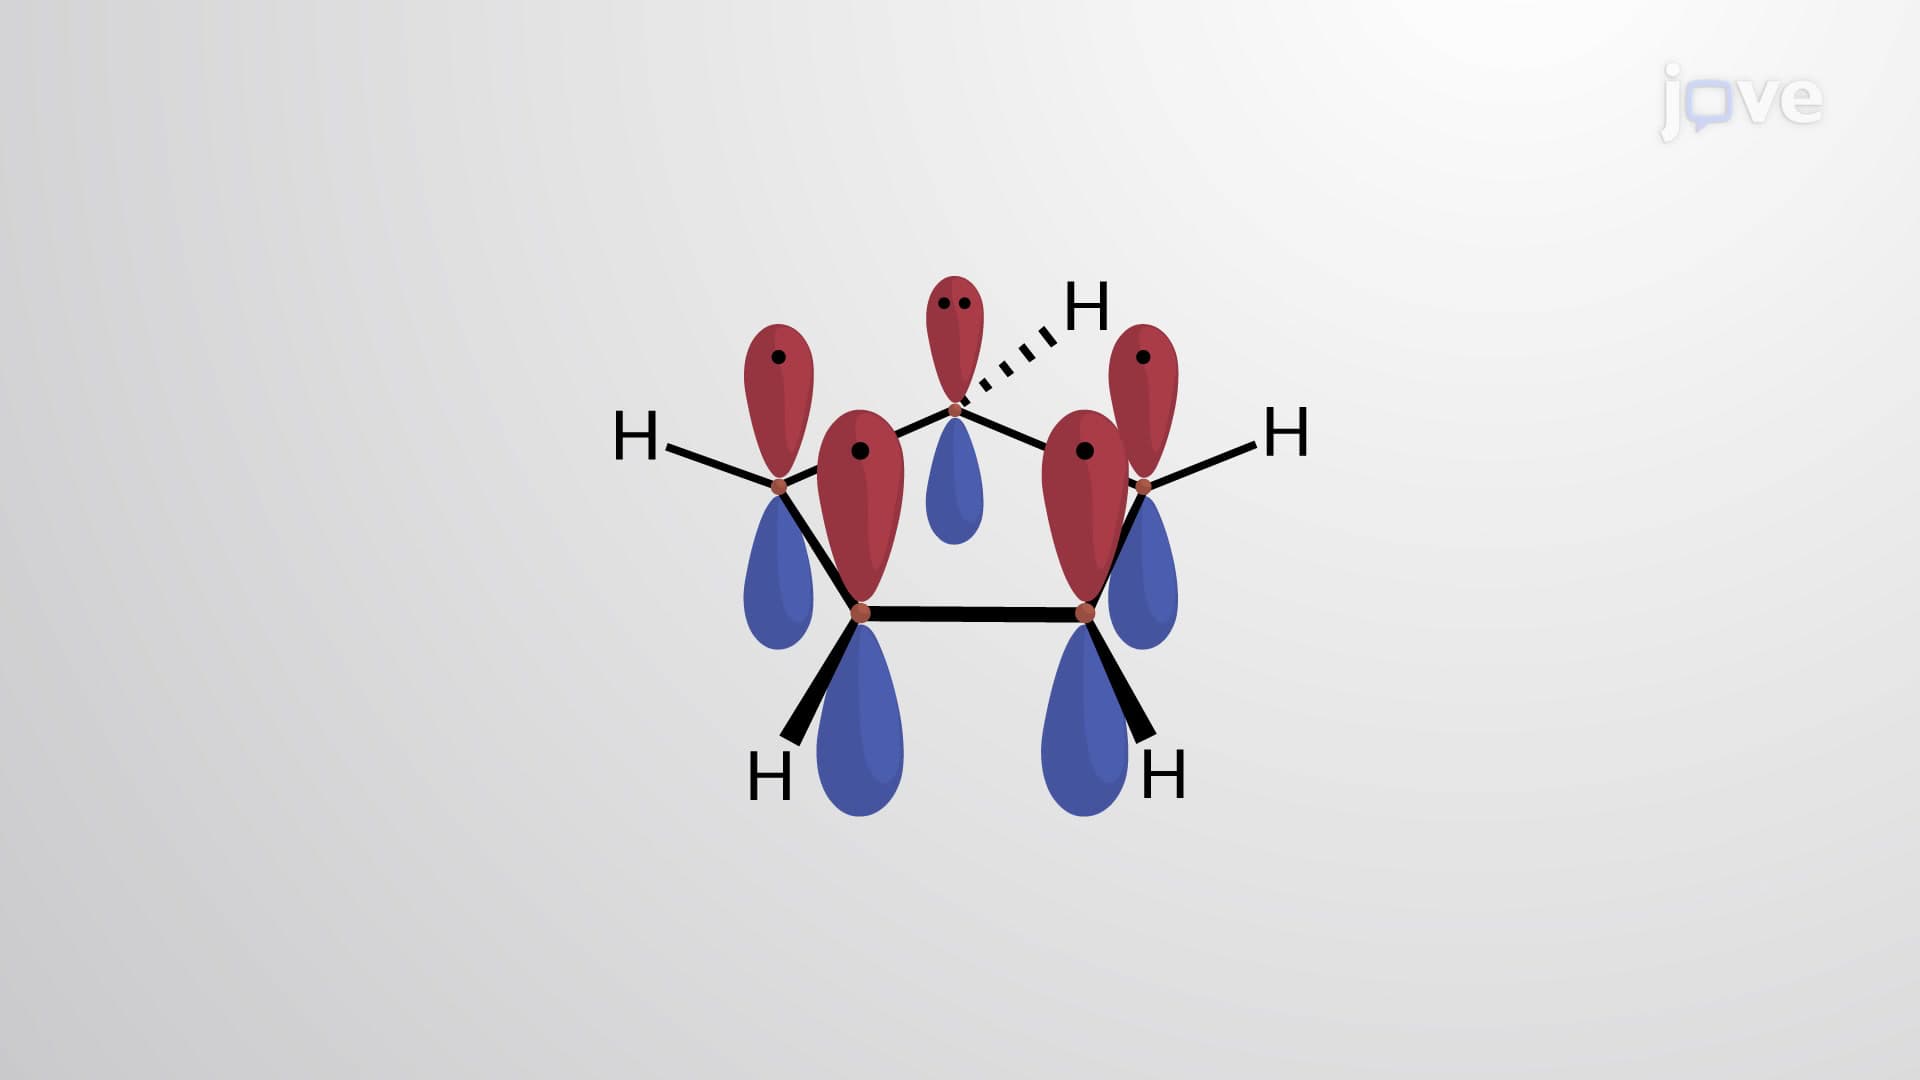
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous overlap of p...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...