Related Experiment Video
Updated: Jun 13, 2026

Real-Time, Semi-Automated Fluorescent Measurement of the Airway Surface Liquid pH of Primary Human Airway Epithelial Cells
Published on: June 13, 2019
Critical role of cytosolic phospholipase A2{alpha} in bronchial mucus hypersecretion in CFTR-deficient mice
1Unité de Défense Innée et Inflammation, Inserm, Paris, France.

Abstract:
Cystic fibrosis (CF) is due to mutations in the CF transmembrane conductance regulator gene CFTR. CF is characterised by mucus dehydration, chronic bacterial infection and inflammation, and increased levels of cytosolic phospholipase A2α (cPLA2α) products in airways. We aimed to examine the role of cPLA2α in the modulation of mucus production and inflammation in CFTR-deficient mice and epithelial cells. Mucus production was assessed using histological analyses, immuno-histochemistry and MUC5AC ELISA. cPLA2α activation was measured using an enzymatic assay and lung inflammation determined by histological analyses and polymorphonuclear neutrophil counts in bronchoalveolar lavages. In lungs from Cftr(-/-) mice, lipopolysaccharide induced mucus overproduction and MUC5AC expression associated with an increased cPLA2α activity. Mucus overproduction was mimicked by instillation of the cPLA2α product arachidonic acid, and abolished by either a cPLA2α null mutation or pharmacological inhibition. An increased cPLA2α activity was observed in bronchial explants from CF patients. CFTR silencing induced cPLA2α activation and MUC5AC expression in bronchial human epithelial cells. This expression was enhanced by arachidonic acid and reduced by cPLA2α inhibition. However, inhibition of CFTR chloride transport function had no effect on MUC5AC expression. Reduction of CFTR expression increased cPLA2α activity. This led to an enhanced mucus production in airway epithelia independent of CFTR chloride transport function. cPLA2α represents a suitable new target for therapeutic intervention in CF.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:24
Chronic Obstructive Pulmonary Disease III: Chronic Bronchitis Features
 01:20
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
Chronic Inflammation
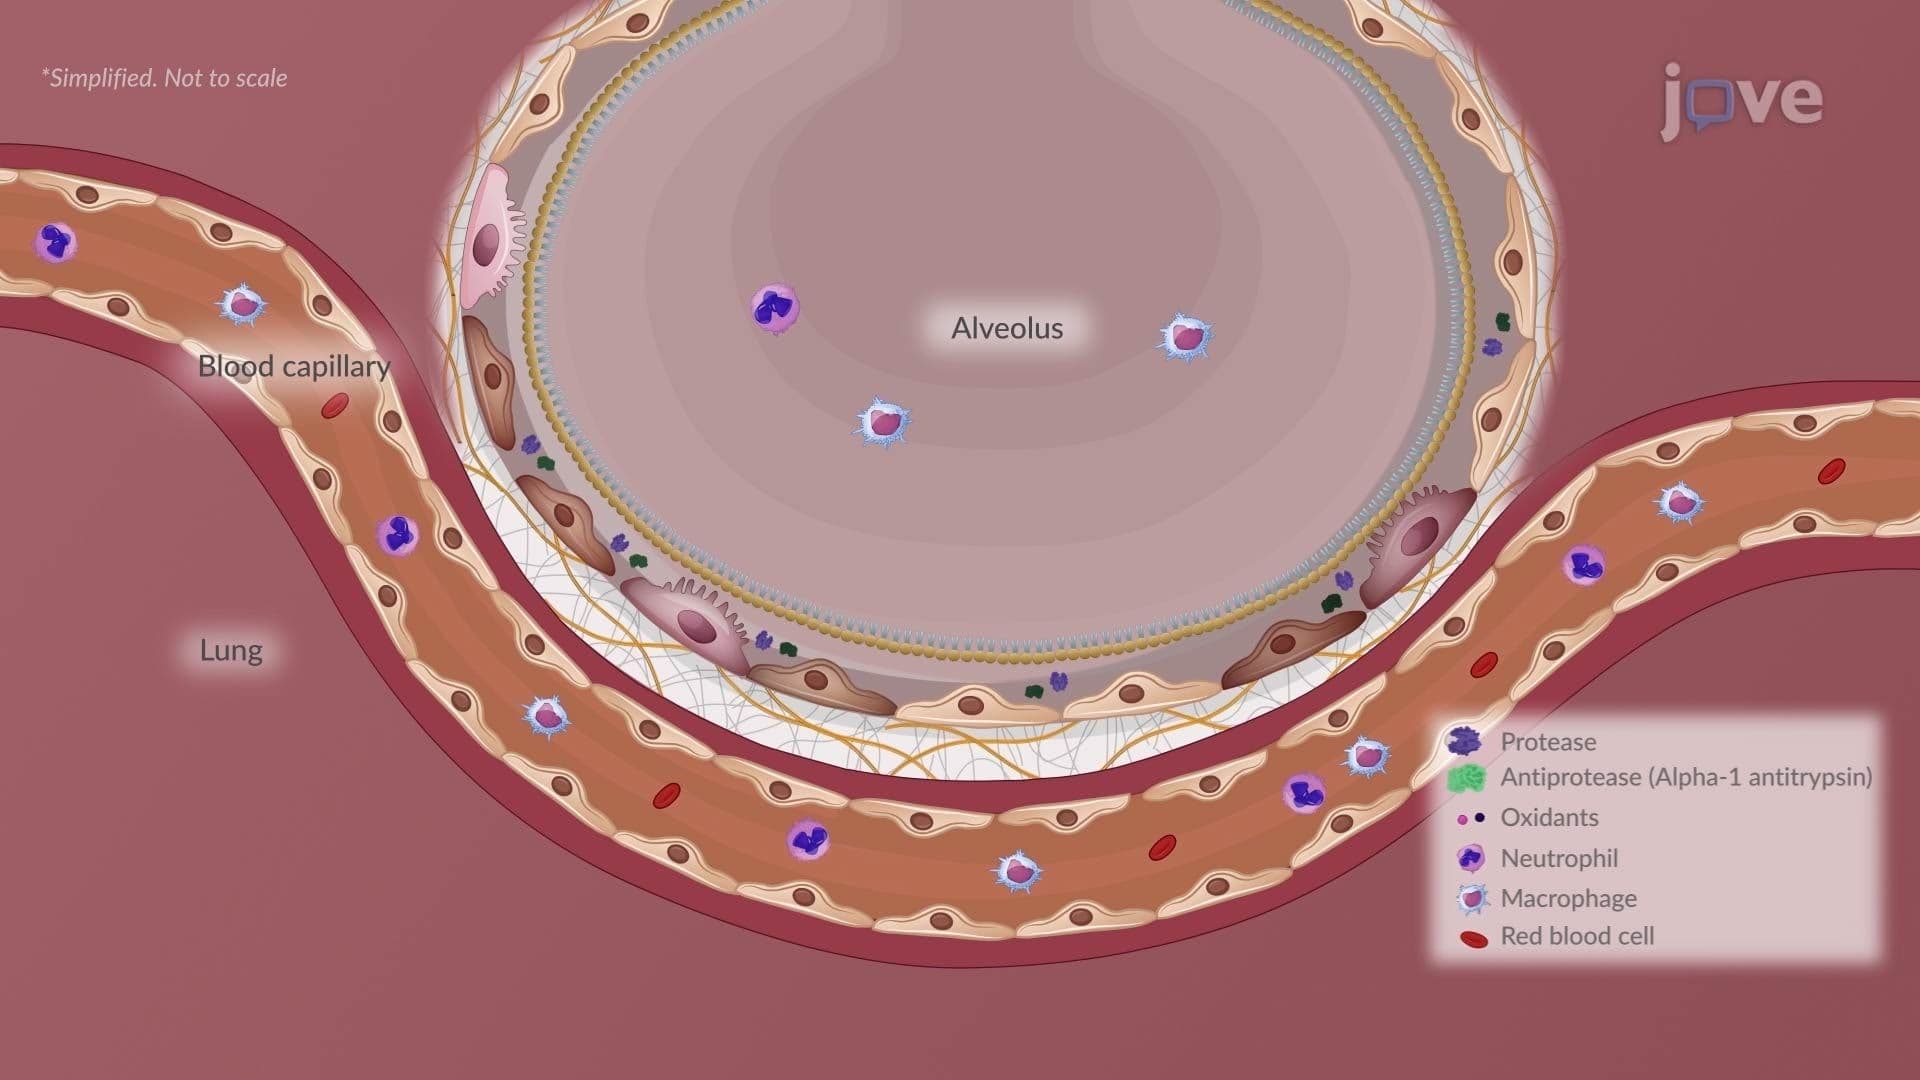 01:23
Chronic Obstructive Pulmonary Disease II: Emphysema
 01:09
GPCRs Regulate Adenylyl Cylase Activity
Two...