Related Experiment Video
Updated: Jun 13, 2026

Microscopic Visualization of Porous Nanographenes Synthesized through a Combination of Solution and On-Surface Chemistry
Published on: March 4, 2021
Role of van der Waals interaction in forming molecule-metal junctions: flat organic molecules on the Au(111) surface
M Mura1, A Gulans, T Thonhauser
1Physics, King's College London, The Strand, WC2R 2LS, London, UK.

Abstract:
The self-assembly of flat organic molecules on metal surfaces is controlled, apart from the kinetic factors, by the interplay between the molecule-molecule and molecule-surface interactions. These are typically calculated using standard density functional theory within the generalized gradient approximation, which significantly underestimates nonlocal correlations, i.e. van der Waals (vdW) contributions, and thus affects interactions between molecules and the metal surface in the junction. In this paper we address this question systematically for the Au(111) surface and a number of popular flat organic molecules which form directional hydrogen bonds with each other. This is done using the recently developed first-principles vdW-DF method which takes into account the nonlocal nature of electron correlation [M. Dion et al., Phys. Rev. Lett. 2004, 92, 246401]. We report here a systematic study of such systems involving completely self-consistent vdW-DF calculations with full geometry relaxation. We find that the hydrogen bonding between the molecules is only insignificantly affected by the vdW contribution, both in the gas phase and on the gold surface. However, the adsorption energies of these molecules on the surface increase dramatically as compared with the ordinary density functional (within the generalized gradient approximation, GGA) calculations, in agreement with available experimental data and previous calculations performed within approximate or semiempirical models, and this is entirely due to the vdW contribution which provides the main binding mechanism. We also stress the importance of self-consistency in calculating the binding energy by the vdW-DF method since the results of non-self-consistent calculations in some cases may be off by up to 20%. Our calculations still support the usually made assumption of the molecule-surface interaction changing little laterally suggesting that single molecules and their small clusters should be quite mobile at room temperature on the surface. These findings support a gas-phase modeling for some flat metal surfaces, such as Au(111), and flat molecules, at least as a first approximation.
More Related Videos
 04:57
Residue-Free Fabrication of van der Waals Heterostructures of Two-Dimensional Materials
Published on: July 18, 2025
 08:29
Au-Interaction of Slp1 Polymers and Monolayer from Lysinibacillus sphaericus JG-B53 - QCM-D, ICP-MS and AFM as Tools for Biomolecule-metal Studies
Published on: January 19, 2016
Related Concept Videos
 01:24
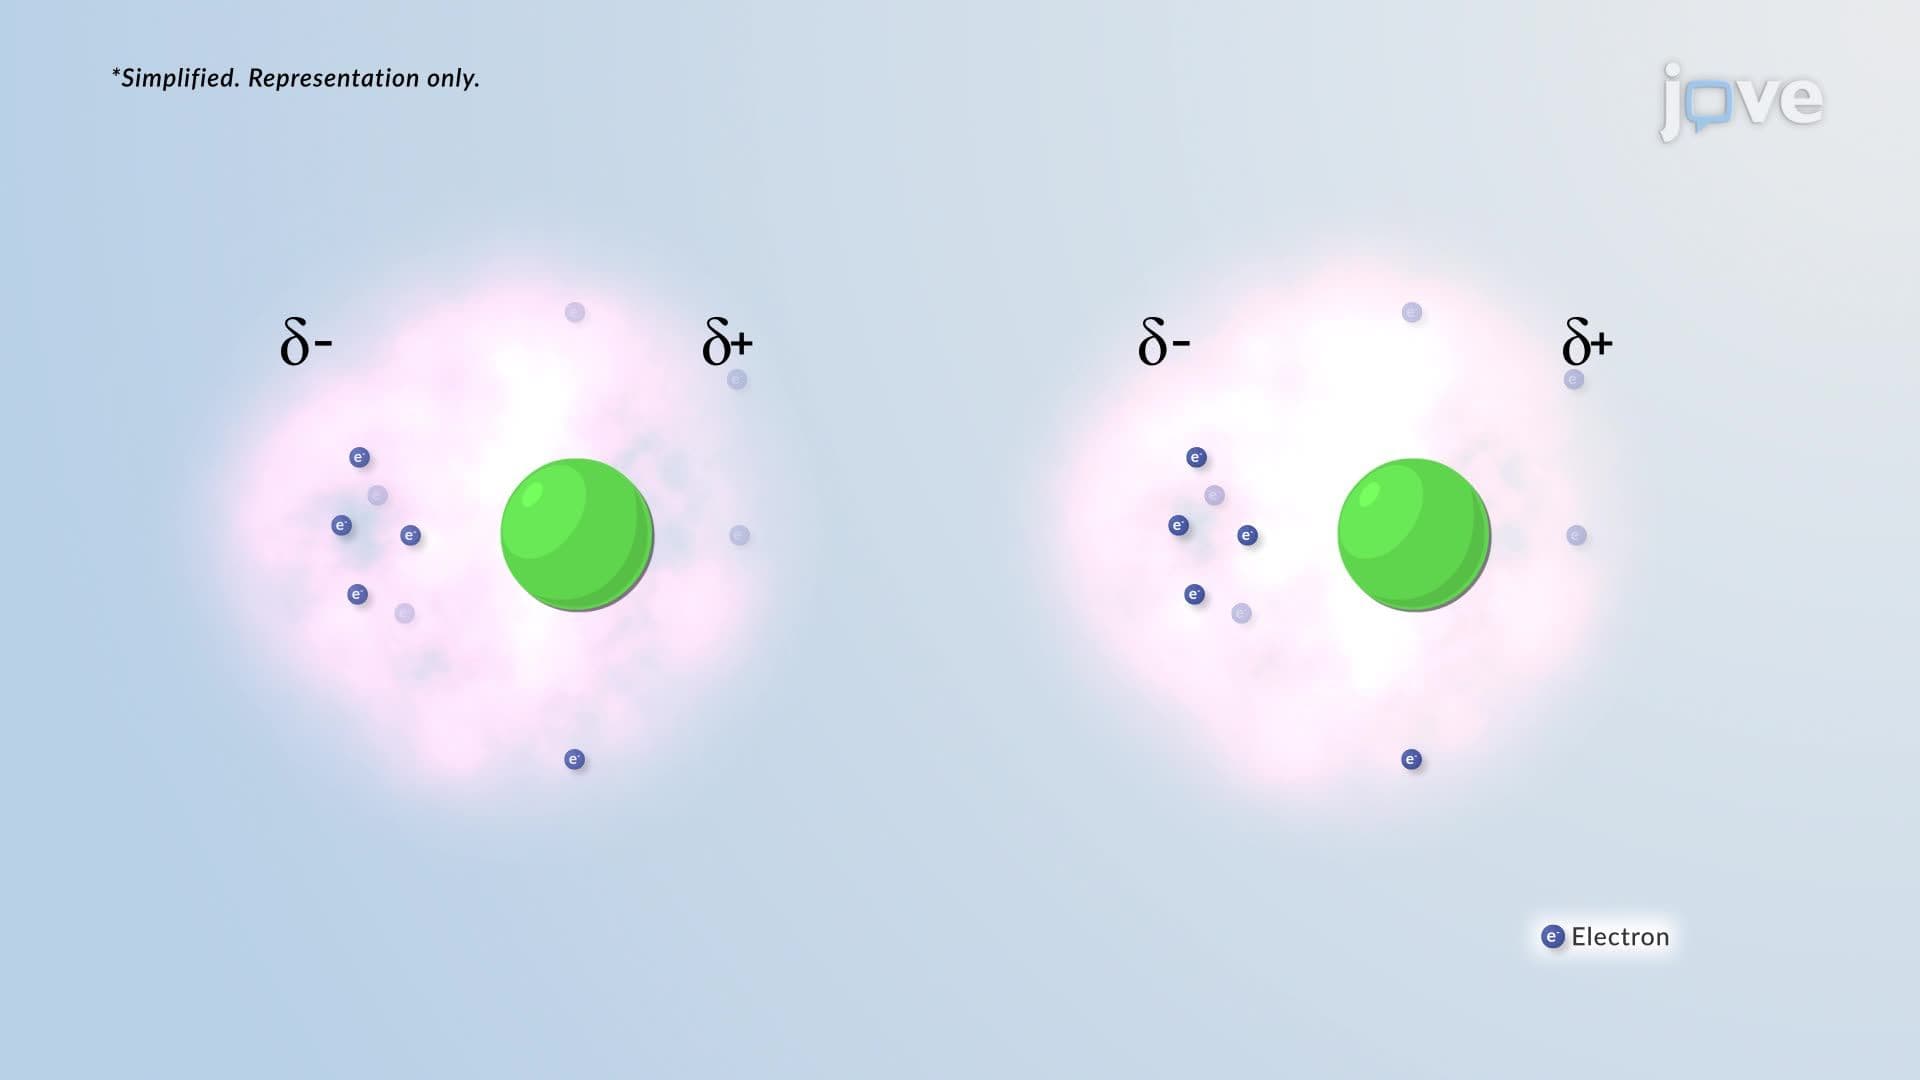
Van der Waals Interactions
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...
 02:42
Valence Bond Theory
 02:45
Valence Bond Theory
 02:32
Bonding in Metals
 02:35
Noncovalent Attractions in Biomolecules
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...