Related Experiment Video
Updated: Jun 13, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Absolutely local occupied and excited molecular orbitals in the third-order single excitation perturbation theory for
1Toyota Physical and Chemical Research Institute, Nagakute, Aichi-gun, Aichi, 480-1192 Japan. riken-iwata@mosk.tytlabs.co.jp

Abstract:
The efficient code to evaluate the molecular interaction energy at the Hartree-Fock level of theory is developed. The method is the third-order single excitation perturbation theory based on the locally projected (LP) molecular orbitals (MO). It is shown that with a proper scaling of the molecular orbital integrals the basis set superposition error can be under controll even with a small size of basis sets. If the basis set contains augmented diffuse functions, the interaction energy, close to the counterpoise corrected energy, can be directly evaluated without the scaling, as fast as a single supermolecule Hartree-Fock calculation. The key of the method is to use the absolutely local occupied and excited MOs. With these MOs, the amount of the charge-transfer (CT) and the energy contribution of the CT terms are evaluated.

Related Concept Videos
 02:35
Molecular Orbital Theory I
 02:40
MO Theory and Covalent Bonding
 03:51
Molecular Orbital Theory II
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:18
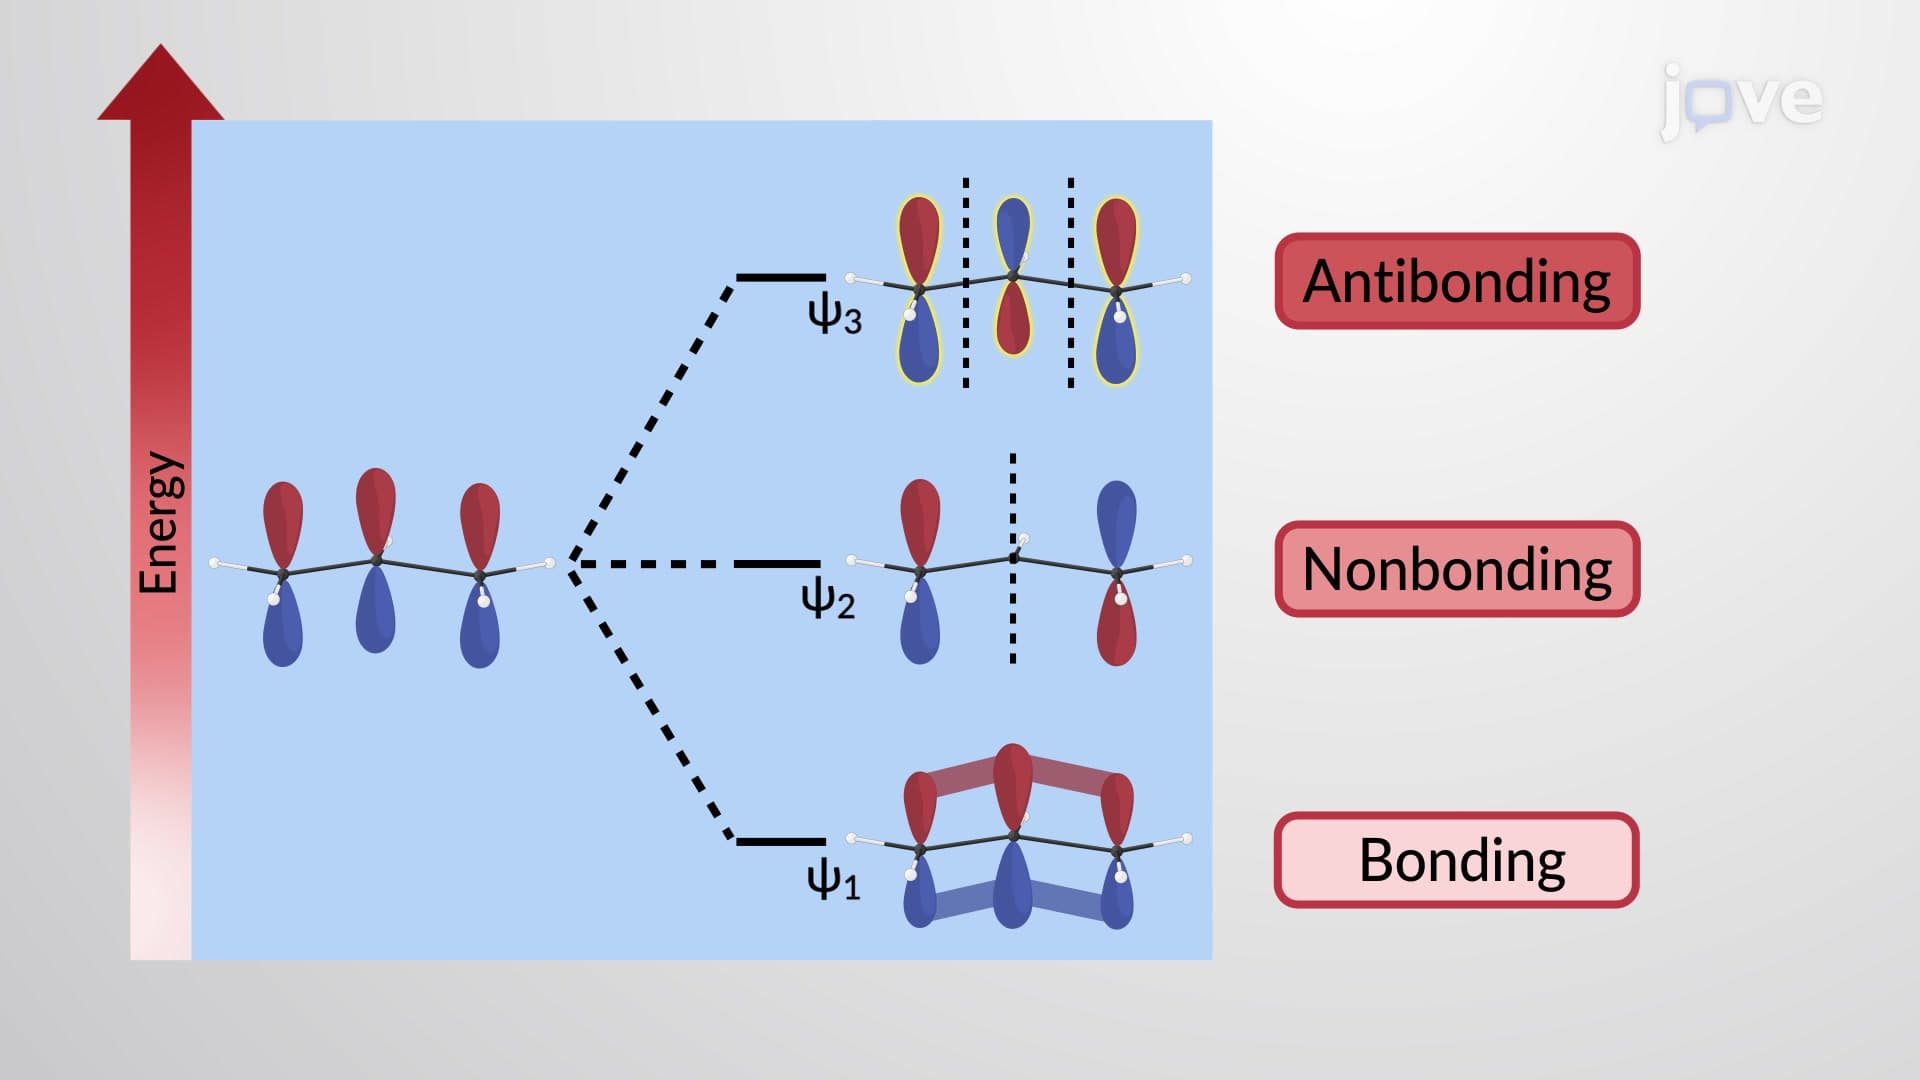
π Molecular Orbitals of the Allyl Cation and Anion
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...