Related Experiment Video
Updated: Jun 13, 2026
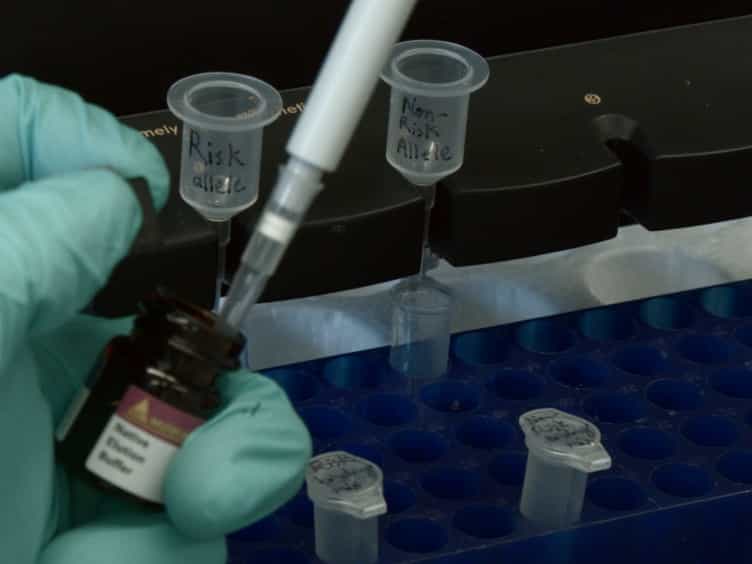
Screening for Functional Non-coding Genetic Variants Using Electrophoretic Mobility Shift Assay (EMSA) and DNA-affinity Precipitation Assay (DAPA)
Published on: August 21, 2016
Interpretation of association signals and identification of causal variants from genome-wide association studies
Kai Wang1, Samuel P Dickson, Catherine A Stolle
1Center for Applied Genomics, Children's Hospital of Philadelphia, Philadelphia, PA 19104, USA. kaichop@gmail.com

Abstract:
GWAS have been successful in identifying disease susceptibility loci, but it remains a challenge to pinpoint the causal variants in subsequent fine-mapping studies. A conventional fine-mapping effort starts by sequencing dozens of randomly selected samples at susceptibility loci to discover candidate variants, which are then placed on custom arrays or used in imputation algorithms to find the causal variants. We propose that one or several rare or low-frequency causal variants can hitchhike the same common tag SNP, so causal variants may not be easily unveiled by conventional efforts. Here, we first demonstrate that the true effect size and proportion of variance explained by a collection of rare causal variants can be underestimated by a common tag SNP, thereby accounting for some of the "missing heritability" in GWAS. We then describe a case-selection approach based on phasing long-range haplotypes and sequencing cases predicted to harbor causal variants. We compare this approach with conventional strategies on a simulated data set, and we demonstrate its advantages when multiple causal variants are present. We also evaluate this approach in a GWAS on hearing loss, where the most common causal variant has a minor allele frequency (MAF) of 1.3% in the general population and 8.2% in 329 cases. With our case-selection approach, it is present in 88% of the 32 selected cases (MAF = 66%), so sequencing a subset of these cases can readily reveal the causal allele. Our results suggest that thinking beyond common variants is essential in interpreting GWAS signals and identifying causal variants.
More Related Videos
 08:27
Large-Scale Multi-Omics Genome-Wide Association Studies (Mo-GWAS): Guidelines for Sample Preparation and Normalization
Published on: July 27, 2021
 07:15
Determining the Likelihood of Variant Pathogenicity Using Amino Acid-level Signal-to-Noise Analysis of Genetic Variation
Published on: January 16, 2019
Related Concept Videos
 01:11
Genome-wide Association Studies-GWAS
GWAS does not require the identification of the target gene involved in...
 01:29
Pharmacogenomics: Identification of New Drug Targets
 02:54
Evolutionary Relationships through Genome Comparisons
 02:46
Genetic Screens
Forward genetic screens
Forward or “classical” genetic screens involve creating random mutations in an organism’s DNA using radiation, mutagens, or insertion of additional bases, which result in visible changes...
 03:36
Genome Annotation and Assembly
 02:26
Comparing Copy Number Variations and SNPs
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...