Related Experiment Video
Updated: Jun 13, 2026

Determining Surface Areas and Pore Volumes of Metal-Organic Frameworks
Published on: March 8, 2024
Theory of nitride oxide adsorption on transition metal (111) surfaces: a first-principles investigation
Zhen-Hua Zeng1, Juarez L F Da Silva, Wei-Xue Li
1State Key Laboratory of Catalysis and Center for Theoretical and Computational Chemistry, Dalian Institute of Chemical Physics, Chinese Academy of Sciences, 116023 Dalian, China.

Abstract:
In this work, we report a density functional theory study of nitric oxide (NO) adsorption on close-packed transition metal (TM) Rh(111), Ir(111), Pd(111) and Pt(111) surfaces in terms of adsorption sites, binding mechanism and charge transfer at a coverage of Theta(NO) = 0.25, 0.50, 0.75 monolayer (ML). Based on our study, an unified picture for the interaction between NO and TM(111) and site preference is established, and valuable insights are obtained. At low coverage (0.25 ML), we find that the interaction of NO/TM(111) is determined by an electron donation and back-donation process via the interplay between NO 5sigma/2pi* and TM d-bands. The extent of the donation and back-donation depends critically on the coordination number (adsorption sites) and TM d-band filling, and plays an essential role for NO adsorption on TM surfaces. DFT calculations shows that for TMs with high d-band filling such as Pd and Pt, hollow-site NO is energetically the most favorable, and top-site NO prefers to tilt away from the normal direction. While for TMs with low d-band filling (Rh and Ir), top-site NO perpendicular to the surfaces is energetically most favorable. Electronic structure analysis show that irrespective of the TM and adsorption site, there is a net charge transfer from the substrate to the adsorbate due to overwhelming back-donation from the TM substrate to the adsorbed NO molecules. The adsorption-induced change of the work function with respect to bare surfaces and dipole moment is however site dependent, and the work function increases for hollow-site NO, but decreases for top-site NO, because of differences in the charge redistribution. The interplay between the energetics, lateral interaction and charge transfer, which is element dependent, rationalizes the structural evolution of NO adsorption on TM(111) surfaces in the submonolayer regime.
More Related Videos
Related Concept Videos
 01:37
1° Amines to Diazonium or Aryldiazonium Salts: Diazotization with NaNO2 Mechanism
 01:28
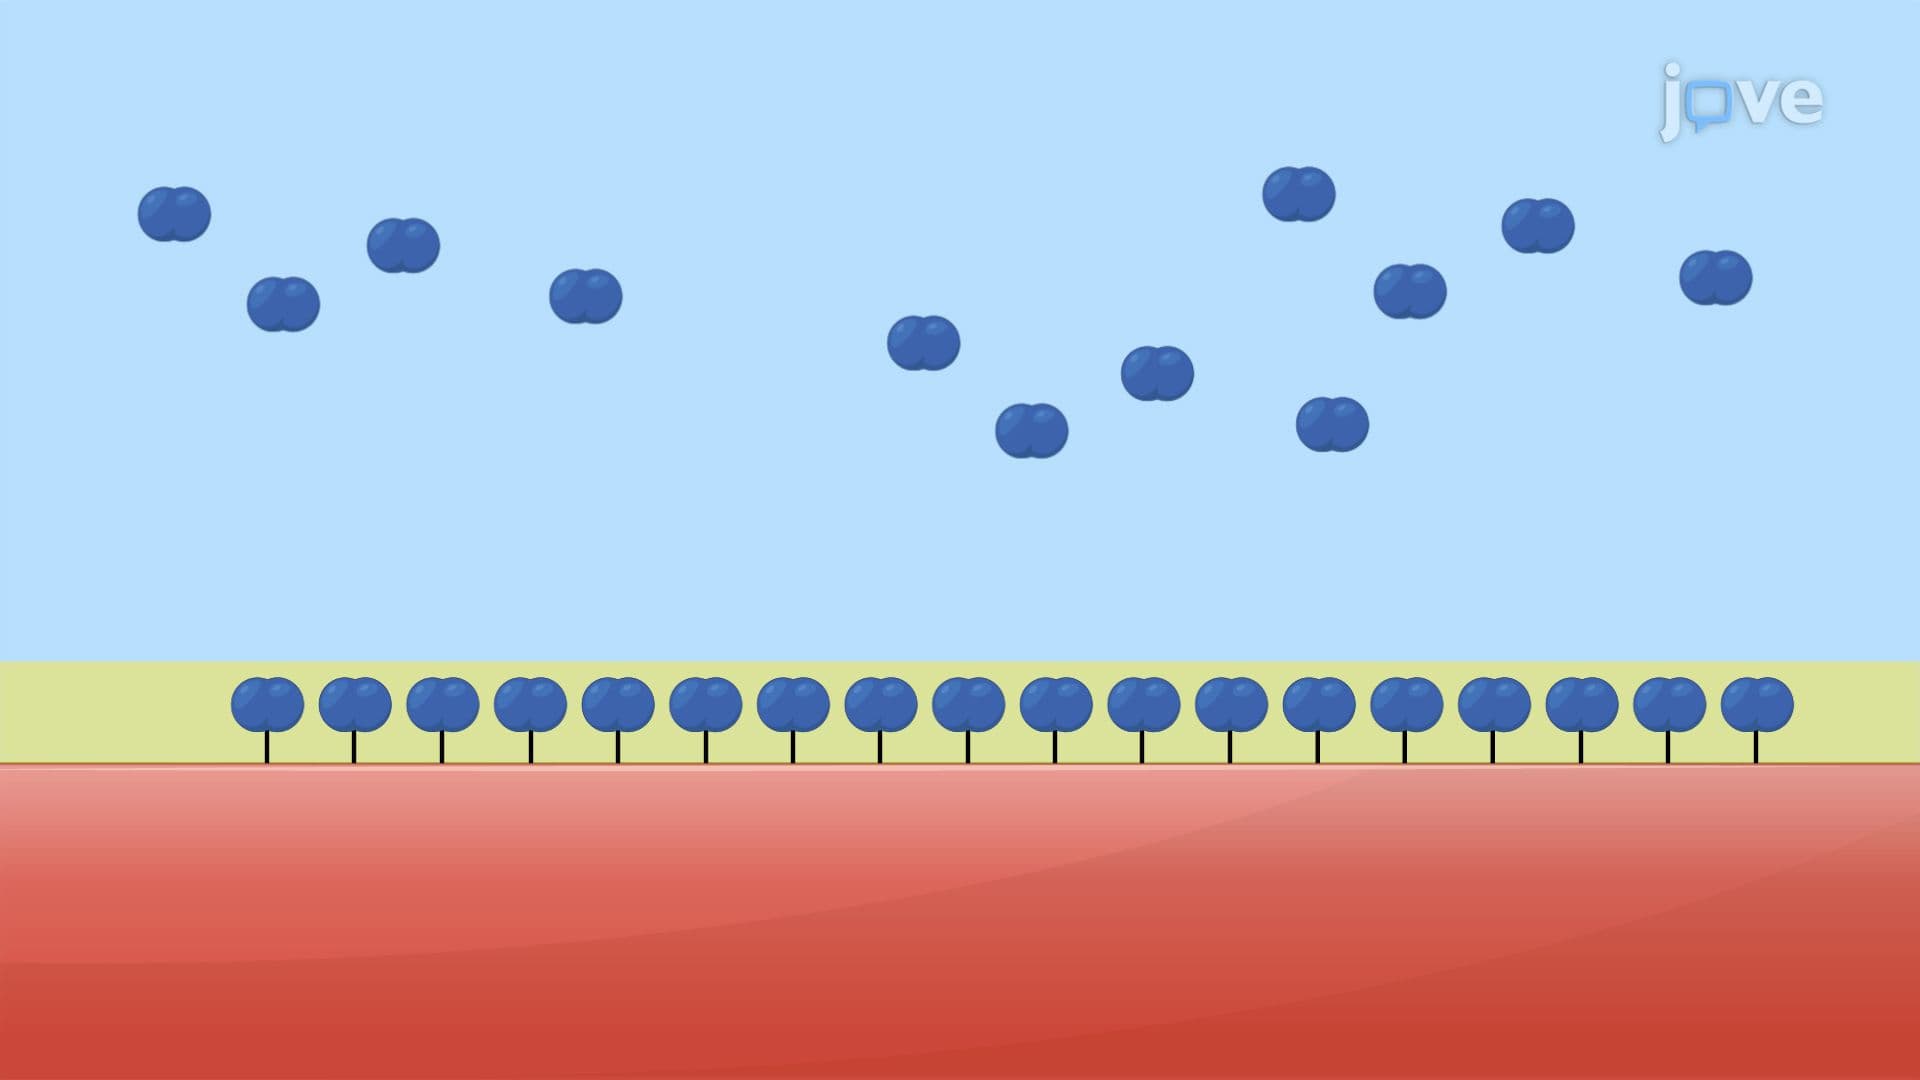
Adsorption of Gases on Solids
 01:25
Adsorption Isotherms II
 02:42
Valence Bond Theory
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...