Improving protein and proteome coverage through data-independent multiplexed peptide fragmentation
Related Experiment Video
Updated: Jun 13, 2026
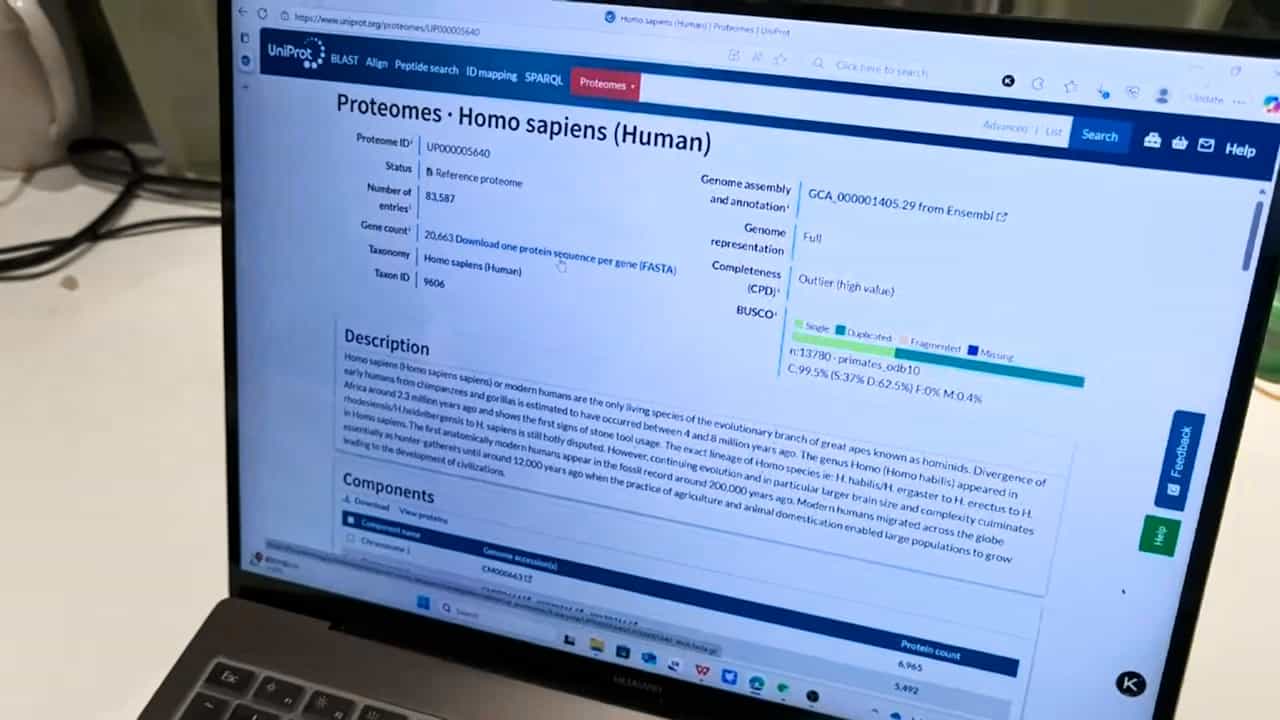
Navigating the Mass Spectrometry-Based Proteomic Data Using Free Computational Tools
Published on: August 19, 2025
Kevin Blackburn1, Flaubert Mbeunkui, Srijeet K Mitra
1Department of Molecular and Structural Biochemistry, North Carolina State University, Raleigh, North Carolina 27695-7622, USA.

Abstract:
Performance differences in protein and proteome characterization achieved by data-independent acquisition (DIA) LC/MS(E) and data-dependent acquisition (DDA) LC/MS/MS approaches were investigated. LC/MS(E) is a novel mode of generating product ion data for all coeluting precursors in parallel as opposed to LC/MS/MS where coeluting precursors must be serially fragmented one at a time. During LC/MS(E) analysis, alternating MS scans of "normal" and "elevated" collision energy are collected at regular intervals, providing nearly a 100% duty cycle for precursor detection and fragmentation because all precursors are fragmented across their full chromatographic elution profile. This is in contrast to DDA-based MS/MS where serial selection of precursor ions is biased toward interrogation and detection of the highest abundance sample components by virtue of the intensity-driven interrogation scheme employed. Both modes of acquisition were applied to a simple four-protein standard mixture with a 16-fold dynamic range in concentration, an in-gel digest of the Arabidopsis thaliana protein FLS2 purified by immunoprecipitation, and a solution-digested tomato leaf proteome sample. Dramatic improvement for individual protein sequence coverage was obtained for all three samples analyzed by the DIA approach, particularly for the lowest abundance sample components. In many instances, precursors readily detected and identified during DIA were either interrogated by MS/MS during DDA at inopportune points in their chromatographic elution profiles resulting in poor quality product ion spectra or not interrogated at all. Detailed evaluation of both DDA and DIA raw data and timing of the MS-to-MS/MS switching events clearly revealed the fundamental limitations of serial MS/MS interrogation and the advantages of parallel fragmentation by DIA for more comprehensive protein identification and characterization which holds promise for enhanced isoform and post-translational modification analysis.
More Related Videos
 10:37
Deep Proteome Profiling by Isobaric Labeling, Extensive Liquid Chromatography, Mass Spectrometry, and Software-assisted Quantification
Published on: November 15, 2017
 09:35
Resolving Affinity Purified Protein Complexes by Blue Native PAGE and Protein Correlation Profiling
Published on: April 1, 2017
Related Concept Videos
 01:33
Peptide Identification Using Tandem Mass Spectrometry
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
 01:33
Proteomics
Proteomics is the study of proteomes' function. It involves the large-scale systematic study of the proteome to denote the protein complement expressed by a genome. Scientist Mark Wilkins coined the term proteomics...