Related Experiment Video
Updated: Jun 13, 2026

Patch Clamp and Perfusion Techniques for Studying Ion Channels Expressed in Xenopus oocytes
Published on: January 10, 2011
μ-conotoxin KIIIA derivatives with divergent affinities versus efficacies in blocking voltage-gated sodium channels
Min-Min Zhang1, Tiffany S Han, Baldomero M Olivera
1Department of Biology, University of Utah, Salt Lake City, Utah 84112, USA.

Abstract:
The possibility of independently manipulating the affinity and efficacy of pore-blocking ligands of sodium channels is of interest for the development of new drugs for the treatment of pain. The analgesic mu-conotoxin KIIIA (KIIIA), a 16-residue peptide with three disulfide bridges, is a pore blocker of voltage-gated sodium channels, including neuronal subtype Na(V)1.2 (K(d) = 5 nM). At saturating concentrations, KIIIA incompletely blocks the sodium current of Na(V)1.2, leaving a 5% residual current (rI(Na)). Lys7 is an important residue: the K7A mutation decreases both the efficacy (i.e., increases rI(Na) to 23%) and the affinity of the peptide (K(d) = 115 nM). In this report, various replacements of residue 7 were examined to determine whether affinity and efficacy were inexorably linked. Because of their facile chemical synthesis, KIIIA analogues that had as a core structure the disulfide-depleted KIIIA[C1A,C2U,C9A,C15U] (where U is selenocysteine) or ddKIIIA were used. Analogues ddKIIIA and ddKIIIA[K7X], where X represents one of nine different amino acids, were tested on voltage-clamped Xenopus oocytes expressing rat Na(V)1.2 or Na(V)1.4. Their affinities ranged from 0.01 to 36 muM and rI(Na) values from 2 to 42%, and these two variables appeared to be uncorrelated. Instead, rI(Na) varied inversely with side chain size, and remarkably charge and hydrophobicity appeared to be inconsequential. The ability to manipulate a mu-conopeptide's affinity and efficacy, as well as its capacity to interfere with subsequent tetrodotoxin binding, greatly expands its scope as a reagent for probing sodium channel structure and function and may also lead to the development of mu-conotoxins as safe analgesics.
Related Concept Videos
 01:17
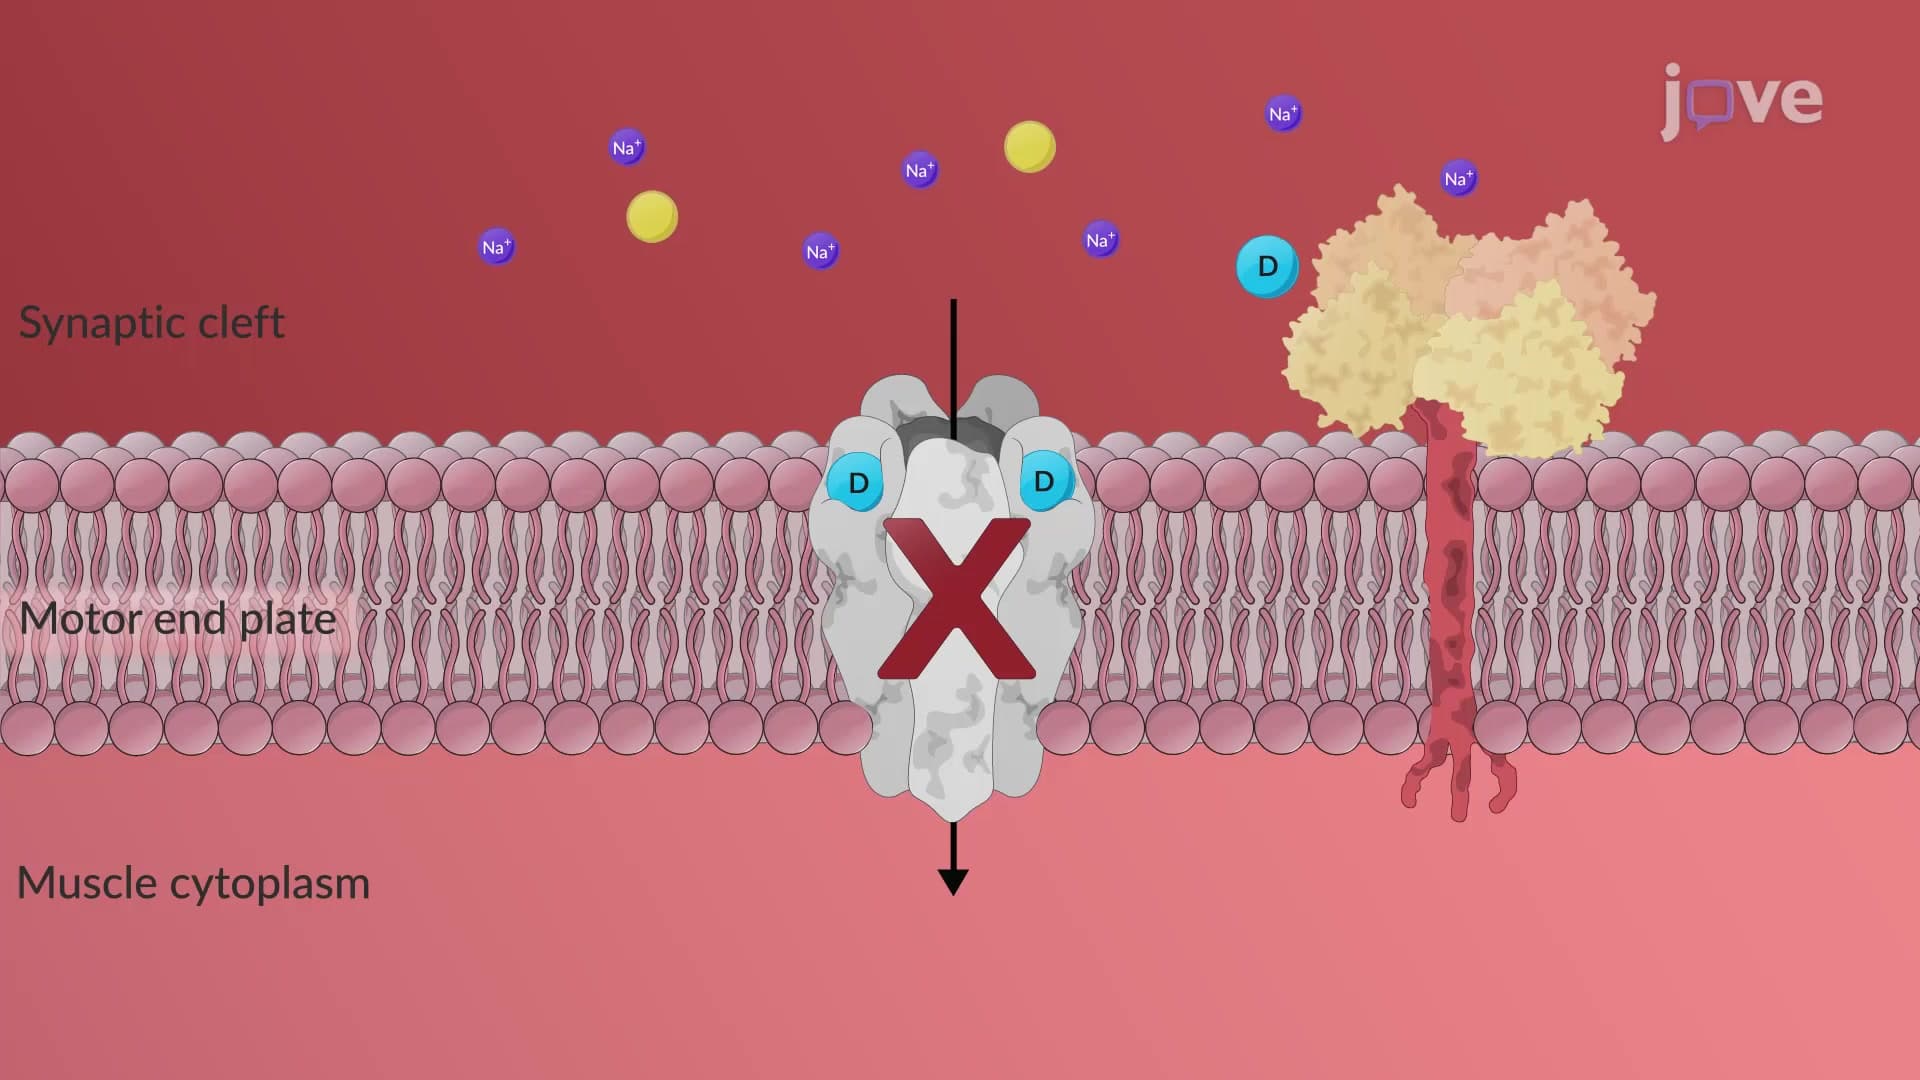
Nondepolarizing (Competitive) Neuromuscular Blockers: Mechanism of Action
Competitive antagonists prevent acetylcholine from binding to its receptor, inhibiting membrane depolarization. Without conformational changes or intrinsic...
 01:22
Antiarrhythmic Drugs: Class I Agents as Sodium Channel Blockers
Class 1A Antiarrhythmic Drugs: These drugs work by moderately blocking sodium channels,...
 01:26
Voltage-gated Ion Channels
Generally, all voltage-gated ion channels have a 'voltage-sensing domain' that spans the lipid bilayer. The charged residues in the sensor move in response to the membrane potential changes that open the channel allowing ions movement. There are several types of...
01:26Voltage-gated Ion Channels
Generally, all voltage-gated ion channels have a 'voltage-sensing domain' that spans the lipid bilayer. The charged residues in the sensor move in response to the membrane potential changes that open the channel allowing ions movement. There are several types of...
 01:28
Depolarizing Blockers: Mechanism of Action
Succinylcholine is the most commonly used depolarizing blocker. Chemically, it constitutes two molecules of acetylcholine joined together by an acetate methyl group. They act on the receptors in the same way as acetylcholine. Because succinylcholine...
 01:30
Ligand-Gated Ion Channel Receptor: Gating Mechanism