Related Experiment Video
Updated: Jun 12, 2026
Monitoring Cell-to-cell Transmission of Prion-like Protein Aggregates in Drosophila Melanogaster
Published on: March 12, 2018
The propagation of prion-like protein inclusions in neurodegenerative diseases
Michel Goedert1, Florence Clavaguera, Markus Tolnay
1MRC Laboratory of Molecular Biology, Hills Road, Cambridge, CB2 0QH, UK. mg@mrc-lmb.cam.ac.uk

Abstract:
The most common neurodegenerative diseases, including Alzheimer's disease and Parkinson's disease, are characterized by the misfolding of a small number of proteins that assemble into ordered aggregates in affected brain cells. For many years, the events leading to aggregate formation were believed to be entirely cell-autonomous, with protein misfolding occurring independently in many cells. Recent research has now shown that cell non-autonomous mechanisms are also important for the pathogenesis of neurodegenerative diseases with intracellular filamentous inclusions. The intercellular transfer of inclusions made of tau, alpha-synuclein, huntingtin and superoxide dismutase 1 has been demonstrated, revealing the existence of mechanisms reminiscent of those by which prions spread through the nervous system.
More Related Videos
Related Concept Videos
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
03:03Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
 01:23
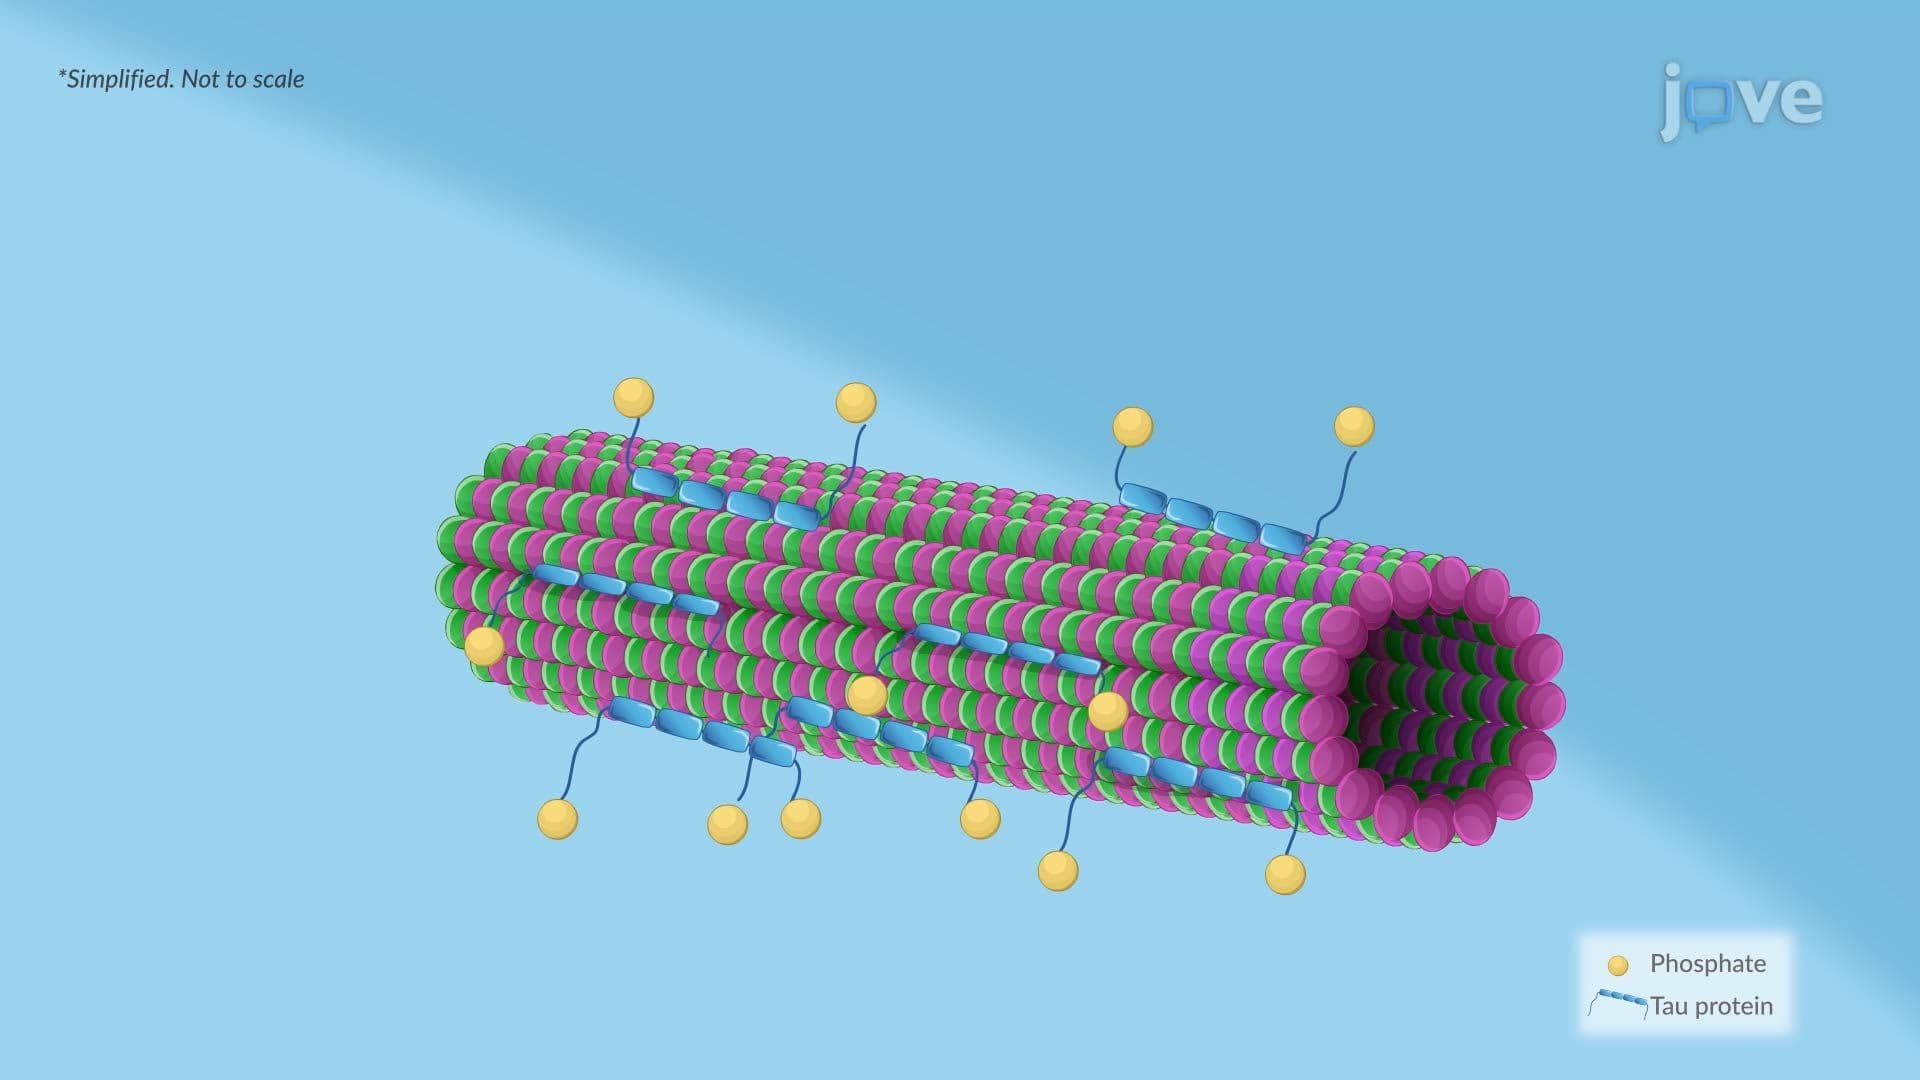
Alzheimer Disease ll: Pathophysiology
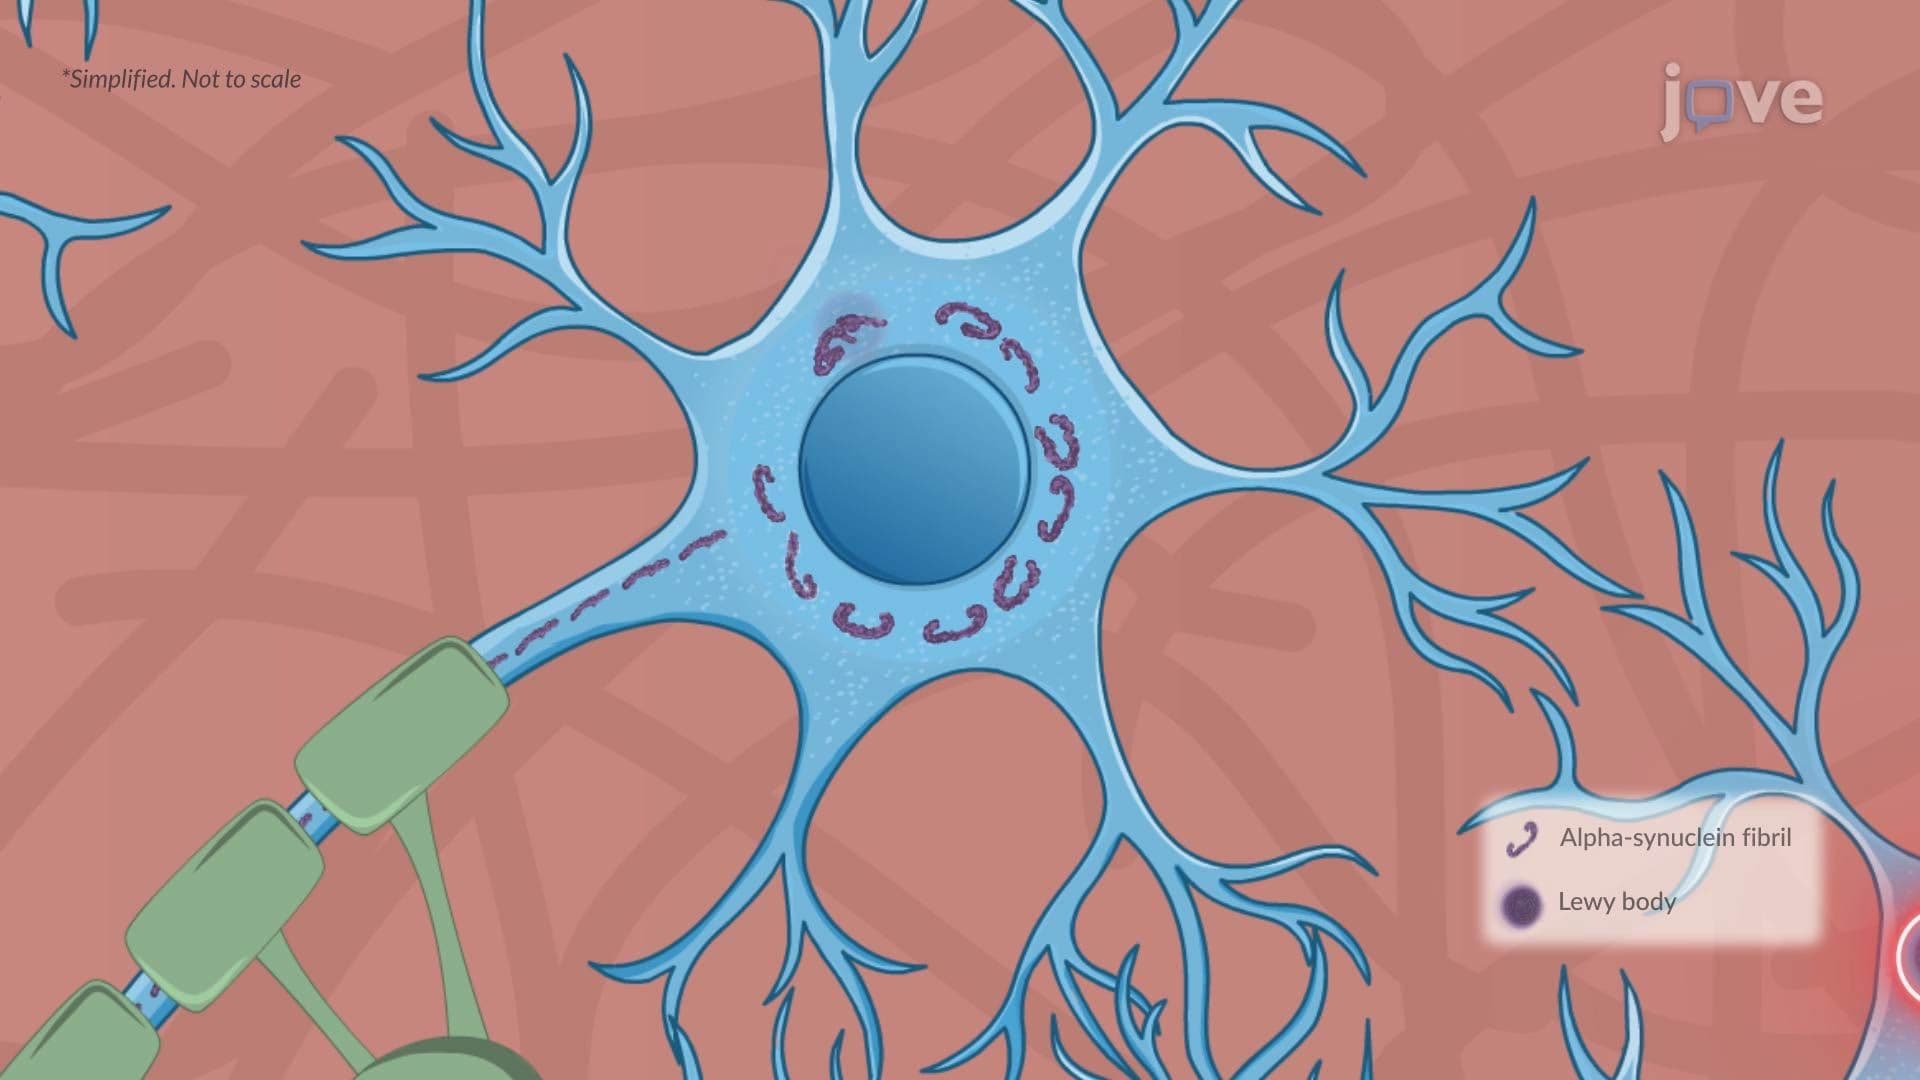 01:24
Parkinson Disease ll: Pathophysiology
 01:37
Neural Regulation
 01:29
Subviral Agents