Related Experiment Video
Updated: Jul 17, 2026

Identification of OTX1 and OTX2 As Two Possible Molecular Markers for Sinonasal Carcinomas and Olfactory Neuroblastomas
Published on: February 28, 2019
Polyalveolar lobe and congenital cystic adenomatoid malformation type II: are they related?
C A Wagenvoort1, P E Zondervan
1Department of Pathology, Erasmus University Rotterdam, The Netherlands.

Insights
Congenital cystic adenomatoid malformation type II (CCAM) can present with a polyalveolar lobe, a previously undescribed complication. This finding in infants suggests a potential causal link between these rare lung anomalies.
Area of Science:
- Pediatric Pathology
- Respiratory Medicine
- Congenital Malformations
Background:
- Congenital cystic adenomatoid malformation type II (CCAM) is a rare pulmonary malformation.
- Polyalveolar lobe, characterized by an increased number of alveoli, is a distinct lung anomaly.
- The coexistence of CCAM and polyalveolar lobe has not been previously reported.
Abstract:
A left lower polyalveolar lobe in a 28-day-old infant and a right lung with congenital cystic adenomatoid malformation type II in a 17-day-old infant are described. The adenomatoid malformation involved all lobes, but the lower lobe appeared to be mostly replaced by a polyalveolar area, a complication hitherto undescribed. The pronounced increase in number of alveoli in both cases was established by two different morphometric methods and was compared with three normal controls of the same age. The simultaneous occurrence of polyalveolar lobe and congenital cystic adenomatoid malformation type II in the same lung suggests a causal relation between both anomalies. In the grossly abnormal lung tissue of an adenomatoid malformation, polyalveolar areas can easily be overlooked.
More Related Videos
 10:21
Primary Sjogren's Syndrome Associated with Lung Adenocarcinoma: Probing the Potential Common Pathogenic Mechanisms and Experimental Verification
Published on: September 20, 2024
 06:14
Isolation of Alveolar Type II Epithelial Cells from Neonatal, Juvenile, and Adult Murine Lungs Adaptable to Infectious Experimental Settings
Published on: December 19, 2025
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:26
Alveoli and Alveolar Ducts
 01:20
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
Chronic Inflammation
 01:23
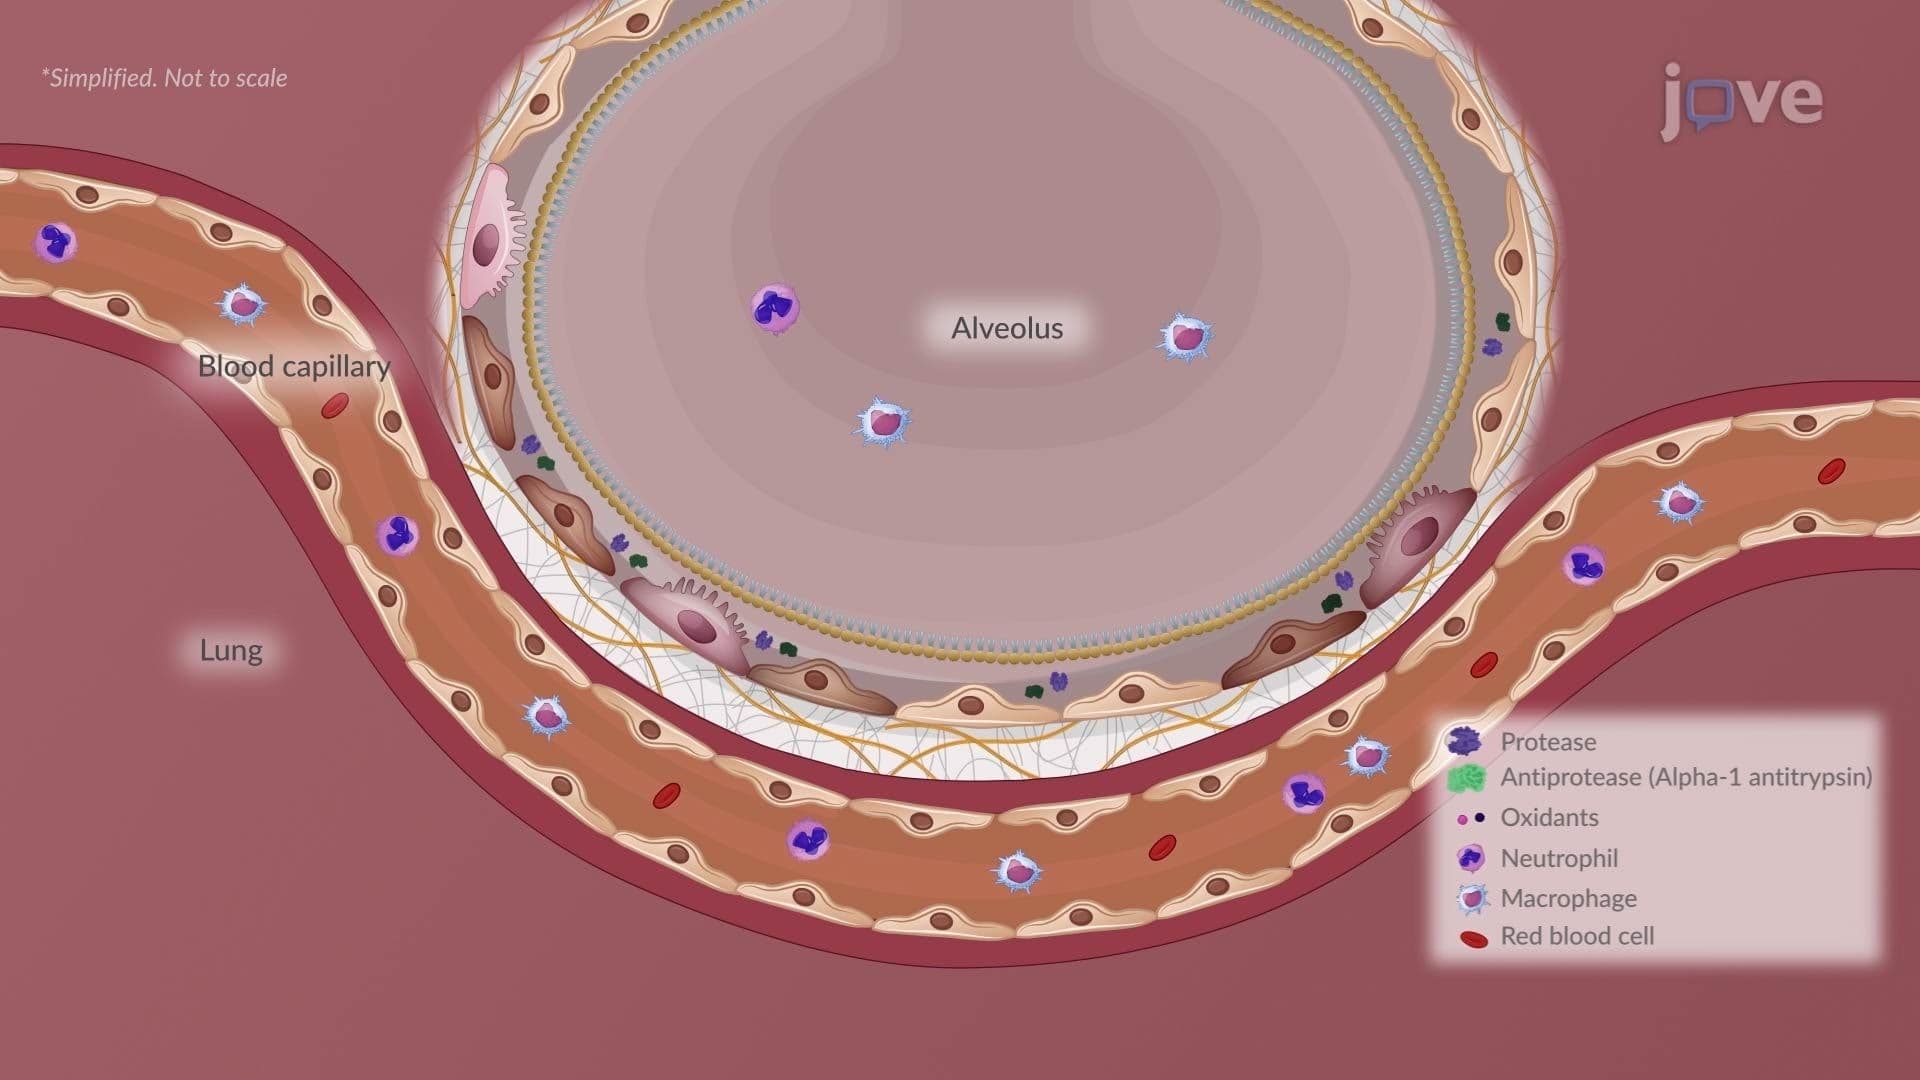
Chronic Obstructive Pulmonary Disease II: Emphysema
 01:24
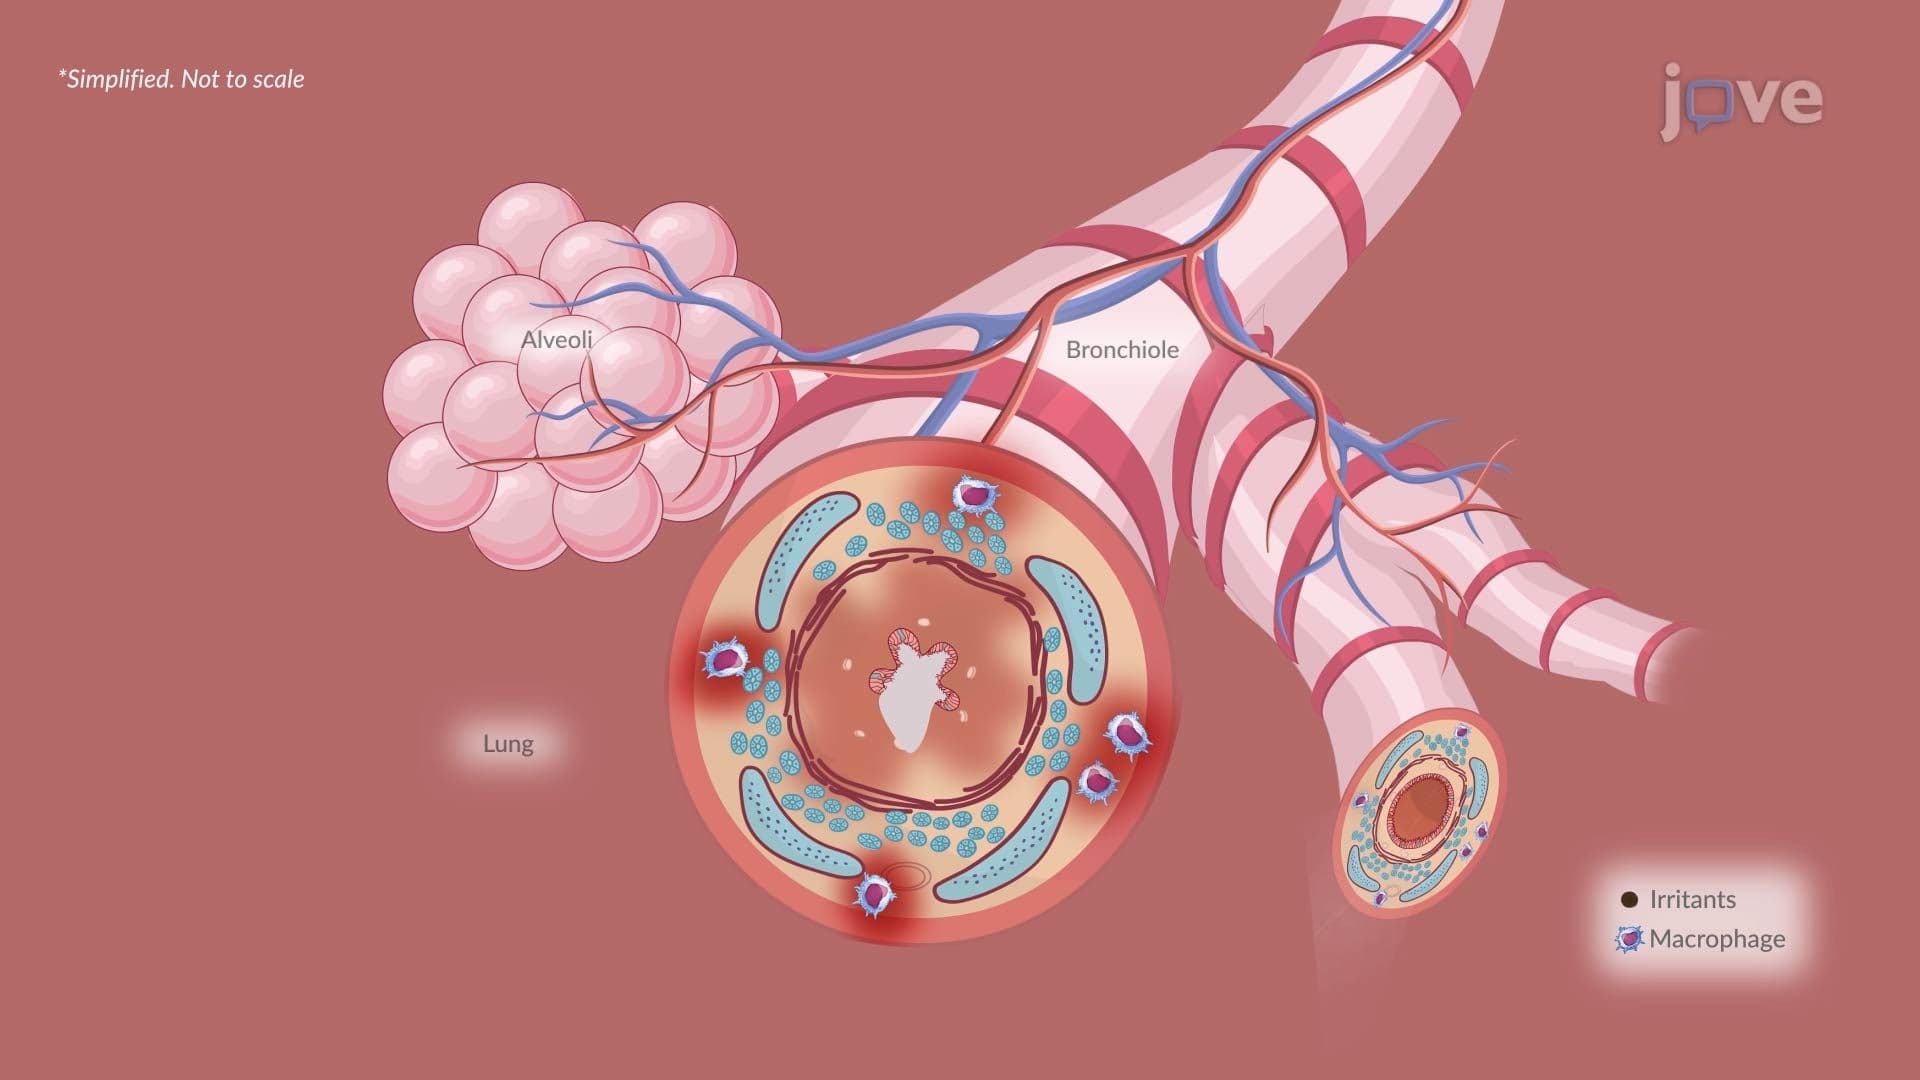
Chronic Obstructive Pulmonary Disease III: Chronic Bronchitis Features