Improving computational protein design by using structure-derived sequence profile
Related Experiment Video
Updated: Jun 12, 2026

Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Liang Dai1, Yuedong Yang, Hyung Rae Kim
1School of Informatics, Indiana University Purdue University, Indianapolis, Indiana 46202, USA.

Abstract:
Designing a protein sequence that will fold into a predefined structure is of both practical and fundamental interest. Many successful, computational designs in the last decade resulted from improved understanding of hydrophobic and polar interactions between side chains of amino acid residues in stabilizing protein tertiary structures. However, the coupling between main-chain backbone structure and local sequence has yet to be fully addressed. Here, we attempt to account for such coupling by using a sequence profile derived from the sequences of five residue fragments in a fragment library that are structurally matched to the five-residue segments contained in a target structure. We further introduced a term to reduce low complexity regions of designed sequences. These two terms together with optimized reference states for amino-acid residues were implemented in the RosettaDesign program. The new method, called RosettaDesign-SR, makes a 12% increase (from 34 to 46%) in fraction of proteins whose designed sequences are more than 35% identical to wild-type sequences. Meanwhile, it reduces 8% (from 22% to 14%) to the number of designed sequences that are not homologous to any known protein sequences according to psi-blast. More importantly, the sequences designed by RosettaDesign-SR have 2-3% more polar residues at the surface and core regions of proteins and these surface and core polar residues have about 4% higher sequence identity to wild-type sequences than by RosettaDesign. Thus, the proteins designed by RosettaDesign-SR should be less likely to aggregate and more likely to have unique structures due to more specific polar interactions.
More Related Videos


Related Concept Videos
 01:24
Protein Organization
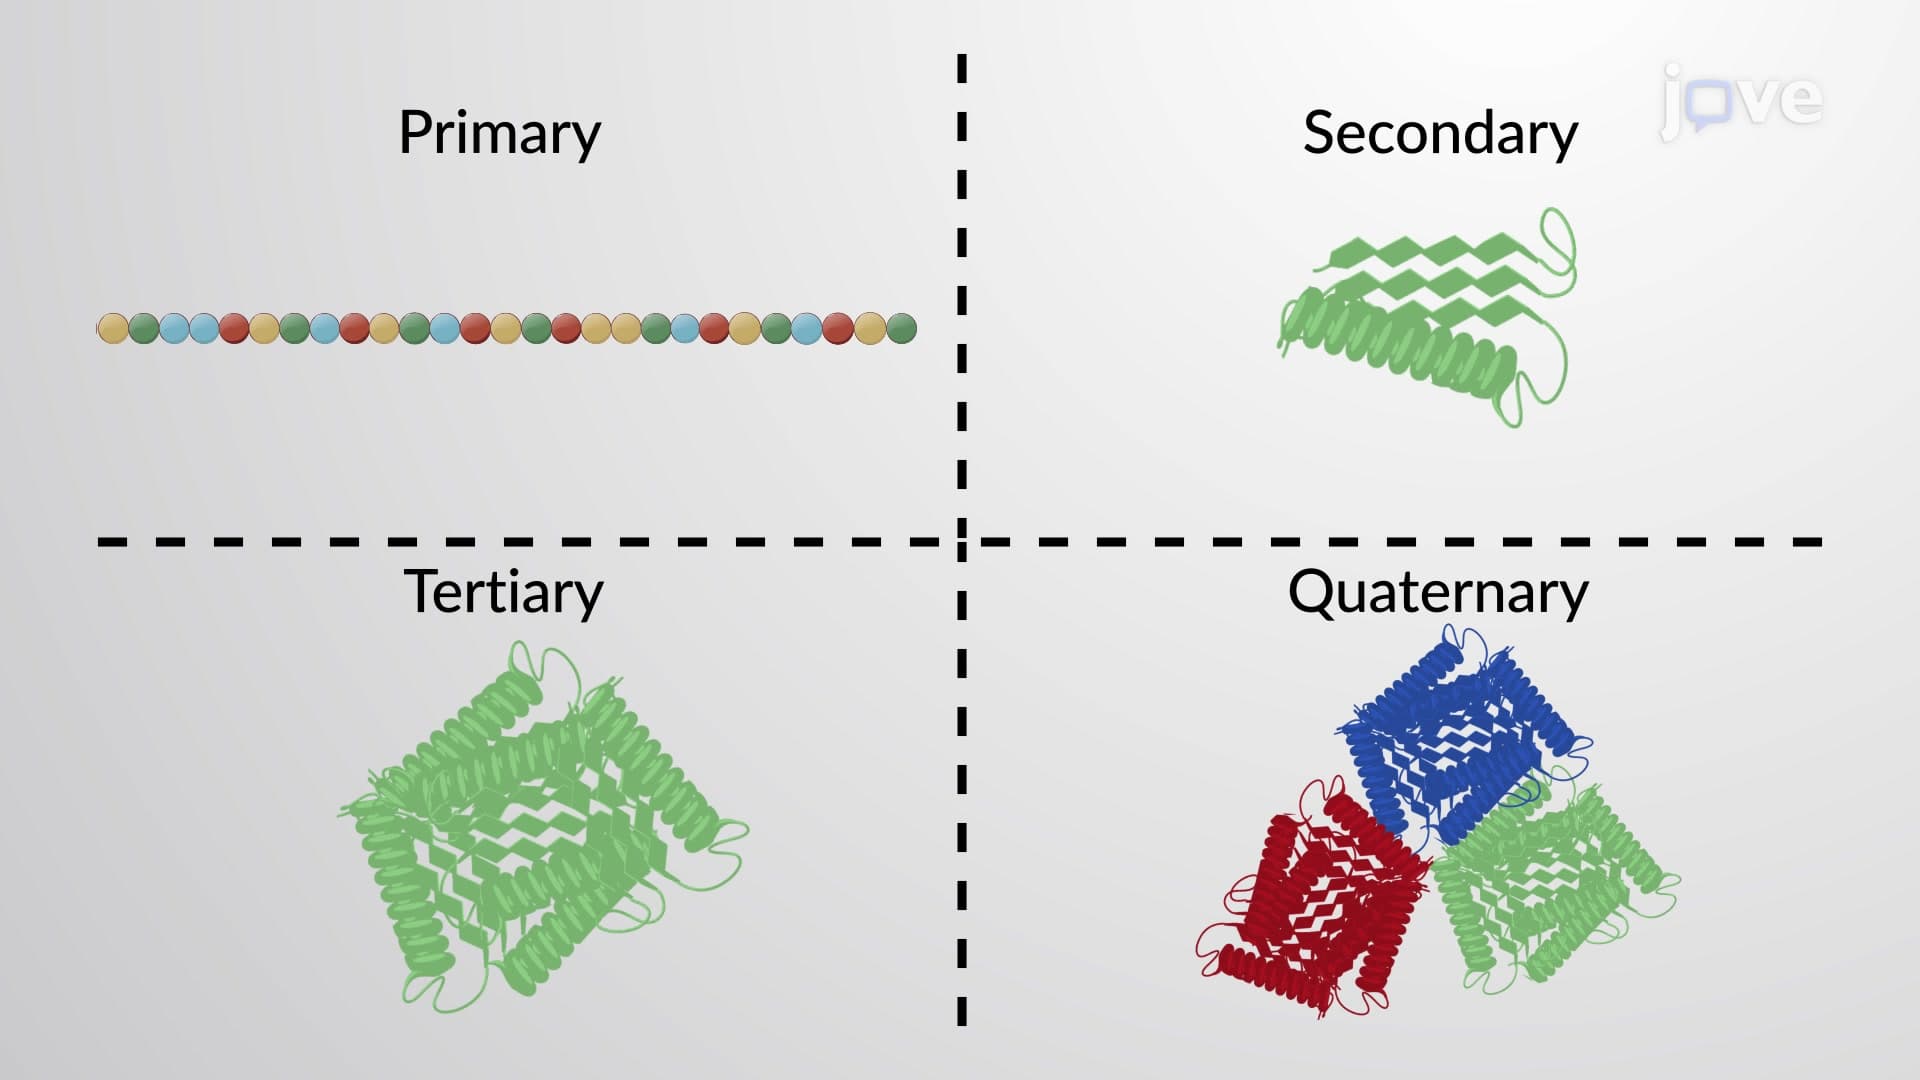
The primary structure of a protein is its amino acid sequence.
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:47
Protein Families
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 02:15
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme can...
 02:04
Protein-protein Interfaces