Related Experiment Video
Updated: Jun 12, 2026

An Intestine/Liver Microphysiological System for Drug Pharmacokinetic and Toxicological Assessment
Published on: December 3, 2020
Prediction of human blood-to-plasma drug concentration ratio
Takahide Uchimura1, Motohiro Kato, Tomohisa Saito
1Clinical Pharmacology Department, Chugai Pharmaceutical Co., Ltd, Kamakura, Kanagawa, Japan. uchimuratkh@chugai-pharm.co.jp <uchimuratkh@chugai-pharm.co.jp>

Abstract:
The objective of this study was to predict R(b) (blood/plasma ratio) in humans using a simple method. Human and rat R(b) and free fraction in plasma (f(p)) values were obtained from the literature. The ratio of total red blood cell concentration to the free concentration in plasma (K(b)) was calculated using f(p) and R(b). Four methods were used for the prediction of R(b): (A) use of rat R(b); (B) use of R(b) calculated from rat K(b) and human f(p); (C) correlation of human log ((1-f(p))/f(p)) and human log K(b); and (D) correlation of log D with human log K(b). The R(b) of 96 compounds in humans ranged from 0.52 to 2.00, with an average of 0.89. A significant correlation was observed among human log K(b), human log ((1-f(p))/f(p)), and log D; however, no obvious correlation was observed among human R(b), human log ((1-f(p))/f(p)), and log D. The errors within 1.25-fold for methods A-D were 68.3%, 77.6%, 61.5% and 64.8%, respectively. All predictive methods considered here were superior to the use of the average value of human R(b) or R(b)=1. Rat R(b) corrected by human f(p) improved the accuracy of the prediction. Method B was the most accurate of the four methods.
Related Concept Videos
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:29
Drug Distribution: Plasma Protein Binding
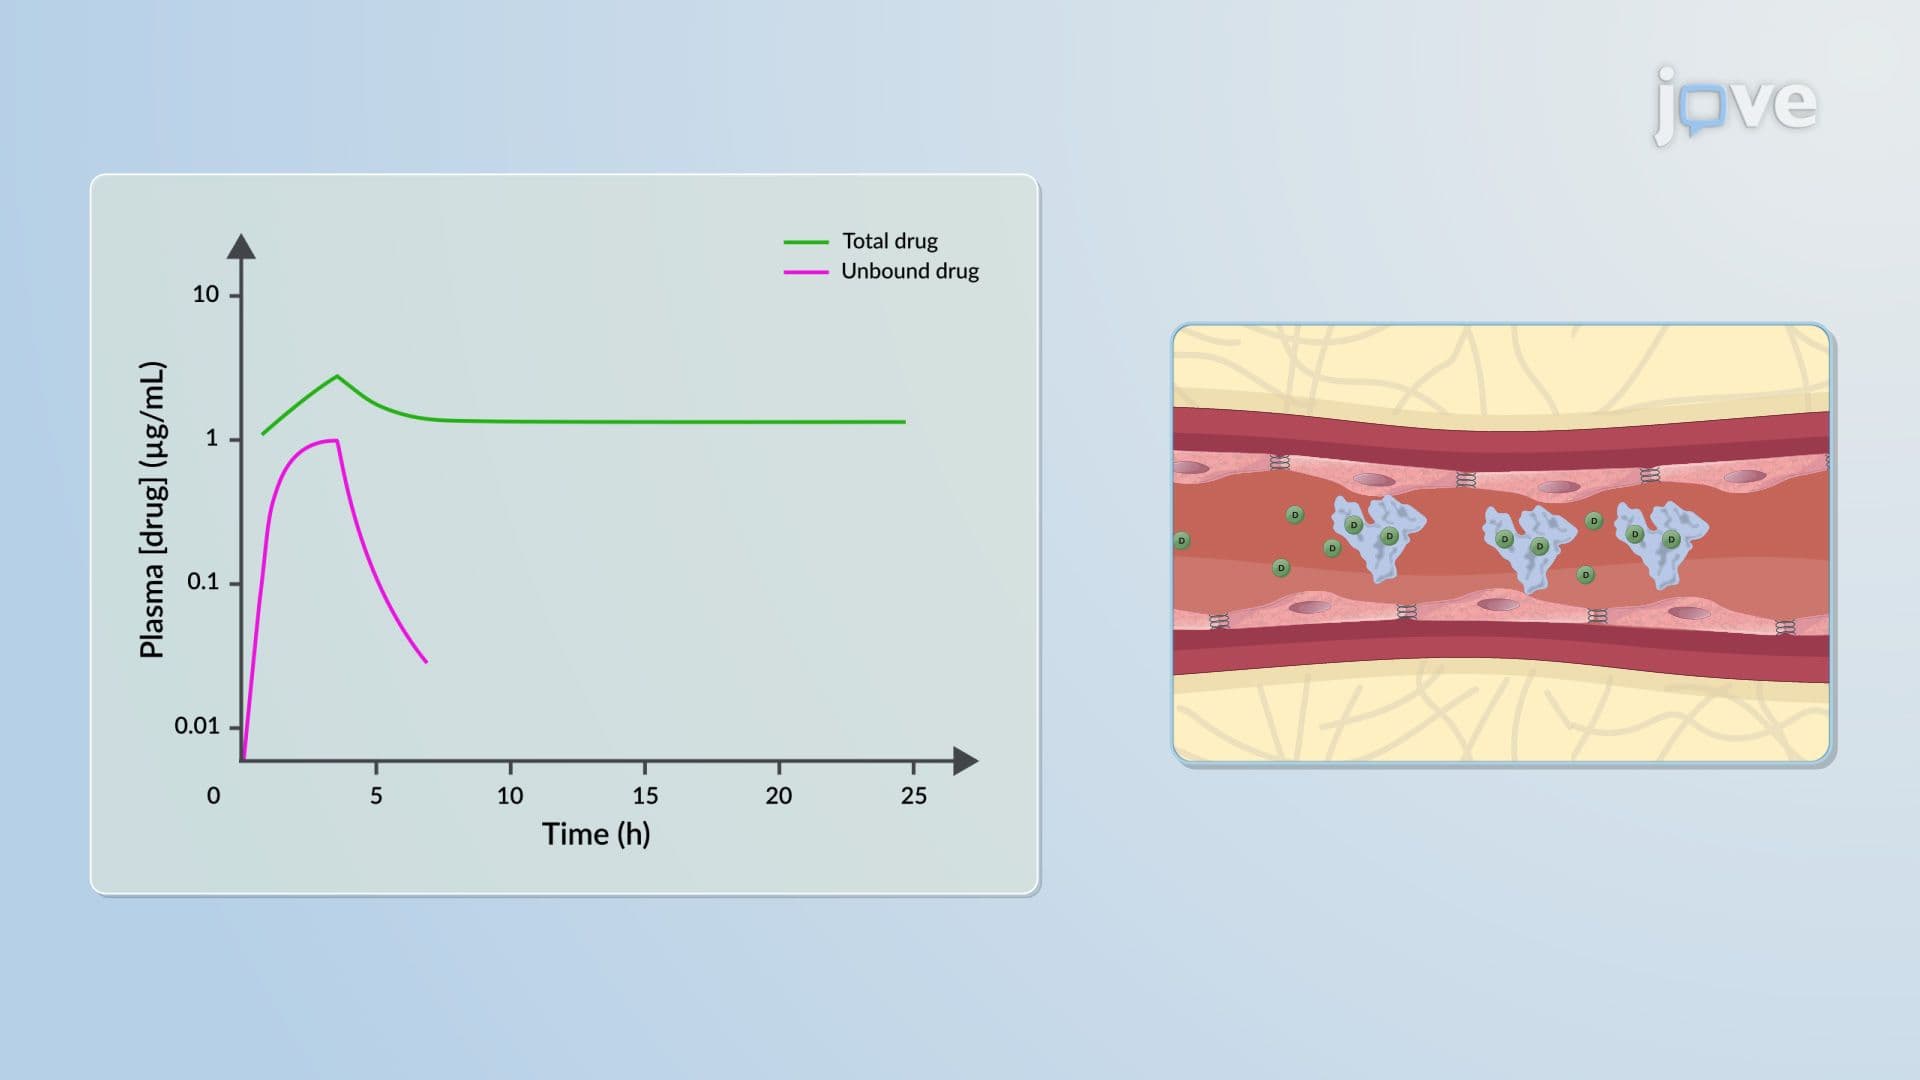 01:24
Pharmacokinetic–Pharmacodynamic Relationship: Problems
 01:01
Dosage Regimens: Partial Pharmacokinetic Parameters
 01:14
Pharmacokinetic–Pharmacodynamic Relationship: Model Components
 01:15
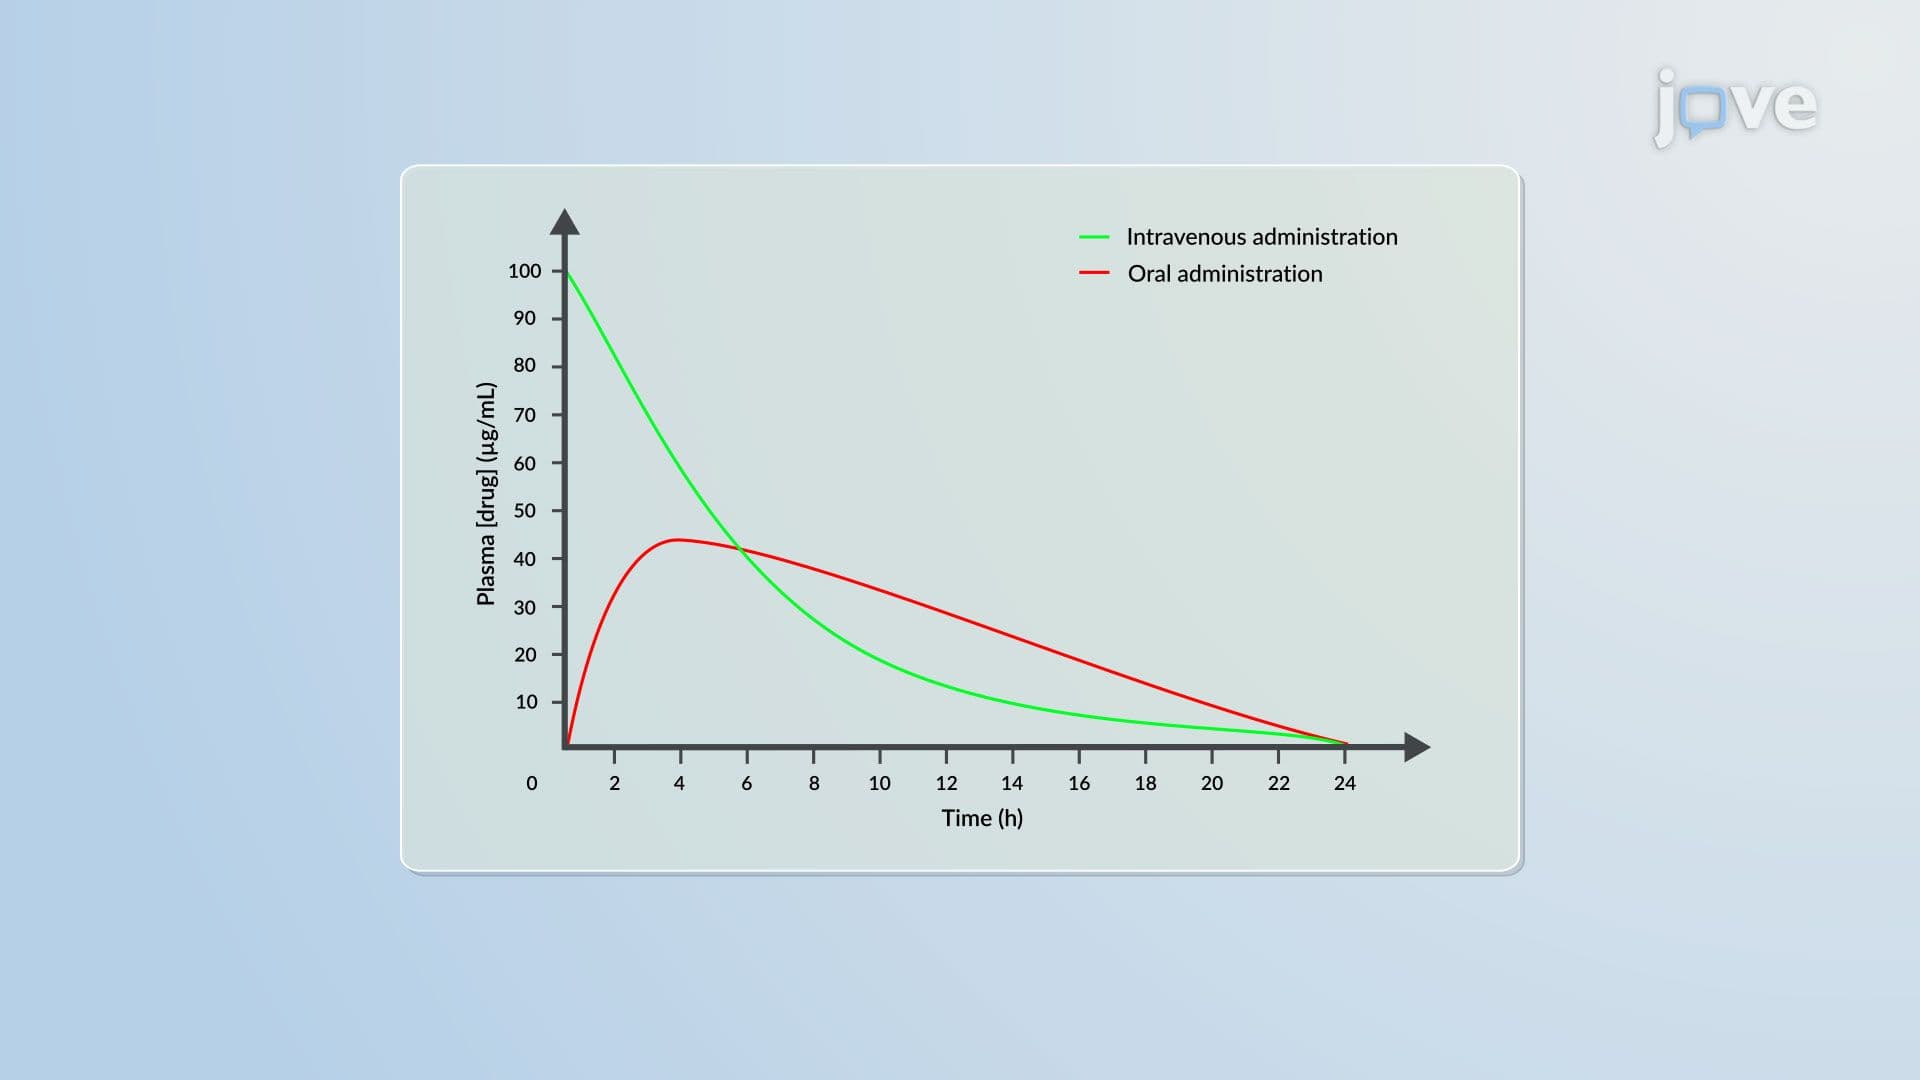
Drug Concentration Versus Time Correlation
Two pivotal parameters are the minimum effective concentration (MEC) and the minimum toxic concentration (MTC). The MEC is the lowest drug...