Related Experiment Video
Updated: Jun 12, 2026
Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
Stable tRNA-based phylogenies using only 76 nucleotides
Jeremy Widmann1, J Kirk Harris, Catherine Lozupone
1Department of Chemistry and Biochemistry, University of Colorado, Boulder, CO 80309, USA

Abstract:
tRNAs are among the most ancient, highly conserved sequences on earth, but are often thought to be poor phylogenetic markers because they are short, often subject to horizontal gene transfer, and easily change specificity. Here we use an algorithm now commonly used in microbial ecology, UniFrac, to cluster 175 genomes spanning all three domains of life based on the phylogenetic relationships among their complete tRNA pools. We find that the overall pattern of similarities and differences in the tRNA pools recaptures universal phylogeny to a remarkable extent, and that the resulting tree is similar to the distribution of bootstrapped rRNA trees from the same genomes. In contrast, the trees derived from tRNAs of identical specificity or of individual isoacceptors generally produced trees of lower quality. However, some tRNA isoacceptors were very good predictors of the overall pattern of organismal evolution. These results show that UniFrac can extract meaningful biological patterns from even phylogenies with high level of statistical inaccuracy and horizontal gene transfer, and that, overall, the pattern of tRNA evolution tracks universal phylogeny and provides a background against which we can test hypotheses about the evolution of individual isoacceptors.
Related Concept Videos
 02:54
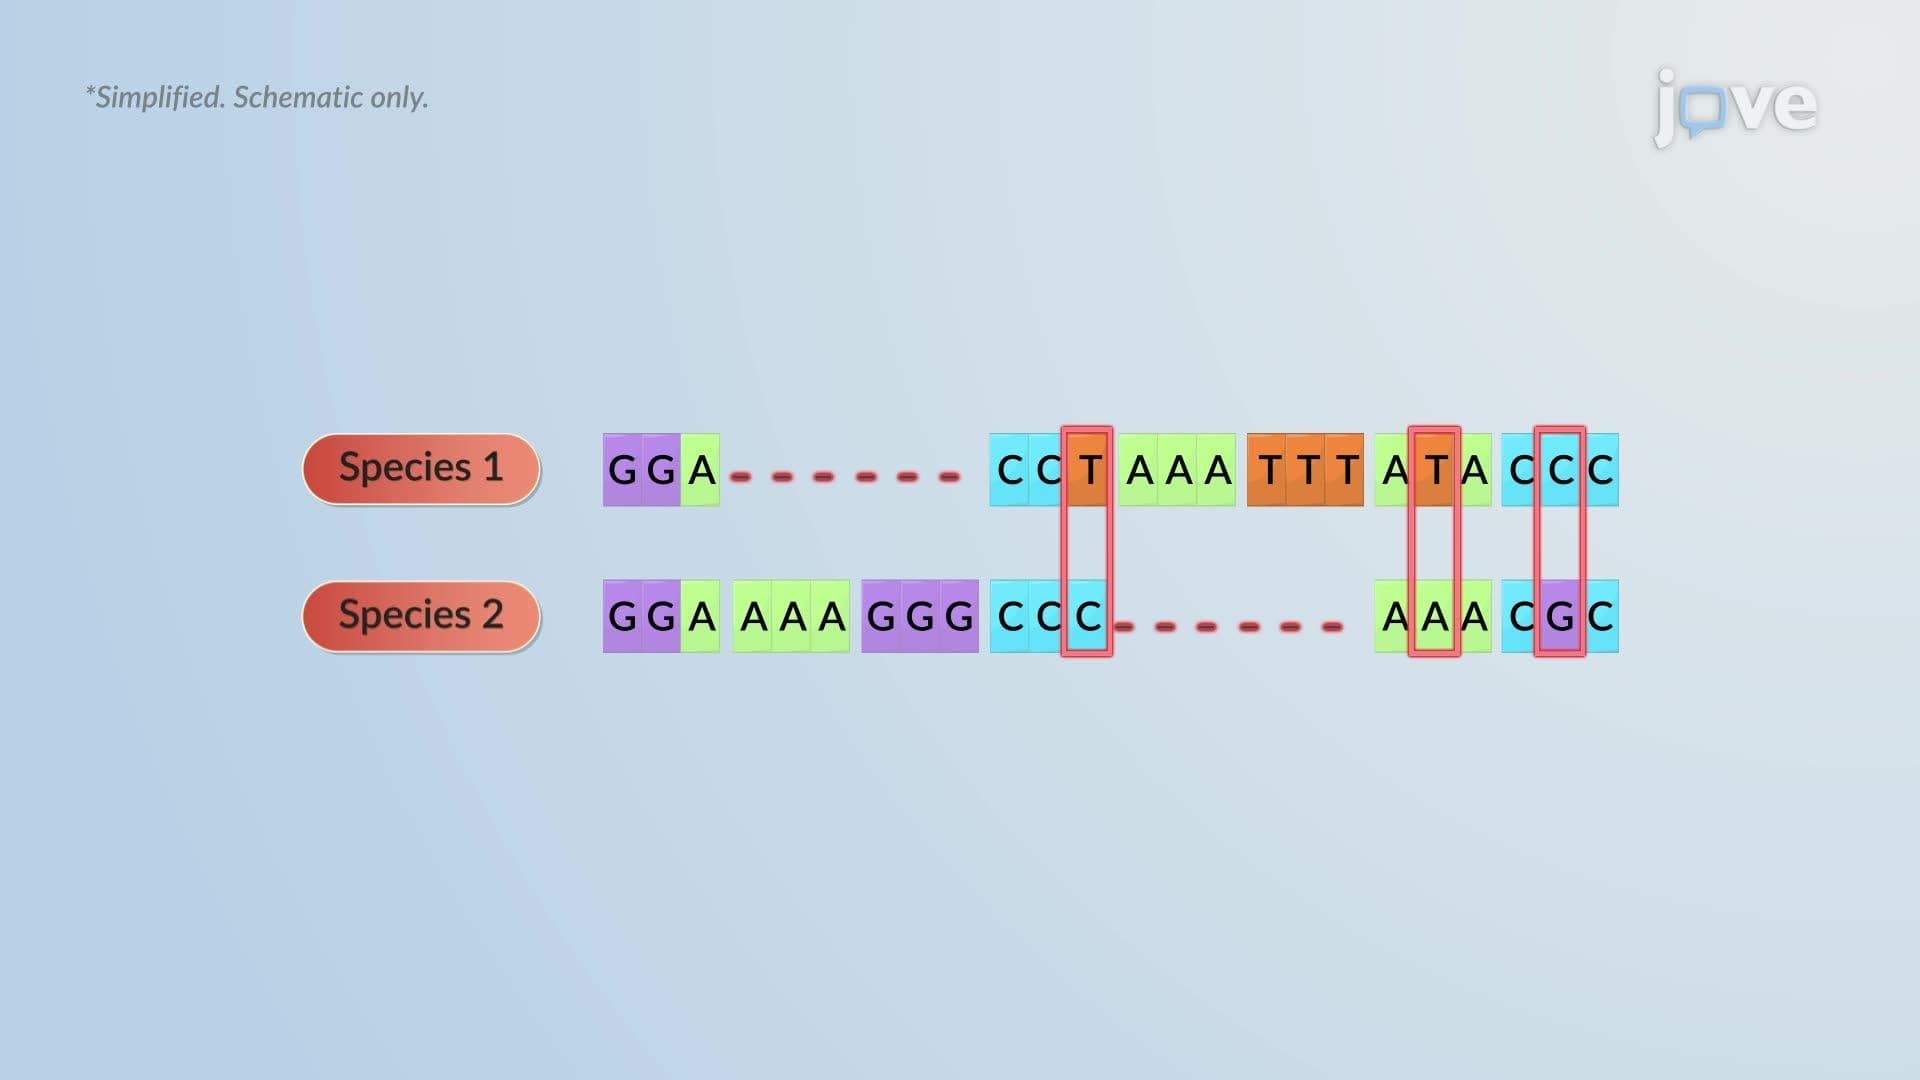
Evolutionary Relationships through Genome Comparisons
 01:28
Microbial Phylogeny
 03:21
Phylogenetic Trees
03:21Phylogenetic Trees
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...
02:05Gene Evolution - Fast or Slow?
In contrast, regions which code...