

Multimode simulation of dimer absorption spectra from first principles calculations: application to the
Julien Guthmuller1, Freddy Zutterman, Benoît Champagne
1Laboratoire de Chimie Théorique, Facultés Universitaires Notre-Dame de la Paix (FUNDP), Rue de Bruxelles 61, B-5000 Namur, Belgium. julien.guthmuller@uni-jena.de

Abstract:
First principles calculations based on density functional theory (DFT) have been combined with the multimode vibronic theory of coupled identical monomers to simulate the absorption spectra of dimers. In comparison to our previous study [J. Guthmuller et al., J. Chem. Theory Comput. 4, 2094 (2008)], where the vibrational excitations strictly accompany the electronic excitations, the vibronic model has been generalized so that the vibronic basis set contains vibrational excitations for both the ground and the excited electronic states. As a matter of illustration, this approach has been applied to a perylenetetracarboxylic diimide dimer employing a fixed dimer geometry. The exciton coupling energy is evaluated with time dependent DFT and random phase approximation calculations and by describing the effects of the solvent with the polarizable continuum model. First, the simulated monomer absorption spectrum is found to be in excellent agreement with experiment. Then, the simulated dimer absorption spectrum presents a strong dependency on the exciton coupling energy and on the inclusion of ground state vibrational excitations in the basis set. It is further shown that considering only fundamental vibrational excitations for the ground electronic state provides almost converged spectra and can therefore be used as a good first approximation. Moreover, the comparison with experiment demonstrates that the dimer absorption spectrum can be successfully reproduced by employing the exciton coupling energy determined at the time dependent DFT level provided that the effects of the solvent are included.
Related Concept Videos
 01:29
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:28
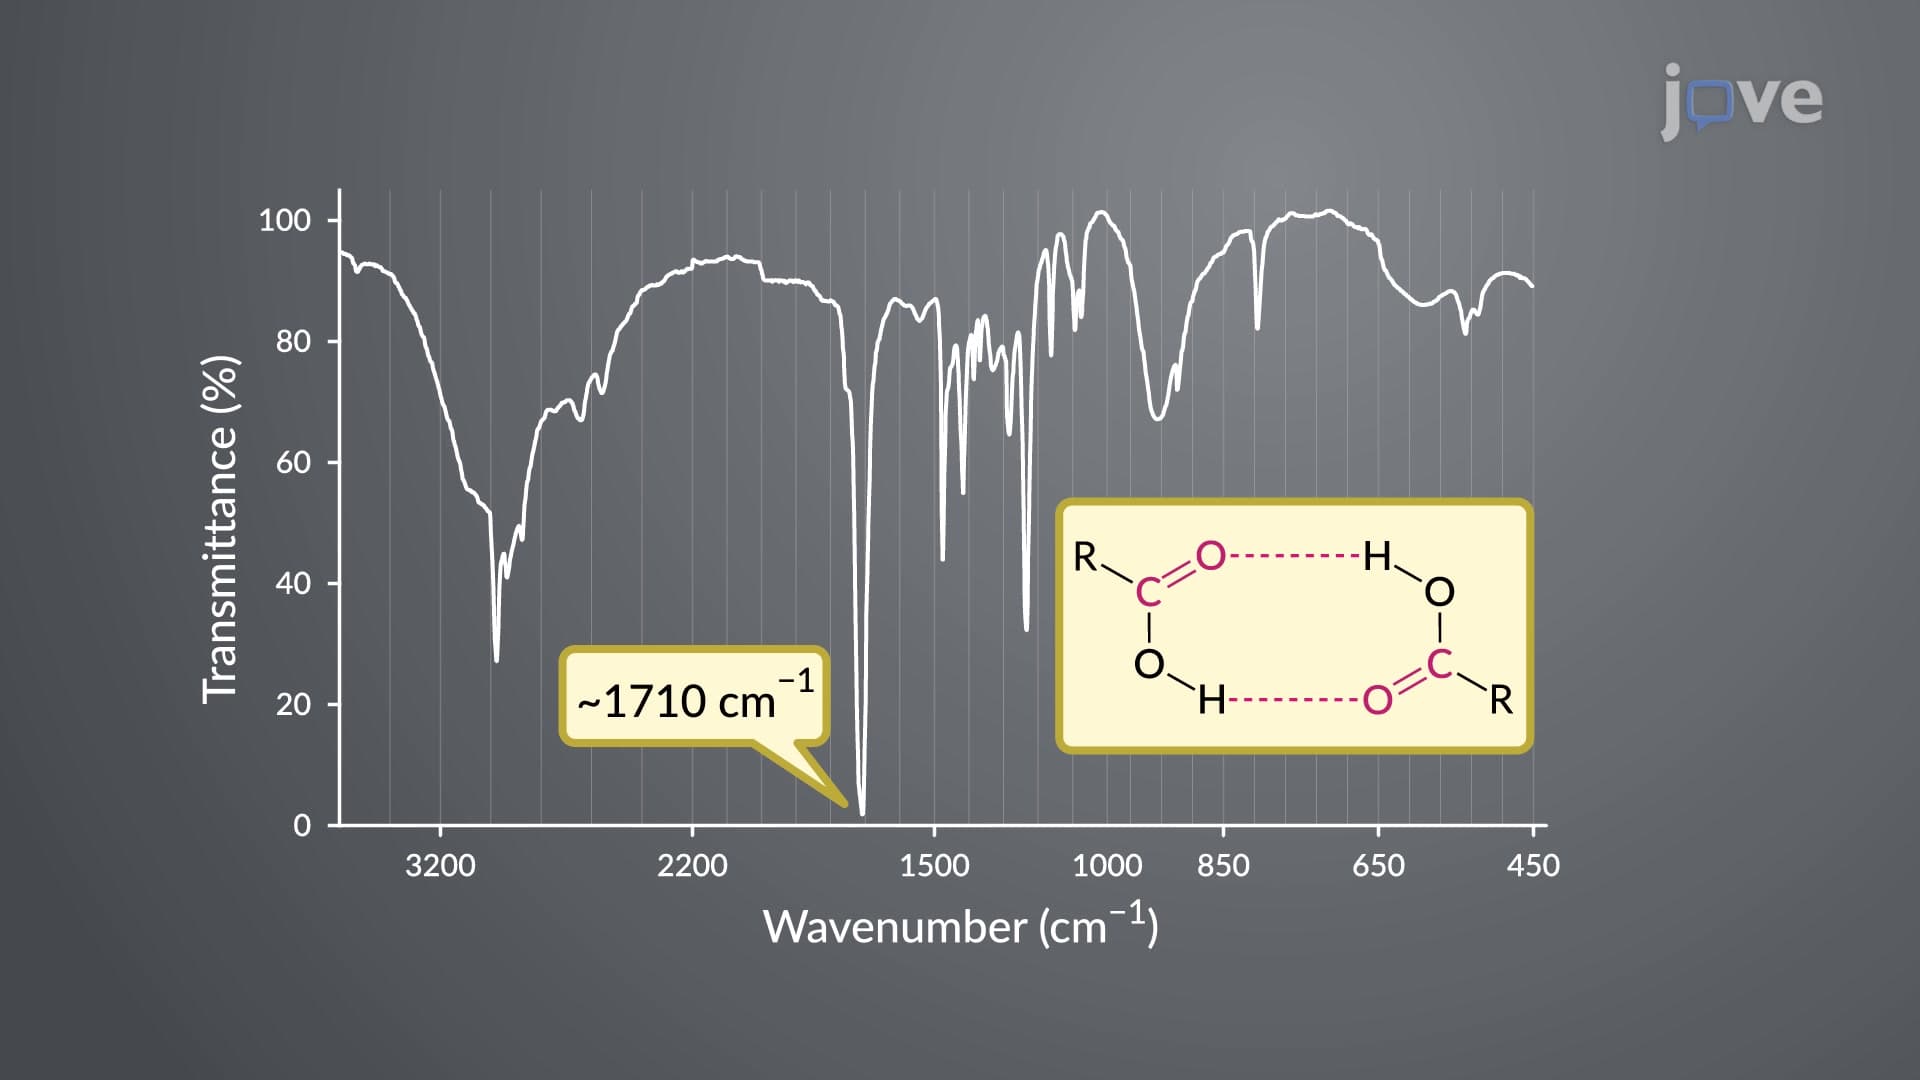
IR and UV–Vis Spectroscopy of Carboxylic Acids
However, the stretching absorptions for the C=O bond vary depending on the structure of carboxylic acids. The C=O bond of the free carboxylic acids shows a higher stretching frequency, 1760 cm−1, while H-bonded carboxylic acids (dimers) exhibit stretching absorptions at a lower frequency, 1710 cm−1. The C=O bond of the...
 01:32
UV–Vis Spectroscopy of Conjugated Systems
One of the factors influencing λmax is the extent of conjugation in the...
 01:20
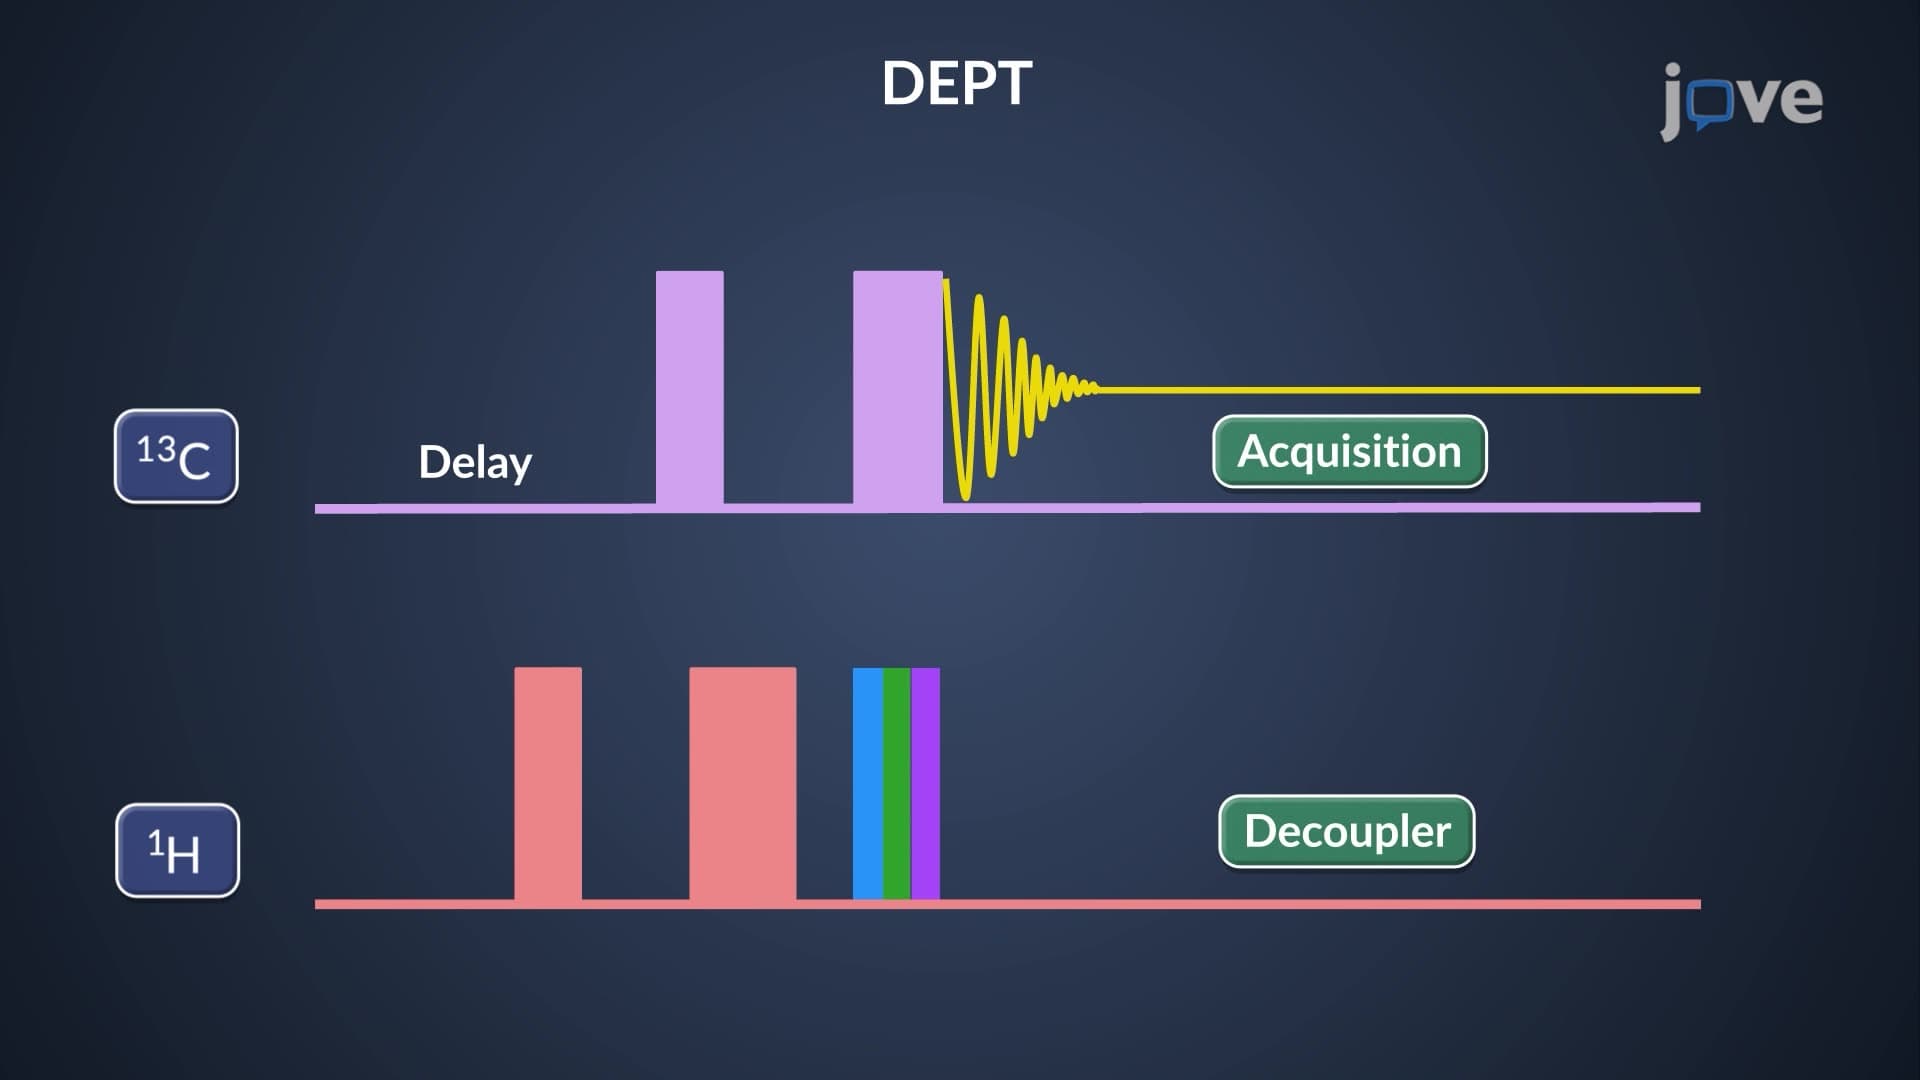
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:19
IR Spectrum
Transmittance is defined as the ratio of the radiant power passing through a sample to that from the radiation's source. Multiplying the transmittance by 100 gives the percent transmittance (%T), which varies between 100% (no absorption) and 0% (complete...
 01:20
IR Spectrum Peak Intensity: Dipole Moment