Related Experiment Video
Updated: Jun 11, 2026


Targeted Next-generation Sequencing and Bioinformatics Pipeline to Evaluate Genetic Determinants of Constitutional Disease
Published on: April 4, 2018
Genetic Creutzfeldt-Jakob disease associated with the E200K mutation: characterization of a complex proteinopathy
Gabor G Kovacs1, Jérémie Seguin, Isabelle Quadrio
1Institute of Neurology, Medical University of Vienna, and Austrian Reference Center for Human Prion Diseases, AKH 4J, Währinger Gürtel 18-20, 1097, Vienna, Austria. gabor.kovacs@meduniwien.ac.at

Abstract:
The E200K mutation is the most frequent prion protein gene (PRNP) mutation detected worldwide that is associated with Creutzfeldt-Jakob disease (CJD) and thought to have overlapping features with sporadic CJD, yet detailed neuropathological studies have not been reported. In addition to the prion protein, deposition of tau, α-synuclein, and amyloid-β has been reported in human prion disease. To describe the salient and concomitant neuropathological alterations, we performed a systematic clinical, neuropathological, and biochemical study of 39 individuals carrying the E200K PRNP mutation originating from different European countries. The most frequent clinical symptoms were dementia and ataxia followed by myoclonus and various combinations of further symptoms, including vertical gaze palsy and polyneuropathy. Neuropathological examination revealed relatively uniform anatomical pattern of tissue lesioning, predominating in the basal ganglia and thalamus, and also substantia nigra, while the deposition of disease-associated PrP was more influenced by the codon 129 constellation, including different or mixed types of PrP(res) detected by immunoblotting. Unique and prominent intraneuronal PrP deposition involving brainstem nuclei was also noted. Systematic examination of protein depositions revealed parenchymal amyloid-β in 53.8%, amyloid angiopathy (Aβ) in 23.1%, phospho-tau immunoreactive neuritic profiles in 92.3%, neurofibrillary degeneration in 38.4%, new types of tau pathology in 33.3%, and Lewy-type α-synuclein pathology in 15.4%. TDP-43 and FUS immunoreactive protein deposits were not observed. This is the first demonstration of intensified and combined neurodegeneration in a genetic prion disease due to a single point mutation, which might become an important model to decipher the molecular interplay between neurodegeneration-associated proteins.
Related Concept Videos
 01:21
Huntington Disease l: Introduction
 02:59
Animal Mitochondrial Genetics
 01:23
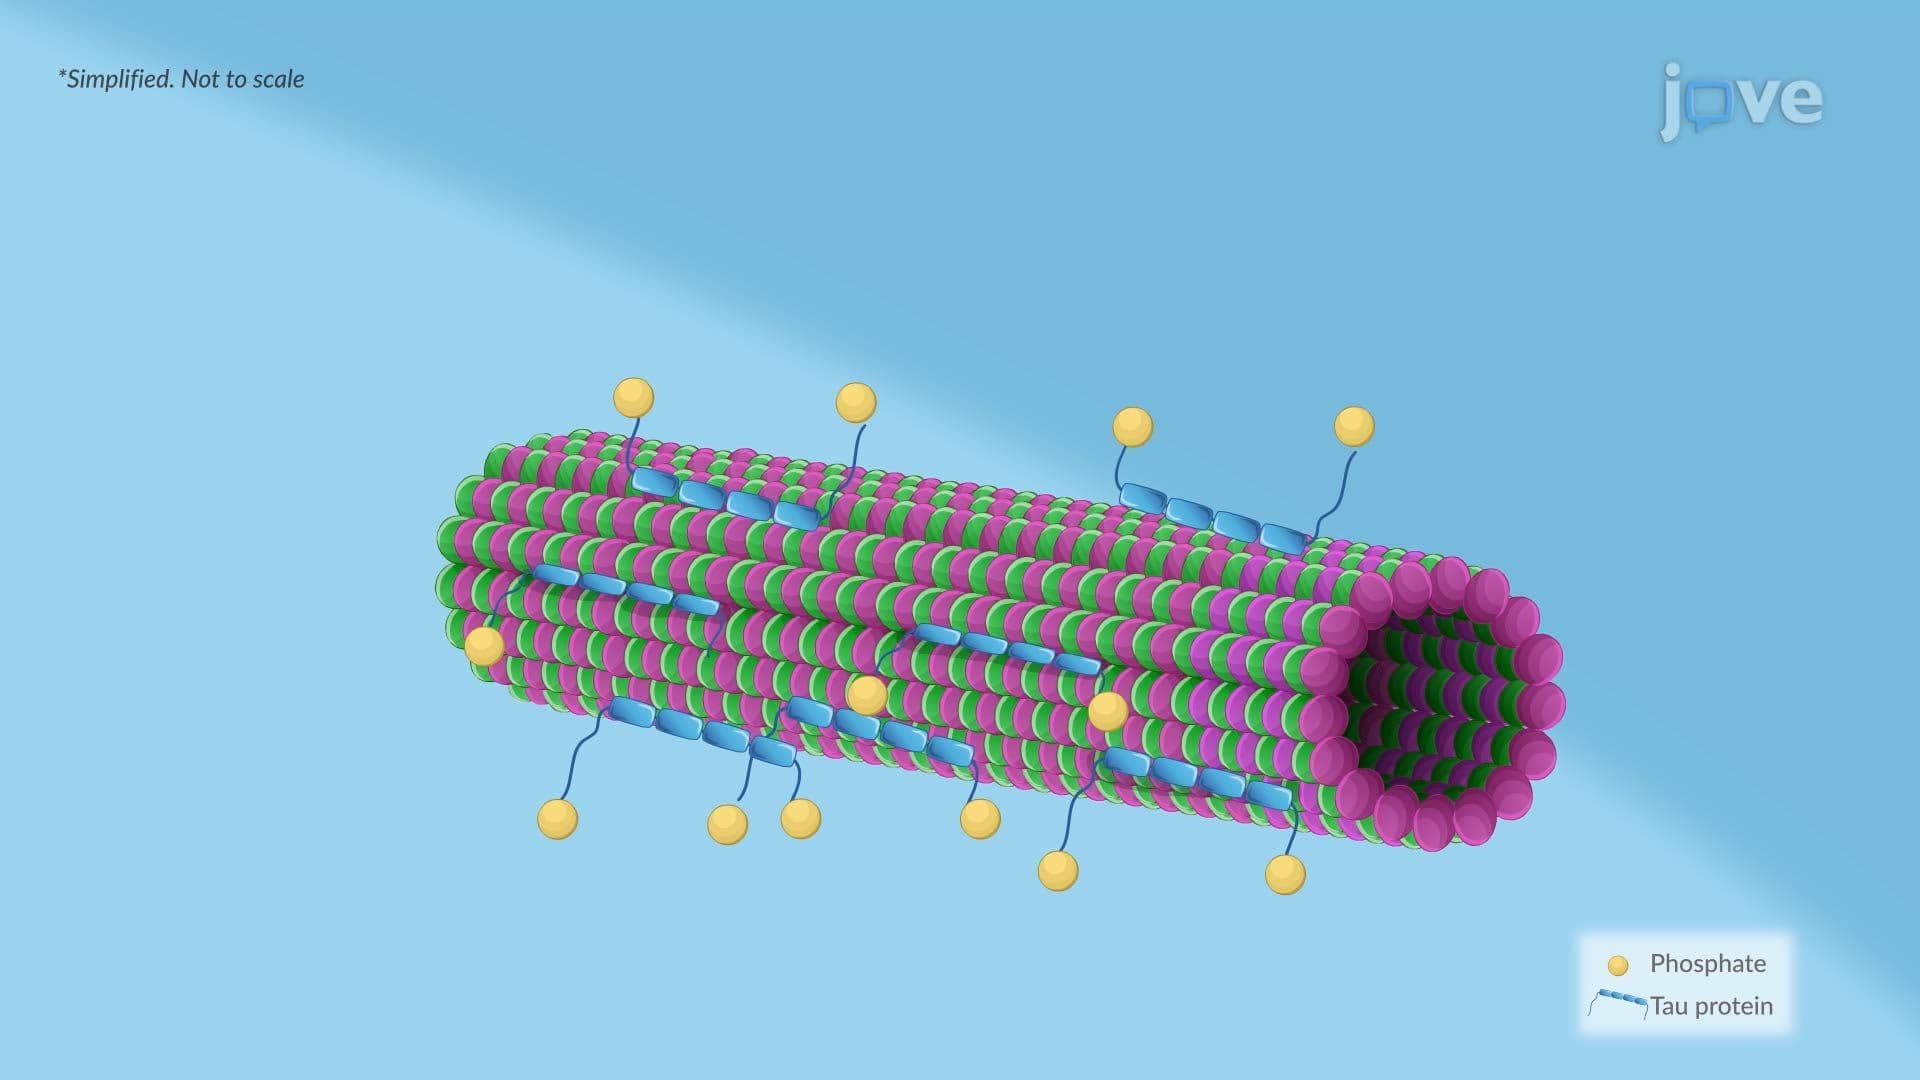
Alzheimer Disease ll: Pathophysiology
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
 01:24
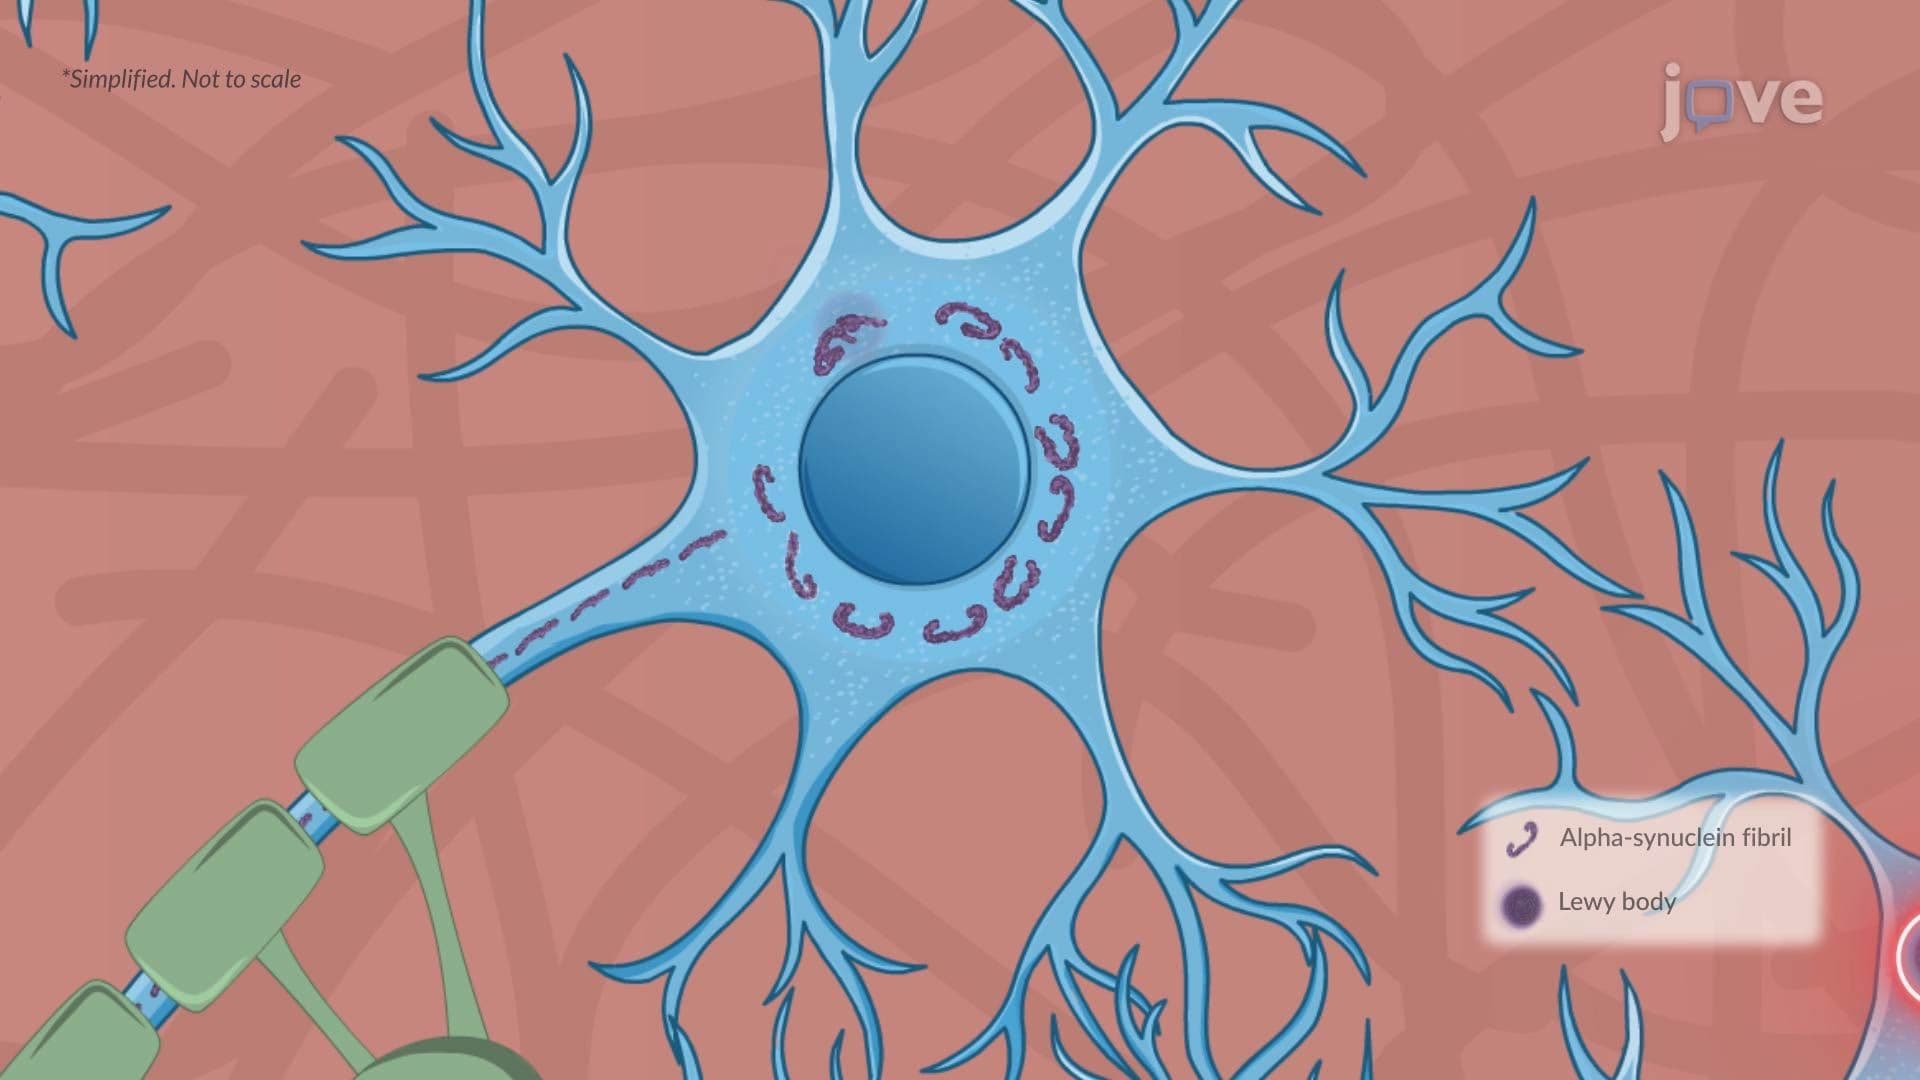
Parkinson Disease ll: Pathophysiology