Related Experiment Video
Updated: Jun 11, 2026

Analyzing Tumor and Tissue Distribution of Target Antigen Specific Therapeutic Antibody
Published on: May 16, 2020
Clinical pharmacokinetics of therapeutic monoclonal antibodies
Ron J Keizer1, Alwin D R Huitema, Jan H M Schellens
1Department of Pharmacy and Pharmacology, Netherlands Cancer Institute/Slotervaart Hospital, Amsterdam, the Netherlands. ron.keizer@slz.nl

Monoclonal antibodies (mAbs) have been used in the treatment of various diseases for over 20 years and combine high specificity with generally low toxicity. Their pharmacokinetic properties differ markedly from those of non-antibody-type drugs, and these properties can have important clinical implications. mAbs are administered intravenously, intramuscularly or subcutaneously. Oral administration is precluded by the molecular size, hydrophilicity and gastric degradation of mAbs. Distribution into tissue is slow because of the molecular size of mAbs, and volumes of distribution are generally low. mAbs are metabolized to peptides and amino acids in several tissues, by circulating phagocytic cells or by their target antigen-containing cells. Antibodies and endogenous immunoglobulins are protected from degradation by binding to protective receptors (the neonatal Fc-receptor [FcRn]), which explains their long elimination half-lives (up to 4 weeks). Population pharmacokinetic analyses have been applied in assessing covariates in the disposition of mAbs. Both linear and nonlinear elimination have been reported for mAbs, which is probably caused by target-mediated disposition. Possible factors influencing elimination of mAbs include the amount of the target antigen, immune reactions to the antibody and patient demographics. Bodyweight and/or body surface area are generally related to clearance of mAbs, but clinical relevance is often low. Metabolic drug-drug interactions are rare for mAbs. Exposure-response relationships have been described for some mAbs. In conclusion, the parenteral administration, slow tissue distribution and long elimination half-life are the most pronounced clinical pharmacokinetic characteristics of mAbs.
Monoclonal antibodies (mAbs) have been used in the treatment of various diseases for over 20 years and combine high specificity with generally low toxicity. Their pharmacokinetic properties differ markedly from those of non-antibody-type drugs, and these properties can have important clinical implications. mAbs are administered intravenously, intramuscularly or subcutaneously. Oral administration is precluded by the molecular size, hydrophilicity and gastric degradation of mAbs. Distribution into tissue is slow because of the molecular size of mAbs, and volumes of distribution are generally low. mAbs are metabolized to peptides and amino acids in several tissues, by circulating phagocytic cells or by their target antigen-containing cells. Antibodies and endogenous immunoglobulins are protected from degradation by binding to protective receptors (the neonatal Fc-receptor [FcRn]), which explains their long elimination half-lives (up to 4 weeks). Population pharmacokinetic analyses have been applied in assessing covariates in the disposition of mAbs. Both linear and nonlinear elimination have been reported for mAbs, which is probably caused by target-mediated disposition. Possible factors influencing elimination of mAbs include the amount of the target antigen, immune reactions to the antibody and patient demographics. Bodyweight and/or body surface area are generally related to clearance of mAbs, but clinical relevance is often low. Metabolic drug-drug interactions are rare for mAbs. Exposure-response relationships have been described for some mAbs. In conclusion, the parenteral administration, slow tissue distribution and long elimination half-life are the most pronounced clinical pharmacokinetic characteristics of mAbs.
More Related Videos
 08:53
In Vivo Immunofluorescence Localization for Assessment of Therapeutic and Diagnostic Antibody Biodistribution in Cancer Research
Published on: September 16, 2019
 11:58
Initial Evaluation of Antibody-conjugates Modified with Viral-derived Peptides for Increasing Cellular Accumulation and Improving Tumor Targeting
Published on: March 8, 2018
Related Concept Videos
 01:20
Measurement of Bioavailability: Pharmacodynamic Methods
 01:29
Drug Distribution: Plasma Protein Binding
 01:22
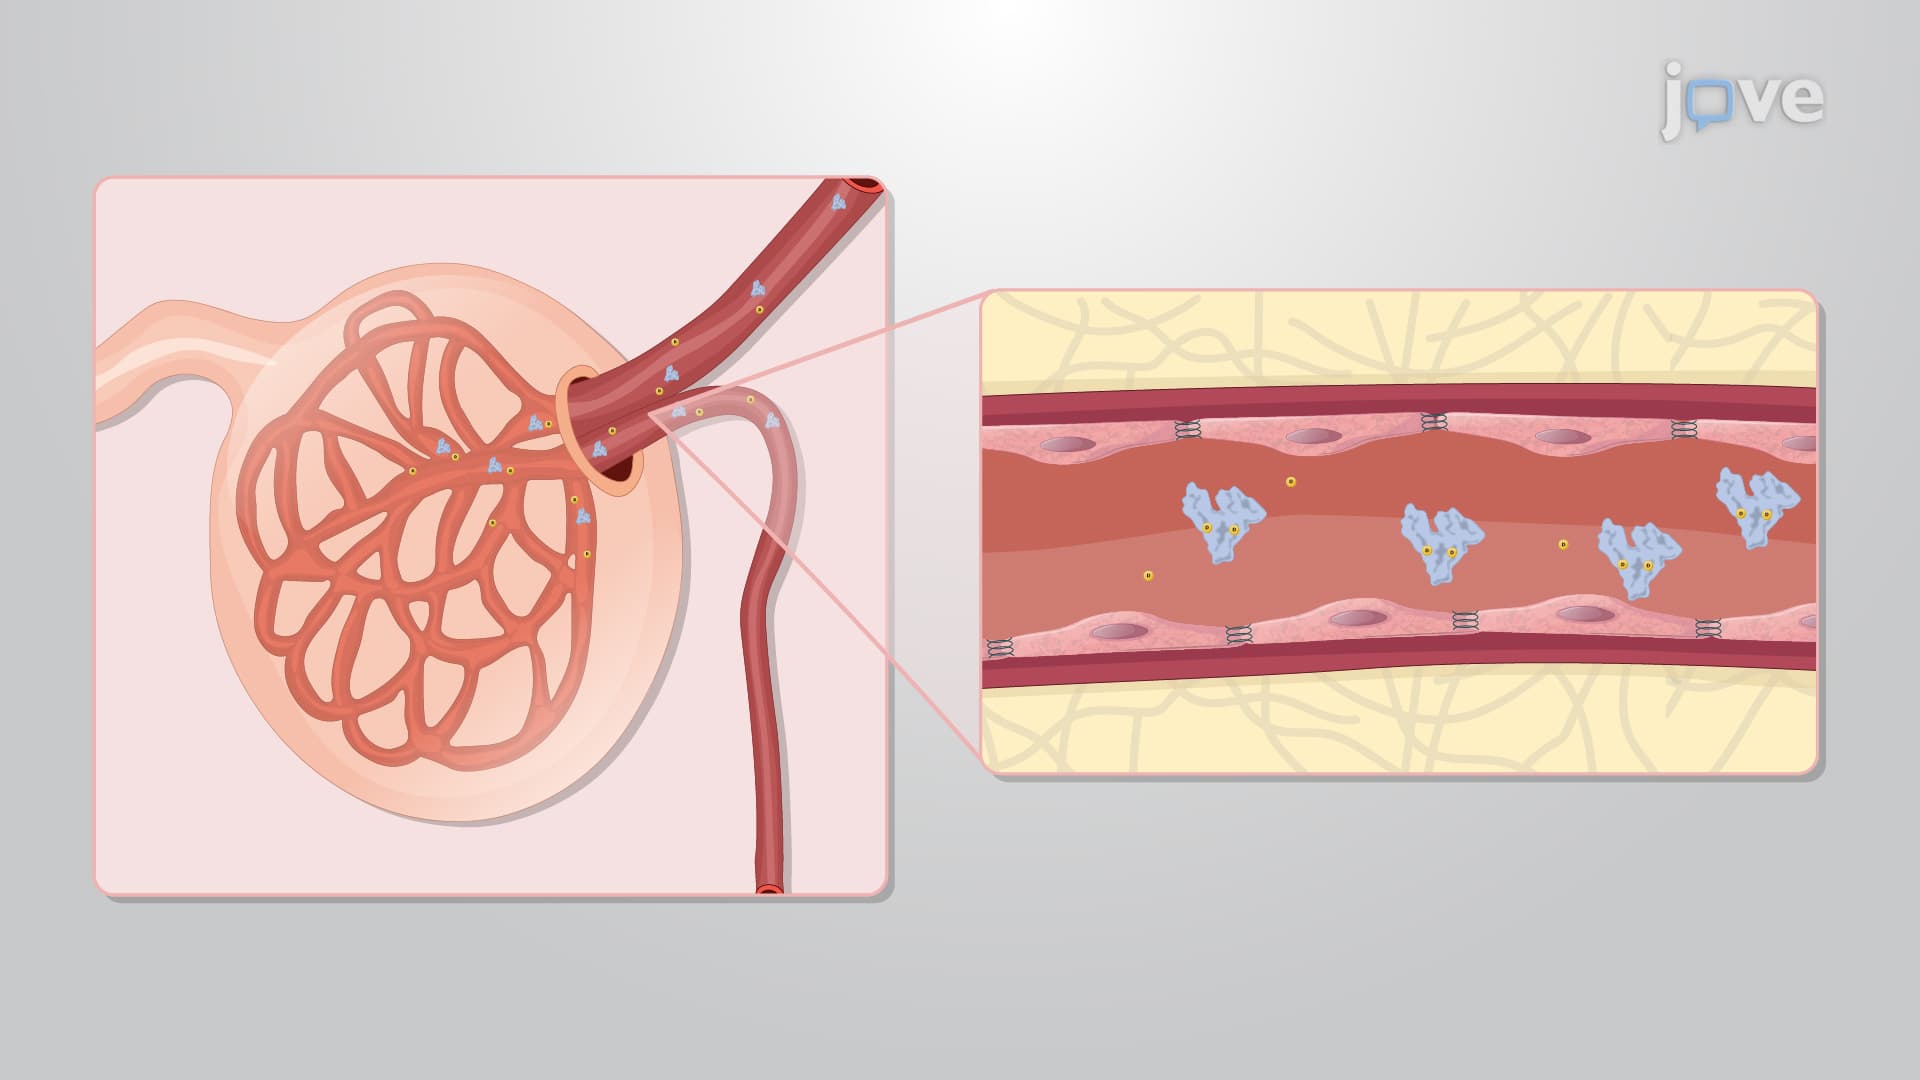
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
 01:24
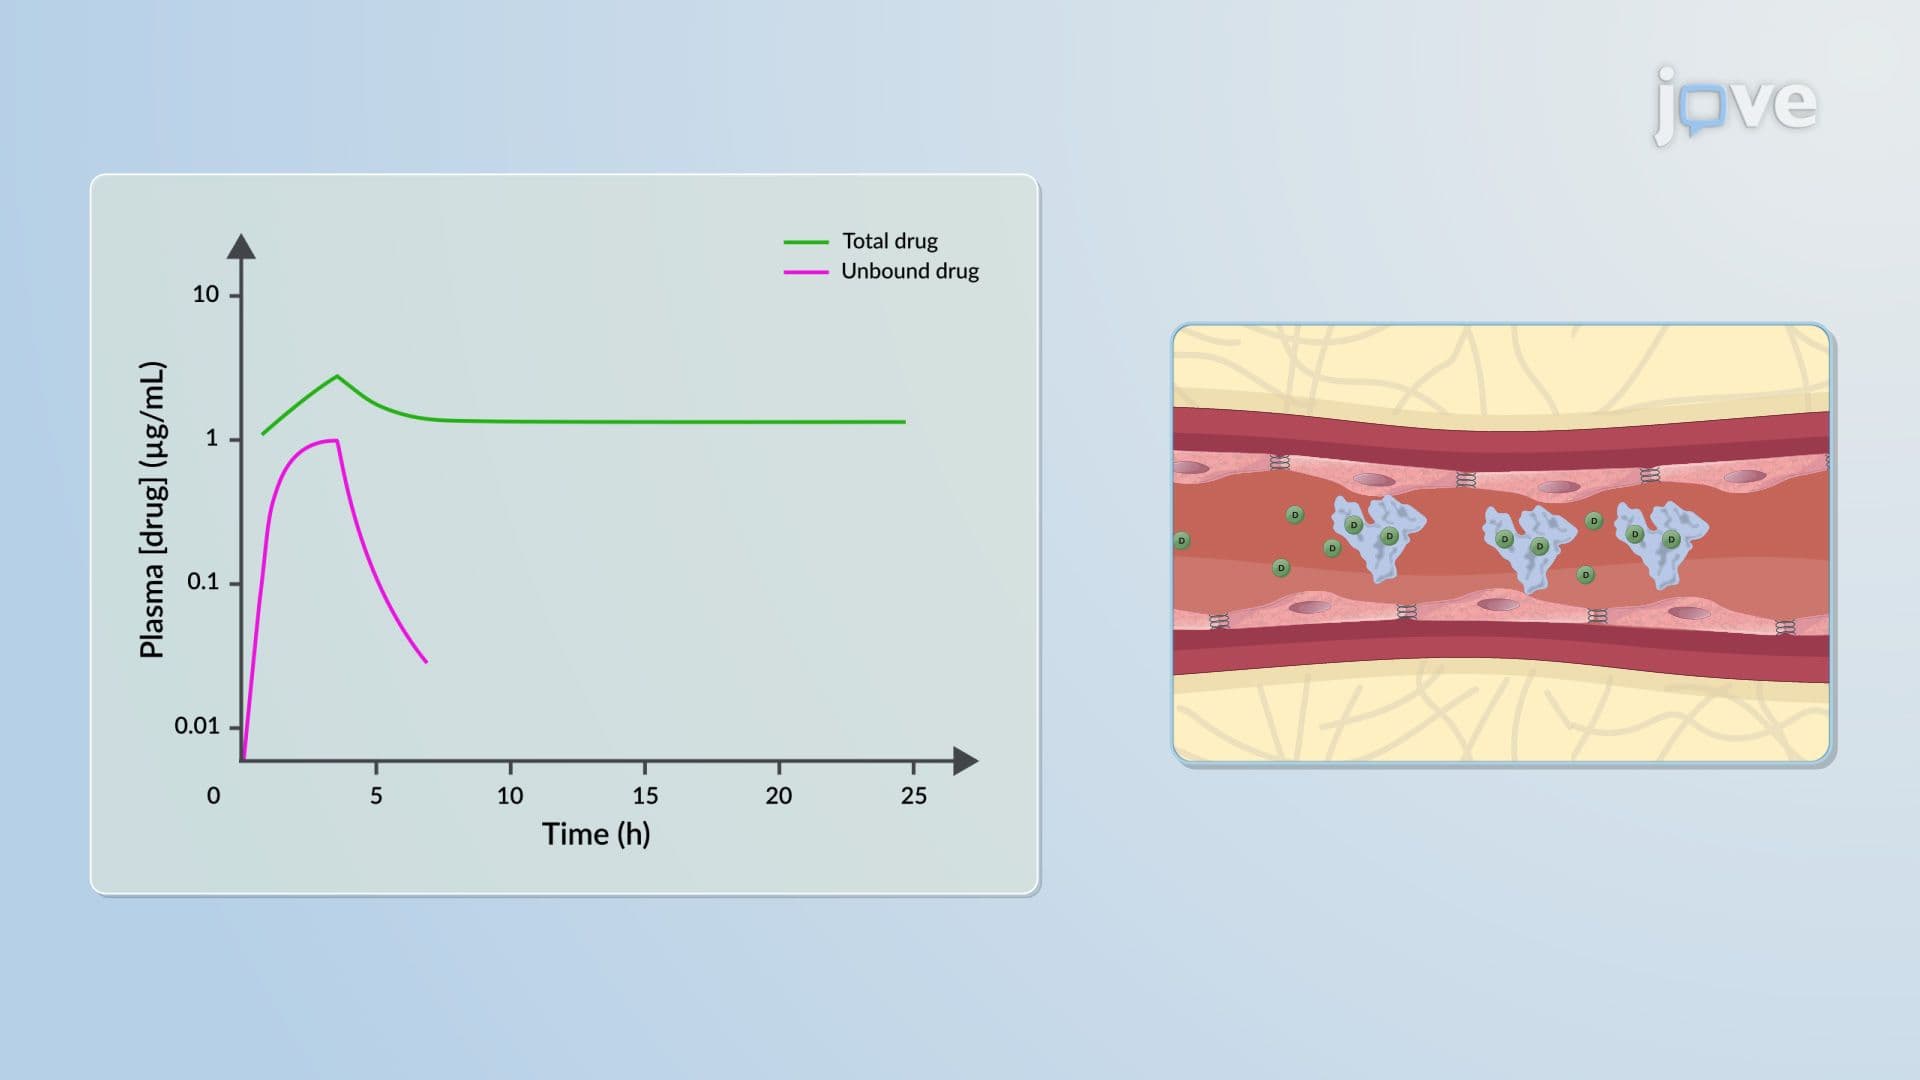
Pharmacokinetic–Pharmacodynamic Relationship: Problems
 01:30
Measurement of Bioavailability: Pharmacokinetic Methods
 01:15
Drug Concentration Versus Time Correlation
Two pivotal parameters are the minimum effective concentration (MEC) and the minimum toxic concentration (MTC). The MEC is the lowest drug...