Estimates of ligand-binding affinities supported by quantum mechanical methods
Related Experiment Video
Updated: Jun 10, 2026
Elucidation of the Material Basis of Yiqi Qingjie Formula Against IgA Nephropathy Using UHPLC-Q-Orbitrap HRMS Integrated with Network Pharmacology
Published on: May 19, 2026
Pär Söderhjelm1, Jacob Kongsted, Samuel Genheden
1Department of Theoretical Chemistry, Lund University, Chemical Centre, Lund, Sweden.

Abstract:
In this paper, we review our efforts to use quantum mechanical (QM) methods to improve free-energy estimates of the binding of drug candidates to their receptor proteins. First, we have tested the influence of various implicit solvation models on predictions of the ligand-binding affinity. The accuracy of implicit solvation models strongly depend on the parameterisation, but also on the magnitude of the solvation energy (i.e. their accuracy should be discussed in relative terms). However, if only relative solvation energies within a series of similar drug molecules with the same net charge are considered, nearly all methods tested give a comparable accuracy of 2-5 kJ/mol. Second, we have studied the conformational dependence of QM charges and their influence on ligand-binding affinities. The conformational dependence is significant, but it is to a large extent cancelled by solvation energies. Third, we have estimated the effect and range of electrostatic interactions beyond a point-charge model. The results show that multipoles up to octupoles and anisotropic polarisabilities have a significant influence on energies for residues up to 10-15 A from the ligand and that different sets of point-charge models may give strongly varying results. However, if only relative energies are considered, the effect is to a large extent cancelled. Fourth, we have tried to develop an accurate QM-based molecular mechanics potential, in which not only the electrostatic terms are improved, but also the dispersion and repulsion. However, even with quite sophisticated expressions, it seems difficult to reduce the average error below 2-3 kJ/mol per interaction (e.g. a hydrogen bond), compared to the full QM treatment. Finally, we have developed a new method, PMISP (polarised multipole interaction with supermolecular pairs), for the calculation of accurate interaction energies. It employs an accurate force field for electrostatics and induction, including multipoles up to octupoles and anisotropic polarisabilities calculated by QM methods on amino-acid fragments of the protein in each conformation observed in snapshots from a molecular dynamics simulation, whereas short-range interactions are estimated by high-level QM calculations for all pairs of the ligand with near-by residues. We show that this approach allows us to go far beyond the current accuracy of molecular mechanics methods, down to an error of 5-10 kJ/mol for a full protein-ligand complex. It can be combined with estimates of solvation, entropy, and dynamic effects to give estimates of binding affinities. However, several problems remain to be solved before any significant improvement in the accuracy can be seen.
Related Concept Videos
 02:18
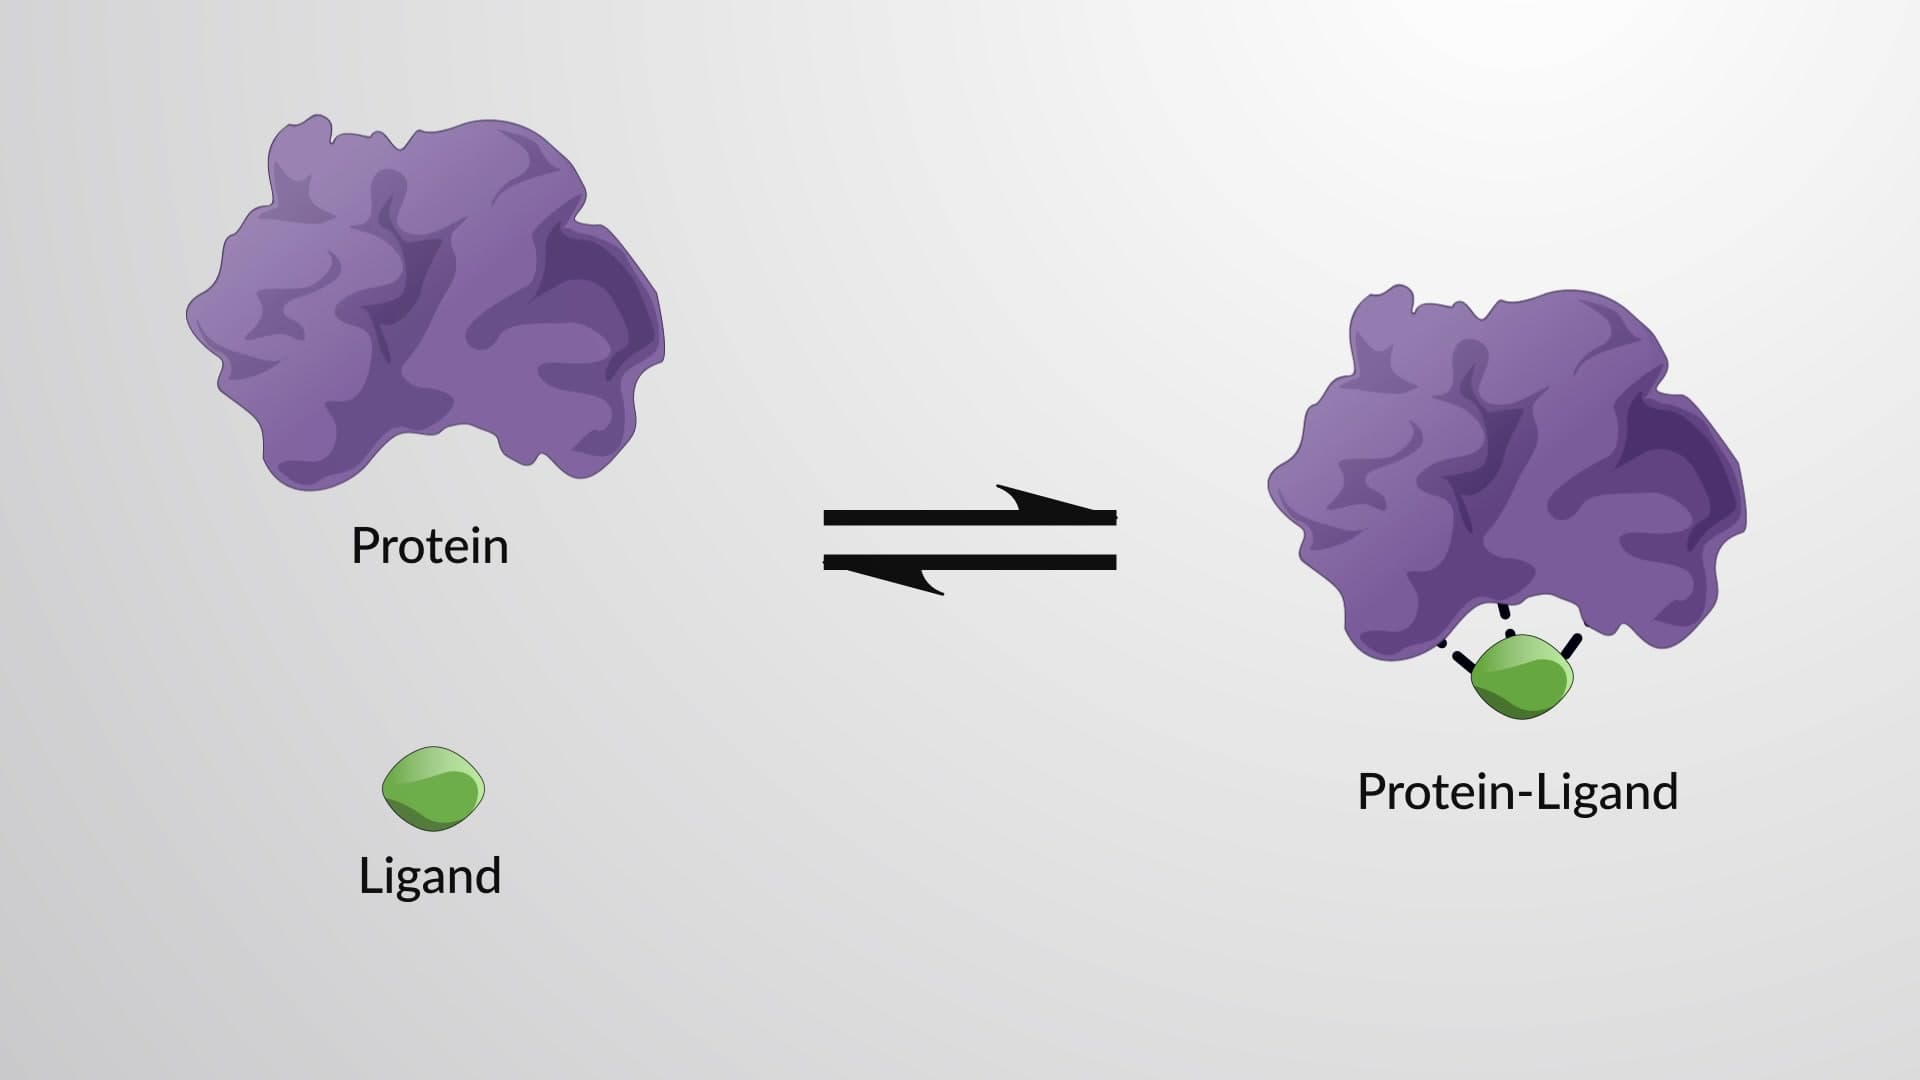
The Equilibrium Binding Constant and Binding Strength
02:18The Equilibrium Binding Constant and Binding Strength
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:30
Quantitative Aspects of Drug-Receptor Interaction