Related Experiment Video
Updated: Jun 10, 2026

Methods of Ex Situ and In Situ Investigations of Structural Transformations: The Case of Crystallization of Metallic Glasses
Published on: June 7, 2018
Calculation of anharmonic OH phonon dispersion curves for the Mg(OH)2 crystal
Pavlin D Mitev1, Kersti Hermansson, Wim J Briels
1Materials Chemistry, The Angström Laboratory, Uppsala University, Box 538, Uppsala S-75121, Sweden. pmitev@mkem.uu.se

Abstract:
Anharmonic OH phonon dispersion curves have been calculated for the Mg(OH)(2) crystal. A crystal Hamiltonian was set up for the vibrational problem, where the coordinates consists of the bond lengths of two hydroxide ions in the central unit cell. Its two-dimensional potential energy surface was constructed from first principle calculations within the density functional theory approximation. Dispersion curves were calculated by diagonalizing the Hamiltonian in a basis of singly excited crystal functions. The single particle functions used to construct the crystal states were taken from a Morse oscillator basis set. These well chosen functions made it possible to restrict calculations to include only very few functions, which greatly contributed to a transparent presentation of the underlying theory. All calculations could be done analytically except for the calculation of a few integrals. We have compared our results with those of a series of harmonic lattice dynamics calculations and have found that the anharmonicity shifts the IR and Raman dispersion curves downward appreciably and slightly changes the energy differences between both curves. From an analysis of the harmonic results we conclude that incorporating the coupling between OH stretching motion and the motion of their centers of mass will appreciably change the overall features of the dispersion curves. Extension of the anharmonic model along these lines will cause no problem to the theoretical approach presented in this paper.
Related Concept Videos
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:27
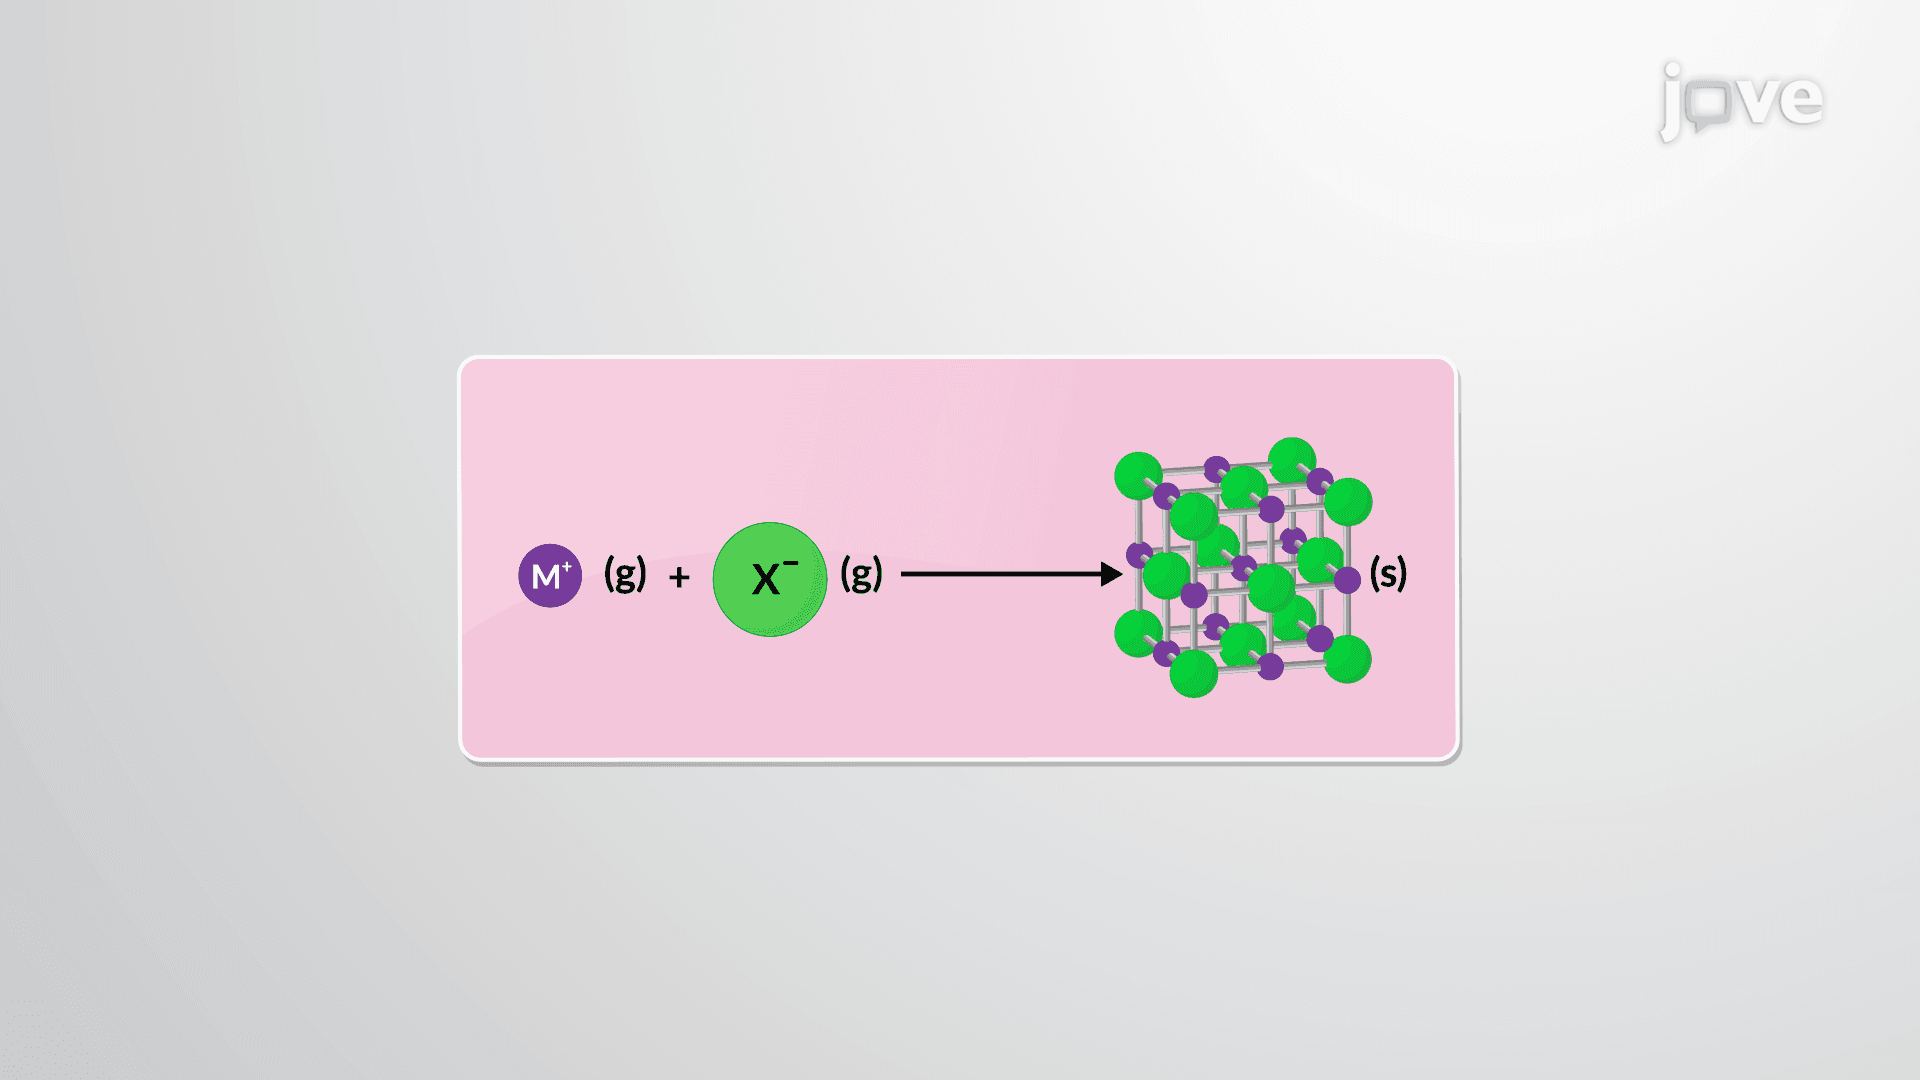
Lattice Energies of Ionic Crystals
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:29
Determination of Crystal Structures
 02:18
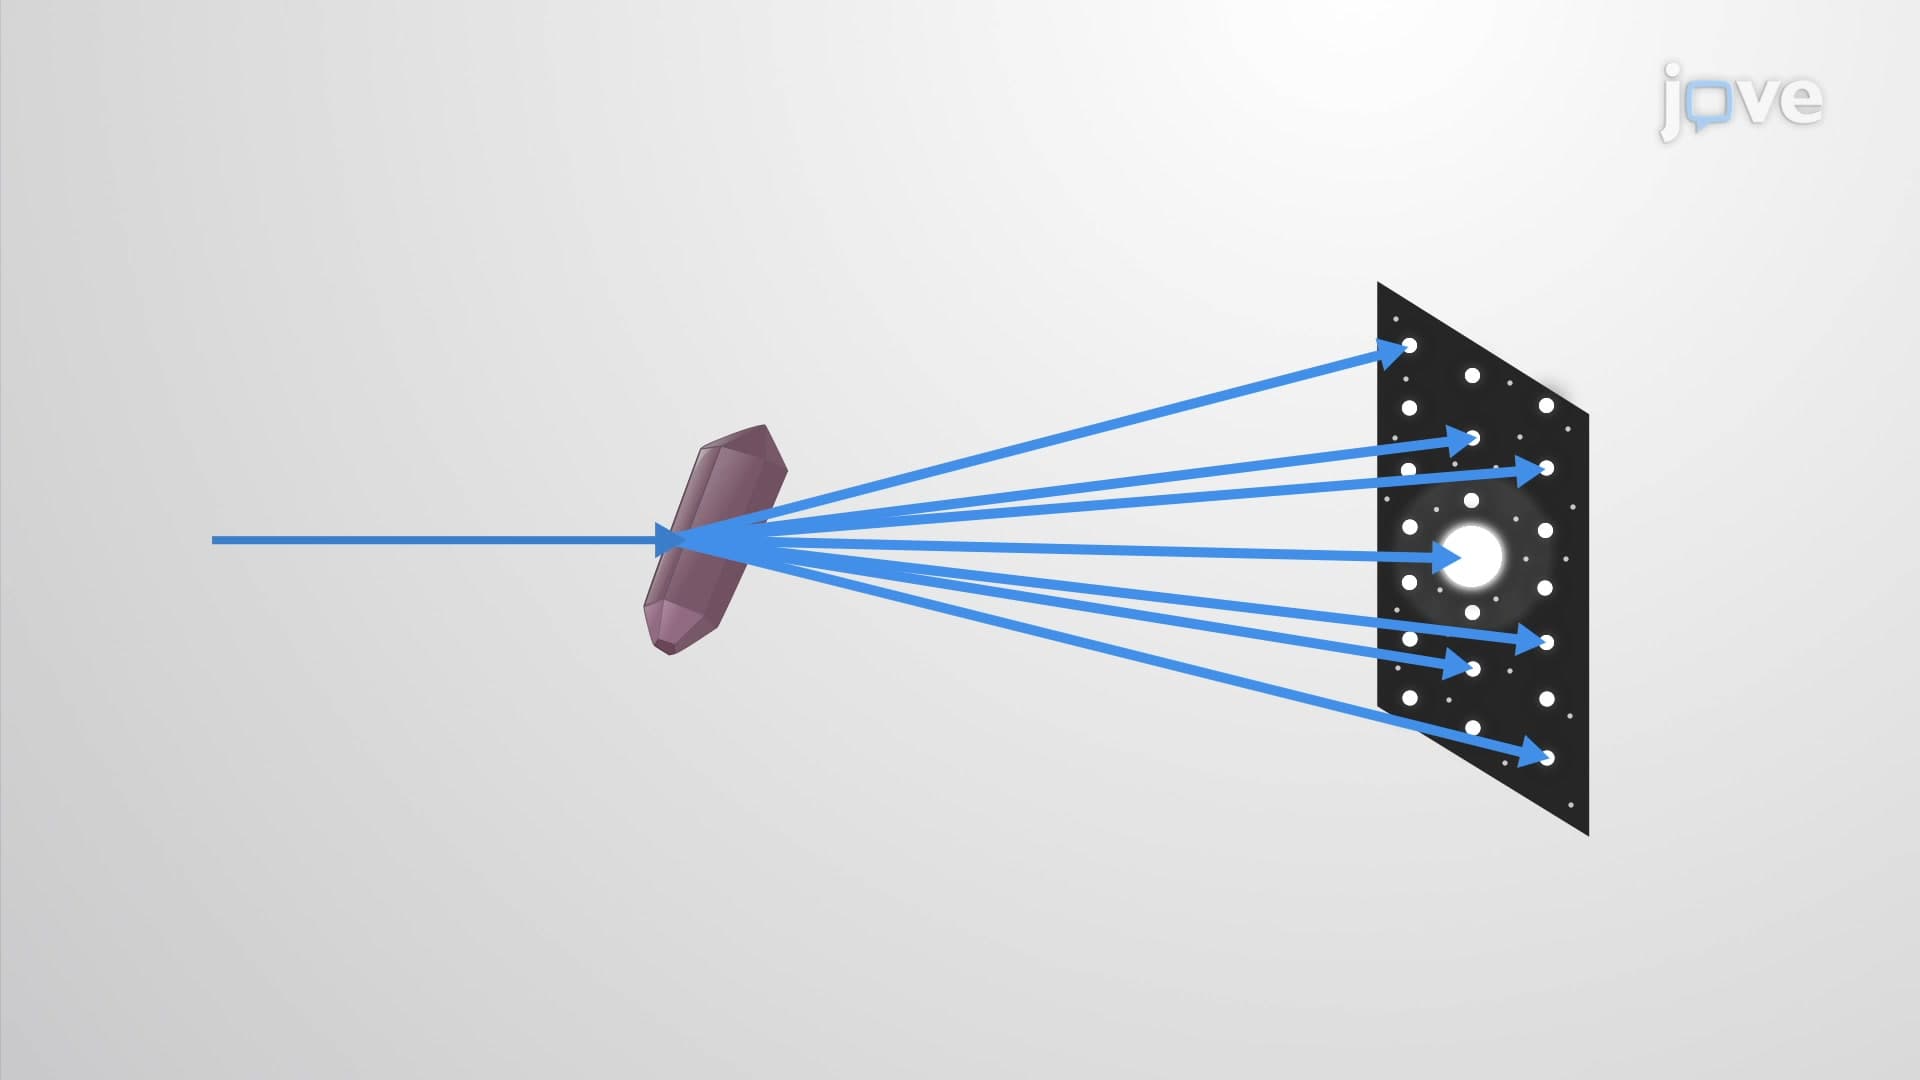
X-ray Crystallography
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...