Related Experiment Video
Updated: Jun 10, 2026
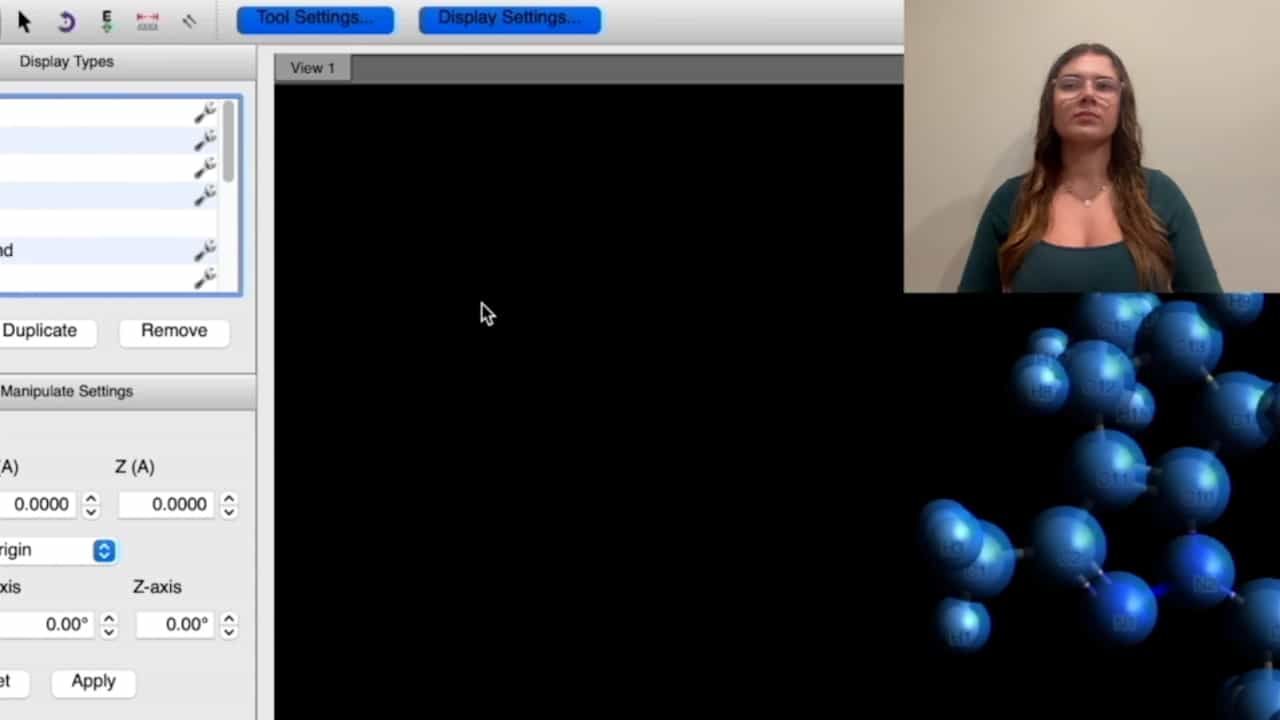

A Bilingual Computational Workflow for Identifying Potential PLK1 Inhibitors in American Sign Language and English
Published on: April 3, 2026
Predicting polypharmacology by binding site similarity: from kinases to the protein universe
Francesca Milletti1, Anna Vulpetti
1CADD, Global Discovery Chemistry, Novartis Institutes for Biomedical Research, CH4002 Basel, Switzerland.

Abstract:
Polypharmacology is receiving increasing attention in the pharmaceutical industry, since finding new targets of a compound is useful not only for anticipating possible side effects but also for opening new therapeutic opportunities. Thus, while system biology and personalized medicine are becoming increasingly important, there is an urgent need to map the inhibition profile of a compound on a large panel of targets by using both experimental and computational methods. This is especially important for kinase inhibitors, given the high similarity at the binding site level for the 518 kinases in the human genome. In this paper, we propose and validate a new method to predict the inhibition map of a compound by comparison of binding pockets. We used a subset of the Ambit panel for the validation-17 inhibitors with K(d) measured on 189 kinases-and found that on average 37% of kinases inhibited with K(d) < 10 microM were retrieved at 10% ROC enrichment. These results make this method particularly suitable to rationalize and optimize the selectivity profile of a compound. In addition, the method was extended to explore all the proteins in the PDB by using as queries pockets occupied by compounds of biological interest (ATP and various marketed drugs). The profiling of compounds against the protein universe revealed that striking structural similarities at the subpocket level (RMSD < 0.5 A) may also occur among targets with different folds, which can be exploited not only to predict off-target effects but also to design novel inhibitors for the target of interest.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
01:49Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:04
Protein-protein Interfaces
 01:16
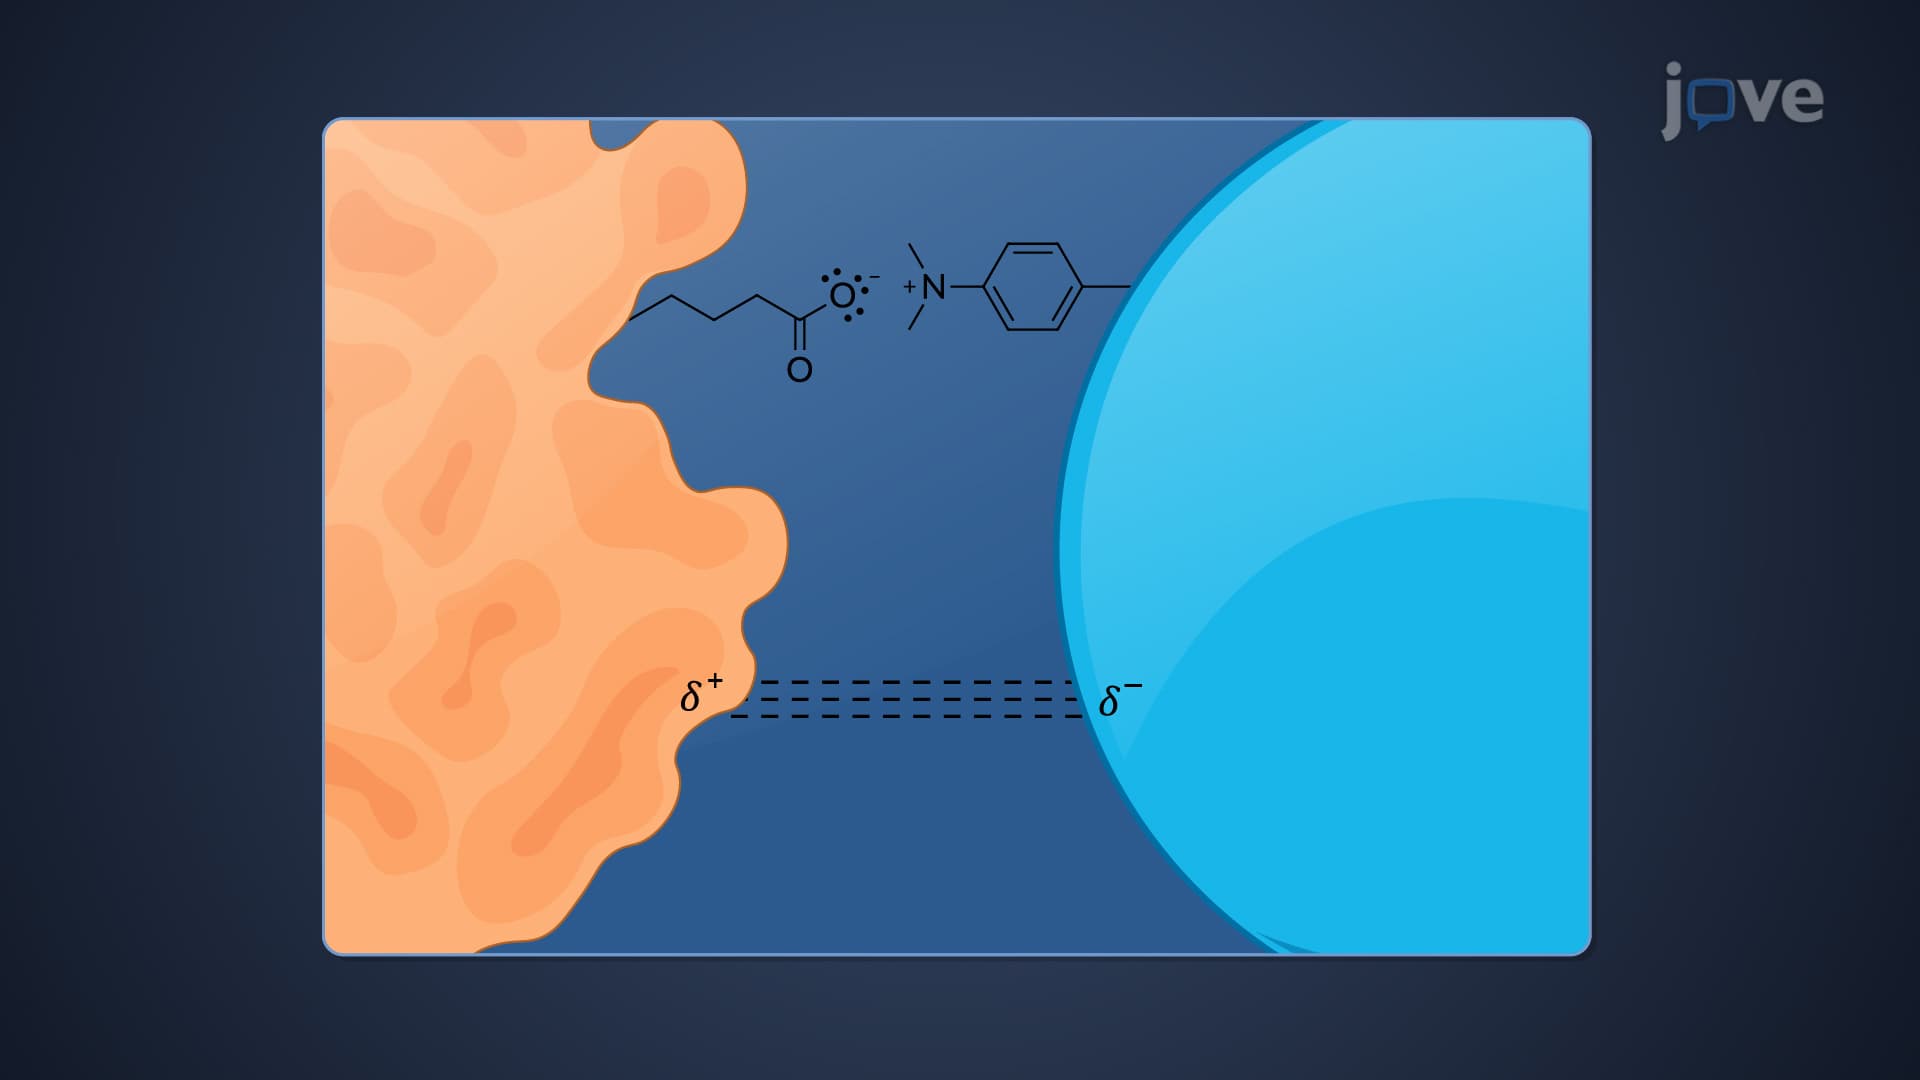
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...