Related Experiment Video
Updated: Jun 10, 2026

Enrichment of Mammalian Tissues and Xenopus Oocytes with Cholesterol
Published on: March 25, 2020
Structural basis of drug binding to CYP46A1, an enzyme that controls cholesterol turnover in the brain
Natalia Mast1, Casey Charvet, Irina A Pikuleva
1Department of Ophthalmology and Visual Sciences, Case Western Reserve University, Cleveland, Ohio 44106, USA.

Abstract:
Cytochrome P450 46A1 (CYP46A1) initiates the major pathway of cholesterol elimination from the brain and thereby controls cholesterol turnover in this organ. We determined x-ray crystal structures of CYP46A1 in complex with four structurally distinct pharmaceuticals; antidepressant tranylcypromine (2.15 Å), anticonvulsant thioperamide (1.65 Å), antifungal voriconazole (2.35 Å), and antifungal clotrimazole (2.50 Å). All four drugs are nitrogen-containing compounds that have nanomolar affinity for CYP46A1 in vitro yet differ in size, shape, hydrophobicity, and type of the nitrogen ligand. Structures of the co-complexes demonstrate that each drug binds in a single orientation to the active site with tranylcypromine, thioperamide, and voriconazole coordinating the heme iron via their nitrogen atoms and clotrimazole being at a 4 Å distance from the heme iron. We show here that clotrimazole is also a substrate for CYP46A1. High affinity for CYP46A1 is determined by a set of specific interactions, some of which were further investigated by solution studies using structural analogs of the drugs and the T306A CYP46A1 mutant. Collectively, our results reveal how diverse inhibitors can be accommodated in the CYP46A1 active site and provide an explanation for the observed differences in the drug-induced spectral response. Co-complexes with tranylcypromine, thioperamide, and voriconazole represent the first structural characterization of the drug binding to a P450 enzyme.
Related Concept Videos
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:26
Targets for Drug Action: Overview
Receptors are either membrane-spanning or intracellular proteins, which upon binding a ligand, get activated and transmit the signal downstream to elicit a response. Drugs bind receptors, either mimicking the action of endogenous ligands or blocking the receptor activity to bring about a modified response. Nearly 35% of approved drugs target the G...
 01:23
Factors Affecting Protein-Drug Binding: Drug Interactions
Displacement interactions can have varying outcomes, ranging from toxicity to virtually...
 01:30
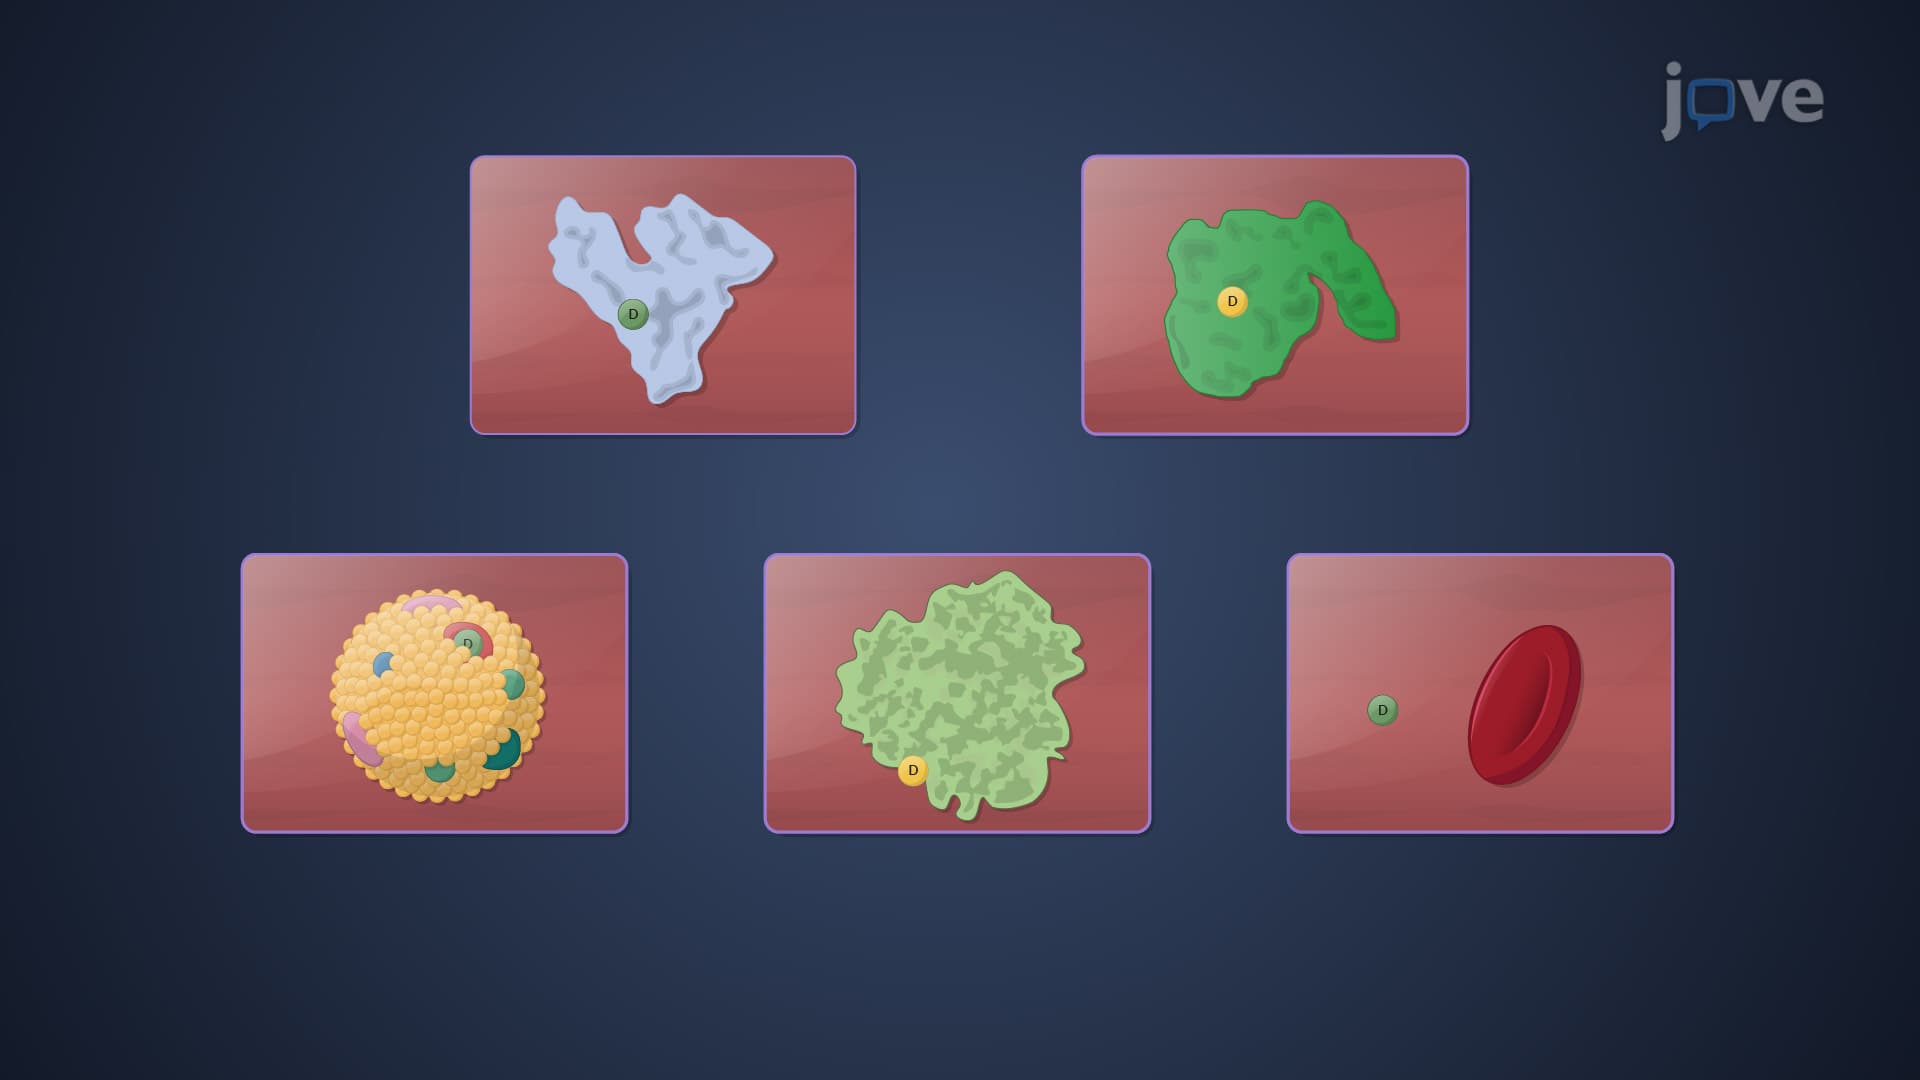
Drug Binding to Blood Components
HSA is the most abundant plasma protein and is vital in drug binding. It contains distinct drug-binding sites, with different drugs exhibiting affinity for specific sites. There are three main drug-binding domains for HSA: sites I, II, and III. These domains are further...
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion, mediated...
 01:16
Drug Biotransformation: Overview