Related Experiment Video
Updated: Jun 10, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Structure-activity relationships of GPR120 agonists based on a docking simulation
Qi Sun1, Akira Hirasawa, Takafumi Hara
1Department of Genomic Drug Discovery Science, Graduate School of Pharmaceutical Sciences, Kyoto University, 46-29 Yoshida Shimoadachi-cho, Sakyo-ku, Kyoto 606-8501, Japan.

Abstract:
GPR120 is a G protein-coupled receptor expressed preferentially in the intestinal tract and adipose tissue, that has been implicated in mediating free fatty acid-stimulated glucagon-like peptide-1 (GLP-1) secretion. To develop GPR120-specific agonists, a series of compounds (denoted as NCG compounds) derived from a peroxisome proliferator-activated receptor γ agonist were synthesized, and their structure-activity relationships as GPR120 agonists were explored. To examine the agonistic activities of these newly synthesized NCG compounds, and of compounds already shown to have GPR120 agonistic activity (grifolic acid and MEDICA16), we conducted docking simulation in a GPR120 homology model that was developed on the basis of a photoactivated model derived from the crystal structure of bovine rhodopsin. We calculated the hydrogen bonding energies between the compounds and the GPR120 model. These energies correlated well with the GPR120 agonistic activity of the compounds (R(2) = 0.73). NCG21, the NCG compound with the lowest calculated hydrogen bonding energy, showed the most potent extracellular signal-regulated kinase (ERK) activation in a cloned GPR120 system. Furthermore, NCG21 potently activated ERK, intracellular calcium responses and GLP-1 secretion in murine enteroendocrine STC-1 cells that express GPR120 endogenously. Moreover, administration of NCG21 into the mouse colon caused an increase in plasma GLP-1 levels. Taken together, our present study showed that a docking simulation using a GPR120 homology model might be useful to predict the agonistic activity of compounds.
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:15
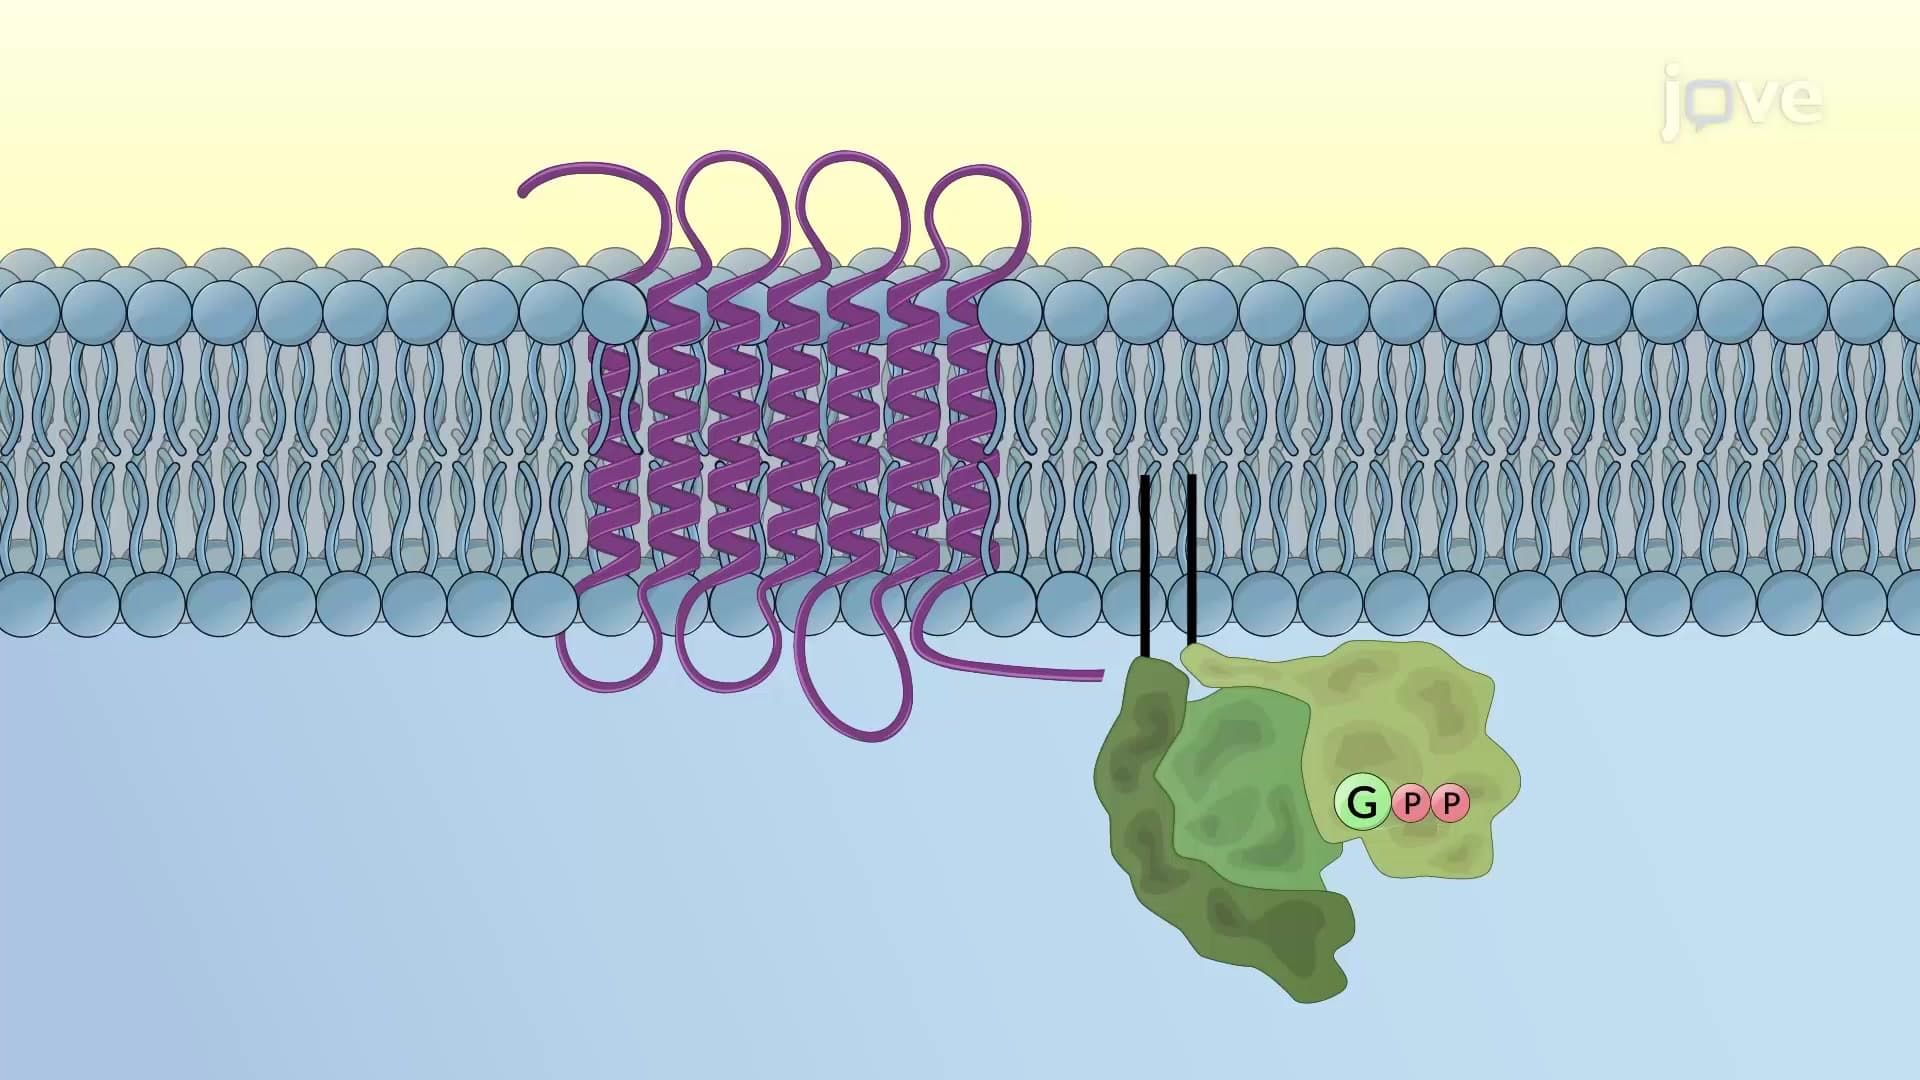
G Protein-coupled Receptors
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with one...
 01:22
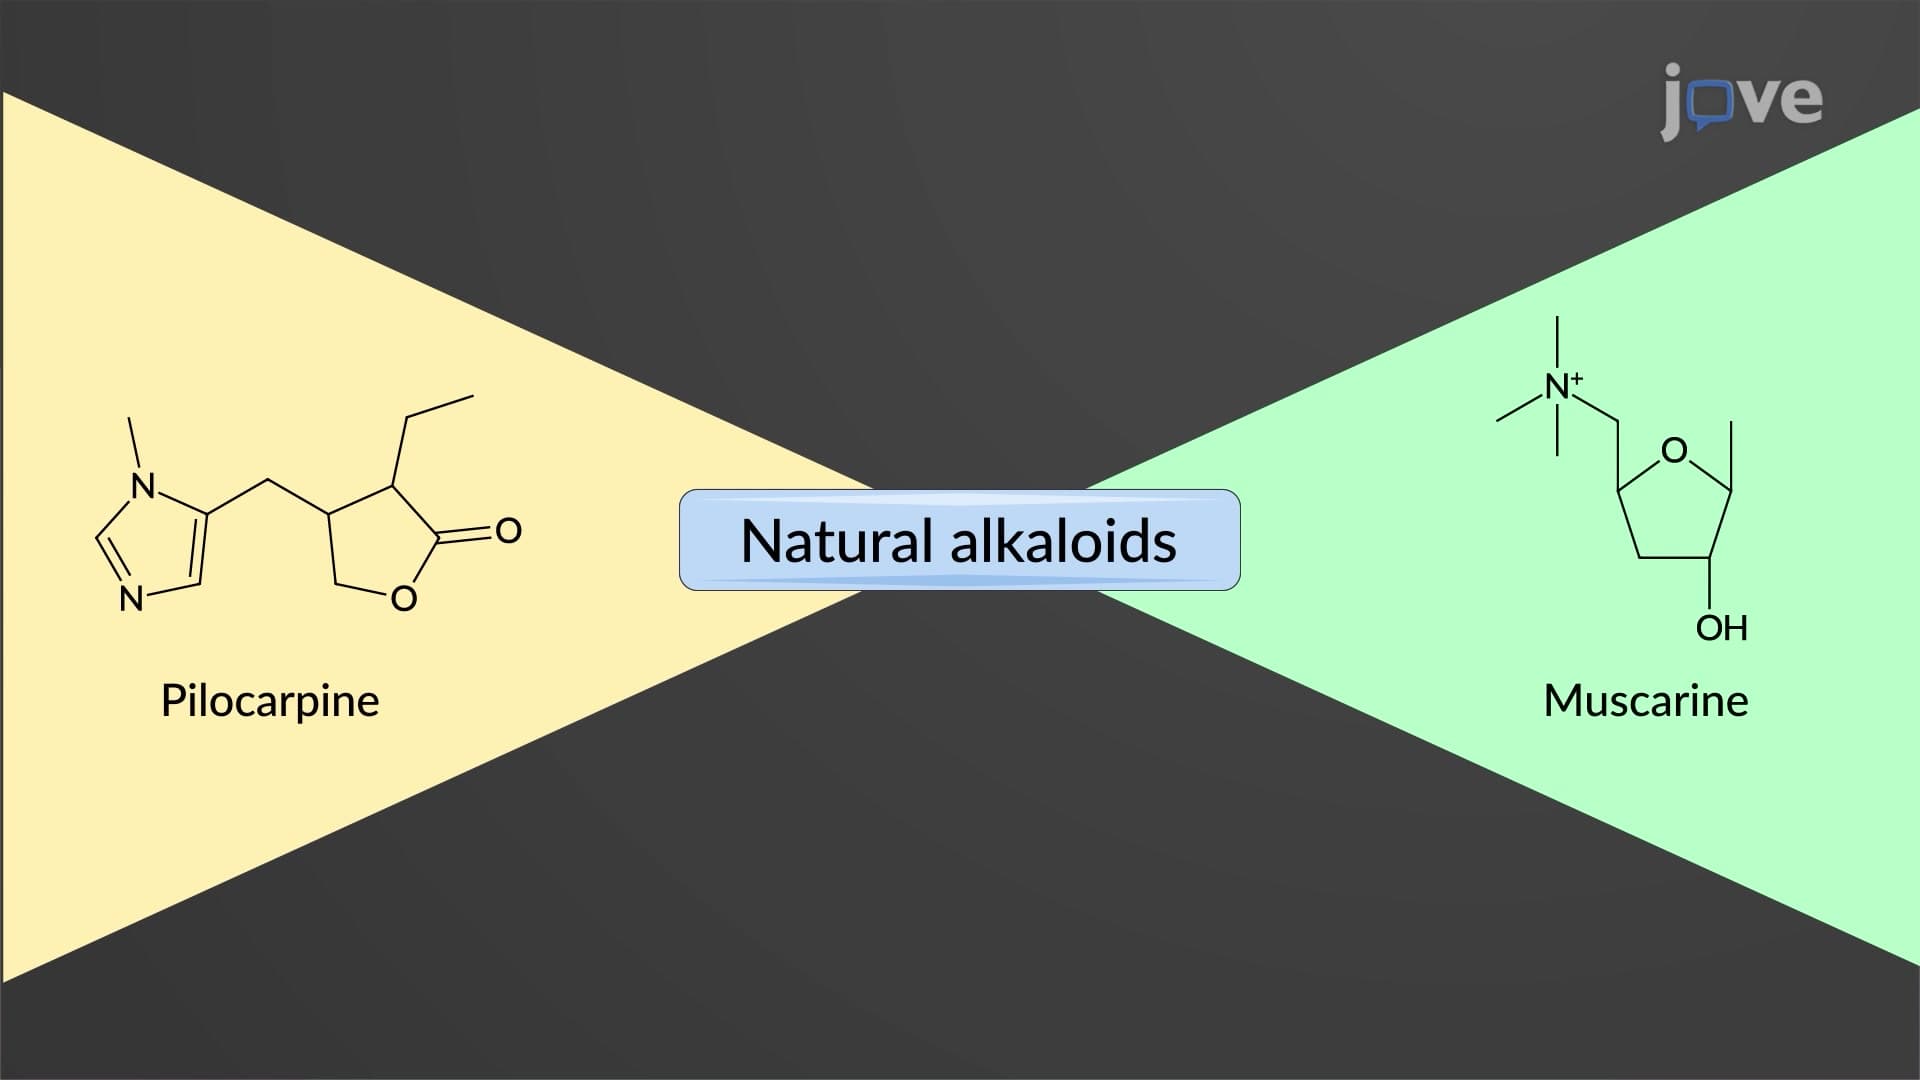
Direct-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
The direct-acting...
 01:25
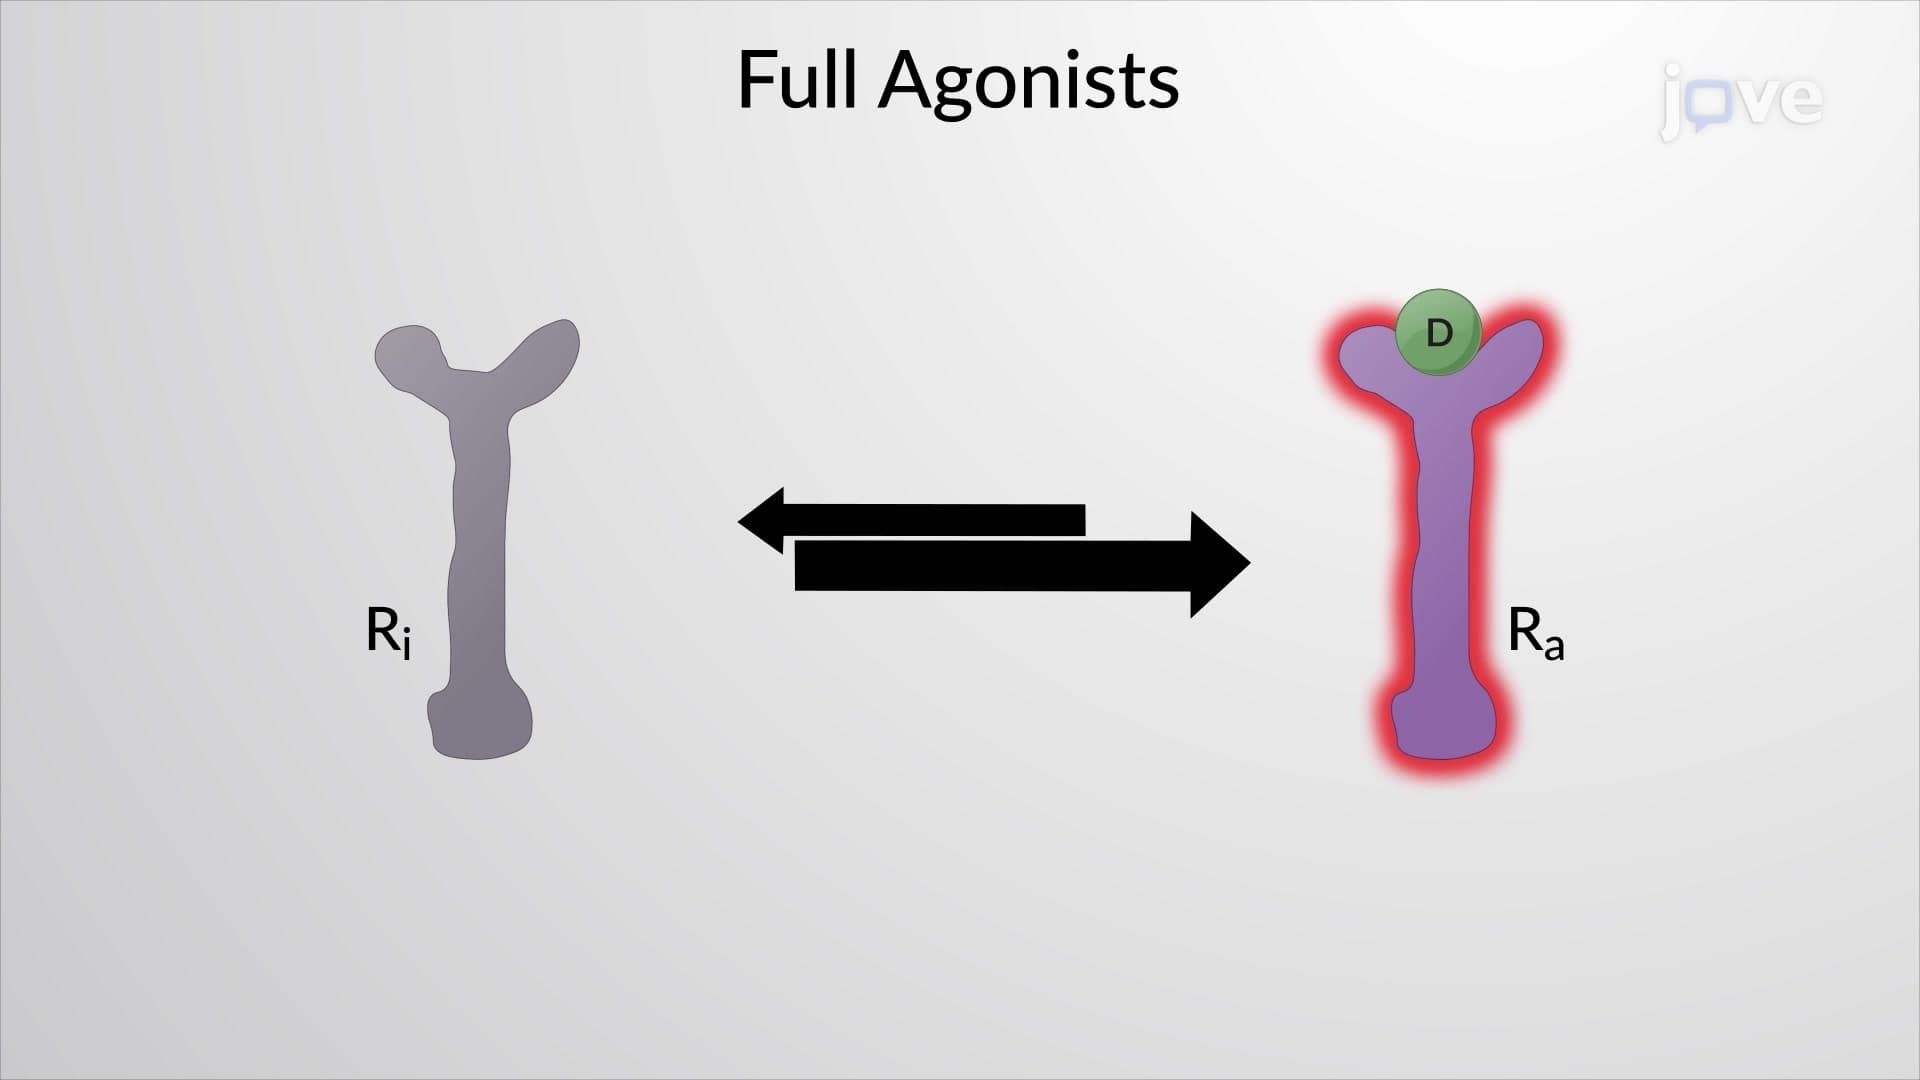
Drug-Receptor Interaction: Agonist
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous ligand's action.