Related Experiment Video
Updated: Jun 10, 2026
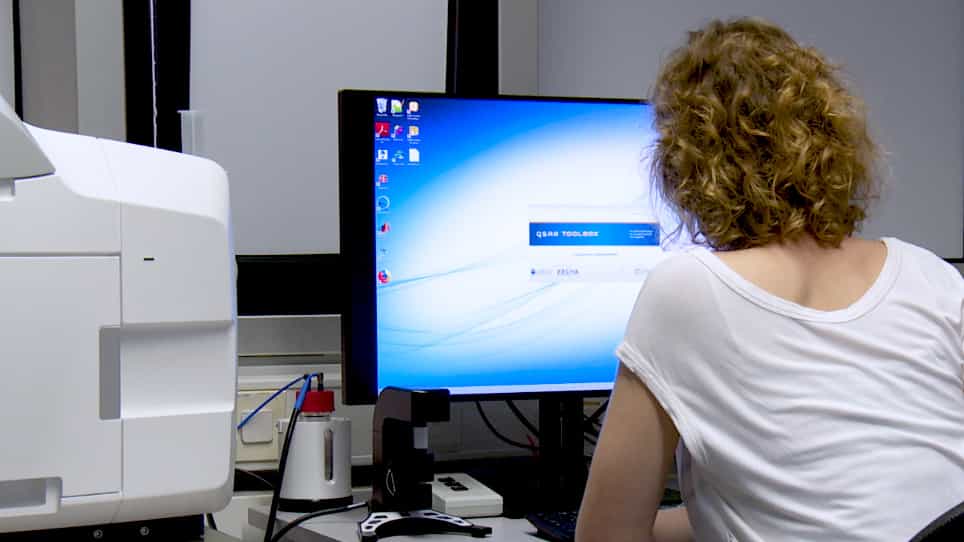

In Silico Modeling Method for Computational Aquatic Toxicology of Endocrine Disruptors: A Software-Based Approach Using QSAR Toolbox
Published on: August 28, 2019
Estrogenic activity of anthraquinone derivatives: in vitro and in silico studies
Fei Li1, Xuehua Li, Jianping Shao
1Key Laboratory of Industrial Ecology and Environmental Engineering, School of Environmental Science and Technology, Dalian University of Technology, Linggong Road 2, Dalian, P. R. China.

Abstract:
Comprehension of the ligand-receptor interactions is a prerequisite for constructing mechanism based quantitative structure-activity relationship (QSAR) models on xenoestrogenic activity. Molecular docking was performed to simulate the interactions between anthraquinone derivative (AQs) molecules and the estrogen receptor alpha (ERalpha). Hydrogen bonding, hydrophobic, and pi-pi interactions were found to be the dominant interactions between AQs and the receptor, which implied the estrogenic activities of the compounds. The recombinant yeast-based assay was employed to determine the estrogenic activities of 20 AQs. On the basis of the observed interactions between the AQs and ERalpha, appropriate molecular structural parameters were computed to develop a QSAR model. The polarizability term, the binding energy, the average molecular polarizability, the most negative formal charge in the molecule, and the average of the negative potentials on the molecular surface were significant parameters explaining the estrogenicity. The developed QSAR model had good robustness, predictive ability, and mechanism interpretability. The interactions between the AQs and ERalpha and the partition ability of the AQs into the biophase are main factors governing the estrogenic activities. Moreover, the applicability domain of the model was described.