Related Experiment Video
Updated: Jun 10, 2026

Preparation Of Gushukang (GSK) Granules for In Vivo and In Vitro Experiments
Published on: May 9, 2019
Time course of vitamin C distribution and absorption after oral administration in SMP30/GNL knockout mice
Mizuki Iwama1, Kentaro Shimokado, Naoki Maruyama
1Department of Geriatrics and Vascular Medicine, Tokyo Medical and Dental University, Tokyo, Japan.

Objective:
Because vitamin C (VC) has multiple metabolic and antioxidant functions, we investigated the movement of VC throughout the tissues of senescence marker protein-30 (SMP30)/gluconolactonase (GNL) knockout (KO) mice.
Methods:
SMP30/GNL KO mice, which cannot synthesize VC in vivo, were divided into two groups: VC sufficient and VC deficient. Starting at 2 mo of age, both groups had free access to water containing 1.5 and 0.0375 g/L of VC for 1 mo.
Results:
The average rate of VC retention in 20 tissues of VC-deficient SMP30/GNL KO mice was only 13.7% of that in VC-sufficient mice. Tissues that retained over 20% of VC were the cerebellum, white fat, testes, eyeballs, and pancreas, and those with less than 5% VC were the kidneys and heart. These results clearly indicate the different VC retention capacities among tissues. Next, we examined the time course of VC distribution and absorption in VC-deficient SMP30/GNL KO mice. After oral VC administration, VC content in the liver and kidney peaked at 3 h and then decreased. VC content in the lungs, adrenal glands, skin, white fat, and pancreas peaked at 6 h and in the cerebellum, cerebrum, skeletal muscles, eyeballs, thyroid gland, and testes at 12 h.
Conclusion:
In this study, we found that exogenous VC administered orally in VC-deficient SMP30/GNL KO mice was distributed at distinctly different rates within individual tissues. The SMP30/GNL KO mice used in this study are a useful animal model that provides unique opportunities for investigating VC movement and metabolism in the entire body.
Insights
Vitamin C (VC) distribution varies significantly across tissues in mice unable to synthesize it. This study highlights differing VC retention and absorption rates in various organs, using a novel knockout mouse model.
Area of Science:
- Biochemistry
- Physiology
- Animal Models
Background:
- Vitamin C (VC) is crucial for metabolic and antioxidant functions.
- Senescence marker protein-30 (SMP30)/gluconolactonase (GNL) knockout (KO) mice cannot synthesize VC endogenously.
- Understanding VC tissue distribution is vital for its physiological role.
Purpose of the Study:
- To investigate the tissue-specific movement and distribution of vitamin C (VC).
- To utilize SMP30/GNL KO mice as a model for studying VC metabolism and transport.
- To compare VC retention in different tissues under sufficient and deficient conditions.
Main Methods:
- Utilized SMP30/GNL KO mice, divided into VC-sufficient and VC-deficient groups.
- Administered controlled doses of VC in drinking water for one month.
- Analyzed VC content across 20 different tissues and tracked temporal distribution post-oral administration.
Main Results:
- VC-deficient mice retained only 13.7% of VC compared to sufficient mice.
- Tissues with high VC retention (>20%) included cerebellum, white fat, testes, eyeballs, and pancreas.
- Kidneys and heart showed very low VC retention (<5%).
- Peak VC levels in liver and kidney occurred at 3 hours, while other tissues like cerebellum and testes peaked at 12 hours post-administration.
Conclusions:
- Exogenous VC exhibits distinct tissue distribution rates in VC-deficient mice.
- SMP30/GNL KO mice serve as a valuable model for in vivo VC research.
- Tissue-specific differences in VC uptake and retention are significant.
Related Concept Videos
 01:09
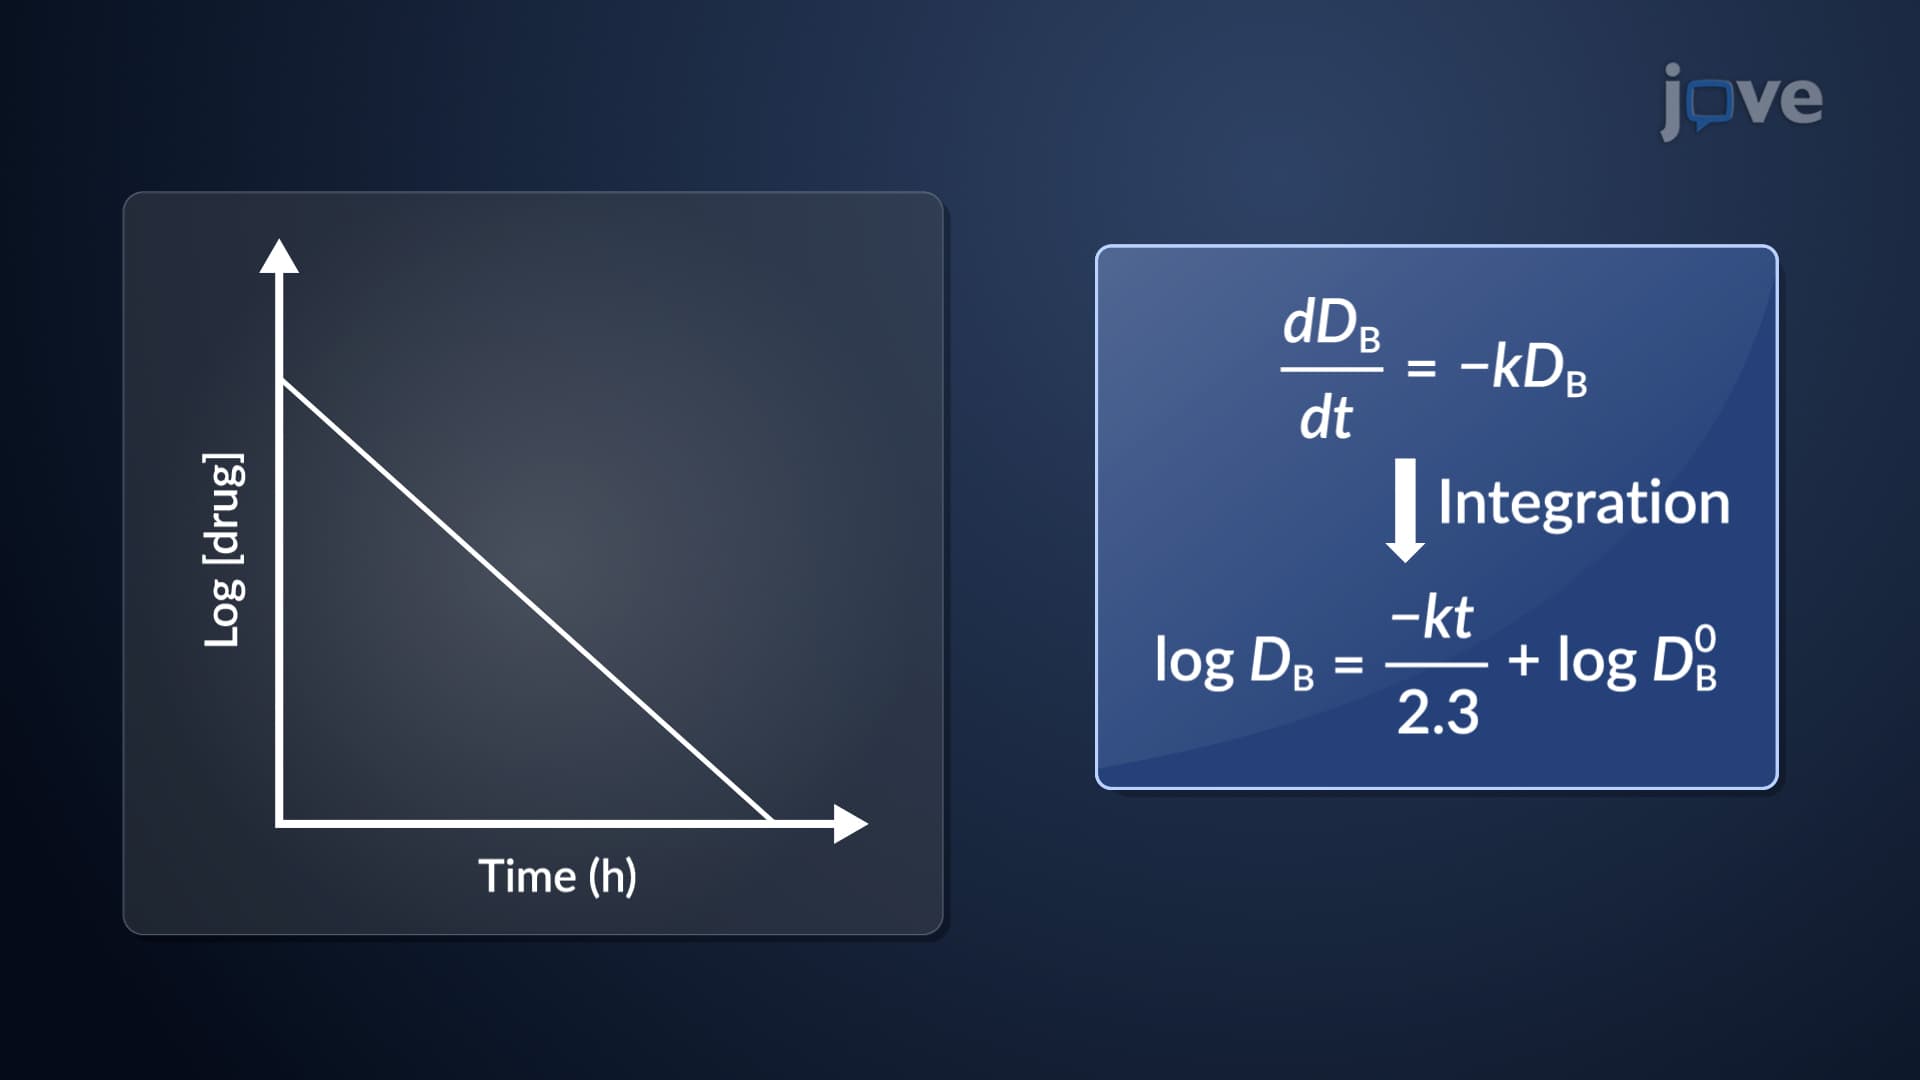
One-Compartment Open Model for IV Bolus Administration: Estimation of Elimination Rate Constant, Half-Life and Volume of Distribution
 01:22
Bioavailability: Influencing Factors
 01:17
Bioavailability: Overview
 01:13
Bioavailability: Overview
 01:14
Time Course of Drug Effect
 01:11
Bioavailability Study Design: Single Versus Multiple Dose Studies