Related Experiment Video
Updated: Jun 9, 2026
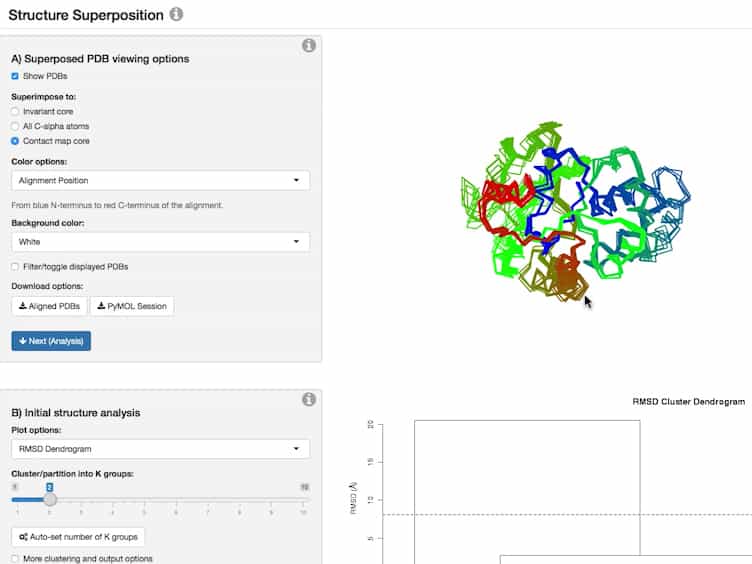
Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Protein functional landscapes, dynamics, allostery: a tortuous path towards a universal theoretical framework
Pavel I Zhuravlev1, Garegin A Papoian
1Department of Chemistry and Biochemistry, Institute for Physical Science and Technology, University of Maryland, College Park, MD 20742, USA.

Abstract:
Energy landscape theories have provided a common ground for understanding the protein folding problem, which once seemed to be overwhelmingly complicated. At the same time, the native state was found to be an ensemble of interconverting states with frustration playing a more important role compared to the folding problem. The landscape of the folded protein - the native landscape - is glassier than the folding landscape; hence, a general description analogous to the folding theories is difficult to achieve. On the other hand, the native basin phase volume is much smaller, allowing a protein to fully sample its native energy landscape on the biological timescales. Current computational resources may also be used to perform this sampling for smaller proteins, to build a 'topographical map' of the native landscape that can be used for subsequent analysis. Several major approaches to representing this topographical map are highlighted in this review, including the construction of kinetic networks, hierarchical trees and free energy surfaces with subsequent structural and kinetic analyses. In this review, we extensively discuss the important question of choosing proper collective coordinates characterizing functional motions. In many cases, the substates on the native energy landscape, which represent different functional states, can be used to obtain variables that are well suited for building free energy surfaces and analyzing the protein's functional dynamics. Normal mode analysis can provide such variables in cases where functional motions are dictated by the molecule's architecture. Principal component analysis is a more expensive way of inferring the essential variables from the protein's motions, one that requires a long molecular dynamics simulation. Finally, the two popular models for the allosteric switching mechanism, 'preexisting equilibrium' and 'induced fit', are interpreted within the energy landscape paradigm as extreme points of a continuum of transition mechanisms. Some experimental evidence illustrating each of these two models, as well as intermediate mechanisms, is presented and discussed.
Related Concept Videos
 01:58
Cooperative Allosteric Transitions
01:58Cooperative Allosteric Transitions
01:58Cooperative Allosteric Transitions
 00:49
Ligand Binding and Linkage
 01:19
Allosteric Proteins-ATCase
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis pathway,...
 01:08
Allosteric Regulation