Human C3 mutation reveals a mechanism of dense deposit disease pathogenesis and provides insights into complement
Rubén Martínez-Barricarte1, Meike Heurich, Francisco Valdes-Cañedo
1Centro de Investigaciones Biológicas (CIB), Consejo Superior de Investigaciones Científicas, Centro de Investigación Biomédica en Enfermedades Raras and Instituto Reina Sofía de Investigaciones Nefrológicas, Madrid, Spain.

Insights
Dense deposit disease (DDD) is linked to a novel complement factor 3 (C3) mutation. This mutation causes uncontrolled alternative pathway (AP) complement activation, leading to severe kidney damage and DDD.
Area of Science:
- Nephrology
- Immunology
- Complement System Biology
Background:
- Dense deposit disease (DDD) is a severe kidney disorder marked by electron-dense material accumulation in the glomeruli.
- Previously, DDD was linked to factor H (fH) deficiency, causing massive complement factor 3 (C3) activation via the alternative pathway (AP).
- The precise molecular mechanisms driving DDD pathogenesis, particularly the role of C3 itself, remain incompletely understood.
Purpose of the Study:
- To investigate the genetic basis and pathogenic mechanisms of DDD in a unique patient pedigree.
- To characterize the functional consequences of a novel C3 mutation on complement system regulation.
- To elucidate the structural requirements of C3 for proper complement activation and regulation.
Main Methods:
- Genetic analysis to identify mutations in a DDD pedigree.
- Biochemical assays to assess the cleavage, activation, and regulatory properties of mutant C3.
- Functional studies using complement regulators like factor H (fH), decay accelerating factor (DAF), and membrane cofactor protein (MCP).
Main Results:
- A novel mutation in the C3 gene, C(3923ΔDG), was identified as the cause of DDD in the studied family.
- Mutant C3 resisted cleavage by the AP C3-convertase but formed a protease-activated convertase resistant to fH regulation.
- Activated mutant C3 was resistant to factor I (fI)/fH but sensitive to fI/MCP, leading to fluid-phase AP dysregulation and C3 consumption.
Conclusions:
- This study reveals critical structural determinants in C3 essential for AP C3-convertase recognition and regulation by fH, DAF, and MCP.
- The findings highlight a novel mechanism of DDD pathogenesis driven by a C3 mutation causing dysregulated fluid-phase AP activation.
- Understanding these C3 structural-functional relationships offers new insights for therapeutic strategies targeting complement-mediated kidney diseases.
Abstract:
Dense deposit disease (DDD) is a severe renal disease characterized by accumulation of electron-dense material in the mesangium and glomerular basement membrane. Previously, DDD has been associated with deficiency of factor H (fH), a plasma regulator of the alternative pathway (AP) of complement activation, and studies in animal models have linked pathogenesis to the massive complement factor 3 (C3) activation caused by this deficiency. Here, we identified a unique DDD pedigree that associates disease with a mutation in the C3 gene. Mutant C(3923ΔDG), which lacks 2 amino acids, could not be cleaved to C3b by the AP C3-convertase and was therefore the predominant circulating C3 protein in the patients. However, upon activation to C3b by proteases, or to C3(H₂O) by spontaneous thioester hydrolysis, C(3923ΔDG) generated an active AP C3-convertase that was regulated normally by decay accelerating factor (DAF) but was resistant to decay by fH. Moreover, activated C(3b923ΔDG) and C3(H₂O)(923ΔDG) were resistant to proteolysis by factor I (fI) in the presence of fH, but were efficiently inactivated in the presence of membrane cofactor protein (MCP). These characteristics cause a fluid phase-restricted AP dysregulation in the patients that continuously activated and consumed C3 produced by the normal C3 allele. These findings expose structural requirements in C3 that are critical for recognition of the substrate C3 by the AP C3-convertase and for the regulatory activities of fH, DAF, and MCP, all of which have implications for therapeutic developments.
Related Concept Videos
 01:27
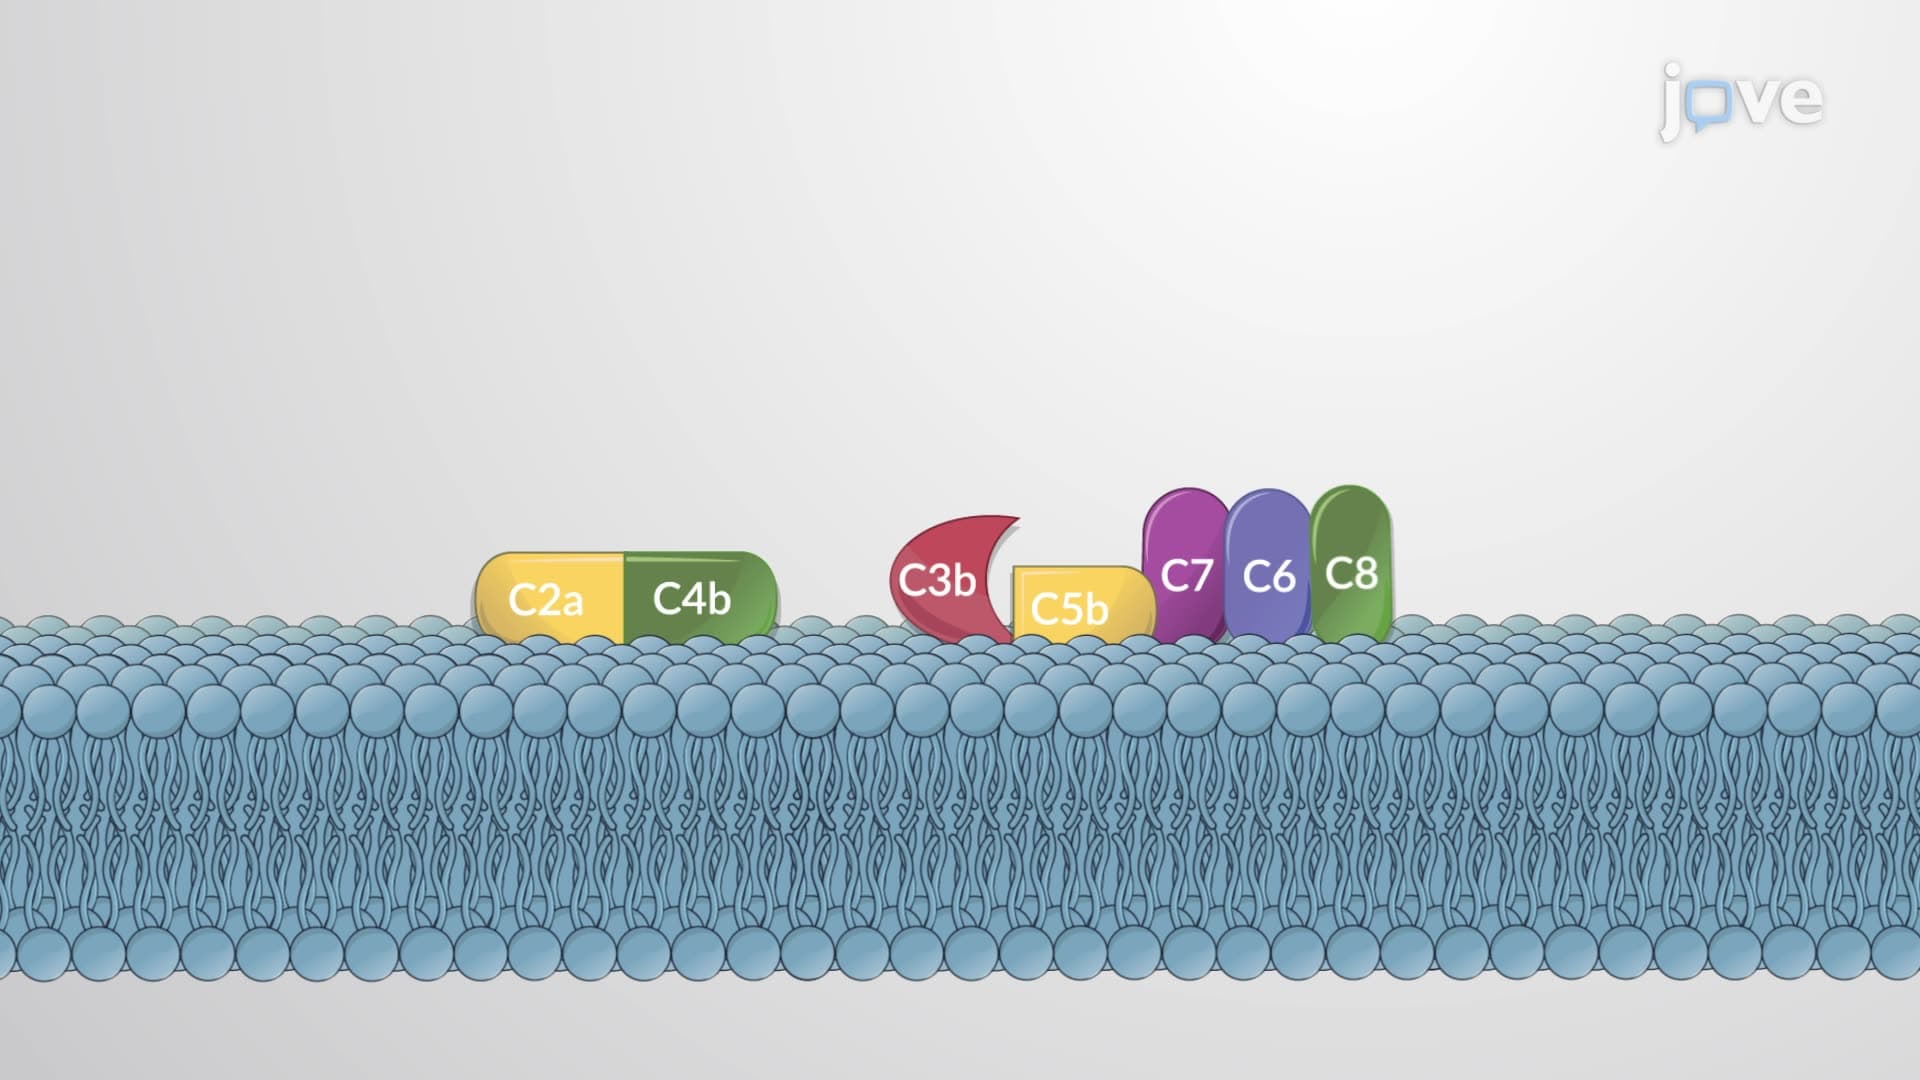
Complement System
 01:29
Cardiomyopathy III: Hypertrophic Cardiomyopathy
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:27
Cytomegalovirus Disease